Related Experiment Video
Updated: May 14, 2026

Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
Quasiclassical trajectory and statistical quantum calculations for the C + OH → CO + H reaction on the first excited
M Jorfi1, T González-Lezana, A Zanchet
1Institut de Chimie des Milieux et des Matériaux de Poitiers, UMR CNRS 6503, Université de Poitiers, 86022 Poitiers Cedex, France.

Abstract:
We report quasiclassical trajectory dynamical calculations for the C((3)P) + OH(X(2)Π) → CO(a(3)Π) + H((2)S) using a recently developed ab initio potential energy surface for the first electronic state of HCO of 1(2)A″ symmetry. The dependence of integral cross sections on the collision energy was determined. Product energy and angular distributions have also been calculated. Integral cross sections show no energy threshold and decrease as the collision energy increases. The comparison with results obtained from a statistical quantum method seems to confirm that the reaction is mainly dominated by an indirect mechanism in which a long-lived intermediate complex is involved.
More Related Videos


Related Concept Videos
 03:24
Hybridization of Atomic Orbitals I
 03:35
Hybridization of Atomic Orbitals II
 01:25
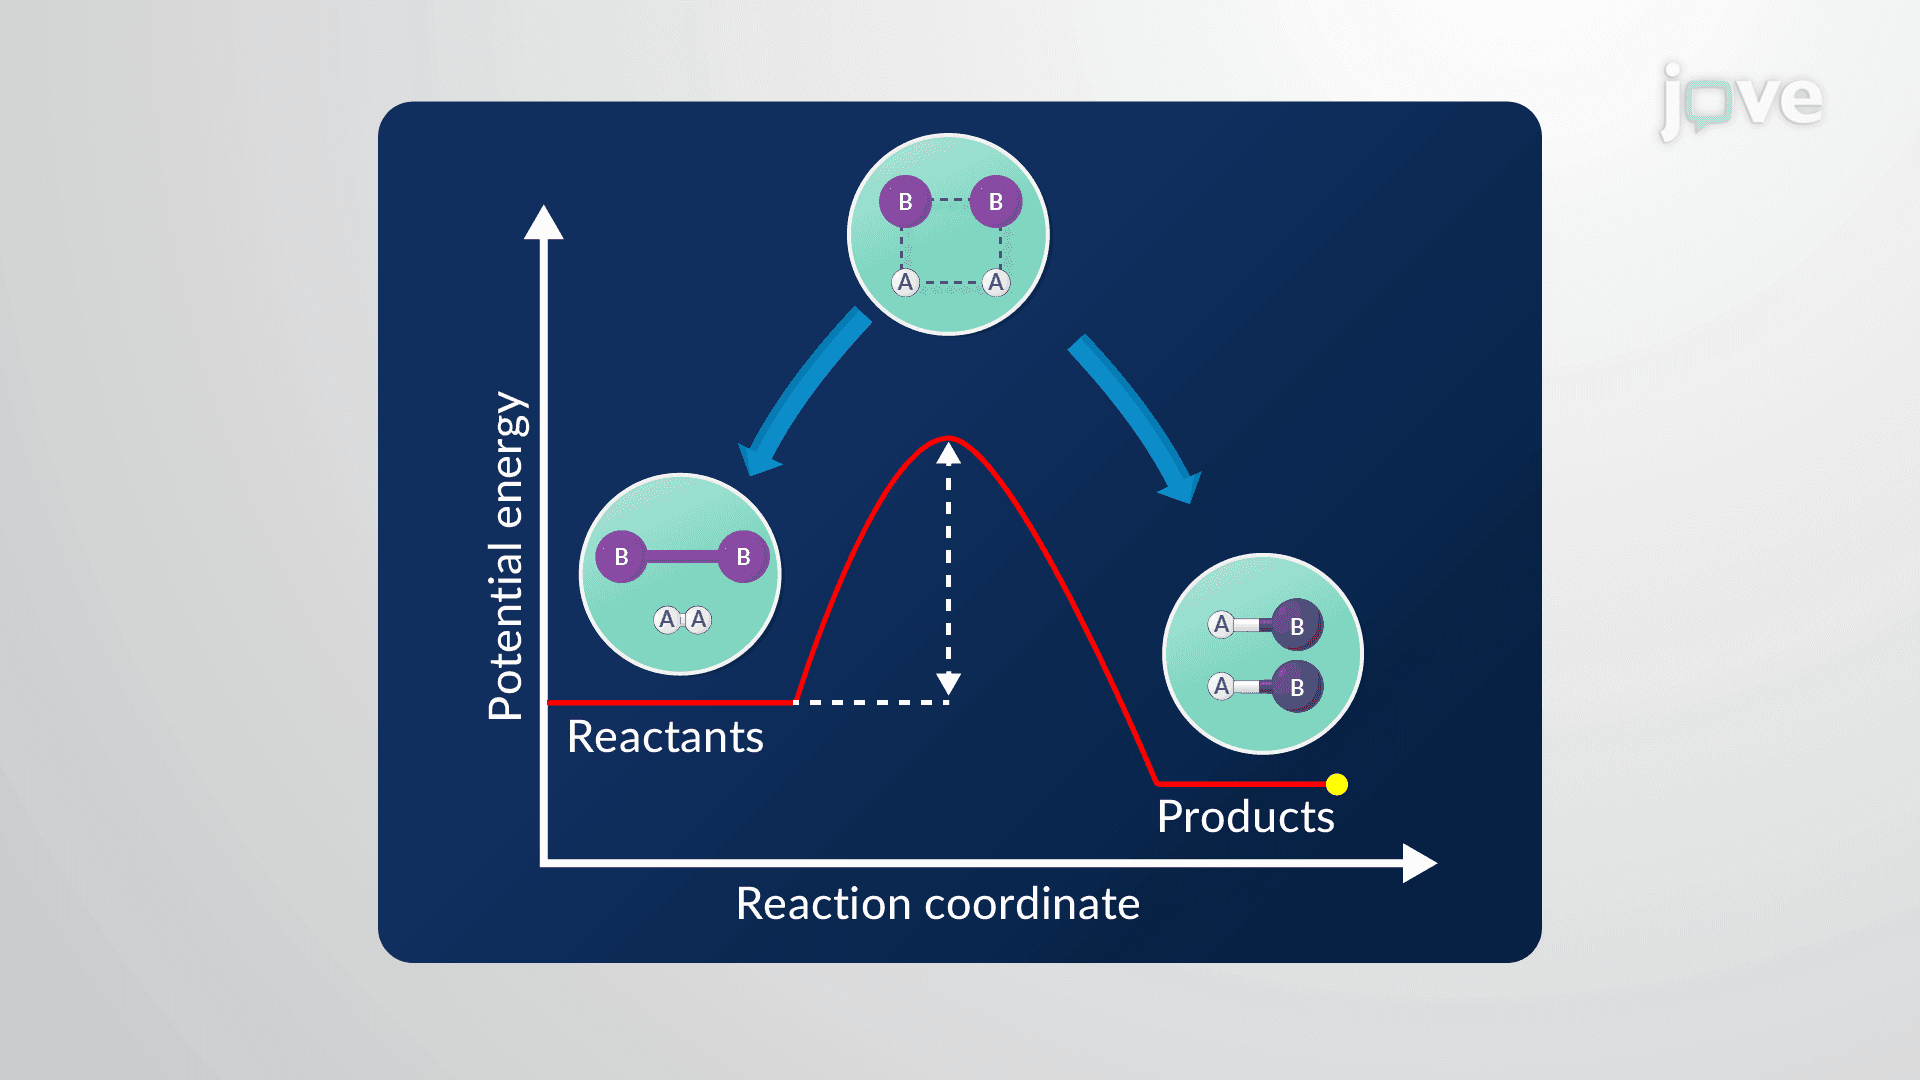
Transition State Theory
 01:26
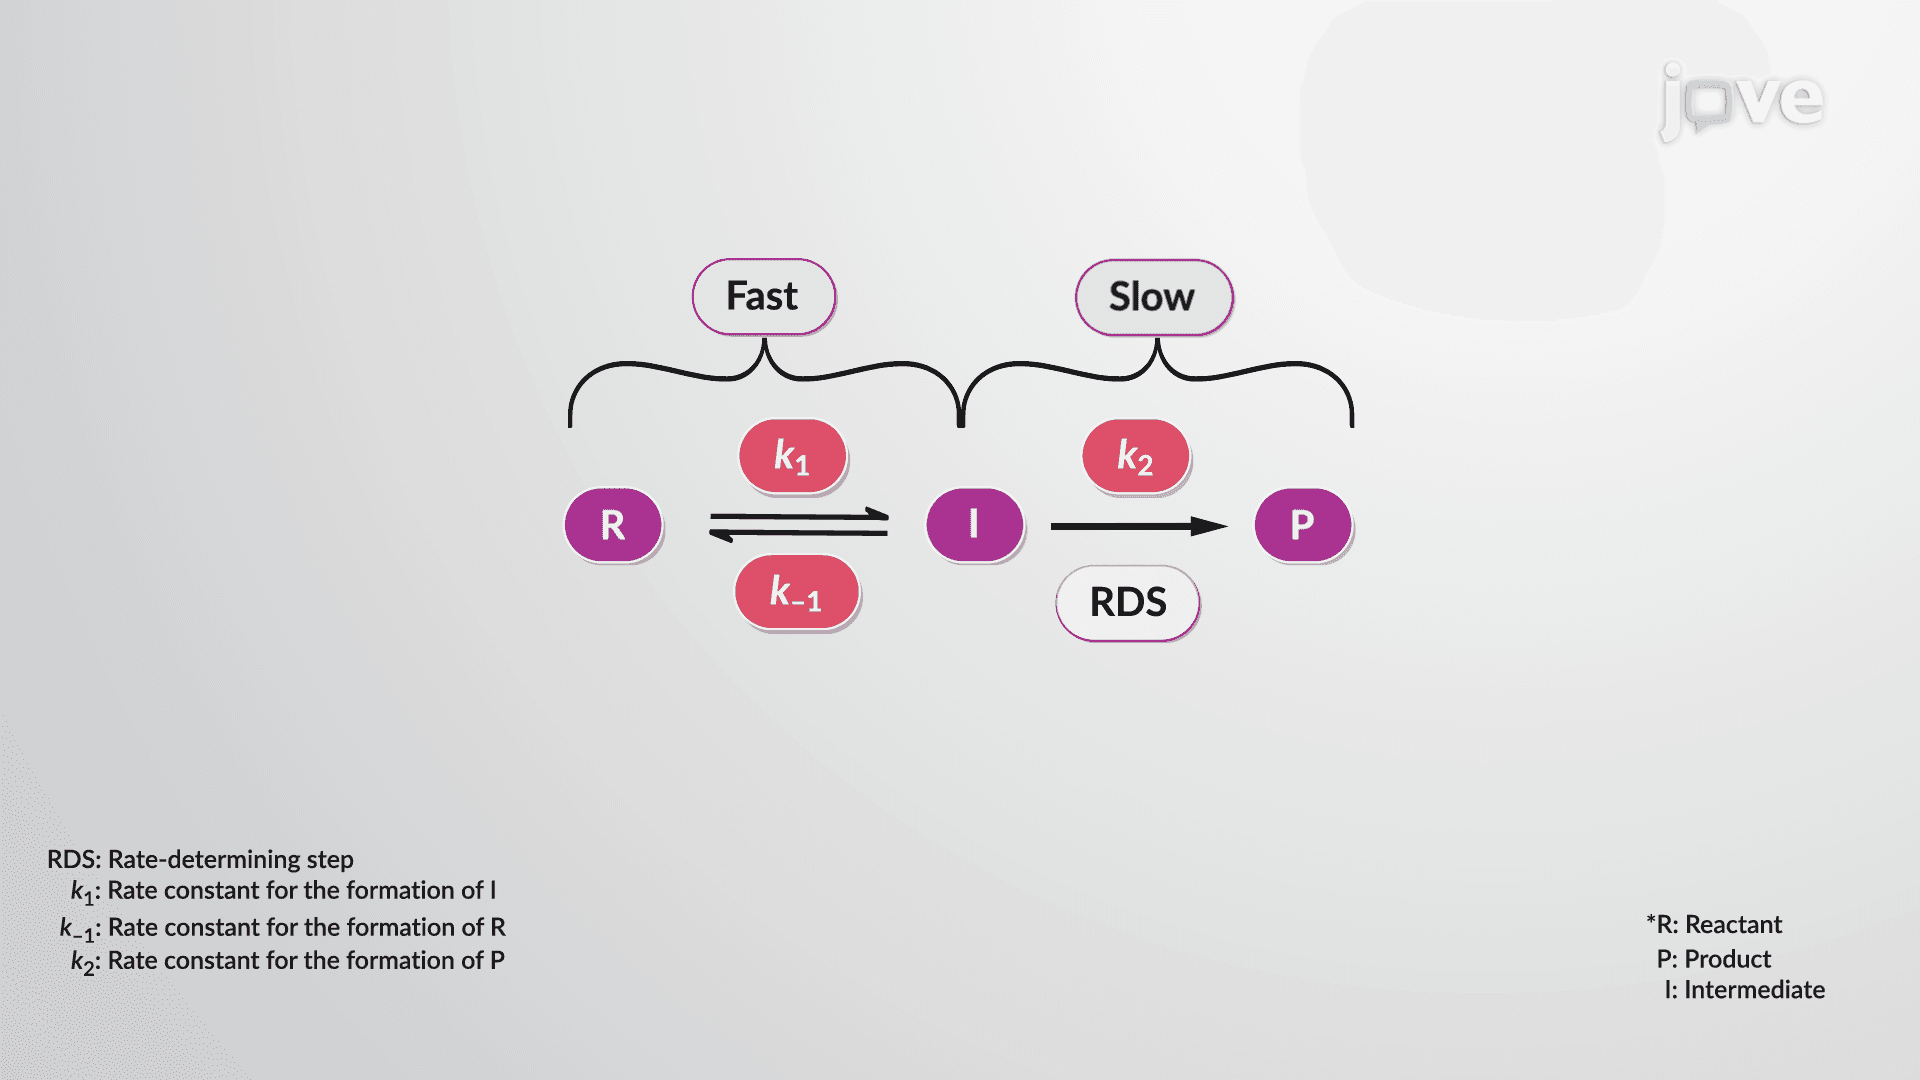
Reaction Mechanisms: The Steady-State Approximation
 02:35
Molecular Orbital Theory I
 03:51
Molecular Orbital Theory II