Related Experiment Video
Updated: May 14, 2026

Comprehensive Autopsy Program for Individuals with Multiple Sclerosis
Published on: July 19, 2019
The natural history of multiple system atrophy: a prospective European cohort study
Gregor K Wenning1, Felix Geser, Florian Krismer
1Department of Neurology, Innsbruck Medical University, Innsbruck, Austria. gregor.wenning@i-med.ac.at

Background:
Multiple system atrophy (MSA) is a fatal and still poorly understood degenerative movement disorder that is characterised by autonomic failure, cerebellar ataxia, and parkinsonism in various combinations. Here we present the final analysis of a prospective multicentre study by the European MSA Study Group to investigate the natural history of MSA.
Methods:
Patients with a clinical diagnosis of MSA were recruited and followed up clinically for 2 years. Vital status was ascertained 2 years after study completion. Disease progression was assessed using the unified MSA rating scale (UMSARS), a disease-specific questionnaire that enables the semiquantitative rating of autonomic and motor impairment in patients with MSA. Additional rating methods were applied to grade global disease severity, autonomic symptoms, and quality of life. Survival was calculated using a Kaplan-Meier analysis and predictors were identified in a Cox regression model. Group differences were analysed by parametric tests and non-parametric tests as appropriate. Sample size estimates were calculated using a paired two-group t test.
Findings:
141 patients with moderately severe disease fulfilled the consensus criteria for MSA. Mean age at symptom onset was 56·2 (SD 8·4) years. Median survival from symptom onset as determined by Kaplan-Meier analysis was 9·8 years (95% CI 8·1-11·4). The parkinsonian variant of MSA (hazard ratio [HR] 2·08, 95% CI 1·09-3·97; p=0·026) and incomplete bladder emptying (HR 2·10, 1·02-4·30; p=0·044) predicted shorter survival. 24-month progression rates of UMSARS activities of daily living, motor examination, and total scores were 49% (9·4 [SD 5·9]), 74% (12·9 [8·5]), and 57% (21·9 [11·9]), respectively, relative to baseline scores. Autonomic symptom scores progressed throughout the follow-up. Shorter symptom duration at baseline (OR 0·68, 0·5-0·9; p=0·006) and absent levodopa response (OR 3·4, 1·1-10·2; p=0·03) predicted rapid UMSARS progression. Sample size estimation showed that an interventional trial with 258 patients (129 per group) would be able to detect a 30% effect size in 1-year UMSARS motor examination decline rates at 80% power.
Interpretation:
Our prospective dataset provides new insights into the evolution of MSA based on a follow-up period that exceeds that of previous studies. It also represents a useful resource for patient counselling and planning of multicentre trials.
Related Concept Videos
 01:19
Multiple Sclerosis l: Introduction
 01:23
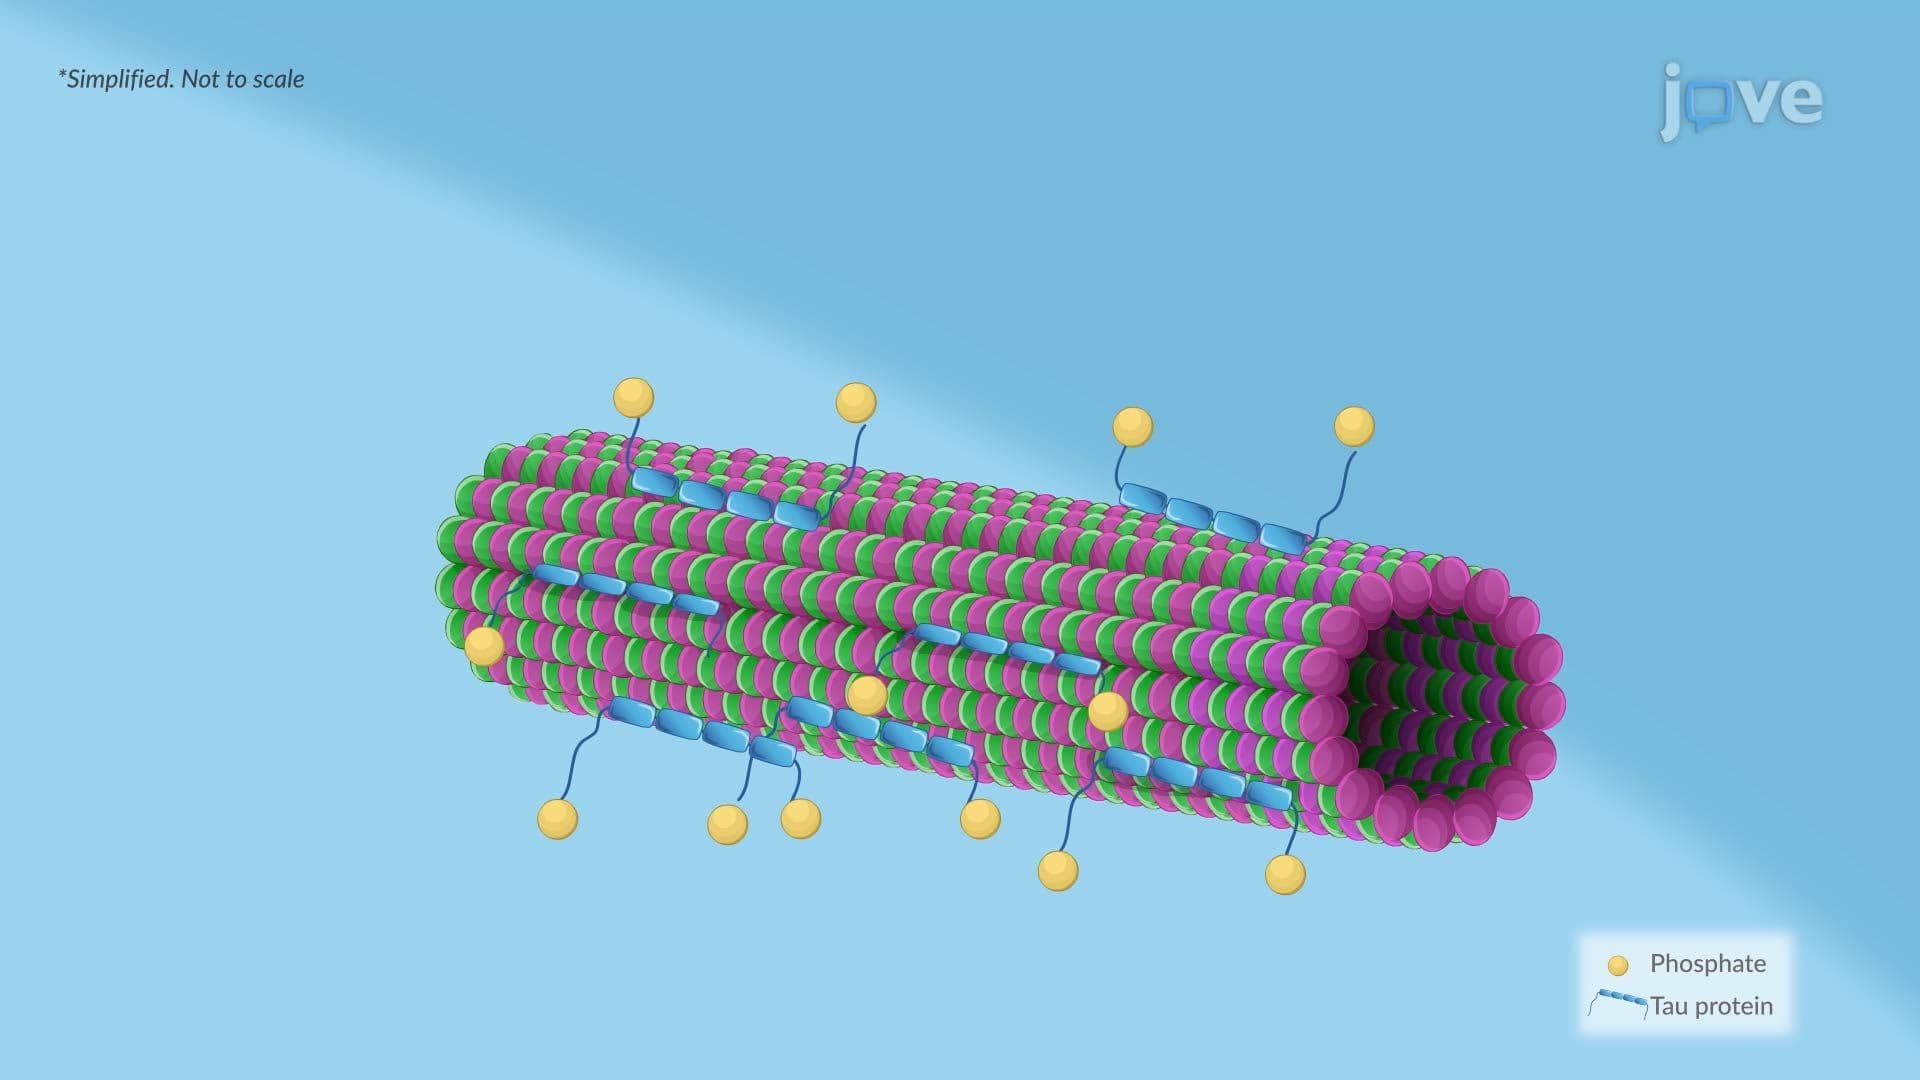
Alzheimer Disease ll: Pathophysiology
 01:29
Alzheimer Disease l: Introduction
 01:21
Huntington Disease l: Introduction
 01:26
Alzheimer's Disease: Overview
The clinical diagnosis of AD hinges on the presence of memory and other cognitive impairments. Biomarkers, such as changes in Aβ and tau...
 01:24
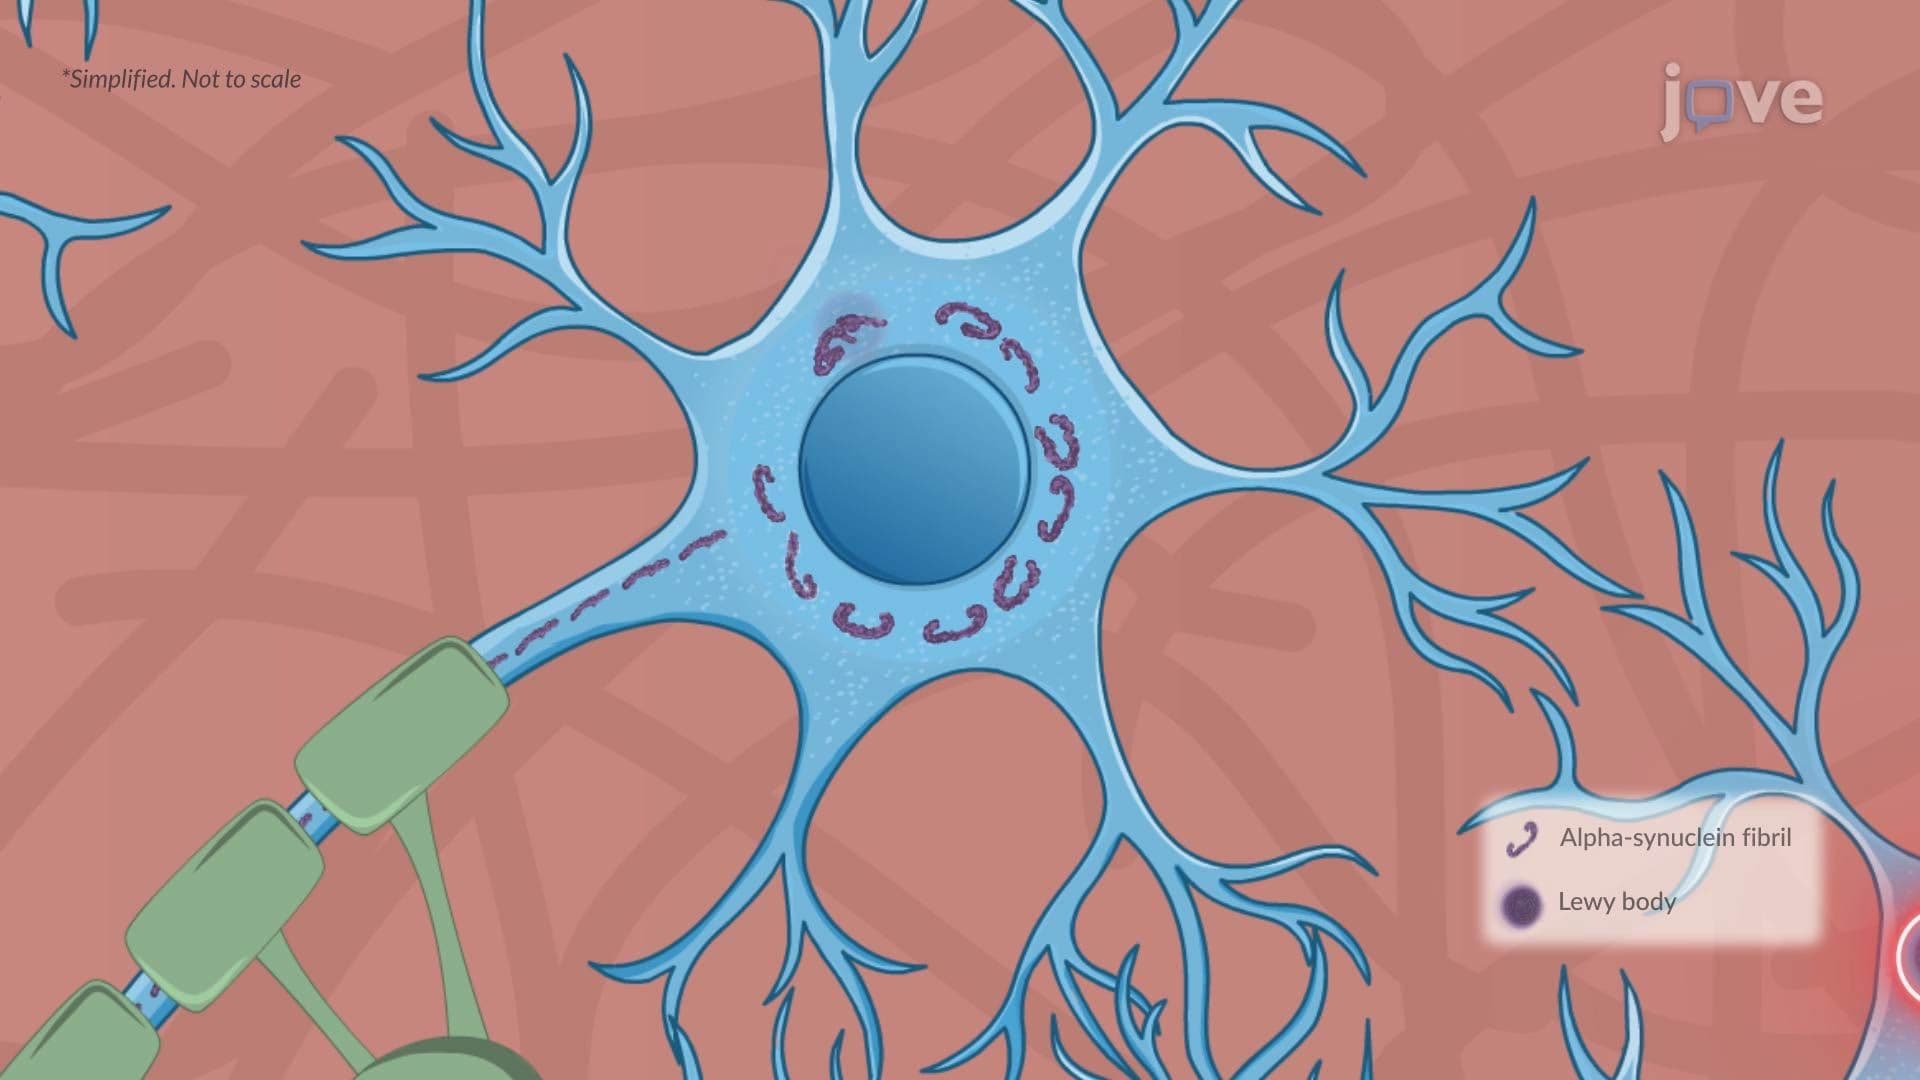
Parkinson Disease ll: Pathophysiology