Related Experiment Video
Updated: May 14, 2026


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
An accurate and linear-scaling method for calculating charge-transfer excitation energies and diabatic couplings
Michele Pavanello1, Troy Van Voorhis, Lucas Visscher
1Department of Chemistry, Rutgers University, Newark, New Jersey 07102-1811, USA. m.pavanello@rutgers.edu

Abstract:
Quantum-mechanical methods that are both computationally fast and accurate are not yet available for electronic excitations having charge transfer character. In this work, we present a significant step forward towards this goal for those charge transfer excitations that take place between non-covalently bound molecules. In particular, we present a method that scales linearly with the number of non-covalently bound molecules in the system and is based on a two-pronged approach: The molecular electronic structure of broken-symmetry charge-localized states is obtained with the frozen density embedding formulation of subsystem density-functional theory; subsequently, in a post-SCF calculation, the full-electron Hamiltonian and overlap matrix elements among the charge-localized states are evaluated with an algorithm which takes full advantage of the subsystem DFT density partitioning technique. The method is benchmarked against coupled-cluster calculations and achieves chemical accuracy for the systems considered for intermolecular separations ranging from hydrogen-bond distances to tens of Ångstroms. Numerical examples are provided for molecular clusters comprised of up to 56 non-covalently bound molecules.
More Related Videos
Related Concept Videos
 01:28
Debye–Huckel–Onsager Conductance Equation
 01:24
Electrochemical Systems
 01:21
Energy Associated With a Charge Distribution
 01:00
Coulometry: Overview
 02:49
Calculating Standard Free Energy Changes
 02:34
Arrhenius Plots
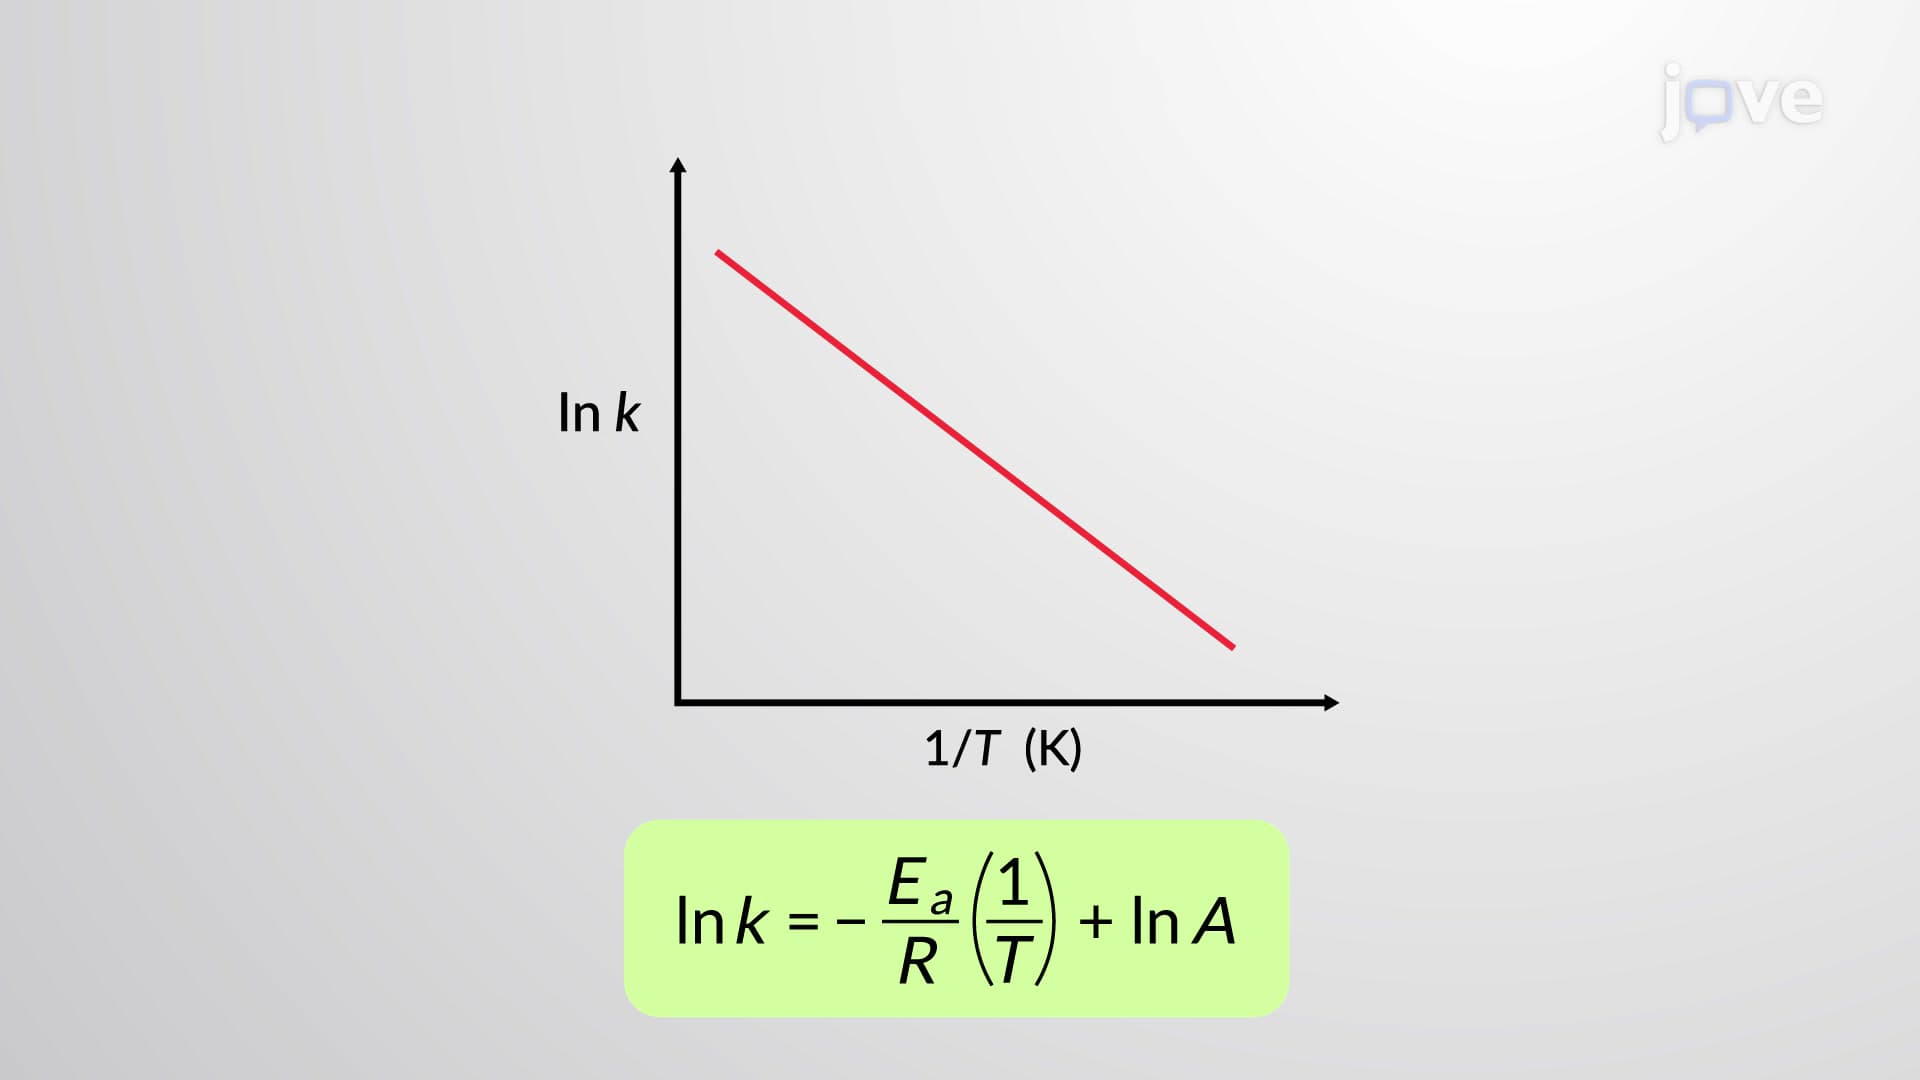
The Arrhenius equation can be used to...