Related Experiment Video
Updated: May 14, 2026

Accelerated Type 1 Diabetes Induction in Mice by Adoptive Transfer of Diabetogenic CD4+ T Cells
Published on: May 6, 2013
The nonconventional MHC class II molecule DM governs diabetes susceptibility in NOD mice
Marc A J Morgan1, Pari S S Muller, Arne Mould
1Sir William Dunn School of Pathology, University of Oxford, Oxford, United Kingdom.

Abstract:
The spontaneous destruction of insulin producing pancreatic beta cells in non-obese diabetic (NOD) mice provides a valuable model of type 1 diabetes. As in humans, disease susceptibility is controlled by the classical MHC class II genes that guide CD4(+) T cell responses to self and foreign antigens. It has long been suspected that the dedicated class II chaperone designated HLA-DM in humans or H-2M in mice also makes an important contribution, but due to tight linkage within the MHC, a possible role played by DM peptide editing has not been previously tested by conventional genetic approaches. Here we exploited newly established germ-line competent NOD ES cells to engineer a loss of function allele. DM deficient NOD mice display defective class II peptide occupancy and surface expression, and are completely protected against type 1 diabetes. Interestingly the mutation results in increased proportional representation of CD4(+)Foxp3(+) regulatory T cells and the absence of pathogenic CD4(+) T effectors. Overall, this striking phenotype establishes that DM-mediated peptide selection plays an essential role in the development of autoimmune diabetes in NOD mice.
Related Concept Videos
 01:26
Type II Diabetes I: Introduction
 01:26
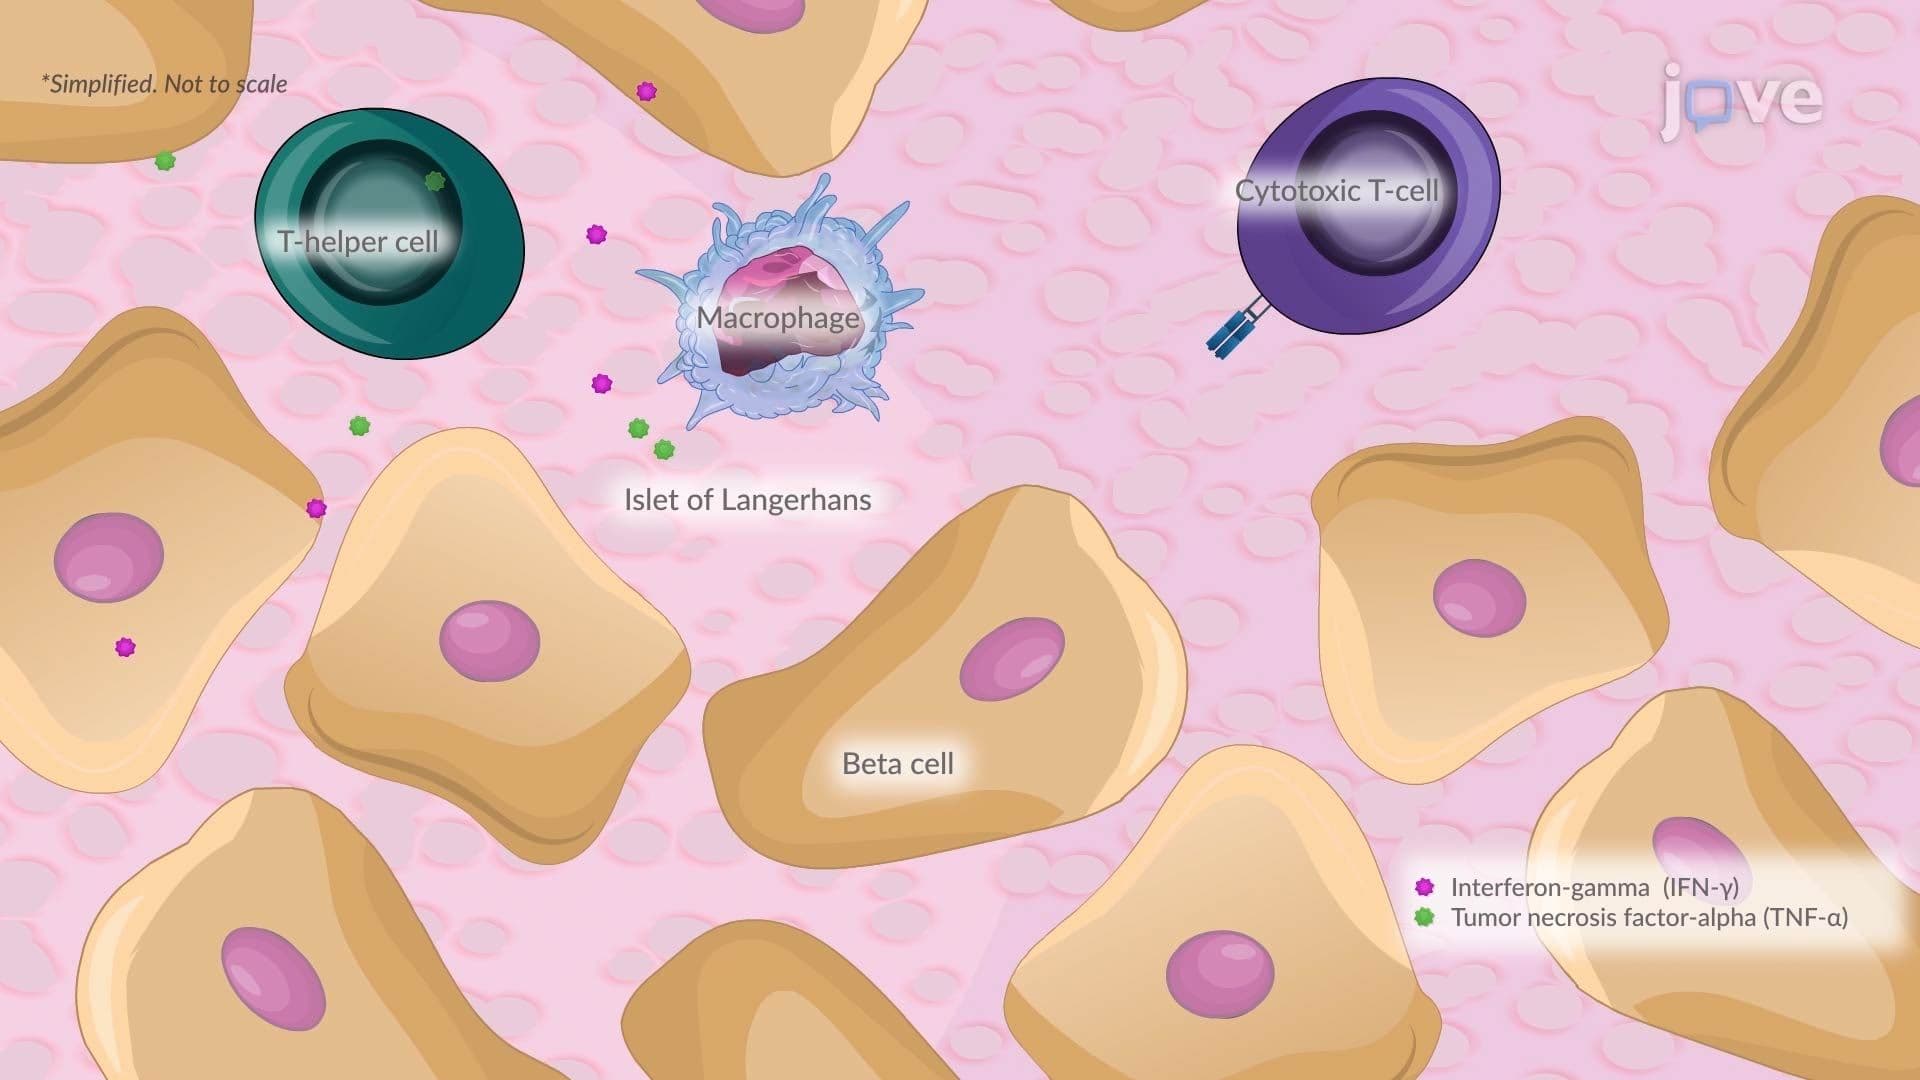
Type I Diabetes II: Pathophysiology
 01:22
Diabetes Mellitus: Type 2 and Gestational
 01:22
Diabetes Mellitus: Overview and Type I Subtype
Type 1 diabetes is an autoimmune disease in which the immune system mistakenly attacks and destroys the insulin-producing beta cells in the pancreas. As a result, the body is unable to produce sufficient insulin, and individuals with...
 01:24
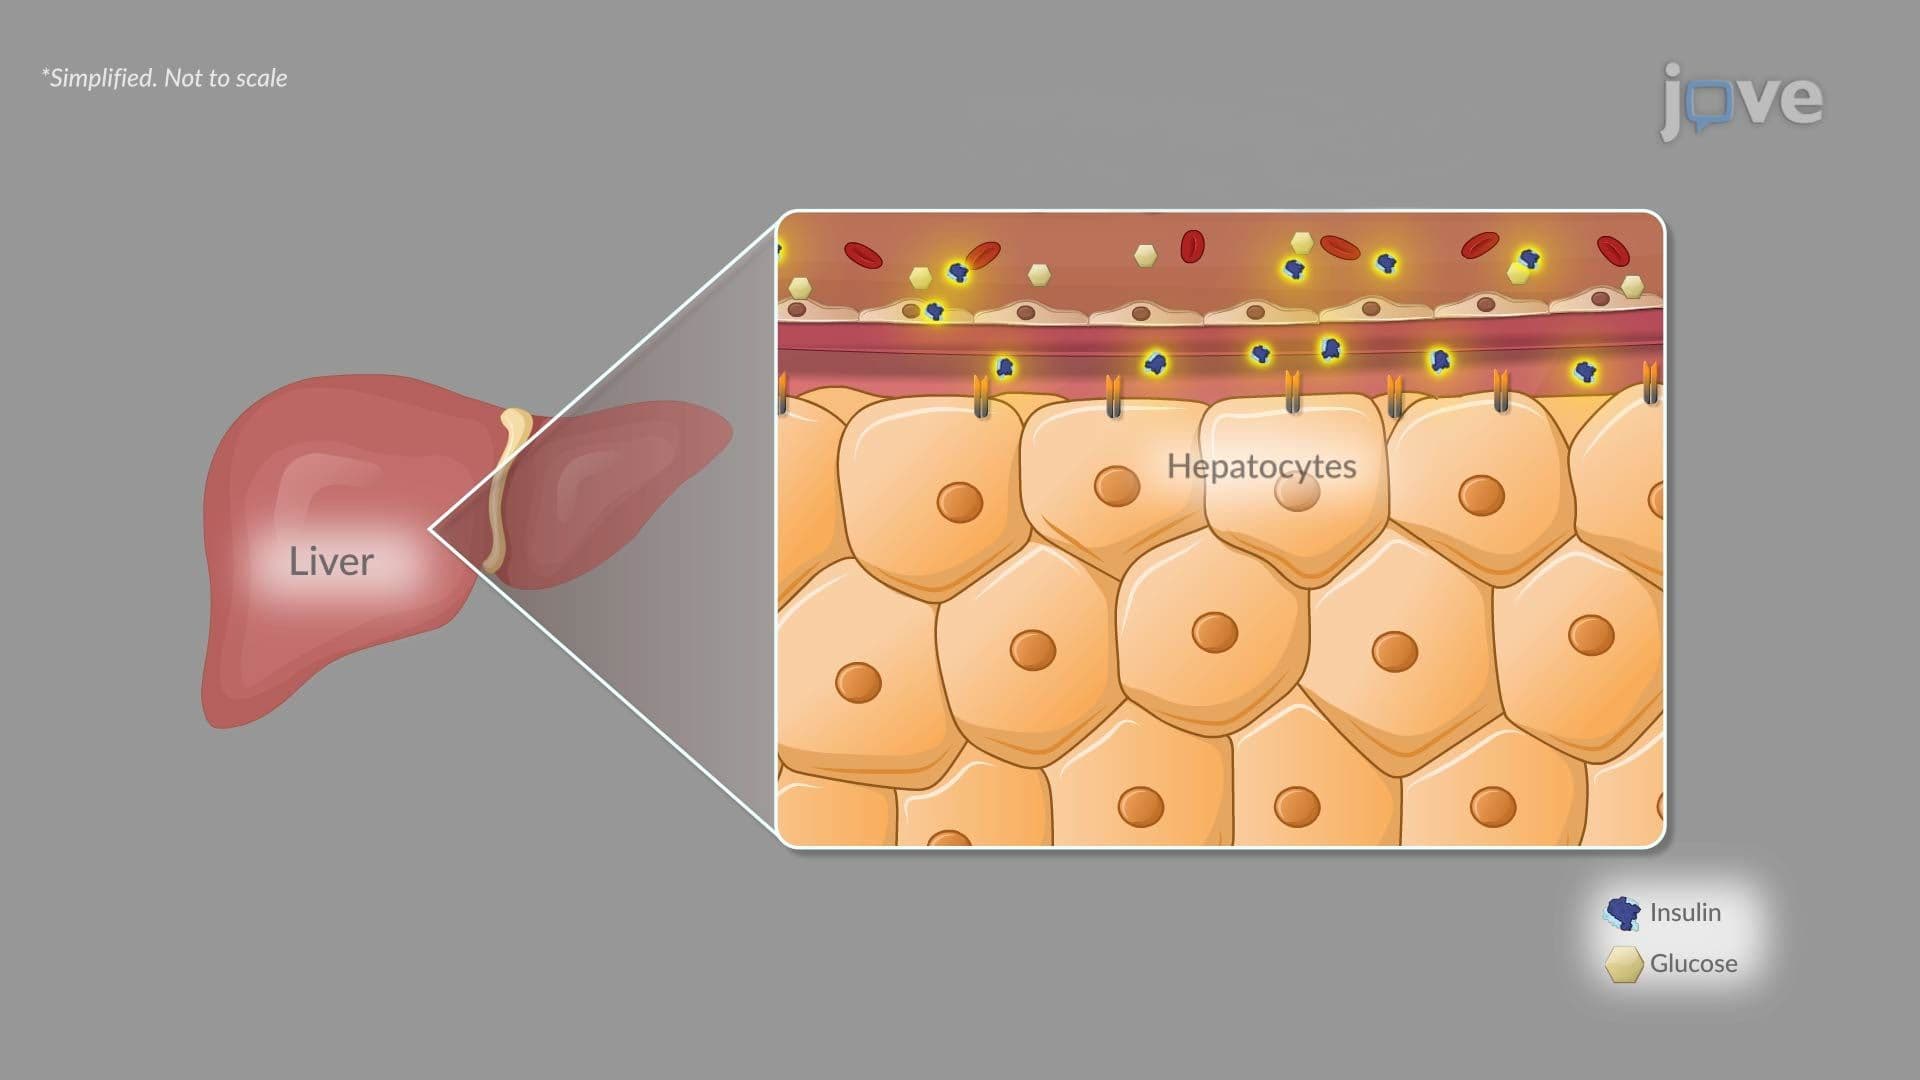
Type II Diabetes II: Pathophysiology
 01:12
Type I Diabetes I: Introduction