Related Experiment Video
Updated: May 14, 2026

Photoelectron Imaging of Anions Illustrated by 310 Nm Detachment of F−
Published on: July 27, 2018
Lowest triplet (n, π*) electronic state of acrolein: determination of structural parameters by cavity ringdown
Nikolaus C Hlavacek1, Michael O McAnally, Stephen Drucker
1Department of Chemistry, University of Wisconsin-Eau Claire, Eau Claire, Wisconsin 54702-4004, USA.

Abstract:
The cavity ringdown absorption spectrum of acrolein (propenal, CH(2)=CH-CH=O) was recorded near 412 nm, under bulk-gas conditions at room temperature and in a free-jet expansion. The measured spectral region includes the 0(0)(0) band of the T(1)(n, π*) ← S(0) system. We analyzed the 0(0)(0) rotational contour by using the STROTA computer program [R. H. Judge et al., J. Chem. Phys. 103, 5343 (1995)], which incorporates an asymmetric rotor Hamiltonian for simulating and fitting singlet-triplet spectra. We used the program to fit T(1)(n, π*) inertial constants to the room-temperature contour. The determined values (cm(-1)), with 2σ confidence intervals, are A = 1.662 ± 0.003, B = 0.1485 ± 0.0006, C = 0.1363 ± 0.0004. Linewidth analysis of the jet-cooled spectrum yielded a value of 14 ± 2 ps for the lifetime of isolated acrolein molecules in the T(1)(n, π*), v = 0 state. We discuss the observed lifetime in the context of previous computational work on acrolein photochemistry. The spectroscopically derived inertial constants for the T(1)(n, π*) state were used to benchmark a variety of computational methods. One focus was on complete active space methods, such as complete active space self-consistent field (CASSCF) and second-order perturbation theory with a CASSCF reference function (CASPT2), which are applicable to excited states. We also examined the equation-of-motion coupled-cluster and time-dependent density function theory excited-state methods, and finally unrestricted ground-state techniques, including unrestricted density functional theory and unrestricted coupled-cluster theory with single and double and perturbative triple excitations. For each of the above methods, we or others [O. S. Bokareva et al., Int. J. Quantum Chem. 108, 2719 (2008)] used a triple zeta-quality basis set to optimize the T(1)(n, π*) geometry of acrolein. We find that the multiconfigurational methods provide the best agreement with fitted inertial constants, while the economical unrestricted Perdew-Burke-Ernzerhof exchange-correlation hybrid functional (UPBE0) technique performs nearly as well.
Related Concept Videos
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:29
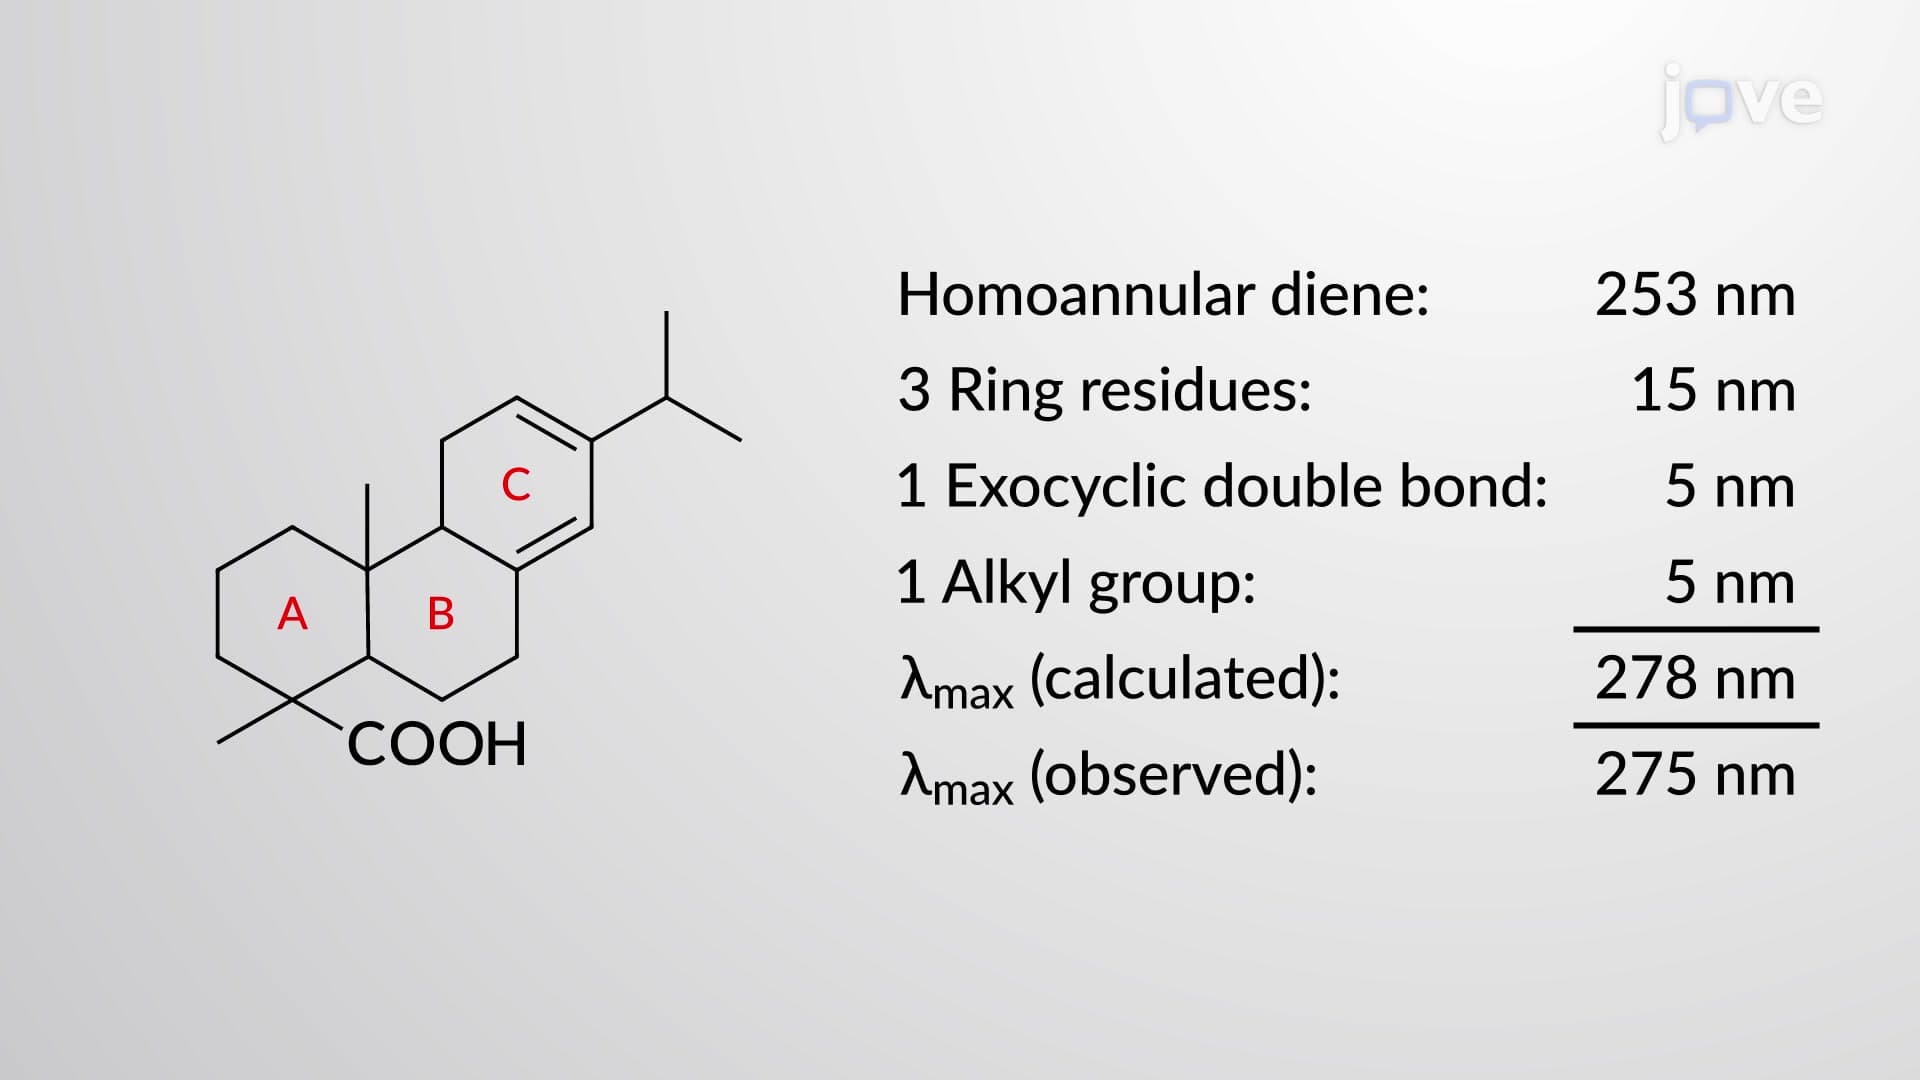
UV–Vis Spectroscopy: Woodward–Fieser Rules
 01:07
Radicals: Electronic Structure and Geometry
Accordingly, the structure of a trivalent radical lies between the geometries of carbocations and carbanions. An sp2-hybridized carbocation is trigonal planar, while an sp3-hybridized carbanion is trigonal pyramidal. Here, the difference in geometry is...
 01:13
¹H NMR: Complex Splitting
Splitting diagrams or splitting tree diagrams are routinely used to depict such complex couplings. While drawing splitting diagrams, the splitting with the larger coupling constant is usually applied first.
 01:26
Spectroscopy of Carboxylic Acid Derivatives
In the...
 01:37
NMR Spectroscopy of Benzene Derivatives