Related Experiment Video
Updated: May 14, 2026
Catalytic Reactions at Amine-Stabilized and Ligand-Free Platinum Nanoparticles Supported on Titania During Hydrogenation of Alkenes and Aldehydes
Published on: June 24, 2022
DFT studies on doping effect of Al12X: adsorption and dissociation of H2O on Al12X clusters
Jian-Ying Zhao1, Feng-Qi Zhao, Si-Yu Xu
1Key Laboratory of Soft Chemistry and Functional Materials of MOE, School of Chemical Engineering, Nanjing University of Science and Technology, Nanjing 210094, PR China.

Abstract:
The adsorption and reaction of H2O molecule on neutral X-centered icosahedronal Al12X clusters (X = Al, Mg, Zn, Ga, Ni, Fe, B, C, Si, P) were investigated by PW91, PBE, and PWC methods. Reaction energies and reaction barriers were determined. The spin states and the doped atoms have important influences on the Al12X geometries, density, electronic properties, and energy density of reaction between Al12X with a single H2O molecule. The energies of the neutral X-centered Al12X are lower than that of surface X-replaced Al12X with the exception of Al12Mg. The H2O dissociation on the Al12X (X = Mg, Zn, Ga, Ni, Fe) clusters have relatively low activation barriers, but large activation barriers for Al12X (X = B, C, Si, P). The activation barrier of water dissociation on the singlet Al12Fe cluster is the lowest, whereas the highest barrier is with the Al12C. The reaction of H2O with Al12Fe is the most exothermic. The center-Fe atom can move out to the surface after the adsorption and dissociation of H2O with an energy barrier of 172 kJ/mol. The results showed that the water dissociation on the Al12X cluster can be tuned by controllable X doping.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 07:20
Discovery and Synthesis Optimization of Isoreticular Al(III) Phosphonate-Based Metal-Organic Framework Compounds Using High-Throughput Methods
Published on: October 6, 2023
Related Concept Videos
 01:27
π Electron Effects on Chemical Shift: Overview
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 00:48
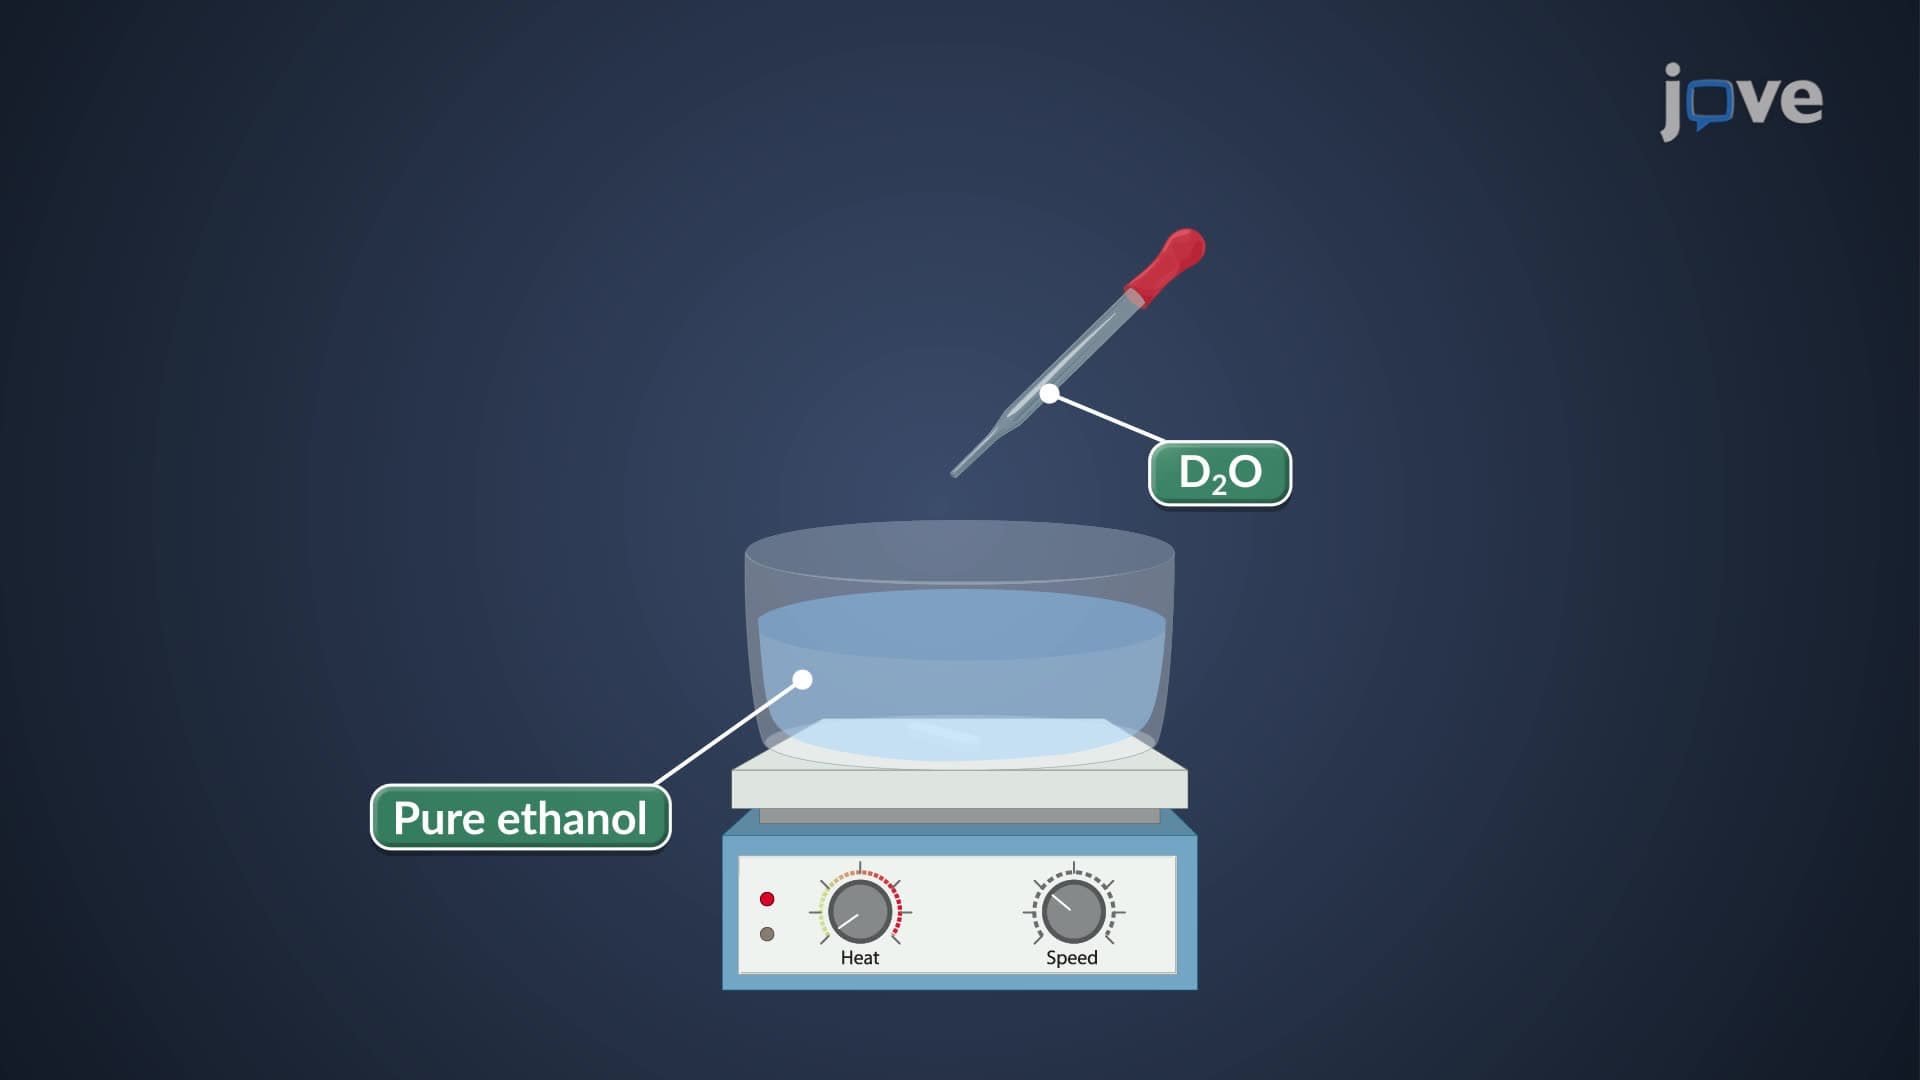
¹H NMR of Labile Protons: Deuterium (²H) Substitution
 01:18
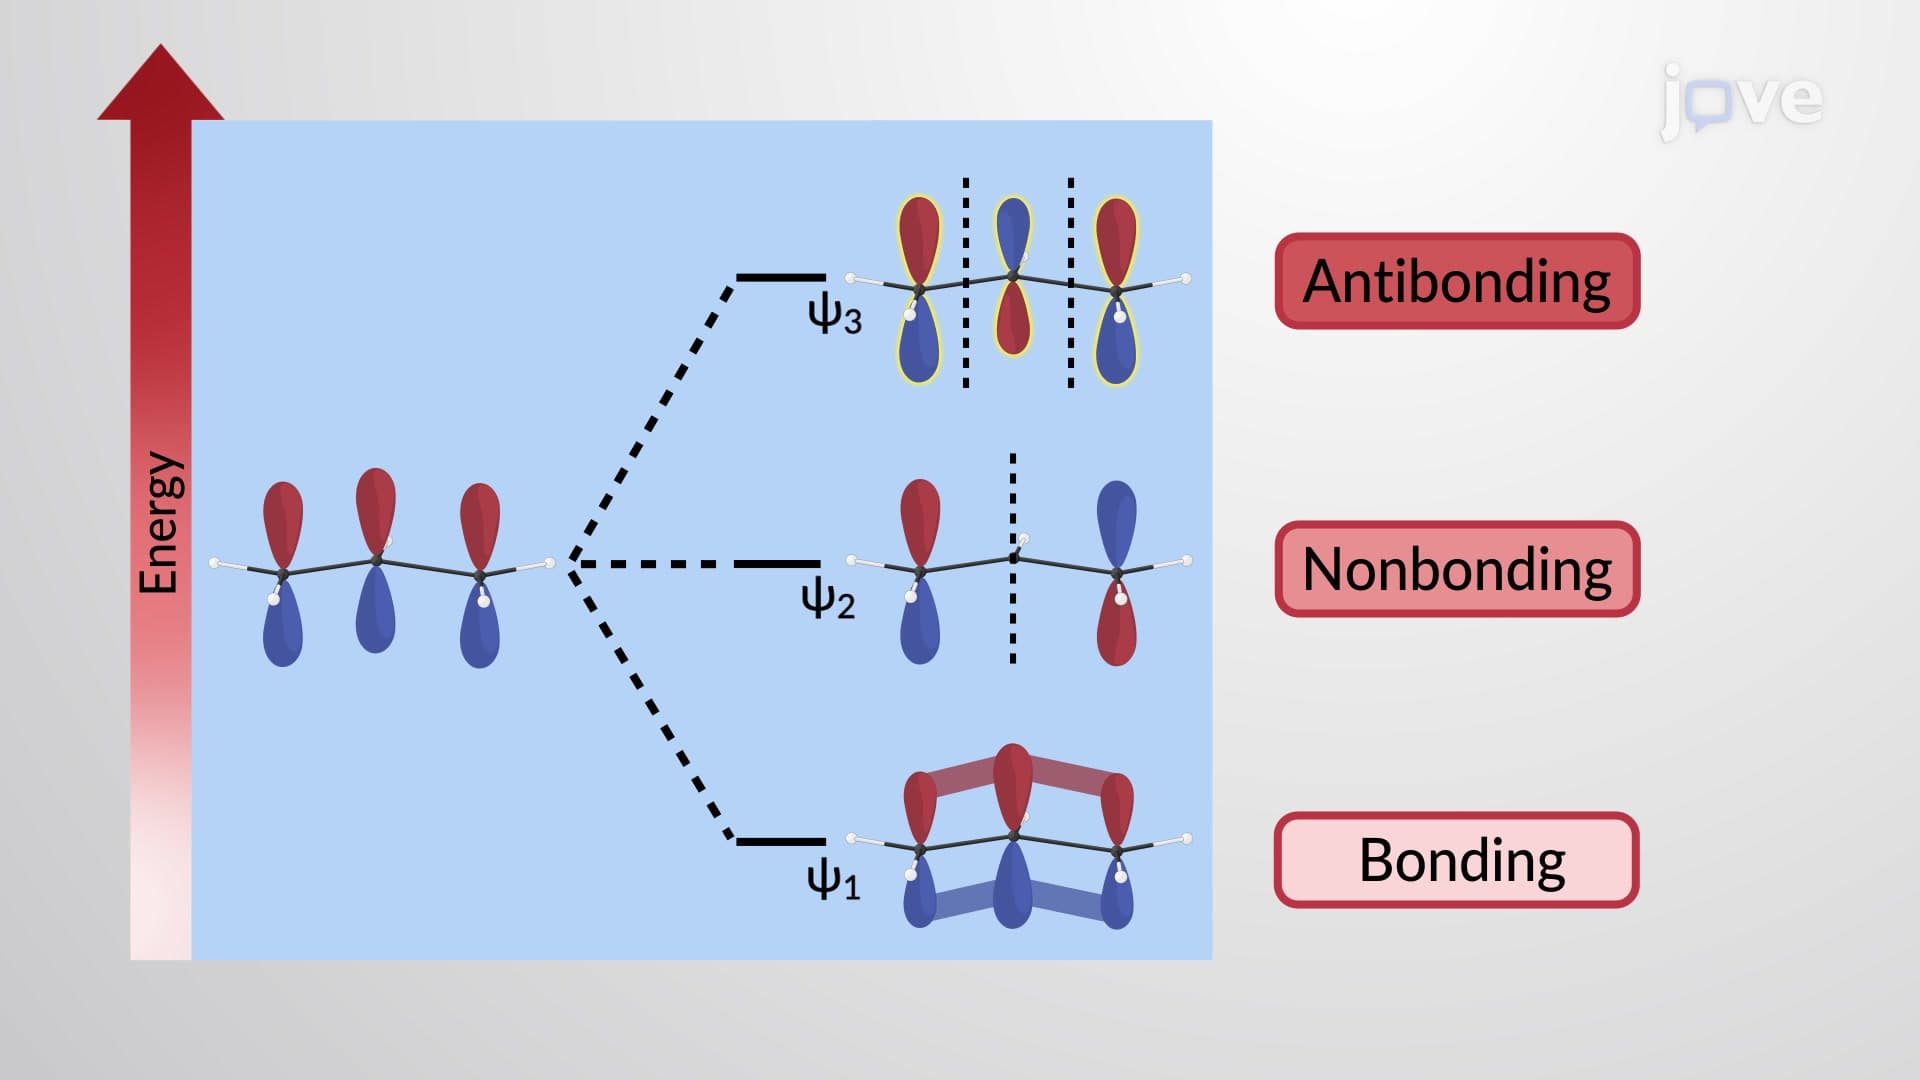
π Molecular Orbitals of the Allyl Cation and Anion
 01:20
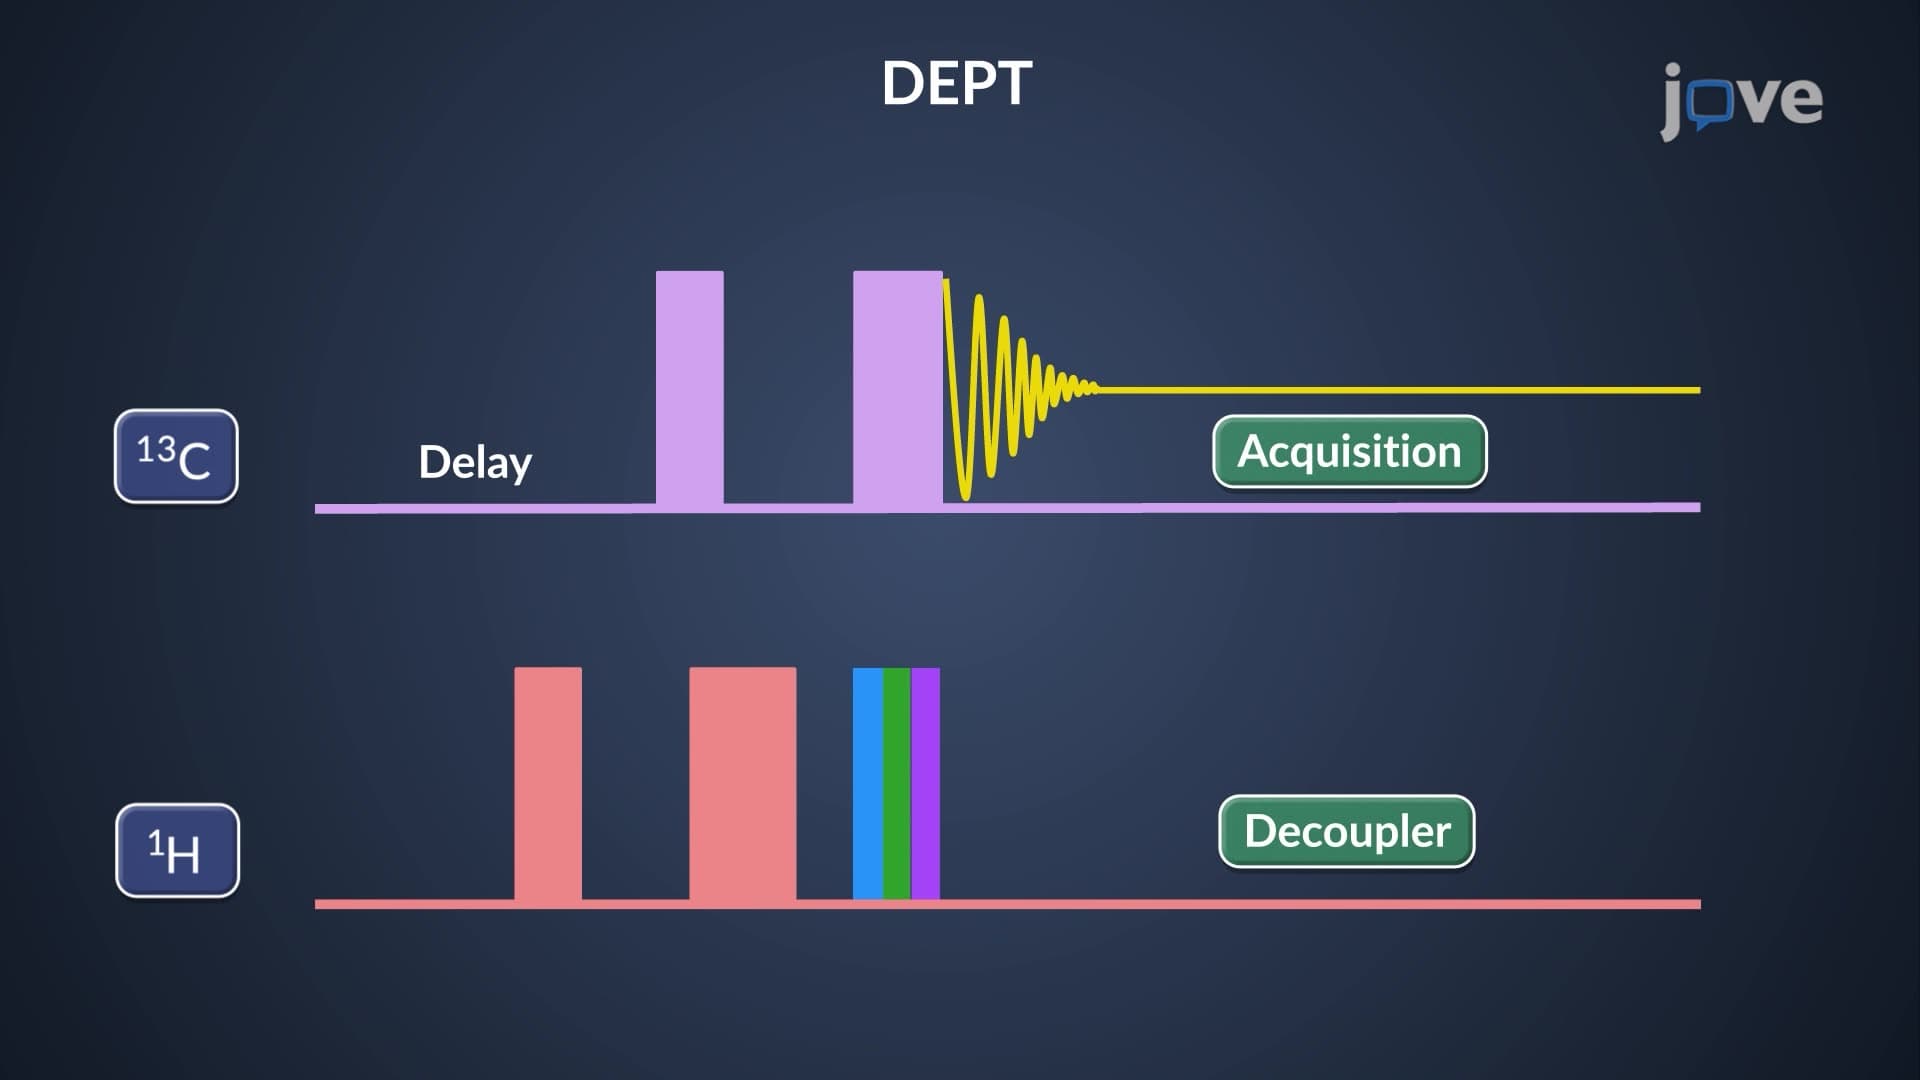
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 01:29
Adsorption Isotherms I