Related Experiment Video
Updated: May 14, 2026
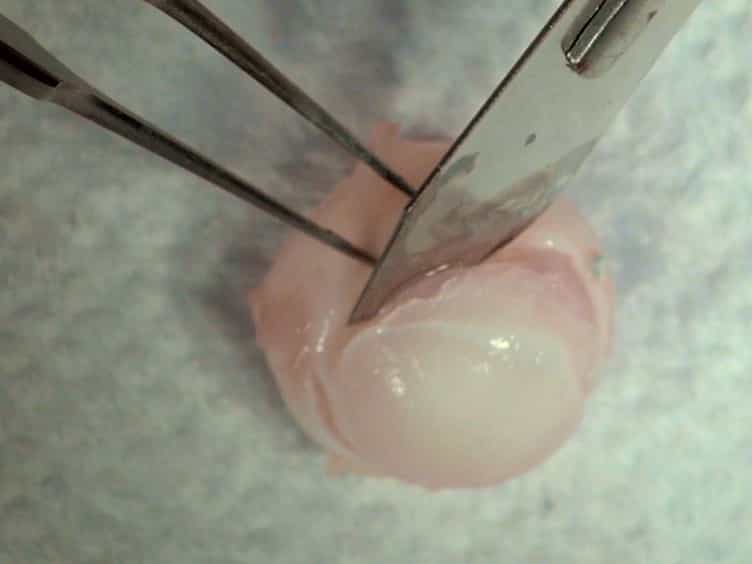
Use of Rabbit Eyes in Pharmacokinetic Studies of Intraocular Drugs
Published on: July 23, 2016
The antibody drug absorption following subcutaneous or intramuscular administration and its mathematical description
Liang Zhao1, Ping Ji, Zhihong Li
1Office of Clinical Pharmacology, Office of Translational Sciences, Center for Drug Evaluation and Research, US Food and Drug Administration, Silver Spring, MD, USA. liang.zhao@fda.hhs.gov

The main objective of this paper is to propose a quantitative model to describe the absorption process for monoclonal antibody (mAb) following subcutaneous (SC) or intramuscular (IM) administration. A hybrid model was established by coupling the physiologically based absorption process with a conventional pharmacokinetic-pharmacodynamic (PK-PD) or PK model associated with intravenous infusion. Key physiological parameters evaluated include the volume distribution before systemic absorption, mAb drug clearance during lymphatic transport, the neonatal Fc receptor (FcRn) capacity, the intrinsic drug clearance via lysosomal proteolytic process, and the lymphatic flow rate. Sensitivity analyses were performed to identify those physiological parameters significantly impacting time to peak concentration (T(max)) or drug bioavailability. Simulation results showed that lymphatic flow rate is the only influential factor to T(max). Lymphatic transit time and drug clearance during lymphatic transport are most influential to bioavailability but not identifiable in their effects. Bioavailability is positively related to the lymphatic flow only at its low range (i.e., at <0.5-fold of the selected value of 0.043 mL/min). The rest of physiological parameters only have marginal effects on either drug bioavailability or T(max). Finally, simulation results confirmed lymphatic transport as the major route of mAb delivery.
The main objective of this paper is to propose a quantitative model to describe the absorption process for monoclonal antibody (mAb) following subcutaneous (SC) or intramuscular (IM) administration. A hybrid model was established by coupling the physiologically based absorption process with a conventional pharmacokinetic-pharmacodynamic (PK-PD) or PK model associated with intravenous infusion. Key physiological parameters evaluated include the volume distribution before systemic absorption, mAb drug clearance during lymphatic transport, the neonatal Fc receptor (FcRn) capacity, the intrinsic drug clearance via lysosomal proteolytic process, and the lymphatic flow rate. Sensitivity analyses were performed to identify those physiological parameters significantly impacting time to peak concentration (T(max)) or drug bioavailability. Simulation results showed that lymphatic flow rate is the only influential factor to T(max). Lymphatic transit time and drug clearance during lymphatic transport are most influential to bioavailability but not identifiable in their effects. Bioavailability is positively related to the lymphatic flow only at its low range (i.e., at <0.5-fold of the selected value of 0.043 mL/min). The rest of physiological parameters only have marginal effects on either drug bioavailability or T(max). Finally, simulation results confirmed lymphatic transport as the major route of mAb delivery.
Related Concept Videos
 01:15
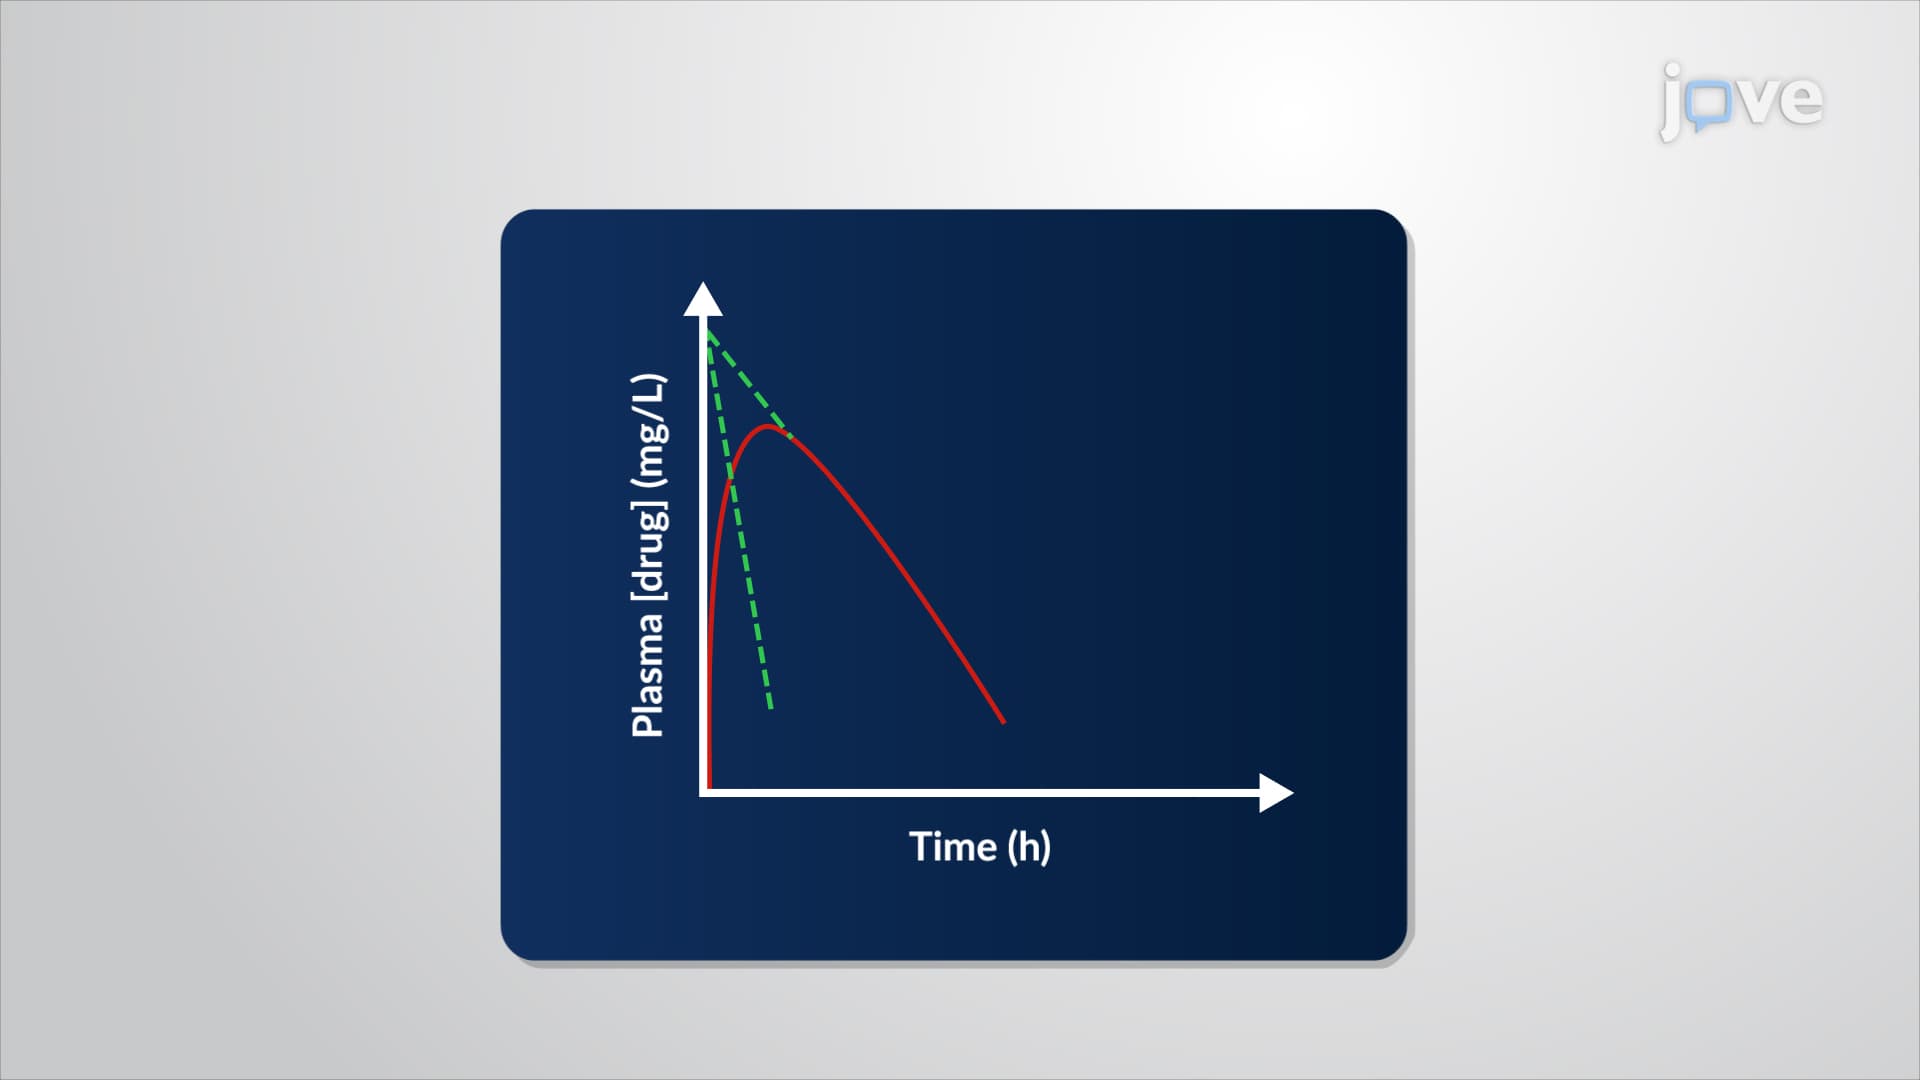
One-Compartment Open Model for Extravascular Administration: First-Order Absorption Model
 01:12
One-Compartment Open Model for Extravascular Administration: Zero-Order Absorption Model
Zero-order absorption maintains a steady rate irrespective of the amount of drug left to be absorbed, making it a constant process. In the...
 01:14
Model Approaches for Pharmacokinetic Data: Compartment Models
Two primary types of compartment models are recognized: mammillary and catenary. The more...
 01:14
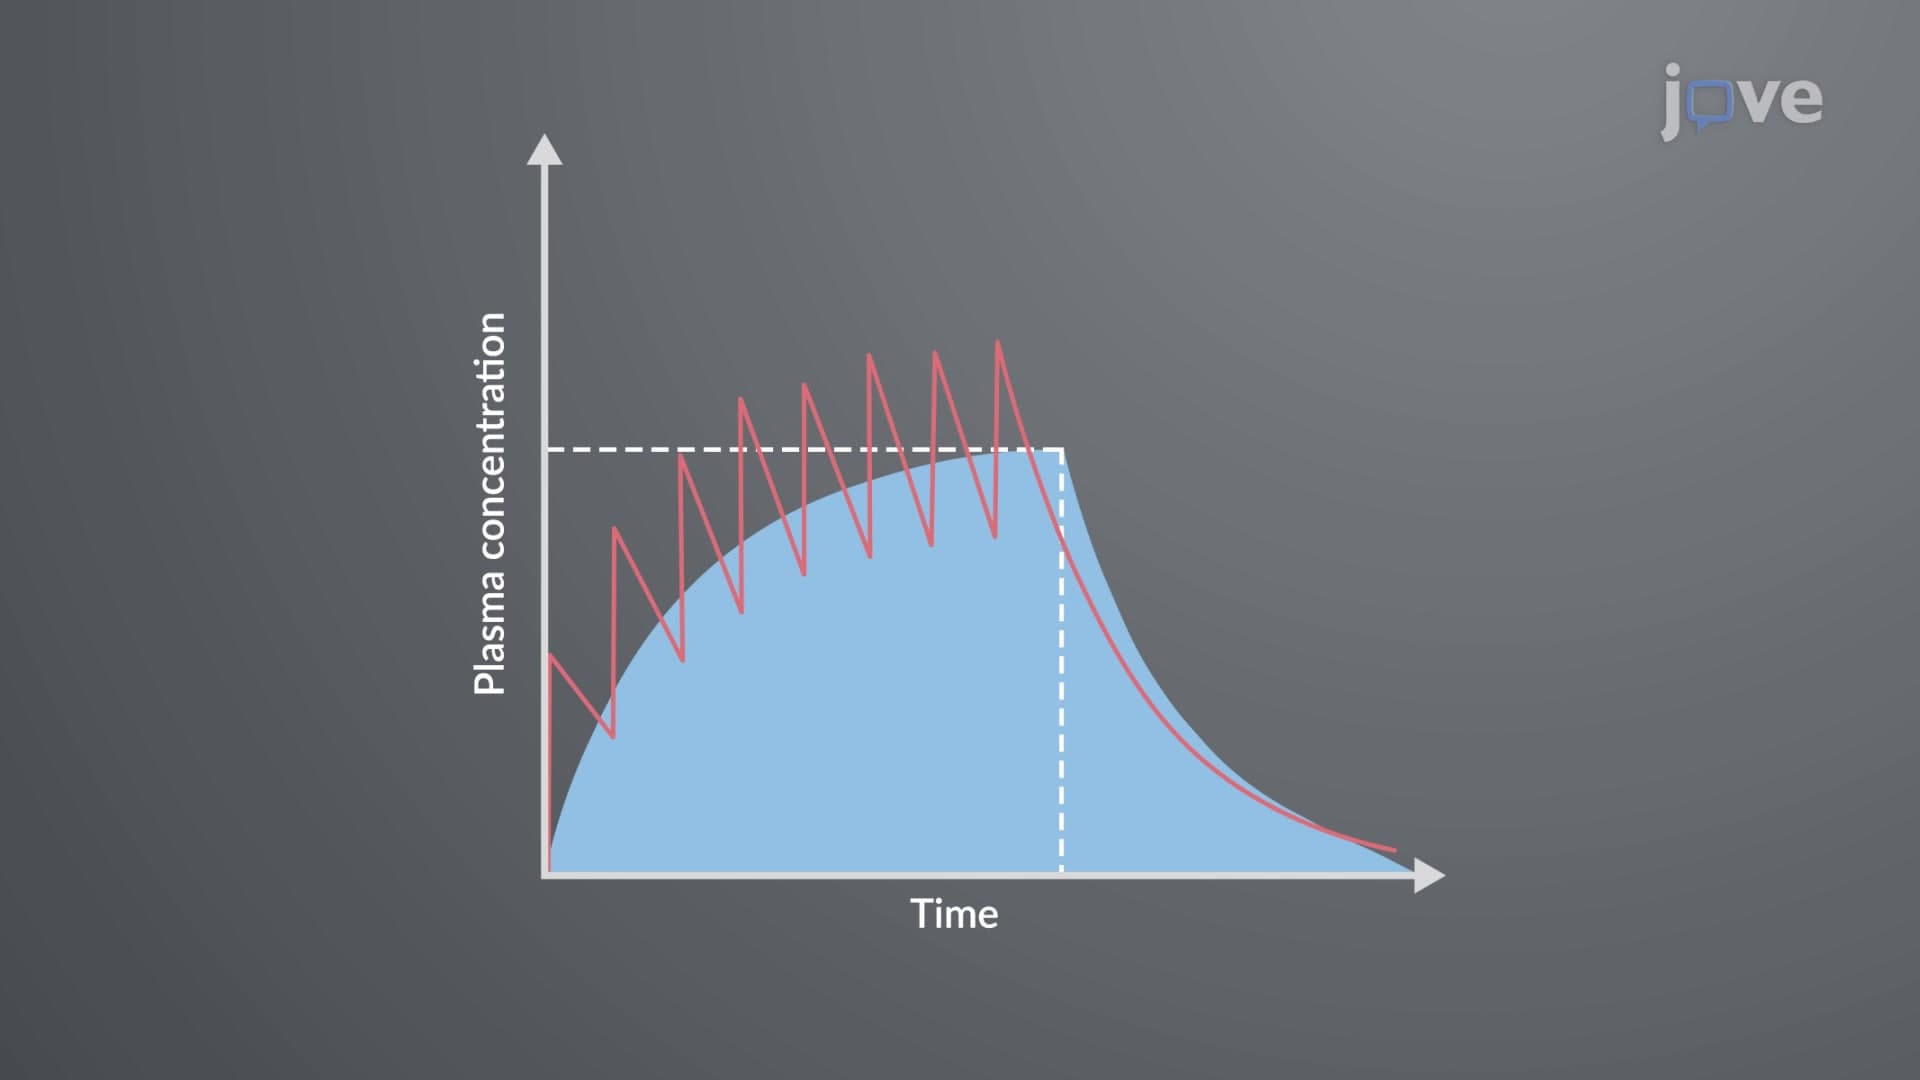
Compartment Models: Single-Compartment Model
 01:12
Two-Compartment Open Model: Extravascular Administration
The absorption exponent (ka) indicates the speed at which the drug is...
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding