Related Experiment Video
Updated: May 13, 2026
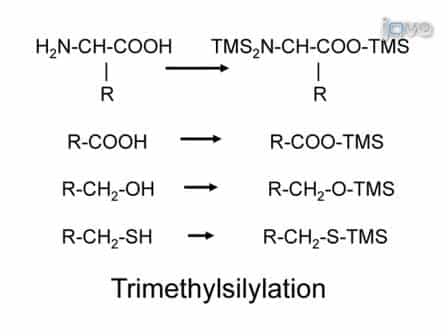
One-step Metabolomics: Carbohydrates, Organic and Amino Acids Quantified in a Single Procedure
Published on: June 25, 2010
Newborn screening for lysosomal storage disorders in hungary
Judit Wittmann1, Eszter Karg, Sàndor Turi
1Centogene GmbH, Laboratories for Biochemical Genetics and Newborn Screening, Vienna, Austria.

Abstract:
Even though lysosomal storage disorders (LSDs) are considered to be orphan diseases, they pose a highly relevant cause for morbidity and mortality as their cumulative prevalence is estimated to be 1:4,000. This is especially important as treatment in form of enzyme replacement therapy, substrate reduction therapy or stem cell transplantation is amenable for some LSDs. It is plausible that an early start of treatment might improve the overall prognosis and, even more important, prevent irreversible damage of key organs. To get a more precise insight into the real frequency of some LSDs in the general population, we screened 40,024 samples from the Hungarian newborn screening (NBS) program in Szeged for Fabry disease (FD), Gaucher disease (GD), Pompe disease (PD), and Niemann-Pick A/B (NPB) disease using tandem mass spectrometry. Altogether, 663 samples (1.66%) were submitted for retesting. Genetic confirmation was carried out for 120 samples with abnormal screening results after retesting, which identified three cases of GD, three cases of FD, nine cases of PD, and two cases with NPB. In some cases, we detected up to now unknown mutations - one in NPB and seven in PD - which raise questions about the clinical consequences of a NBS in the sense of late-onset manifestations. Overall, we conclude that screening for LSDs by tandem MS/MS followed by a genetic workup in identified patients is a robust, easy, valid, and feasible technology in newborn screening programs. Furthermore, early diagnosis of LSDs gives a chance to early treatment, but needs more clinical long-term data especially regarding the consequence of private mutations.
More Related Videos
 08:04
Isolation of Primary Mouse Hepatocytes for Nascent Protein Synthesis Analysis by Non-radioactive L-azidohomoalanine Labeling Method
Published on: October 23, 2018
 10:16
In Vitro Enzyme Measurement to Test Pharmacological Chaperone Responsiveness in Fabry and Pompe Disease
Published on: December 20, 2017
Related Concept Videos
 01:22
Lysosomal Hydrolases
 01:20
Inborn Errors of Metabolism
 01:27
Glucose Transporters
Facilitated diffusion-glucose transporters (GLUTs) are encoded by the solute-linked carrier (SLC) family 2, subfamily A gene family, or SLC2A. The 14 GLUT protein members are distributed into three classes: