Protein structure based prediction of catalytic residues
Related Experiment Video
Updated: Jan 19, 2026

Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
J Eduardo Fajardo1, Andras Fiser
1Department of Systems and Computational Biology, Albert Einstein College of Medicine, 1300 Morris Park Avenue, Bronx, NY 10461, USA.

Background:
Worldwide structural genomics projects continue to release new protein structures at an unprecedented pace, so far nearly 6000, but only about 60% of these proteins have any sort of functional annotation.
Results:
We explored a range of features that can be used for the prediction of functional residues given a known three-dimensional structure. These features include various centrality measures of nodes in graphs of interacting residues: closeness, betweenness and page-rank centrality. We also analyzed the distance of functional amino acids to the general center of mass (GCM) of the structure, relative solvent accessibility (RSA), and the use of relative entropy as a measure of sequence conservation. From the selected features, neural networks were trained to identify catalytic residues. We found that using distance to the GCM together with amino acid type provide a good discriminant function, when combined independently with sequence conservation. Using an independent test set of 29 annotated protein structures, the method returned 411 of the initial 9262 residues as the most likely to be involved in function. The output 411 residues contain 70 of the annotated 111 catalytic residues. This represents an approximately 14-fold enrichment of catalytic residues on the entire input set (corresponding to a sensitivity of 63% and a precision of 17%), a performance competitive with that of other state-of-the-art methods.
Conclusions:
We found that several of the graph based measures utilize the same underlying feature of protein structures, which can be simply and more effectively captured with the distance to GCM definition. This also has the added the advantage of simplicity and easy implementation. Meanwhile sequence conservation remains by far the most influential feature in identifying functional residues. We also found that due the rapid changes in size and composition of sequence databases, conservation calculations must be recalibrated for specific reference databases.


Related Concept Videos
 01:24
Protein Organization
The primary structure of a protein is its amino acid sequence....
 01:13
Protein Organization
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 01:22
Protein Folding
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
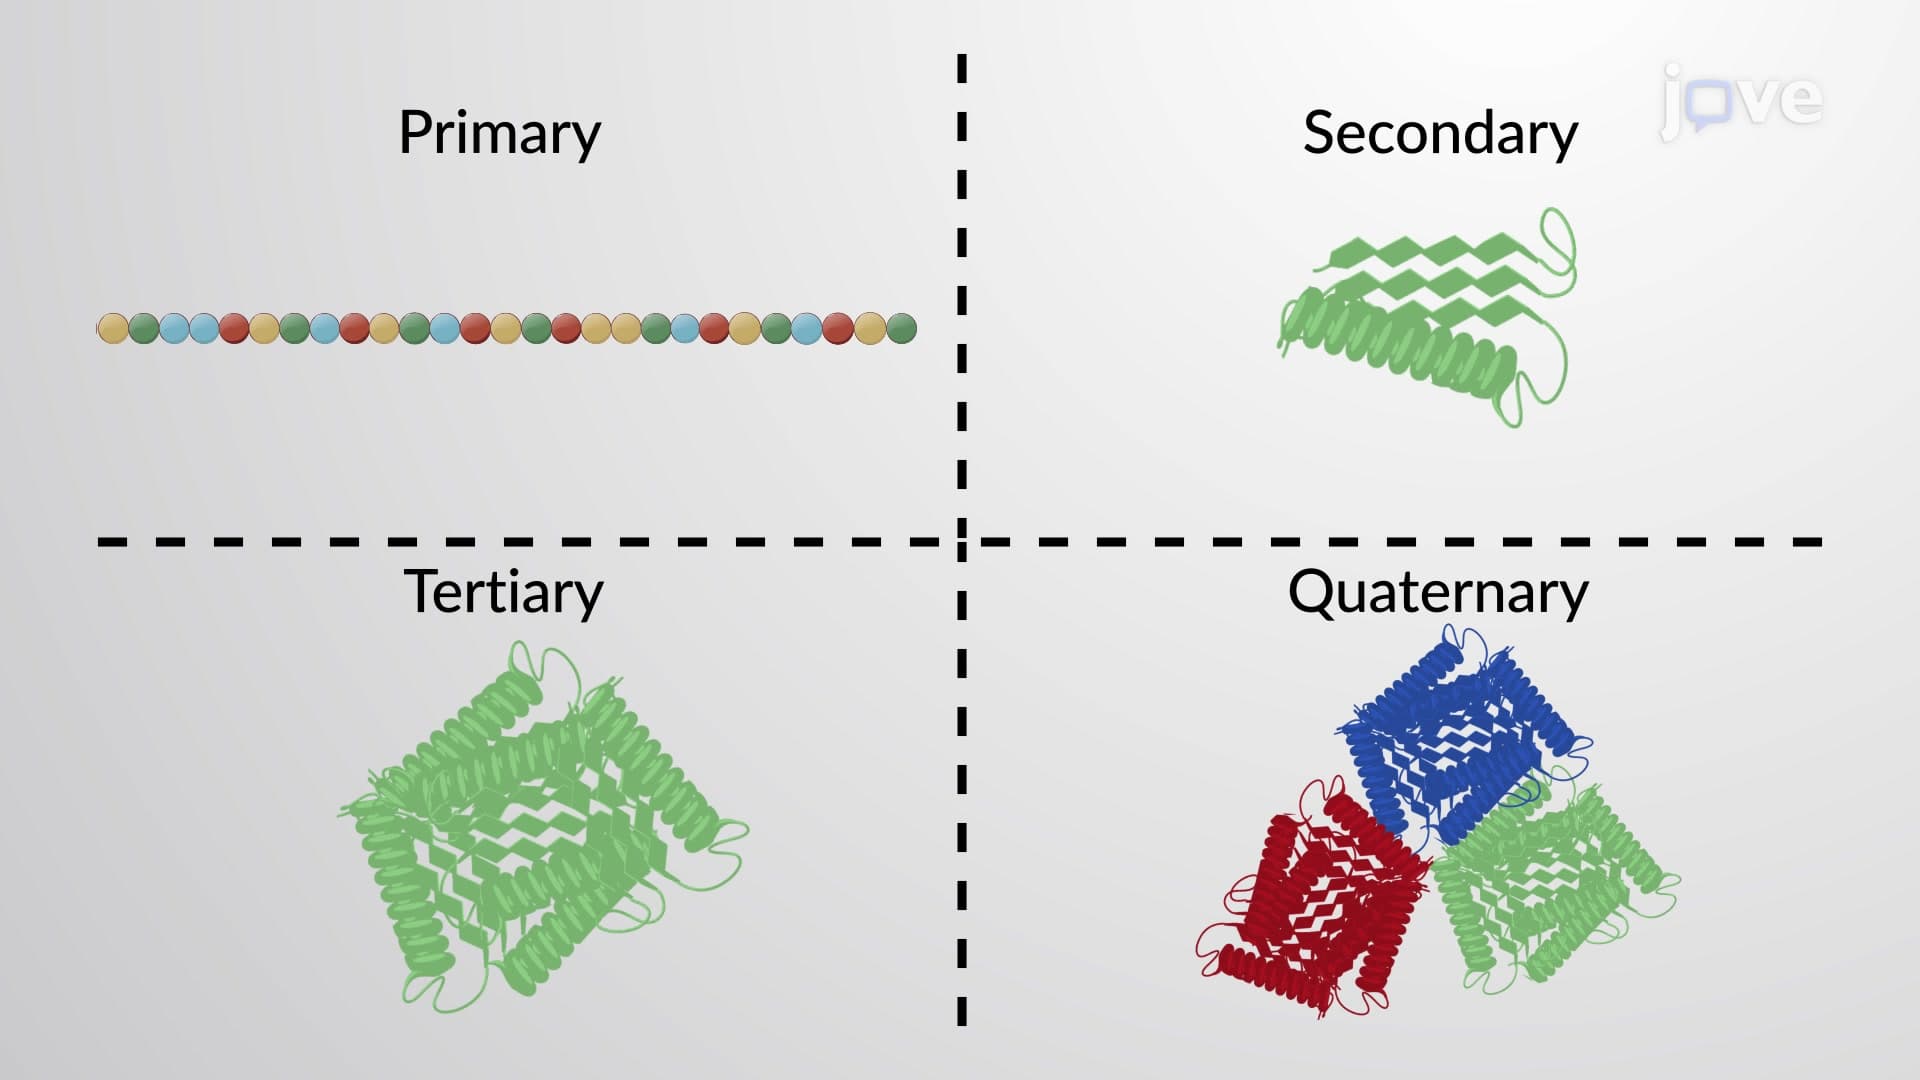
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...
 02:15
Protein and Protein Structure
A protein's shape is critical to its function. For example, an enzyme...