Related Experiment Video
Updated: May 13, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Which density functional is close to CCSD accuracy to describe geometry and interaction energy of small non-covalent
Karunakaran Remya1, Cherumuttathu H Suresh
1Chemical Sciences and Technology Division, CSIR-National Institute for Interdisciplinary, Science and Technology, Trivandrum 695 019, Kerala, India.

Abstract:
A benchmark study on all possible density functional theory (DFT) methods in Gaussian09 is done to locate functionals that agree well with CCSD/aug-cc-pVTZ geometry and Ave-CCSD(T)/(Q-T) interaction energy (Eint) for small non-covalently interacting molecular dimers in "dispersion-dominated" (class 1), "dipole-induced dipole" (class 2), and "dipole-dipole" (class 3) classes. A DFT method is recommended acceptable if the geometry showed close agreement to CCSD result (RMSD < 0.045) and Eint was within 80-120% accuracy. Among 382 tested functionals, 1-46% gave good geometry, 13-44% gave good Eint , while 1-33% satisfied geometry and energy criteria. Further screening to locate the best performing functionals for all the three classes was made by counting the acceptable values of energy and geometry given by each functionals. The meta-generalized gradient approximation (GGA) functional M06L was the best performer with total 14 hits; seven acceptable energies and seven acceptable geometries. This was the only functional "recommended" for at least two dimers in each class. The functionals M05, B2PLYPD, B971, mPW2PLYPD, PBEB95, and CAM-B3LYP gave 11 hits while PBEhB95, PW91B95, Wb97x, BRxVP86, BRxP86, HSE2PBE, HSEh1PBE, PBE1PBE, PBEh1PBE, and PW91TPSS gave 10 hits. Among these, M05, B971, mPW2PLYPD, Wb97x, and PW91TPSS were among the "recommended" list of at least one dimer from each class. Long-range correction (LC) of Hirao and coworkers to exchange-correlation functionals showed massive improvement in geometry and Eint . The best performing LC-functionals were LC-G96KCIS and LC-PKZBPKZB. Our results predict that M06L is the most trustworthy DFT method in Gaussian09 to study small non-covalently interacting systems.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 02:36
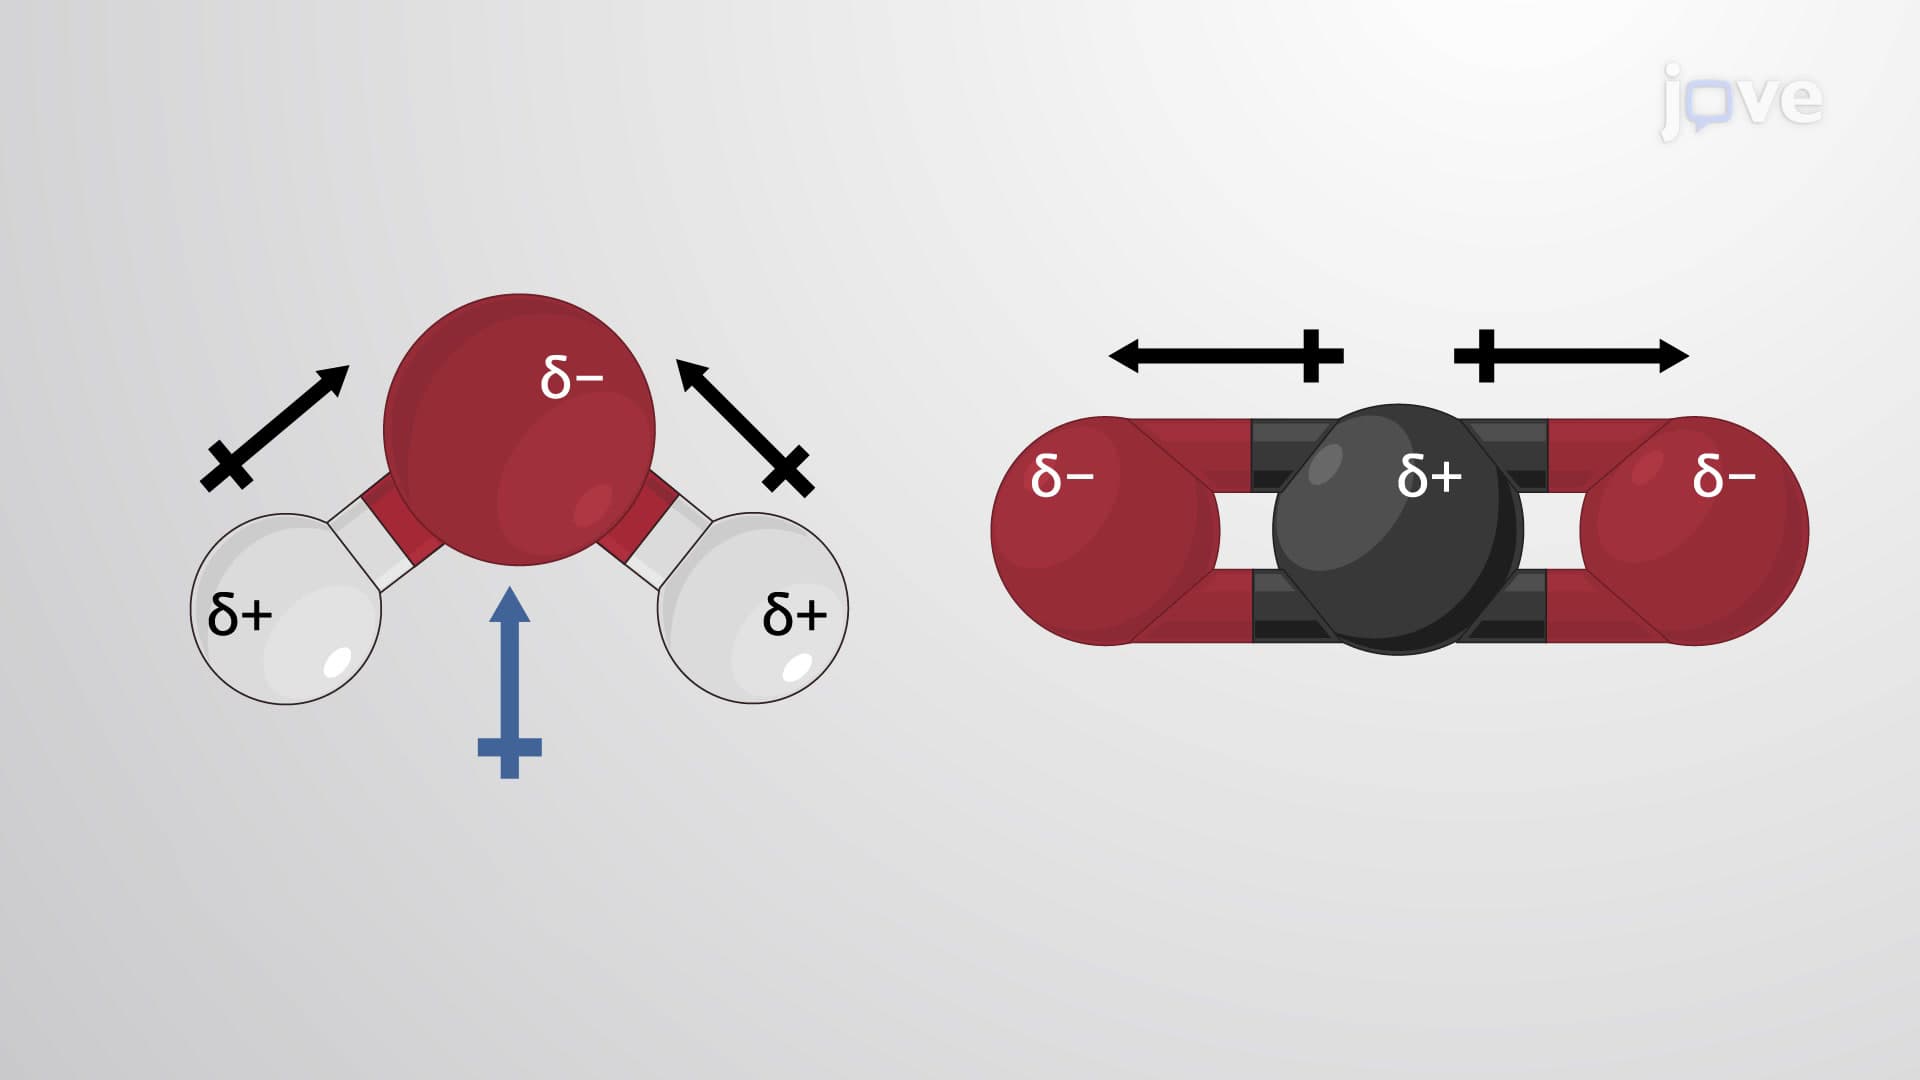
Molecular Geometry and Dipole Moments
 01:19
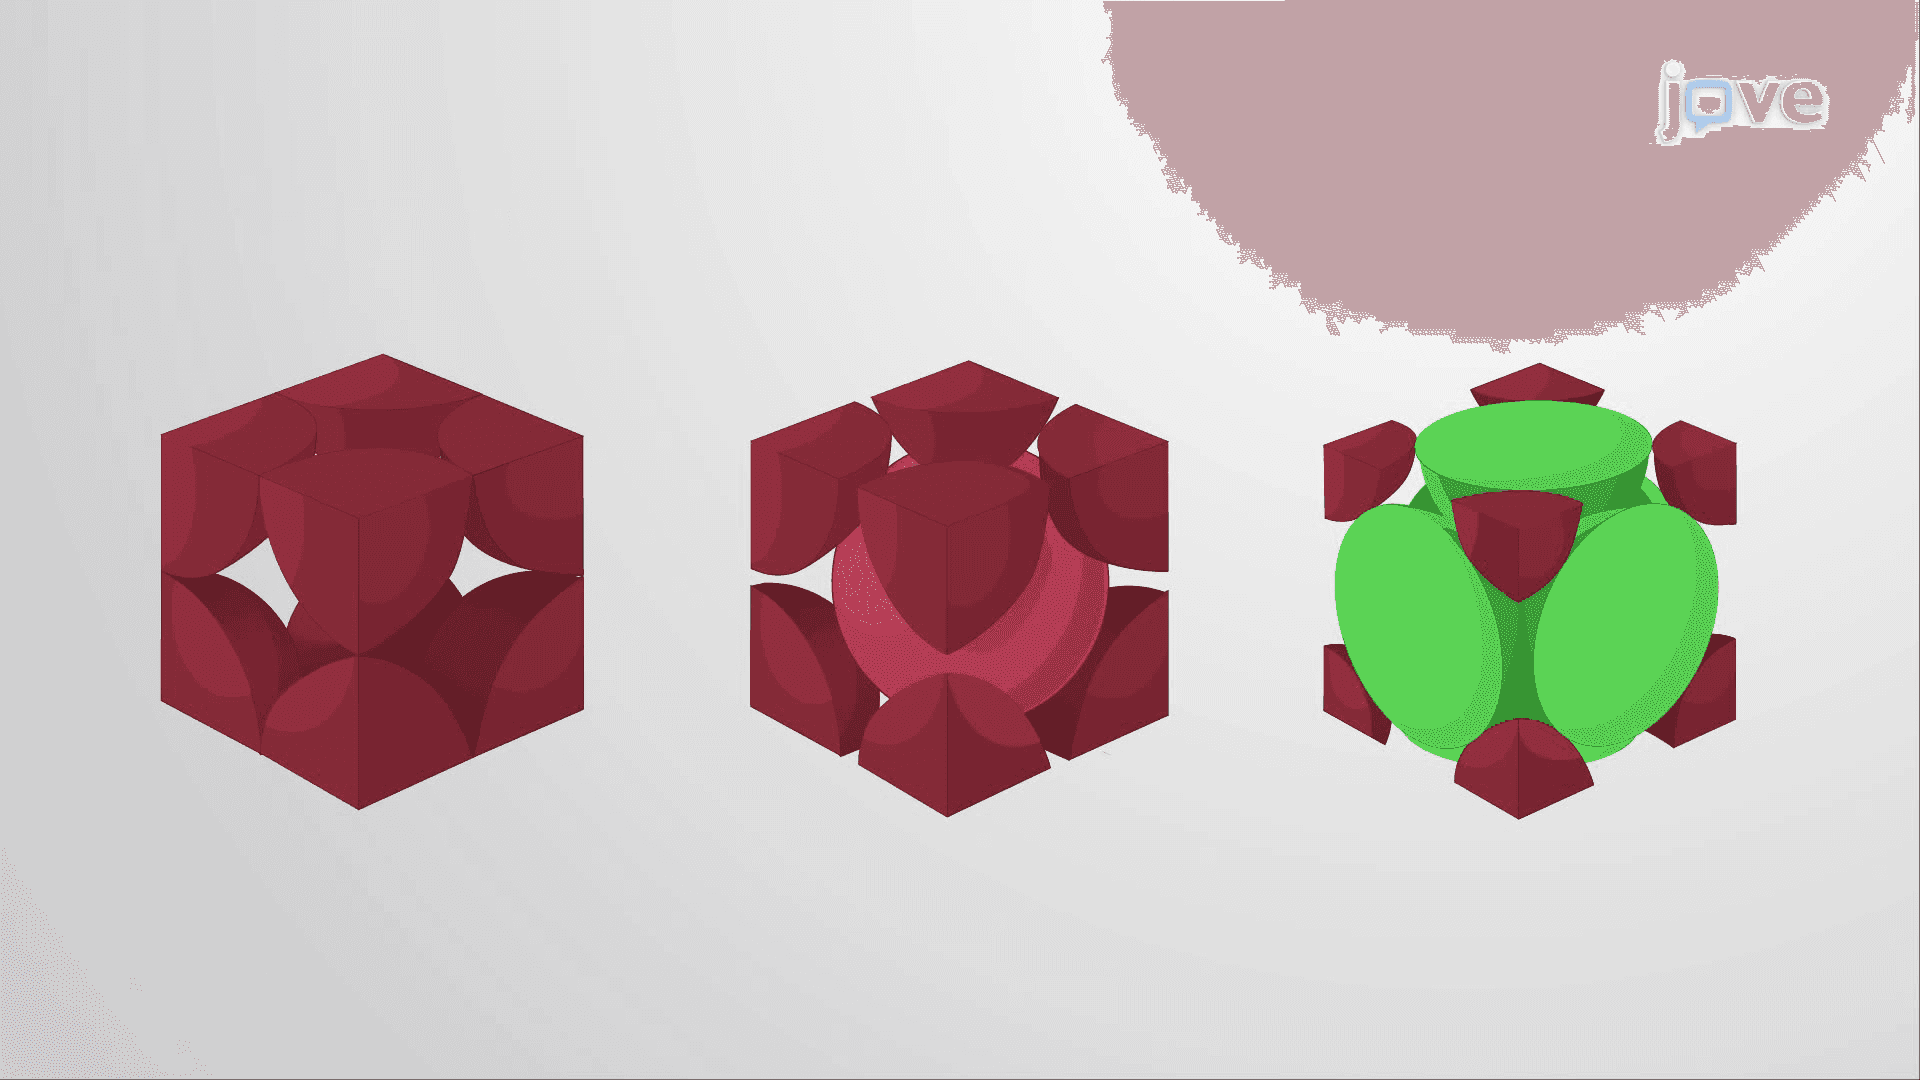
Crystal Density
 01:28
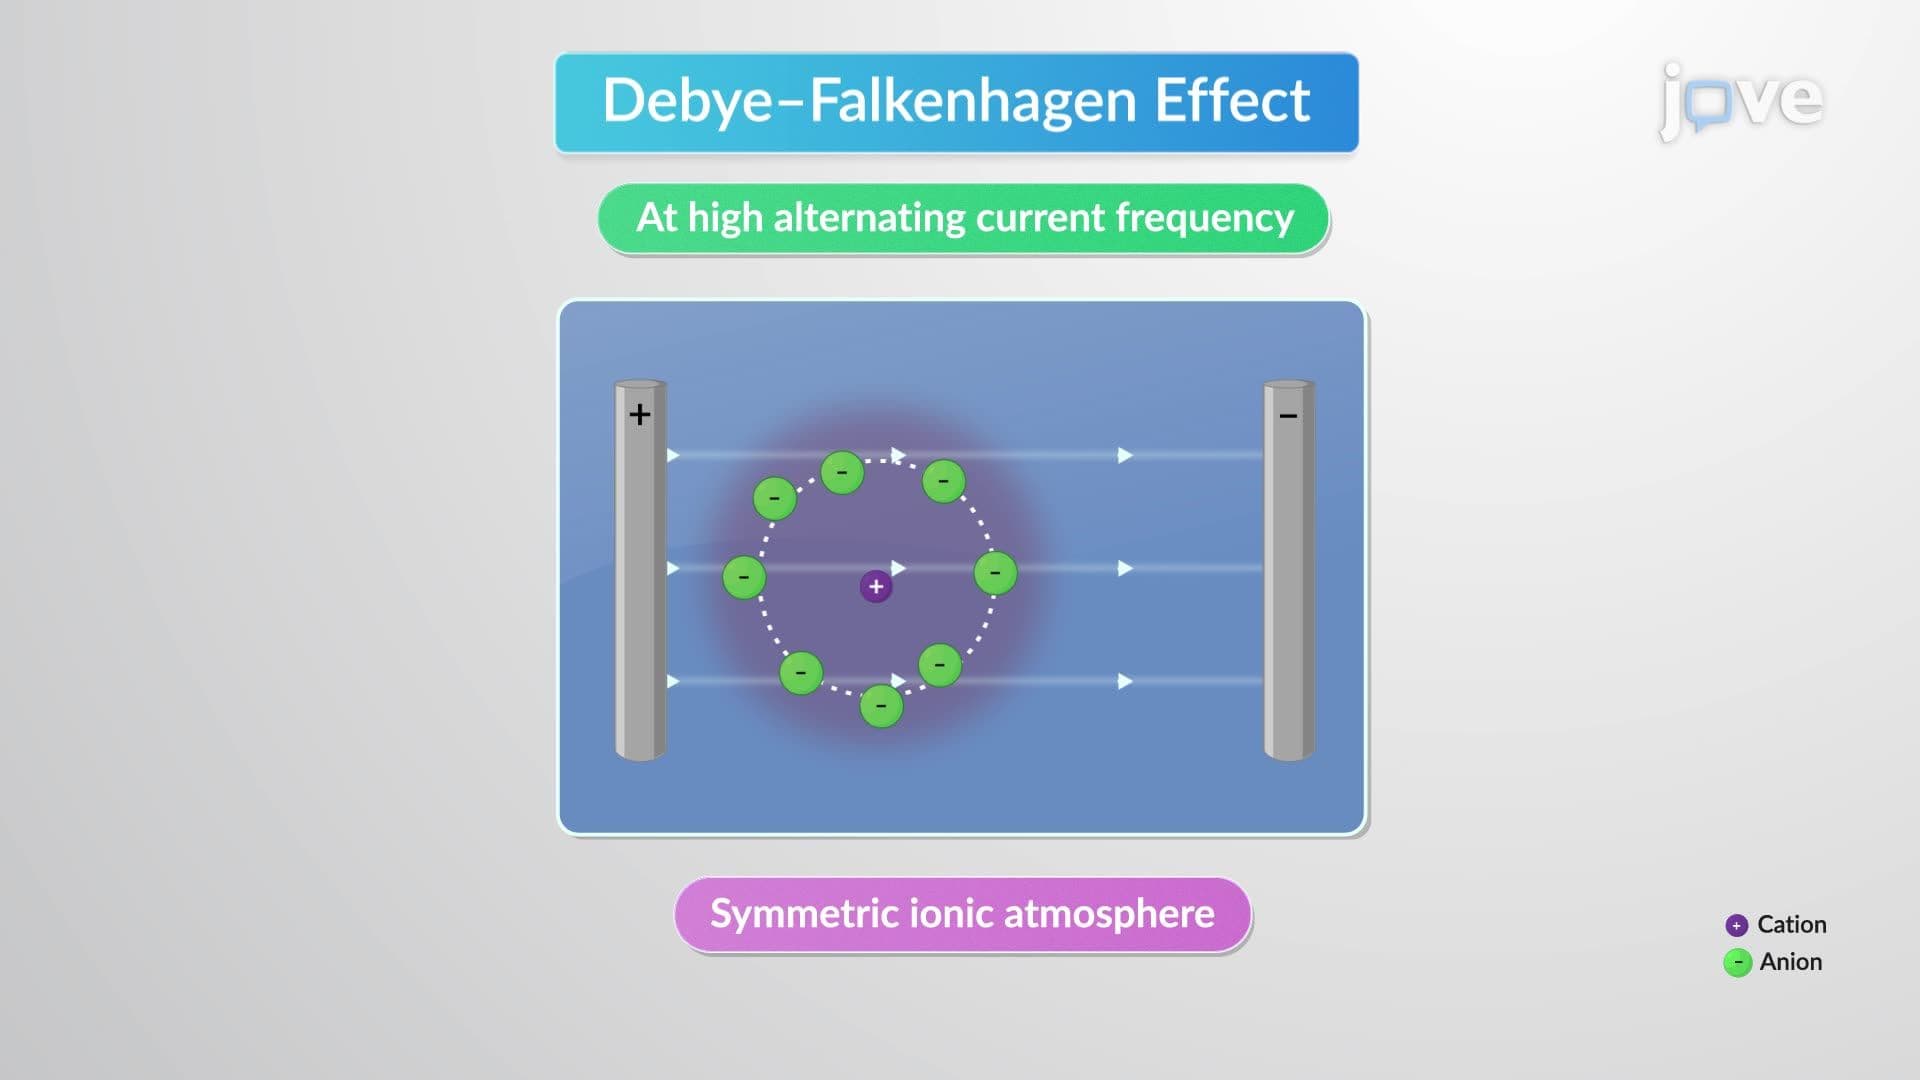
Debye–Huckel–Onsager Conductance Equation
 02:33
Lattice Centering and Coordination Number
Types of Unit Cells
Imagine taking a large number of identical...
 01:10
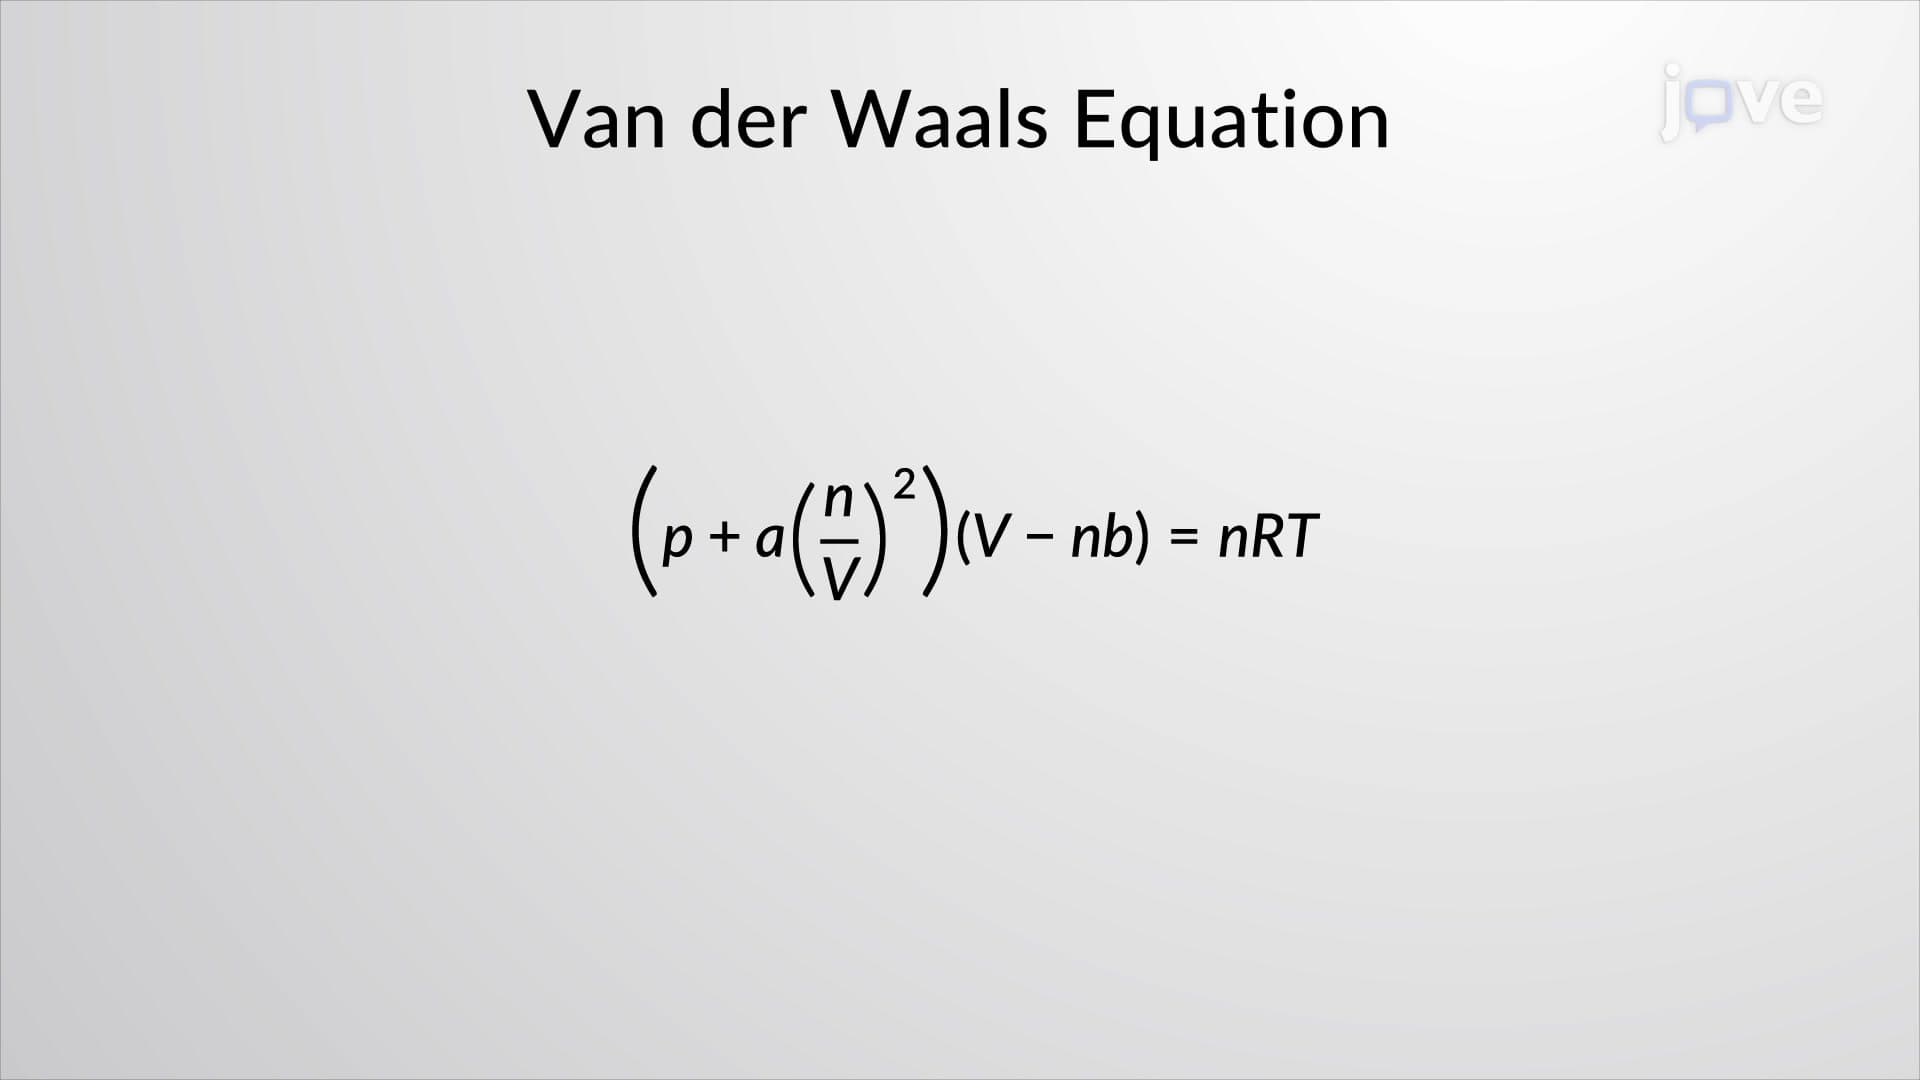
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:24
Van der Waals Interactions