Related Experiment Video
Updated: May 13, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Useful lower limits to polarization contributions to intermolecular interactions using a minimal basis of localized
R Julian Azar1, Paul Richard Horn, Eric Jon Sundstrom
1Department of Chemistry, University of California Berkeley, Berkeley, California 94720, USA. julianazar2323@berkeley.edu

Abstract:
The problem of describing the energy-lowering associated with polarization of interacting molecules is considered in the overlapping regime for self-consistent field wavefunctions. The existing approach of solving for absolutely localized molecular orbital (ALMO) coefficients that are block-diagonal in the fragments is shown based on formal grounds and practical calculations to often overestimate the strength of polarization effects. A new approach using a minimal basis of polarized orthogonal local MOs (polMOs) is developed as an alternative. The polMO basis is minimal in the sense that one polarization function is provided for each unpolarized orbital that is occupied; such an approach is exact in second-order perturbation theory. Based on formal grounds and practical calculations, the polMO approach is shown to underestimate the strength of polarization effects. In contrast to the ALMO method, however, the polMO approach yields results that are very stable to improvements in the underlying AO basis expansion. Combining the ALMO and polMO approaches allows an estimate of the range of energy-lowering due to polarization. Extensive numerical calculations on the water dimer using a large range of basis sets with Hartree-Fock theory and a variety of different density functionals illustrate the key considerations. Results are also presented for the polarization-dominated Na(+)CH4 complex. Implications for energy decomposition analysis of intermolecular interactions are discussed.
Related Concept Videos
 02:40
MO Theory and Covalent Bonding
 03:51
Molecular Orbital Theory II
 01:24
Van der Waals Interactions
 02:36
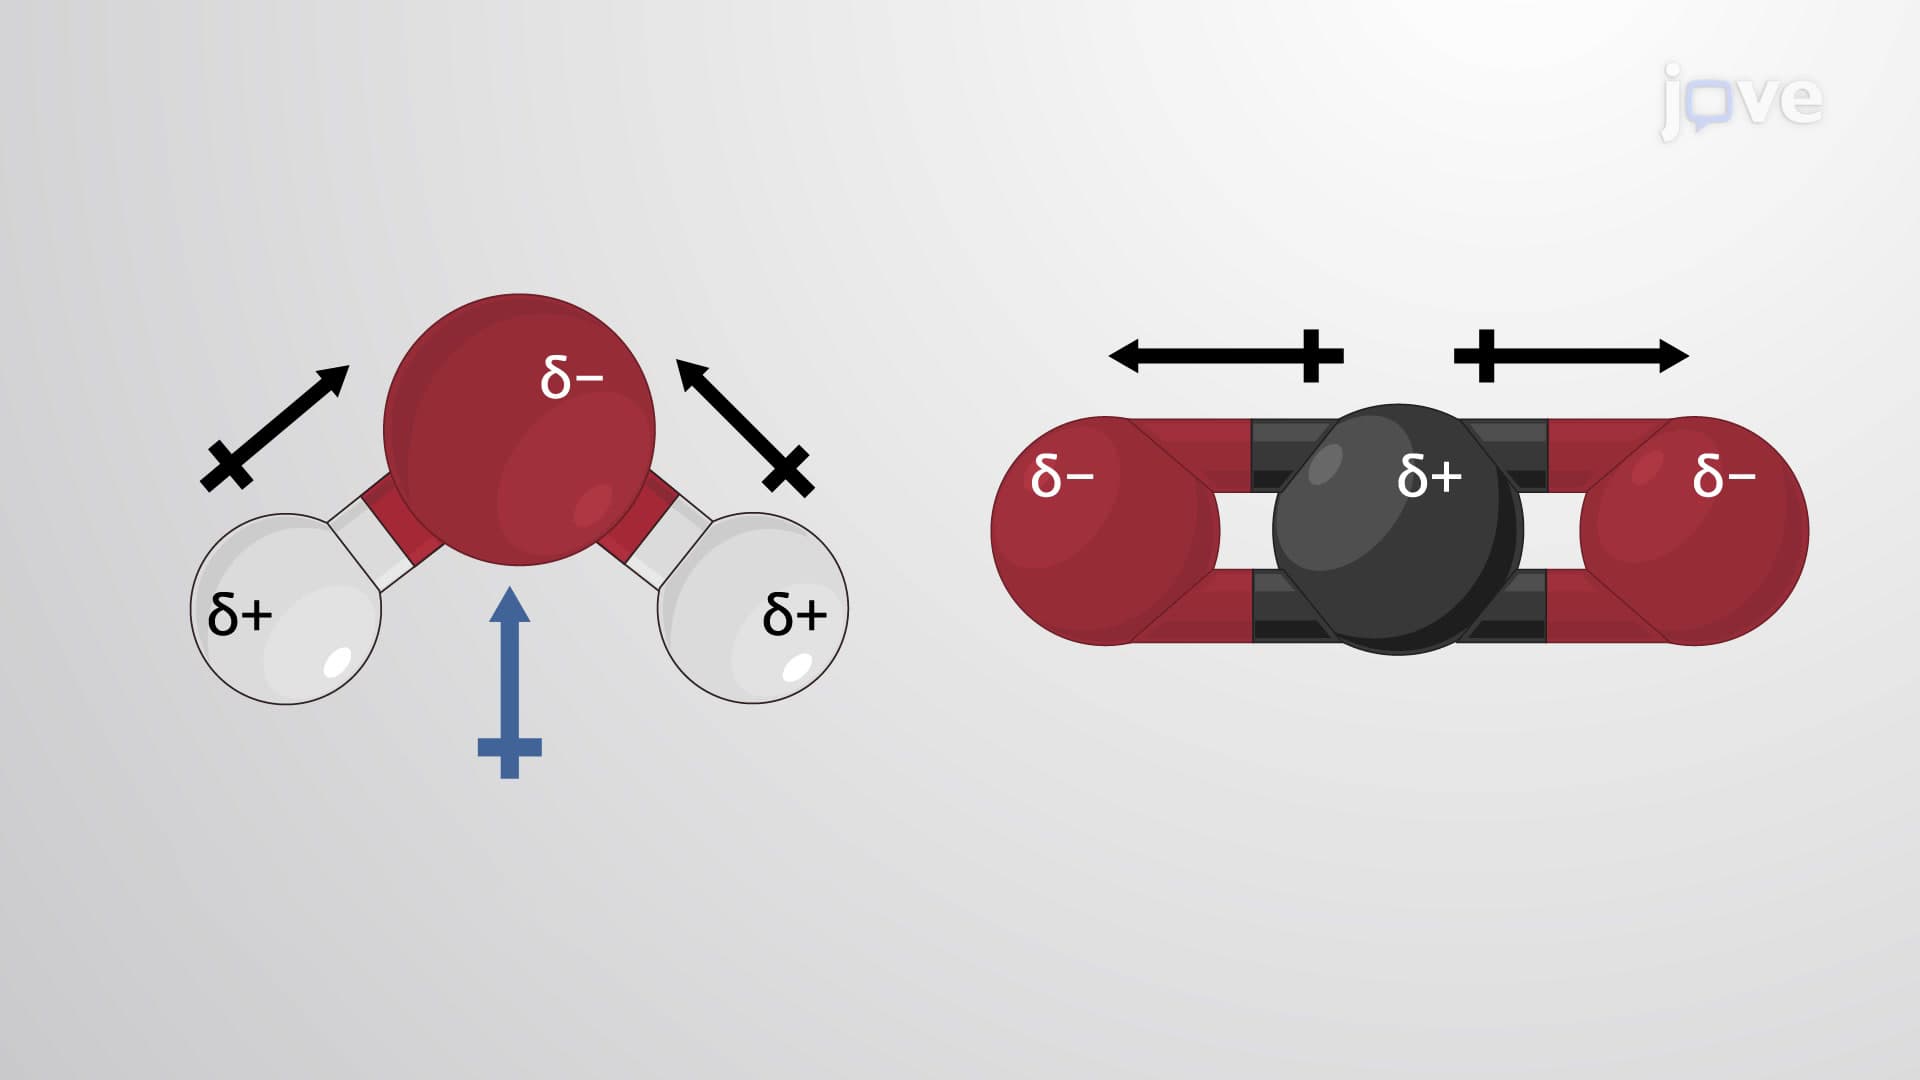
Molecular Geometry and Dipole Moments
 02:35
Molecular Orbital Theory I
 03:13
Intermolecular Forces