Related Experiment Video
Updated: May 13, 2026

Amplicon Sequencing using the Long-Read Sequencing Technologies
Published on: August 29, 2025
Use of whole genome sequencing to determine the microevolution of Mycobacterium tuberculosis during an outbreak
Midori Kato-Maeda1, Christine Ho, Ben Passarelli
1Curry International Tuberculosis Center, Division of Pulmonary and Critical Care Medicine, University of California San Francisco, San Francisco, California, USA. midori.kato-maeda@ucsf.edu

Rationale:
Current tools available to study the molecular epidemiology of tuberculosis do not provide information about the directionality and sequence of transmission for tuberculosis cases occurring over a short period of time, such as during an outbreak. Recently, whole genome sequencing has been used to study molecular epidemiology of Mycobacterium tuberculosis over short time periods.
Objective:
To describe the microevolution of M. tuberculosis during an outbreak caused by one drug-susceptible strain. METHOD AND MEASUREMENTS: We included 9 patients with tuberculosis diagnosed during a period of 22 months, from a population-based study of the molecular epidemiology in San Francisco. Whole genome sequencing was performed using Illumina's sequencing by synthesis technology. A custom program written in Python was used to determine single nucleotide polymorphisms which were confirmed by PCR product Sanger sequencing.
Main Results:
We obtained an average of 95.7% (94.1-96.9%) coverage for each isolate and an average fold read depth of 73 (1 to 250). We found 7 single nucleotide polymorphisms among the 9 isolates. The single nucleotide polymorphisms data confirmed all except one known epidemiological link. The outbreak strain resulted in 5 bacterial variants originating from the index case A1 with 0-2 mutations per transmission event that resulted in a secondary case.
Conclusions:
Whole genome sequencing analysis from a recent outbreak of tuberculosis enabled us to identify microevolutionary events observable during transmission, to determine 0-2 single nucleotide polymorphisms per transmission event that resulted in a secondary case, and to identify new epidemiologic links in the chain of transmission.
More Related Videos
 06:46
Whole-Genome Deoxyribonucleic Acid Extraction from Mycobacterium Species via the Cetyltrimethylammonium Bromide Technique
Published on: December 12, 2025
 15:28
A Microscopic Phenotypic Assay for the Quantification of Intracellular Mycobacteria Adapted for High-throughput/High-content Screening
Published on: January 17, 2014
Related Concept Videos
 01:29
Modern Molecular Taxonomy
 01:08
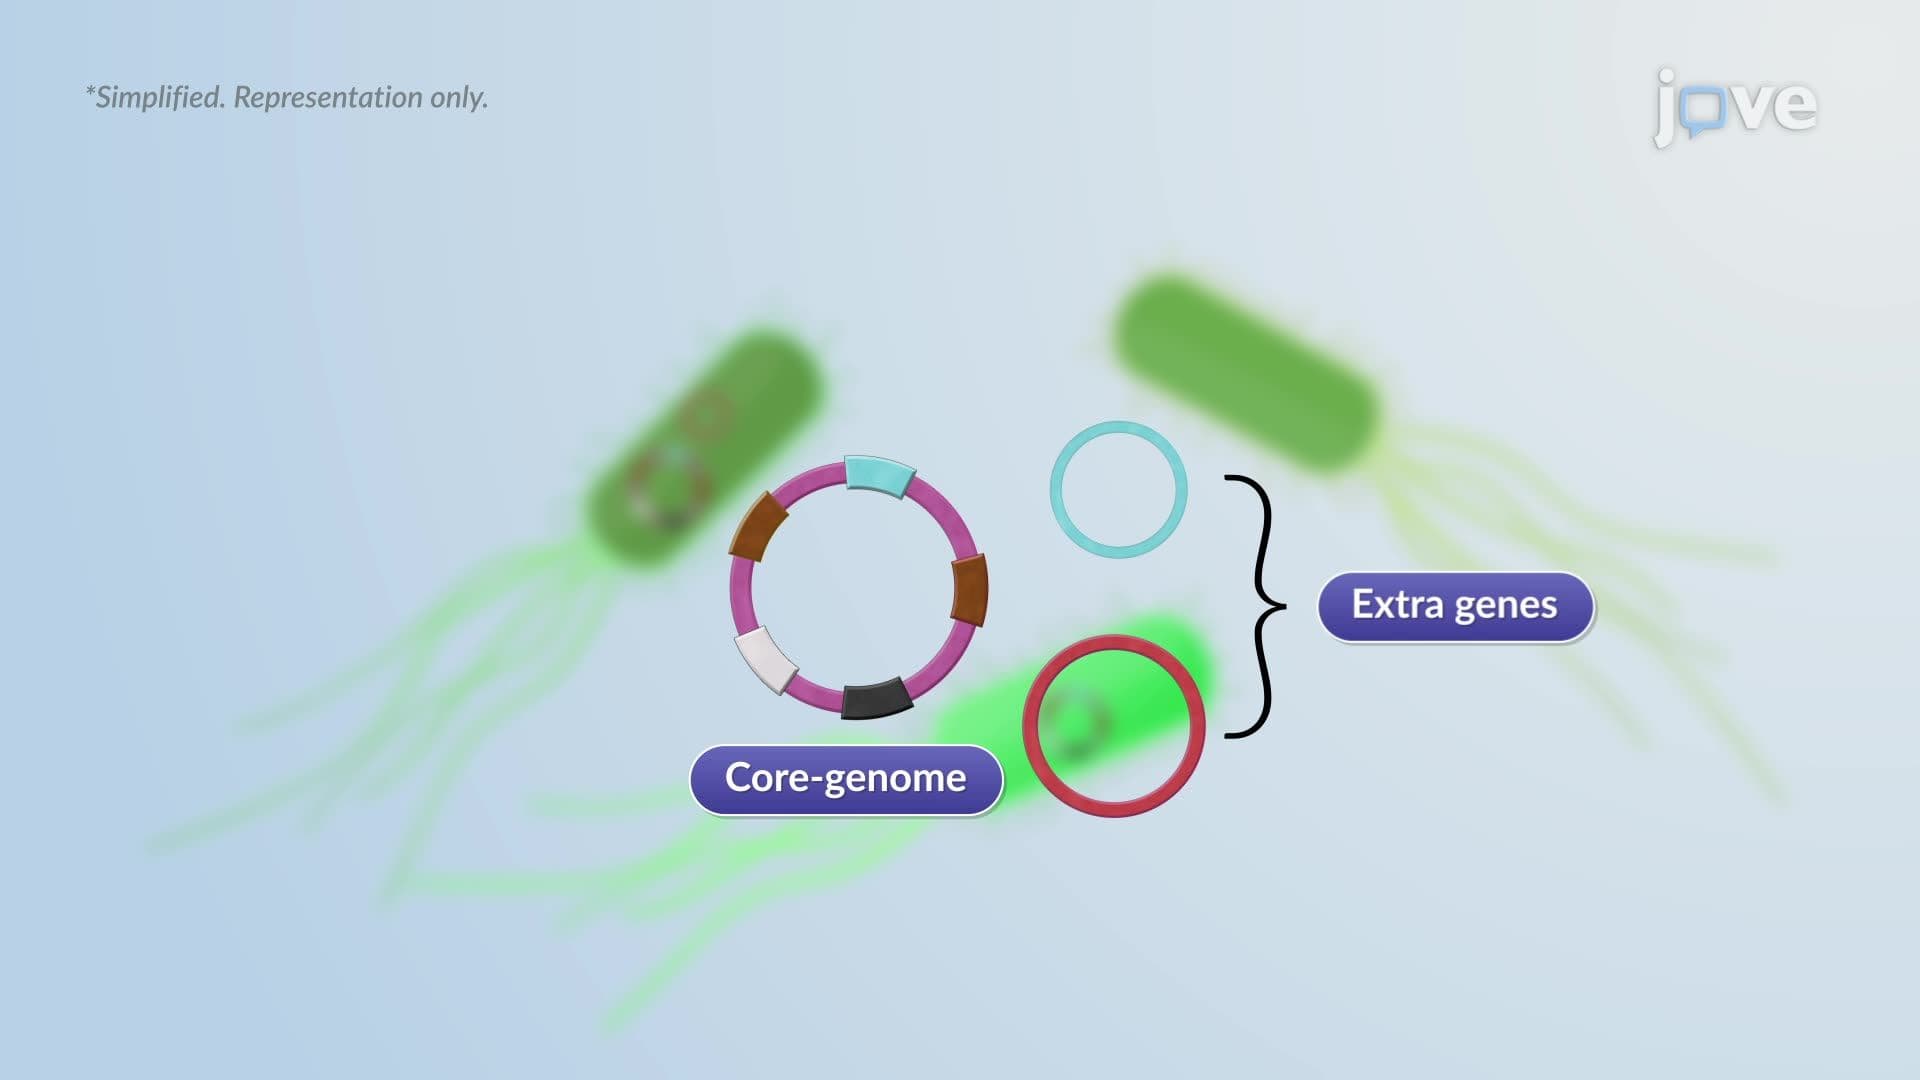
Evolution of Microbial Genome
 02:54
Evolutionary Relationships through Genome Comparisons
 01:23
Investigation of Disease Outbreaks
 02:02
Genomics
 01:20
Applications of Molecular Taxonomy