Related Experiment Video
Updated: May 13, 2026
The Cell-based L-Glutathione Protection Assays to Study Endocytosis and Recycling of Plasma Membrane Proteins
Published on: December 13, 2013
Cystic fibrosis transmembrane conductance regulator recruitment to phagosomes in neutrophils
Yun Zhou1, Kejing Song, Richard G Painter
1Department of Microbiology, Louisiana State University Health Sciences Center, New Orleans, LA 70112, USA.

Abstract:
Optimal microbicidal activity of human polymorphonuclear leukocytes (PMN) relies on the generation of toxic agents such as hypochlorous acid (HOCl) in phagosomes. HOCl formation requires H2O2 produced by the NADPH oxidase, myeloperoxidase derived from azurophilic granules, and chloride ion. Chloride transport from cytoplasm into phagosomes requires chloride channels which include cystic fibrosis transmembrane conductance regulator (CFTR), a cAMP-activated chloride channel. However, the phagosomal targeting of CFTR in PMN has not been defined. Using human peripheral blood PMN, we determined that 95-99% of lysosomal-associated membrane protein 1 (LAMP-1)-positive mature phagosomes were CFTR positive, as judged by immunostaining and flow cytometric analysis. To establish a model cell system to evaluate CFTR phagosomal recruitment, we stably expressed enhanced green fluorescent protein (EGFP) alone, EGFP-wt-CFTR and EGFP-DF508-CFTR fusion proteins in promyelocytic PLB-985 cells, respectively. After differentiation into neutrophil-like cells, CFTR presentation to phagosomes was examined. EGFP-wt-CFTR was observed to associate with phagosomes and colocalize with LAMP-1. Flow cytometric analysis of the isolated phagosomes indicated that such a phagosomal targeting was determined by the CFTR portion of the fusion protein. In contrast, significantly less EGFP-DF508-CFTR was found in phagosomes, indicating a defective targeting of the molecule to the organelle. Importantly, the CFTR corrector compound VRT-325 facilitated the recruitment of DF508-CFTR to phagosomes. These data demonstrate the possibility of pharmacologic correction of impaired recruitment of mutant CFTR, thereby providing a potential means to augment chloride supply to the phagosomes of PMN in patients with cystic fibrosis to enhance their microbicidal function.
Insights
Cystic fibrosis transmembrane conductance regulator (CFTR) is crucial for human polymorphonuclear leukocytes (PMN) microbicidal activity. This study shows CFTR targets phagosomes and its defective recruitment in cystic fibrosis can be pharmacologically corrected.
Area of Science:
- Immunology
- Cell Biology
- Medical Science
Background:
- Optimal microbicidal activity in human polymorphonuclear leukocytes (PMN) depends on toxic agents like hypochlorous acid (HOCl) generated within phagosomes.
- HOCl formation requires hydrogen peroxide (H2O2), NADPH oxidase, myeloperoxidase, and chloride ions, with chloride transport facilitated by chloride channels such as cystic fibrosis transmembrane conductance regulator (CFTR).
- The precise phagosomal targeting of CFTR in PMN has remained undefined.
Purpose of the Study:
- To investigate the phagosomal targeting of CFTR in human peripheral blood PMN.
- To establish a model cell system for evaluating CFTR phagosomal recruitment.
- To assess the potential for pharmacologic correction of impaired CFTR recruitment.
Main Methods:
- Immunostaining and flow cytometry were used to analyze CFTR localization in mature phagosomes of human PMN.
- A stable cell line expressing enhanced green fluorescent protein (EGFP)-tagged wild-type (wt) and mutant (DF508) CFTR was developed in PLB-985 cells.
- Phagosomal association and colocalization with lysosomal-associated membrane protein 1 (LAMP-1) were examined in differentiated neutrophil-like cells.
Main Results:
- Over 95% of mature phagosomes in human PMN were found to be CFTR positive.
- EGFP-wt-CFTR fusion proteins successfully targeted phagosomes and colocalized with LAMP-1 in the model system.
- Significantly reduced phagosomal localization of EGFP-DF508-CFTR indicated defective targeting, which was partially restored by the CFTR corrector VRT-325.
Conclusions:
- CFTR is demonstrably recruited to phagosomes in PMN, a process essential for microbicidal function.
- The DF508 mutation impairs CFTR phagosomal targeting, potentially contributing to reduced microbicidal capacity in cystic fibrosis.
- Pharmacologic agents like VRT-325 show promise in correcting defective CFTR recruitment, offering a potential therapeutic strategy to enhance PMN microbicidal function in cystic fibrosis patients.
More Related Videos
 14:56
Expression and Purification of the Cystic Fibrosis Transmembrane Conductance Regulator Protein in Saccharomyces cerevisiae
Published on: March 10, 2012
 10:07
"Phagosome Closure Assay" to Visualize Phagosome Formation in Three Dimensions Using Total Internal Reflection Fluorescent Microscopy (TIRFM)
Published on: August 26, 2016
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:25
Immune Surveillance by NK Cells and Phagocytes
Natural Killer Cells: The Fast Responders
NK cells are large granular lymphocytes found in the blood and lymphatic system. These...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...
 01:25
Rab Cascades
 01:12
Clathrin Coated Vesicles
 00:41
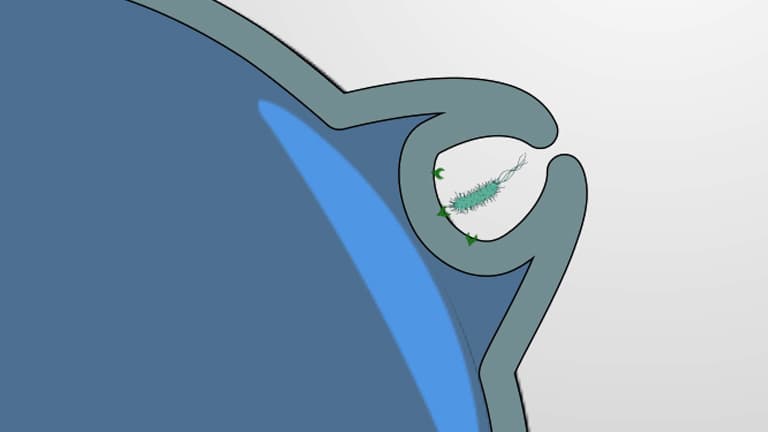
Phagocytosis