Related Experiment Video
Updated: May 13, 2026

Drug-induced Sensitization of Adenylyl Cyclase: Assay Streamlining and Miniaturization for Small Molecule and siRNA Screening Applications
Published on: January 27, 2014
Activation and Desensitization of Rat A3-Adenosine Receptors by Selective Adenosine Derivatives and
Kyung-Sun Park1, Carsten Hoffmann, Hea Ok Kim
1Laboratory of Bioorganic Chemistry, National Institute of Diabetes, and Digestive and Kidney Diseases, National Institutes of Health, Bethesda, Maryland, USA.

Abstract:
Strategy, Management and Health PolicyVenture Capital Enabling TechnologyPreclinical ResearchPreclinical Development Toxicology, Formulation Drug Delivery, PharmacokineticsClinical Development Phases I-III Regulatory, Quality, ManufacturingPostmarketing Phase IV Xanthine and adenosine derivatives, known to bind to recombinant rat A3 adenosine receptors stably expressed in Chinese hamster ovary cells, were characterized in a functional assay consisting of activation of A3 receptor-stimulated binding of [35S]GTPγS in rat RBL-2H3 cell membranes. 1,3-Dibutylxanthine-7-riboside-5'-N-methylcarboxamide (DBXRM, 7b), previously shown to inhibit adenylyl cyclase via rat A3 receptors with full efficacy, appeared to be a partial agonist at the rat A3 receptor of RBL-2H3 cells. Full agonists, such as Cl-IB-MECA or I-AB-MECA, were more potent and effective than the partial agonist DBXRM in causing desensitization of rat A3 receptors, as indicated by loss of [35S]GTPγS binding. At A1 receptors, antagonism of agonist-elicited inhibition of rat adipocyte adenylyl cyclase was observed for several xanthine-7-riboside derivatives that had been shown to be full agonists at rat A3 receptors. A new xanthine riboside (3'-deoxyDBXRM, 7c) was synthesized and found to be a partial agonist at rat A3 receptors and an antagonist at rat A1 receptors. Thus, it is possible for the same compound to stimulate one adenosine receptor subtype (A3) and block another subtype (A1) within the same species.
Related Concept Videos
 01:31
Adrenergic Receptors: ɑ Subtype
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase C—inositol-1,4,5-trisphosphate...
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:30
Adrenergic Agonists: Direct-Acting Agents
These agents can be classified...
 01:22
Adrenergic Antagonists: Pharmacological Actions of ɑ-Receptor Blockers
α1-blockers: These drugs inhibit α1-adrenoceptors on smooth muscle cells, resulting in vasodilation. This vasodilation lowers blood pressure, making α1-blockers valuable in treating hypertension. Additionally, α1-blockers effectively address urinary obstruction...
 01:17
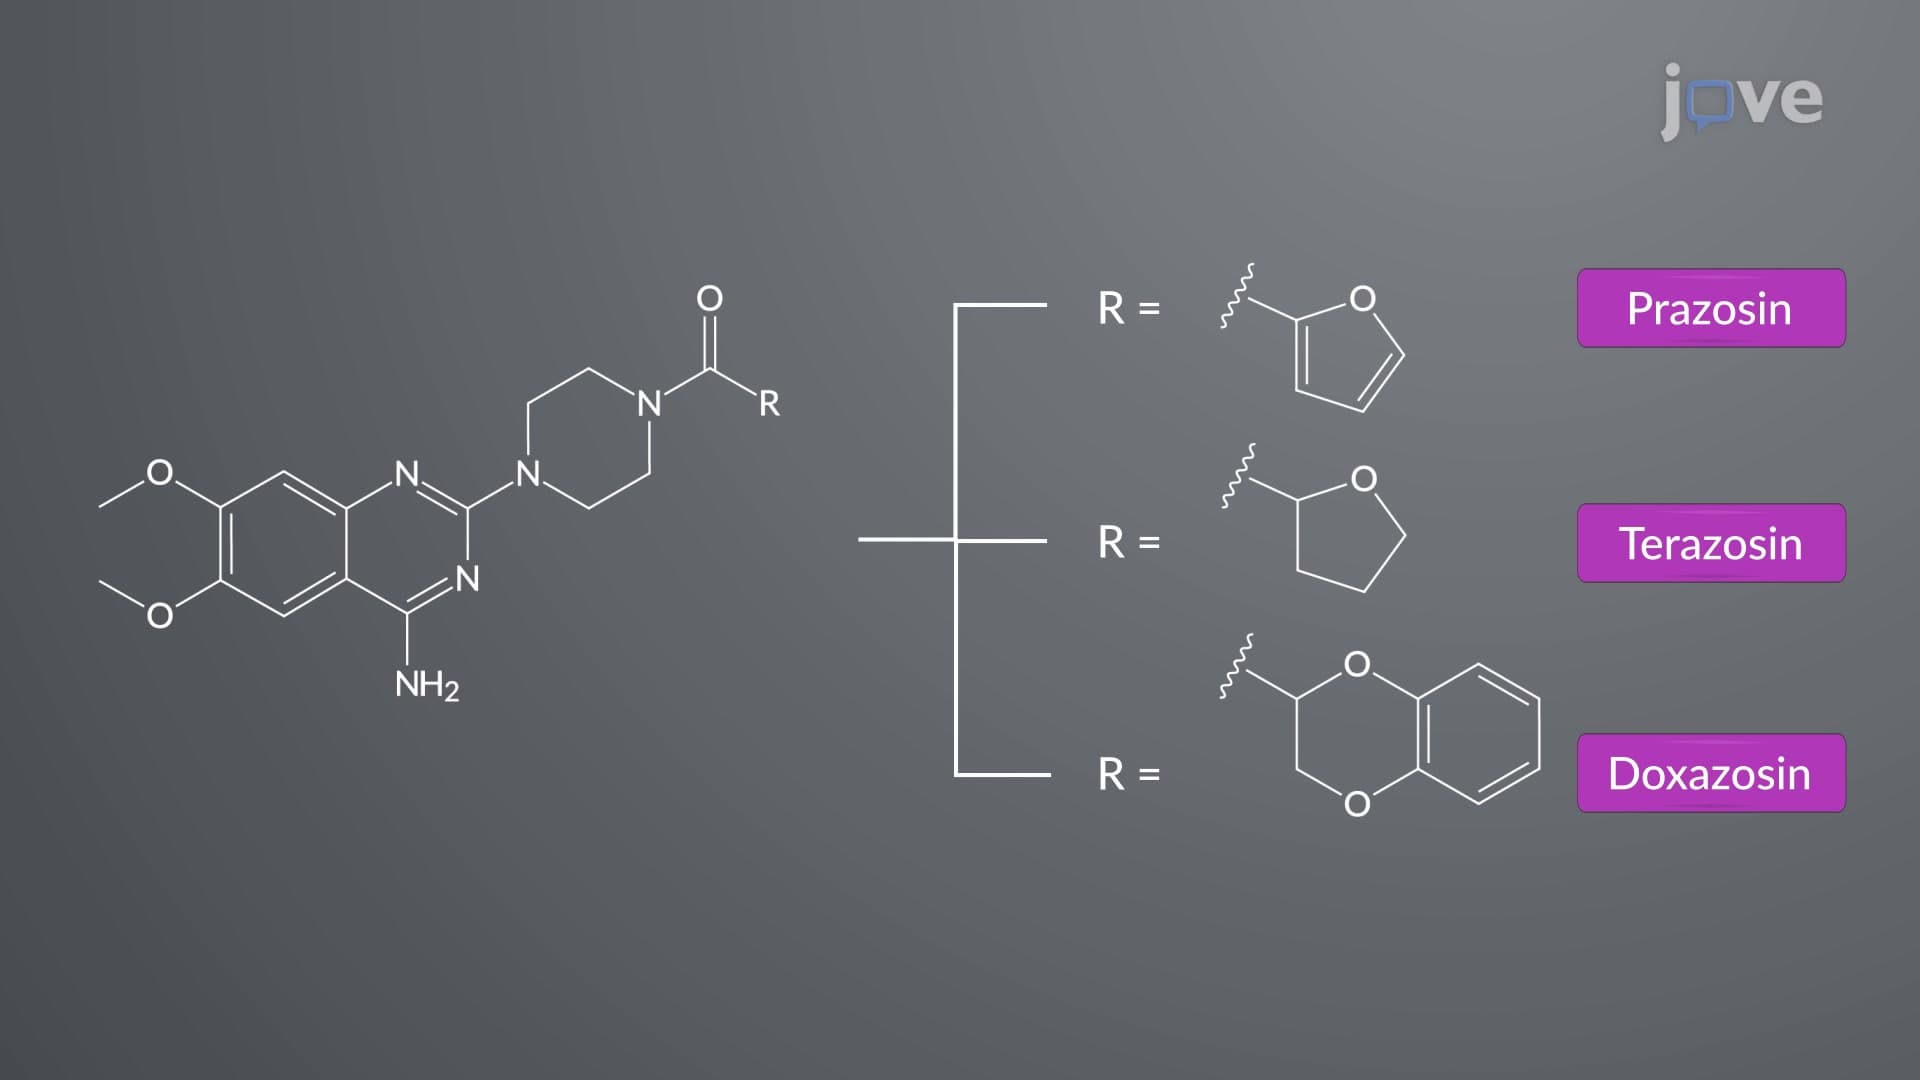
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline moieties. Phenoxybenzamine, with a haloalkylamine...
 01:25
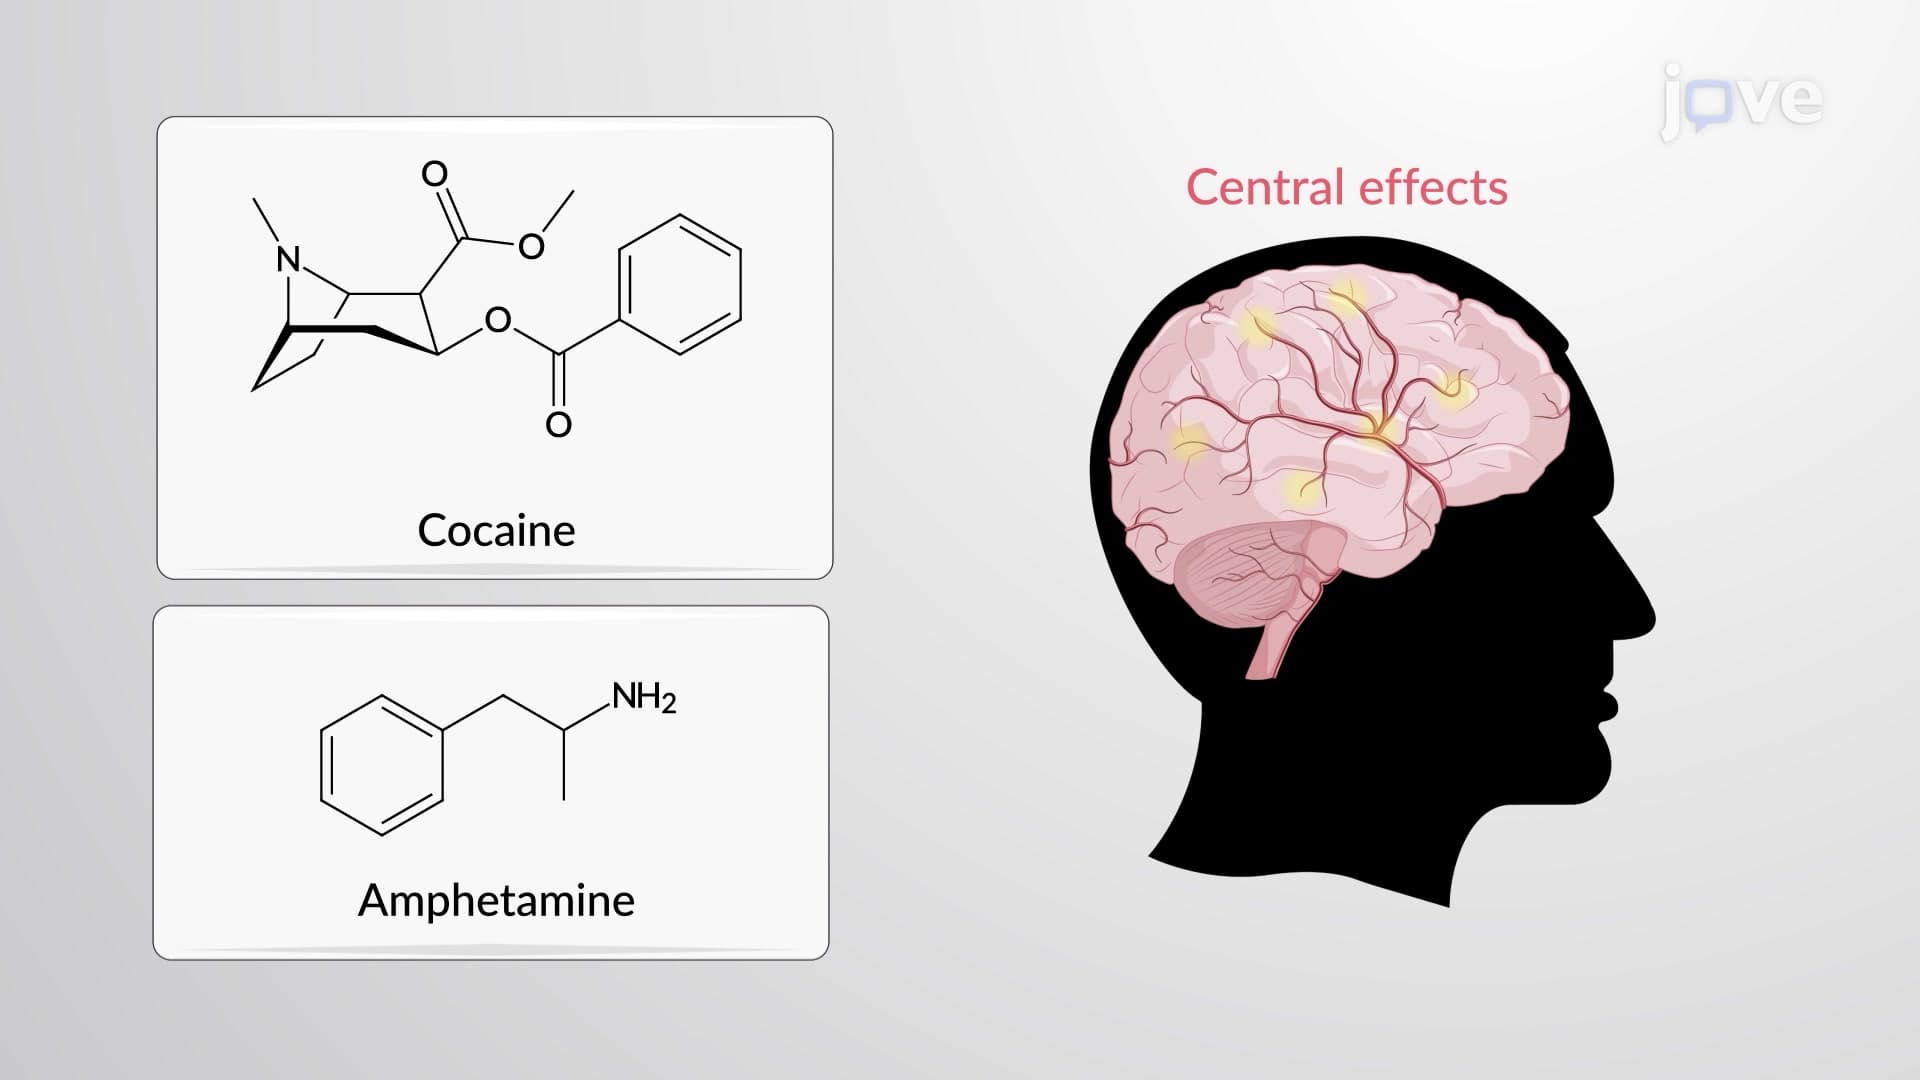
Adrenergic Agonists: Indirect-Acting Agents
One mechanism involves depleting stored catecholamines by displacing them from synaptic vesicles. These agents, known as "displacers," are transported into vesicles at the expense of noradrenaline. Examples include amphetamine and tyramine, which lack a catechol moiety, resulting in prolonged action, improved oral bioavailability, and...