Related Experiment Video
Updated: May 12, 2026

Rapid Screening of HIV Reverse Transcriptase and Integrase Inhibitors
Published on: April 9, 2014
Telaprevir: clinical pharmacokinetics, pharmacodynamics, and drug-drug interactions
Tony K L Kiang1, Kyle J Wilby, Mary H H Ensom
1Faculty of Pharmaceutical Sciences, The University of British Columbia, Vancouver, BC, Canada.

Abstract:
This article provides an unbiased review of the pharmacokinetic, pharmacodynamic, and drug-drug interaction data of telaprevir, an NS3/4A protease inhibitor. Telaprevir is well absorbed with fatty food, moderately protein bound (59-76 %) with a large volume of distribution (~252 L), primarily metabolized by cytochrome P450 (CYP) 3A4 and P-glycoprotein, and is largely excreted into feces. Pharmacokinetic and pharmacodynamic parameters are well described in healthy subjects and individuals infected with hepatitis C virus (HCV), although only limited data are available in specific patient subpopulations. Telaprevir is recommended to be given at 750 mg by mouth every 8 h for 12 weeks, in combination with peginterferon and ribavirin (the standard care). The addition of telaprevir to the standard care regimen results in increased sustained virological response in treatment-naïve patients (30 %) and treatment-experienced patients (up to 50 %), and works synergistically to lower viral resistance. Telaprevir is a substrate and/or inhibitor of CYP3A4 and P-glycoprotein, and drug-drug interaction studies in humans have focused on these pathways. Based on our analysis, a few reported drug-drug interactions may be classified as clinically significant, but more experiments under dosing conditions that resemble those given in the clinic are needed to understand the relevance of some of the reported interactions. Future studies should focus on the pharmacokinetics/pharmacodynamics of telaprevir in special populations or patients with concomitant conditions that will likely co-exist with HCV infection, with an emphasis on establishing pharmacokinetic-pharmacodynamic relationships. In vitro characterization of other phase 1-3 metabolic pathways could assist in elucidating the mechanisms of the drug-drug interactions observed in humans.
More Related Videos
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
 12:03
Evaluation of the Efficacy And Toxicity of RNAs Targeting HIV-1 Production for Use in Gene or Drug Therapy
Published on: September 5, 2016
Related Concept Videos
 01:26
Pharmacokinetics: Drug–Food and Drug–Viral Interactions
 01:25
Pharmacokinetics: Drug–Drug Interactions
 01:30
Drug toxicity: Drug–Drug Interaction
 01:08
Effect of Hepatic Disease on Pharmacokinetics: Dose Adjustments Due to Hepatic Impairment
 01:14
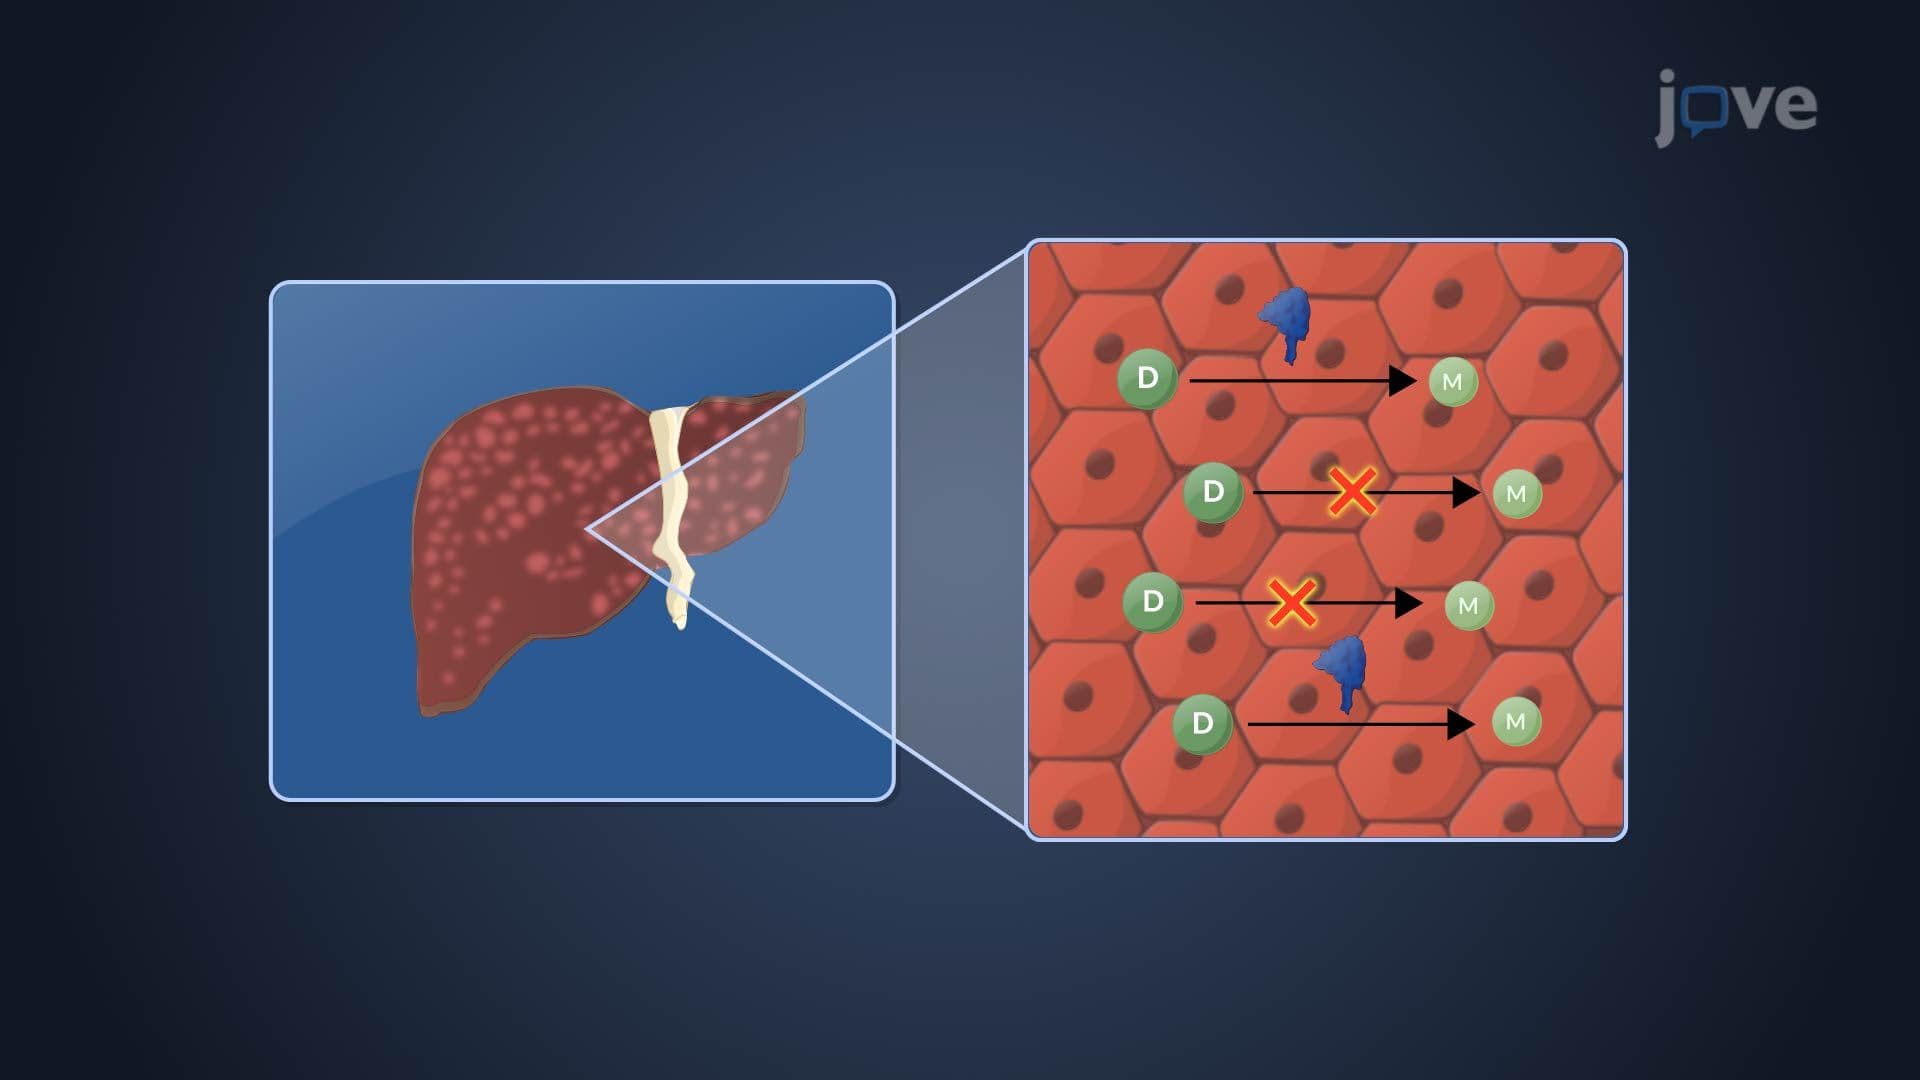
Effect of Hepatic Disease on Pharmacokinetics: Active Drug, Metabolite and Fraction of Metabolized Drug
 01:22
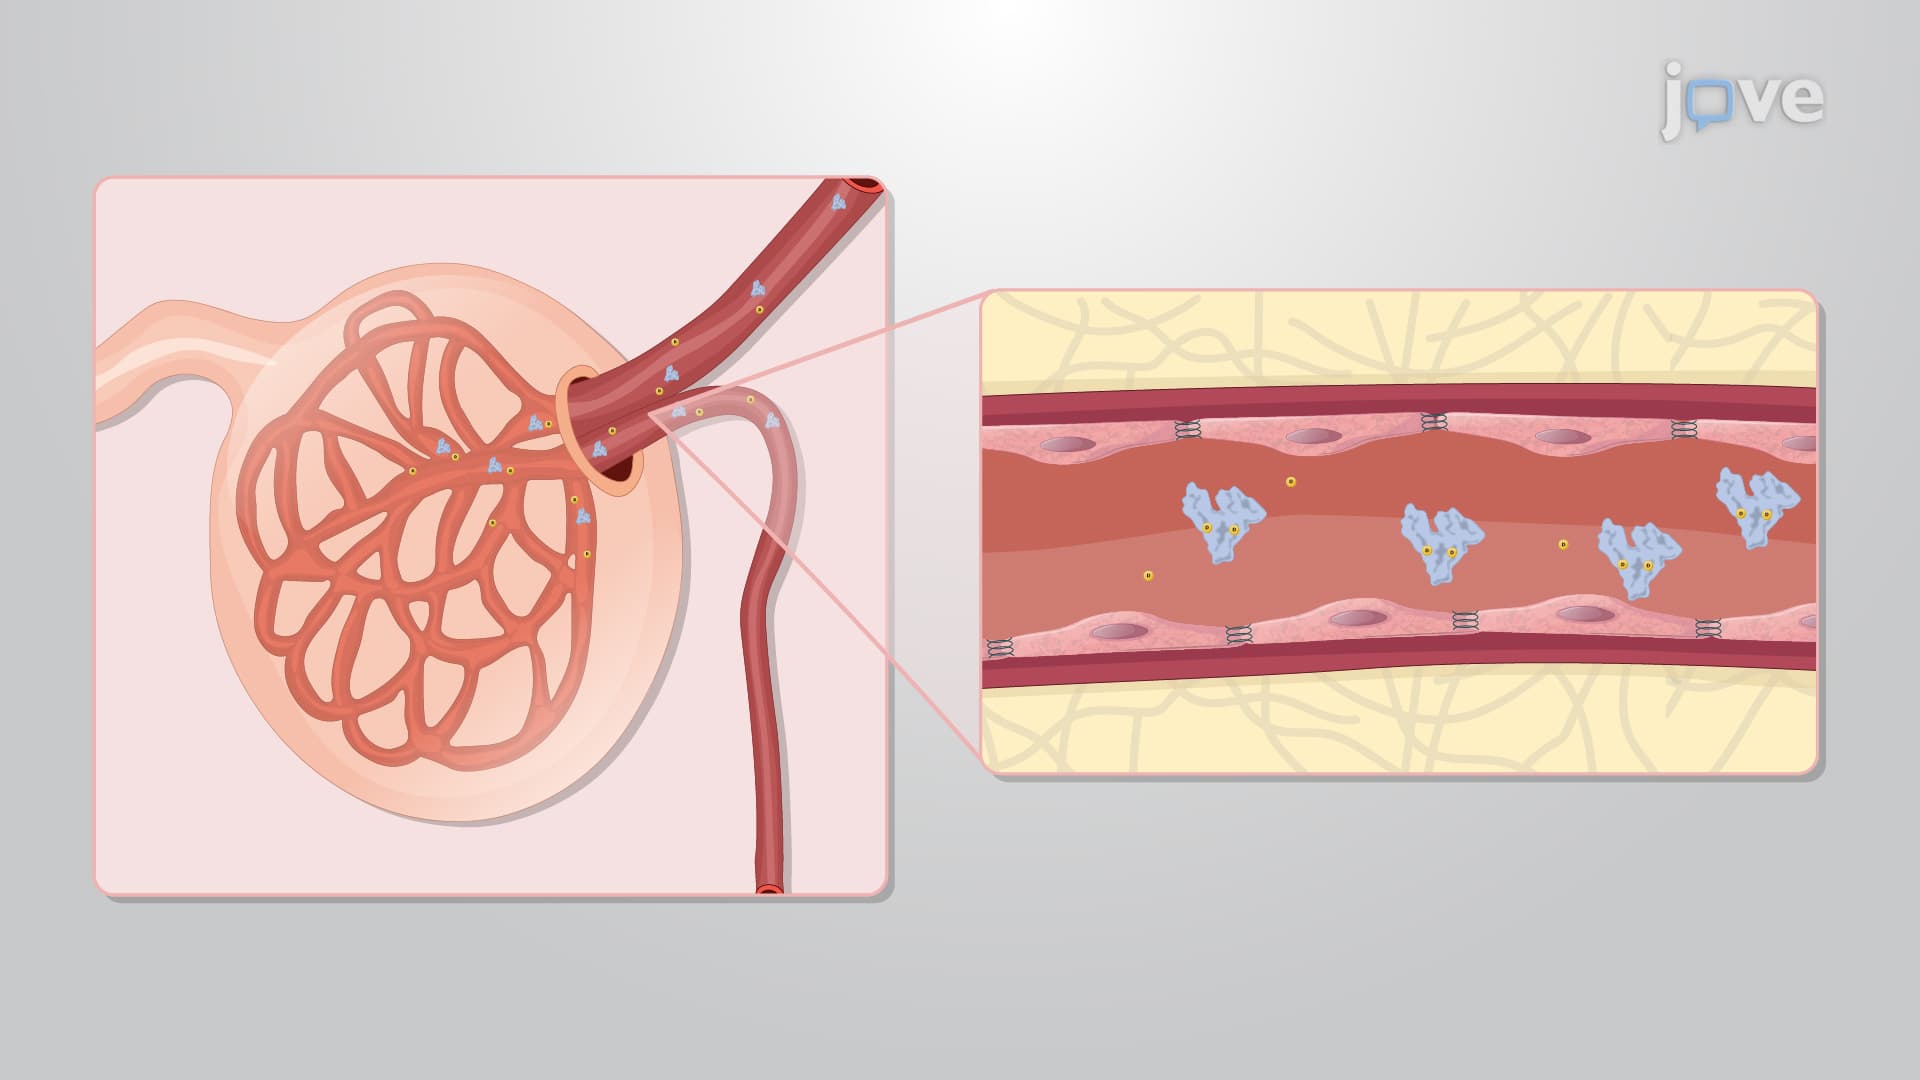
Nonlinear Pharmacokinetics: Bioavailability and Protein-Drug Binding
To quantify the extent of bioavailability, pharmacologists often use a parameter called .