Performance of density functional theory in computing nonresonant vibrational (hyper)polarizabilities
Related Experiment Video
Updated: May 11, 2026
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Ireneusz W Bulik1, Robert Zaleśny, Wojciech Bartkowiak
1Department of Chemistry, Rice University, 6100 Main Street, Houston, Texas 77005-1892, USA.

Abstract:
A set of exchange-correlation functionals, including BLYP, PBE0, B3LYP, BHandHLYP, CAM-B3LYP, LC-BLYP, and HSE, has been used to determine static and dynamic nonresonant (nuclear relaxation) vibrational (hyper)polarizabilities for a series of all-trans polymethineimine (PMI) oligomers containing up to eight monomer units. These functionals are assessed against reference values obtained using the Møller-Plesset second-order perturbation theory (MP2) and CCSD methods. For the smallest oligomer, CCSD(T) calculations confirm the choice of MP2 and CCSD as appropriate for assessing the density functionals. By and large, CAM-B3LYP is the most successful, because it is best for the nuclear relaxation contribution to the static linear polarizability, intensity-dependent refractive index second hyperpolarizability, static second hyperpolarizability, and is close to the best for the electro-optical Pockels effect first hyperpolarizability. However, none of the functionals perform satisfactorily for all the vibrational (hyper)polarizabilities studied. In fact, in the case of electric field-induced second harmonic generation all of them, as well as the Hartree-Fock approximation, yield the wrong sign. We have also found that the Pople 6-31+G(d) basis set is unreliable for computing nuclear relaxation (hyper)polarizabilities of PMI oligomers due to the spurious prediction of a nonplanar equilibrium geometry.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to the...
 01:28
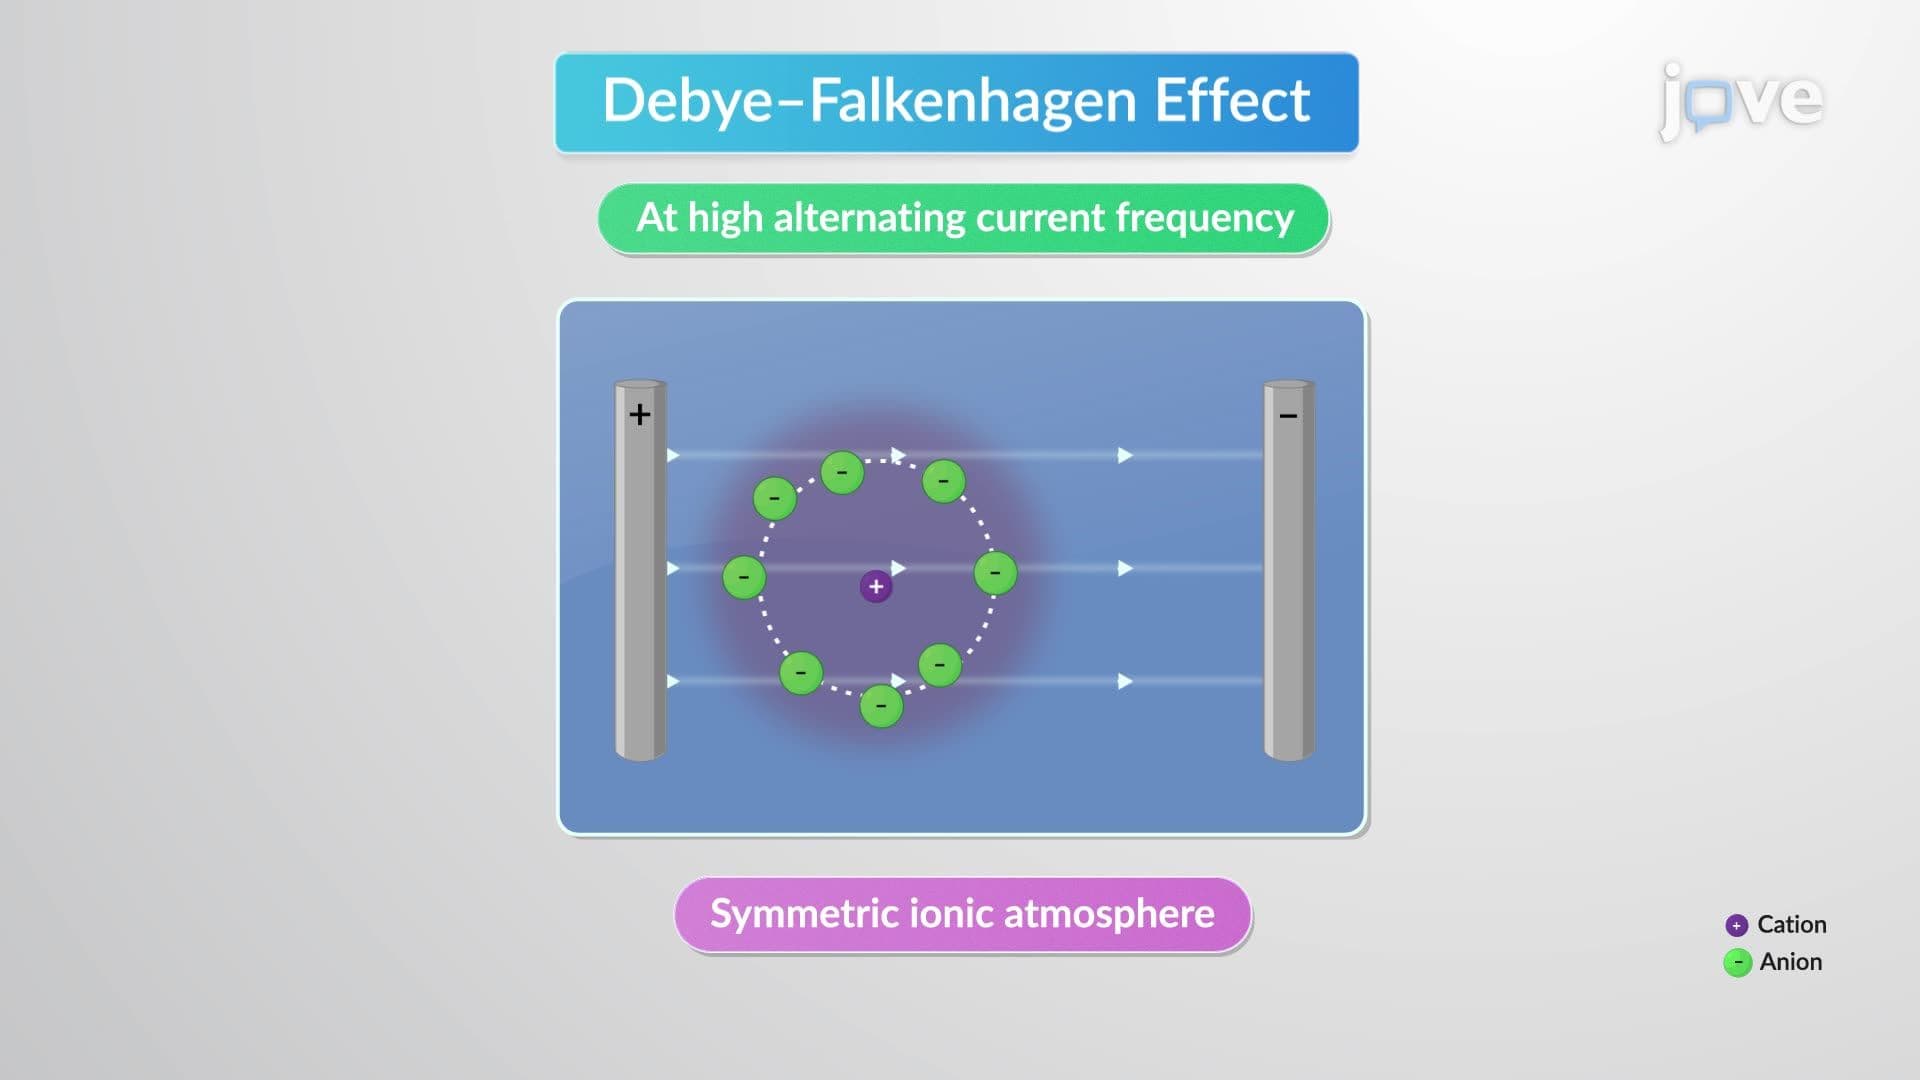
Debye–Huckel–Onsager Conductance Equation
 02:38
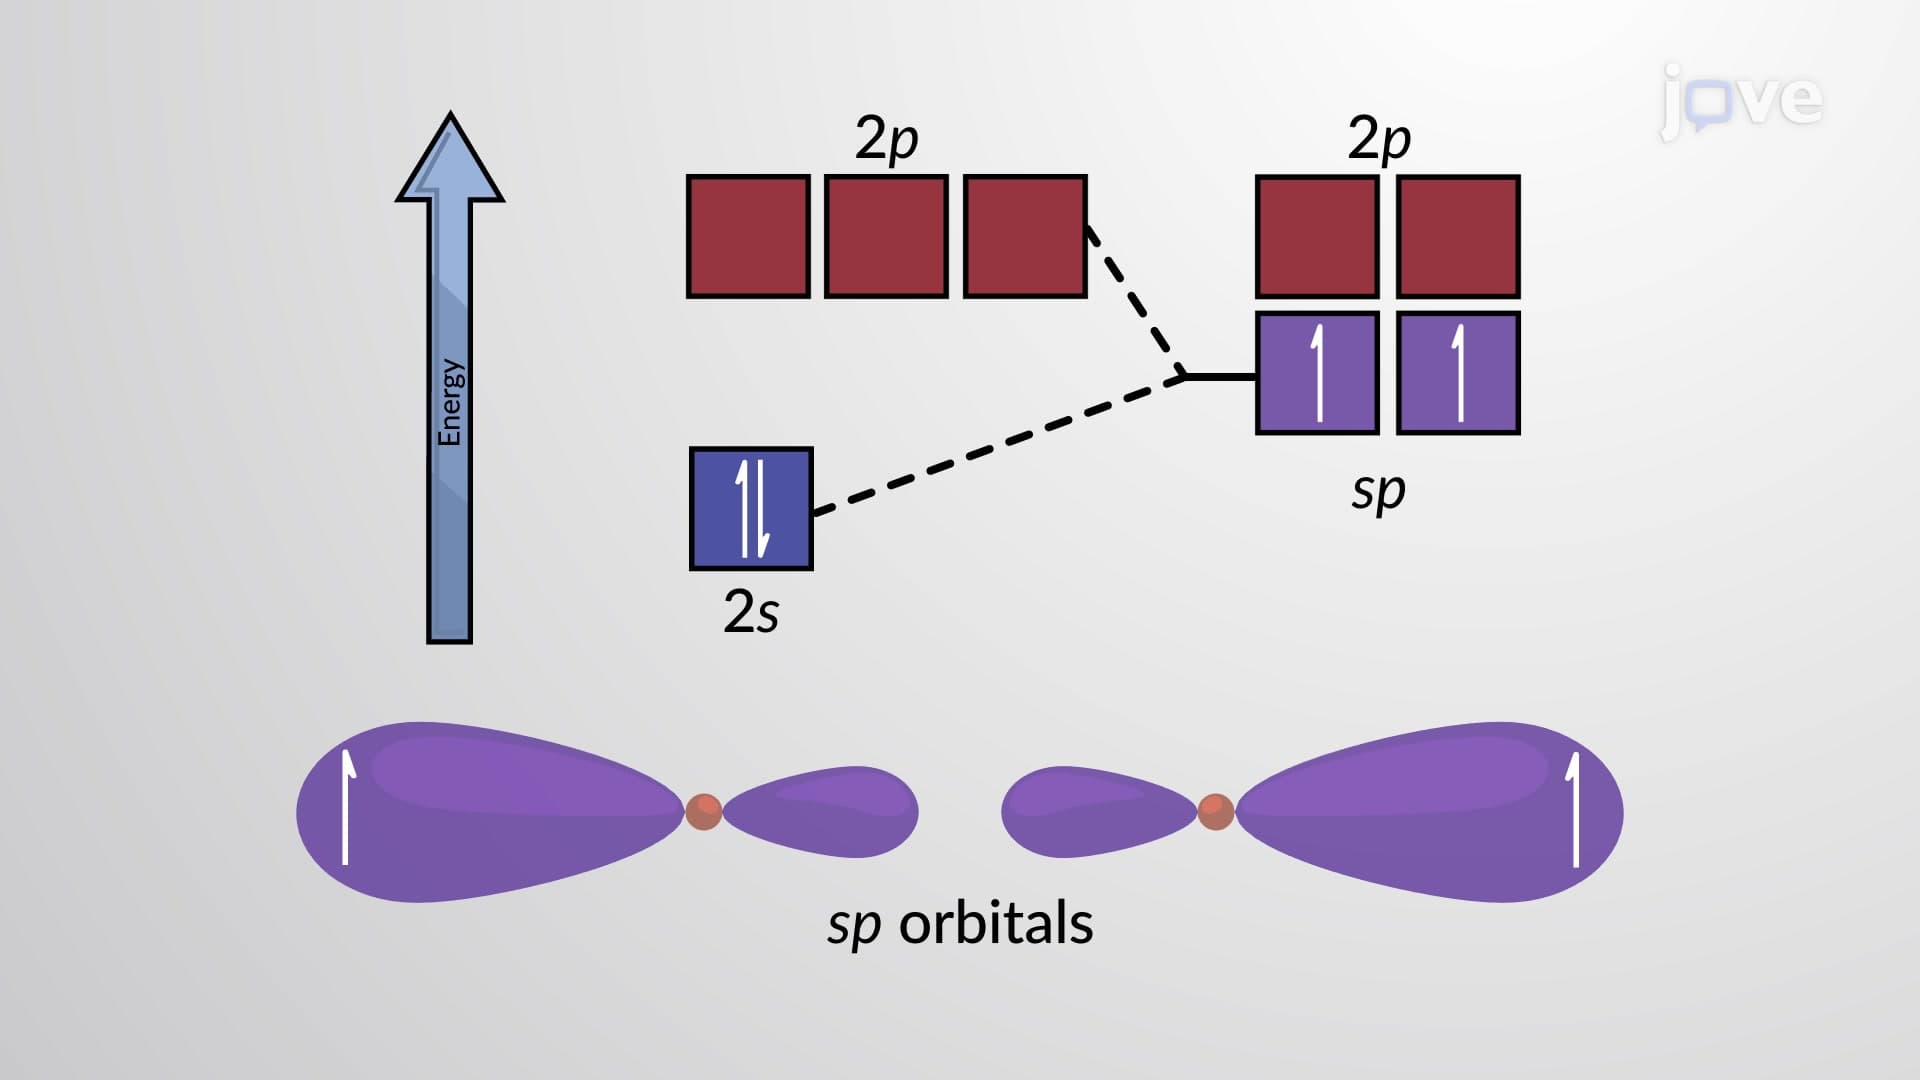
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 02:16
Resonance and Hybrid Structures
Resonance Structures and Resonance Hybrids
The Lewis structure of a nitrite anion (NO2−) may actually be drawn in two different ways, distinguished by the locations of the N–O and N=O bonds.
 02:36
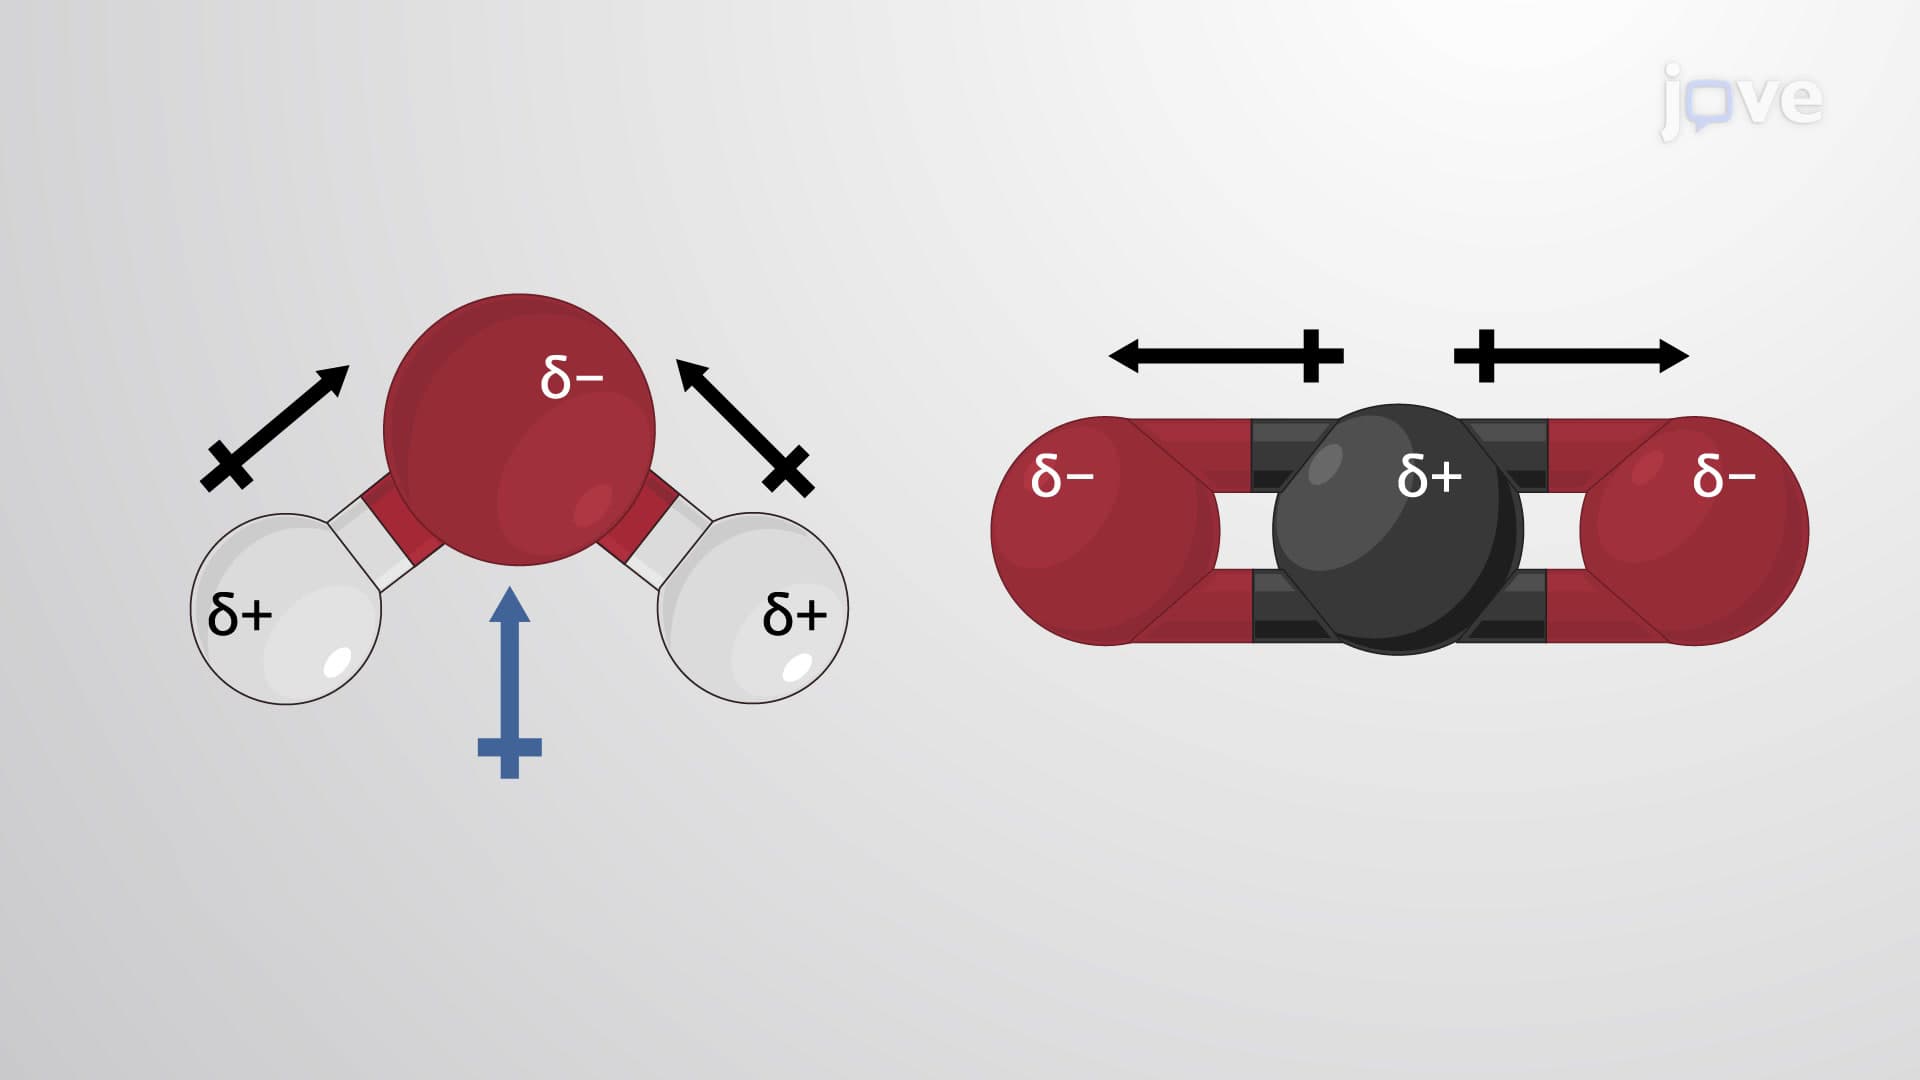
Molecular Geometry and Dipole Moments
 01:10
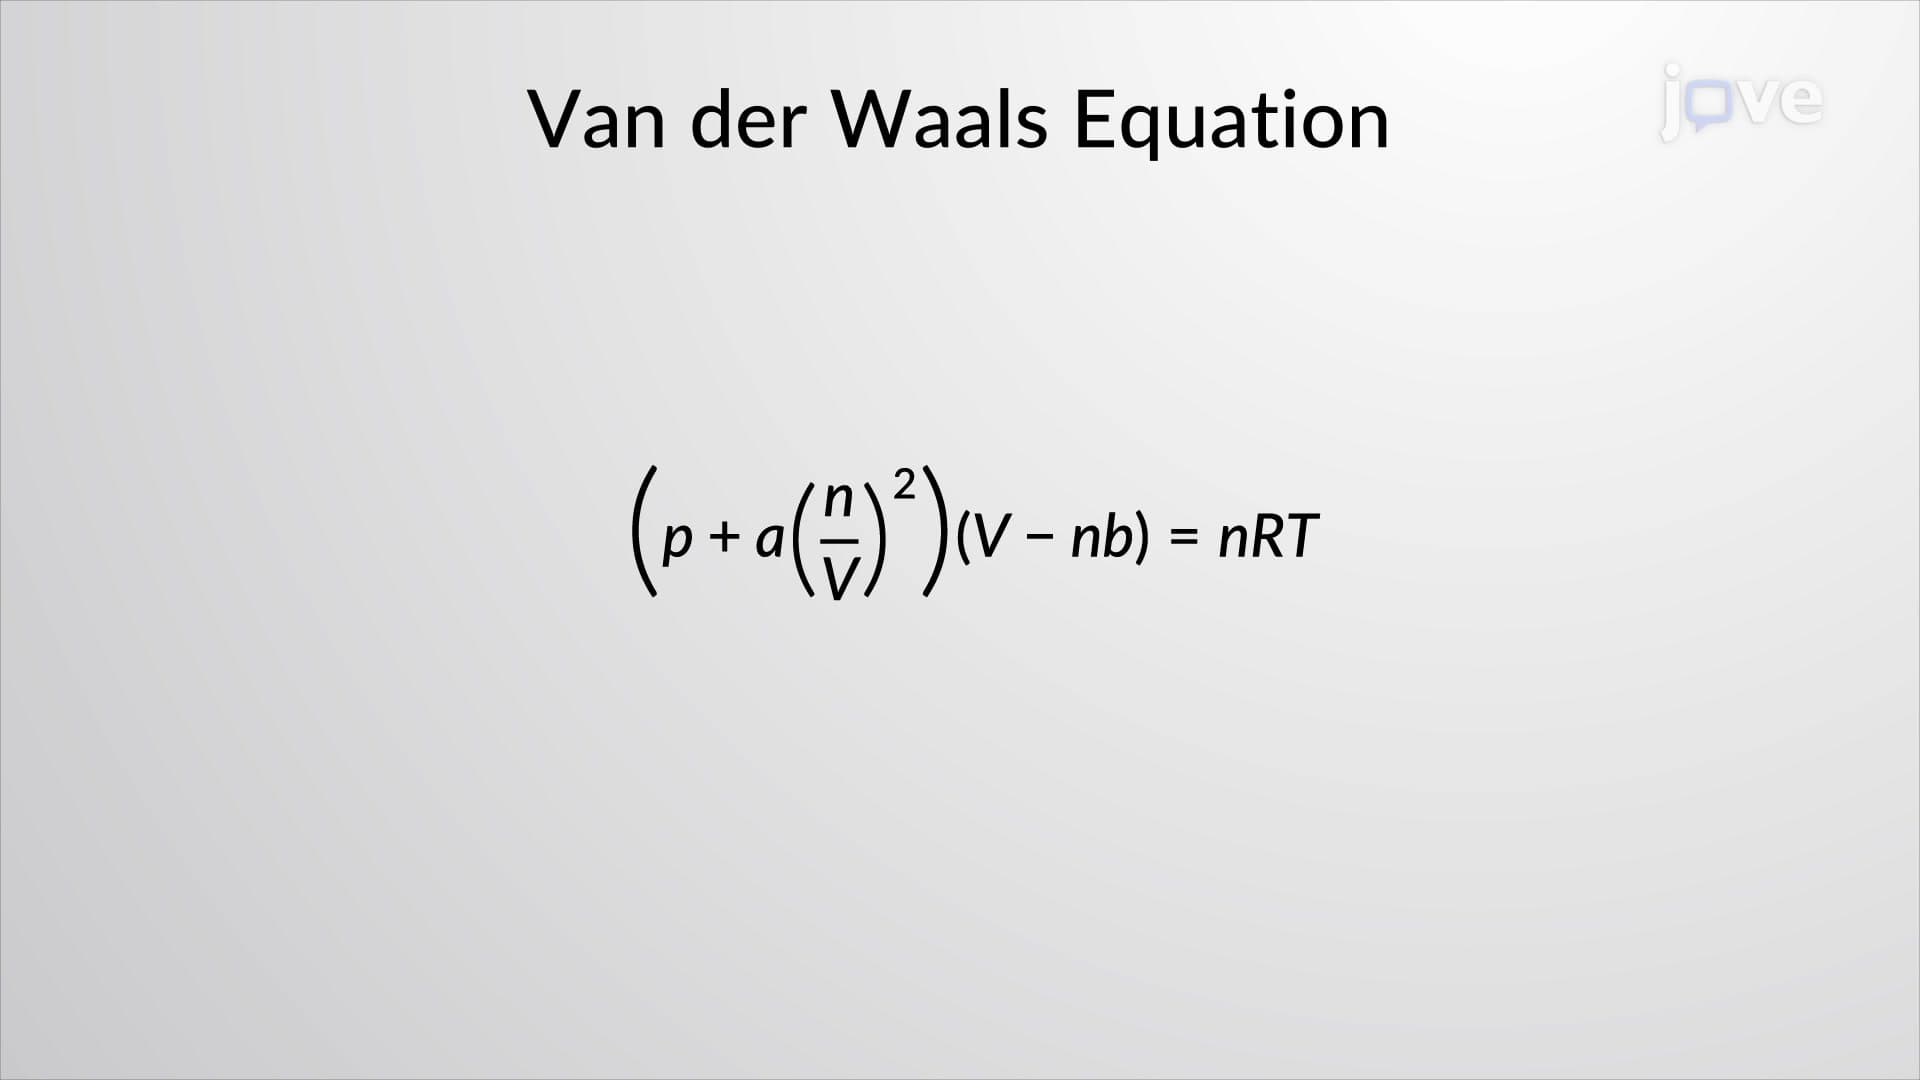
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...