Comparative analysis of RNA sequencing methods for degraded or low-input samples
Related Experiment Video
Updated: May 8, 2026
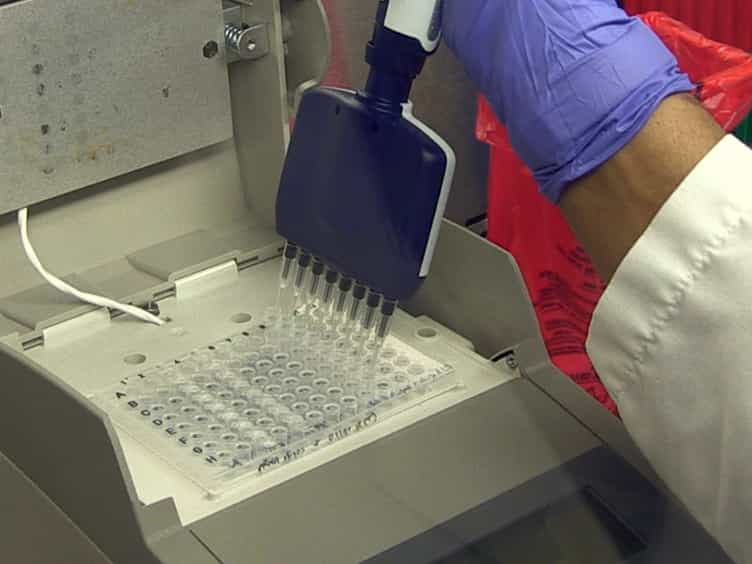
Unbiased Deep Sequencing of RNA Viruses from Clinical Samples
Published on: July 2, 2016
Xian Adiconis1, Diego Borges-Rivera, Rahul Satija
1Broad Institute of MIT and Harvard, Cambridge, Massachusetts, USA.

Abstract:
RNA-seq is an effective method for studying the transcriptome, but it can be difficult to apply to scarce or degraded RNA from fixed clinical samples, rare cell populations or cadavers. Recent studies have proposed several methods for RNA-seq of low-quality and/or low-quantity samples, but the relative merits of these methods have not been systematically analyzed. Here we compare five such methods using metrics relevant to transcriptome annotation, transcript discovery and gene expression. Using a single human RNA sample, we constructed and sequenced ten libraries with these methods and compared them against two control libraries. We found that the RNase H method performed best for chemically fragmented, low-quality RNA, and we confirmed this through analysis of actual degraded samples. RNase H can even effectively replace oligo(dT)-based methods for standard RNA-seq. SMART and NuGEN had distinct strengths for measuring low-quantity RNA. Our analysis allows biologists to select the most suitable methods and provides a benchmark for future method development.
Related Concept Videos
 03:21
RNA-seq
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while microarray-based...
 02:24
Ribosome Profiling
Applications of ribosome profiling
Ribosome profiling has many applications, including in vivo monitoring of translation inside a particular organ or tissue type and quantifying new protein synthesis levels.
The technique helps...