Computational methods for high resolution prediction and refinement of protein structures
Related Experiment Video
Updated: May 11, 2026

Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Richard A Friesner1, Robert Abel, Dahlia A Goldfeld
1Department of Chemistry, Columbia University, New York, NY, USa. rich@chem.columbia.edu

Abstract:
We review advances in implicit solvation and sampling algorithms which have resulted in enhanced capabilities in predicting and refining localized protein structures (e.g. loop regions) to high resolution. Improvements in the generalized Born model and hydrophobicity term yield significantly more accurate energetics; specialized sampling algorithms allow complex local structures, such as a loop-helix-loop region, to be reliably predicted. A novel penalty term is added for loops containing patterns of dihedrals seldom found in experimental structures. We show prediction of diverse sets of large loops, in the native backbone environment, to subångström accuracy. The methodology offers the promise of addressing the refinement problem in homology modeling if an approach can be devised to handle delocalized errors in the structure.

Related Concept Videos
 01:24
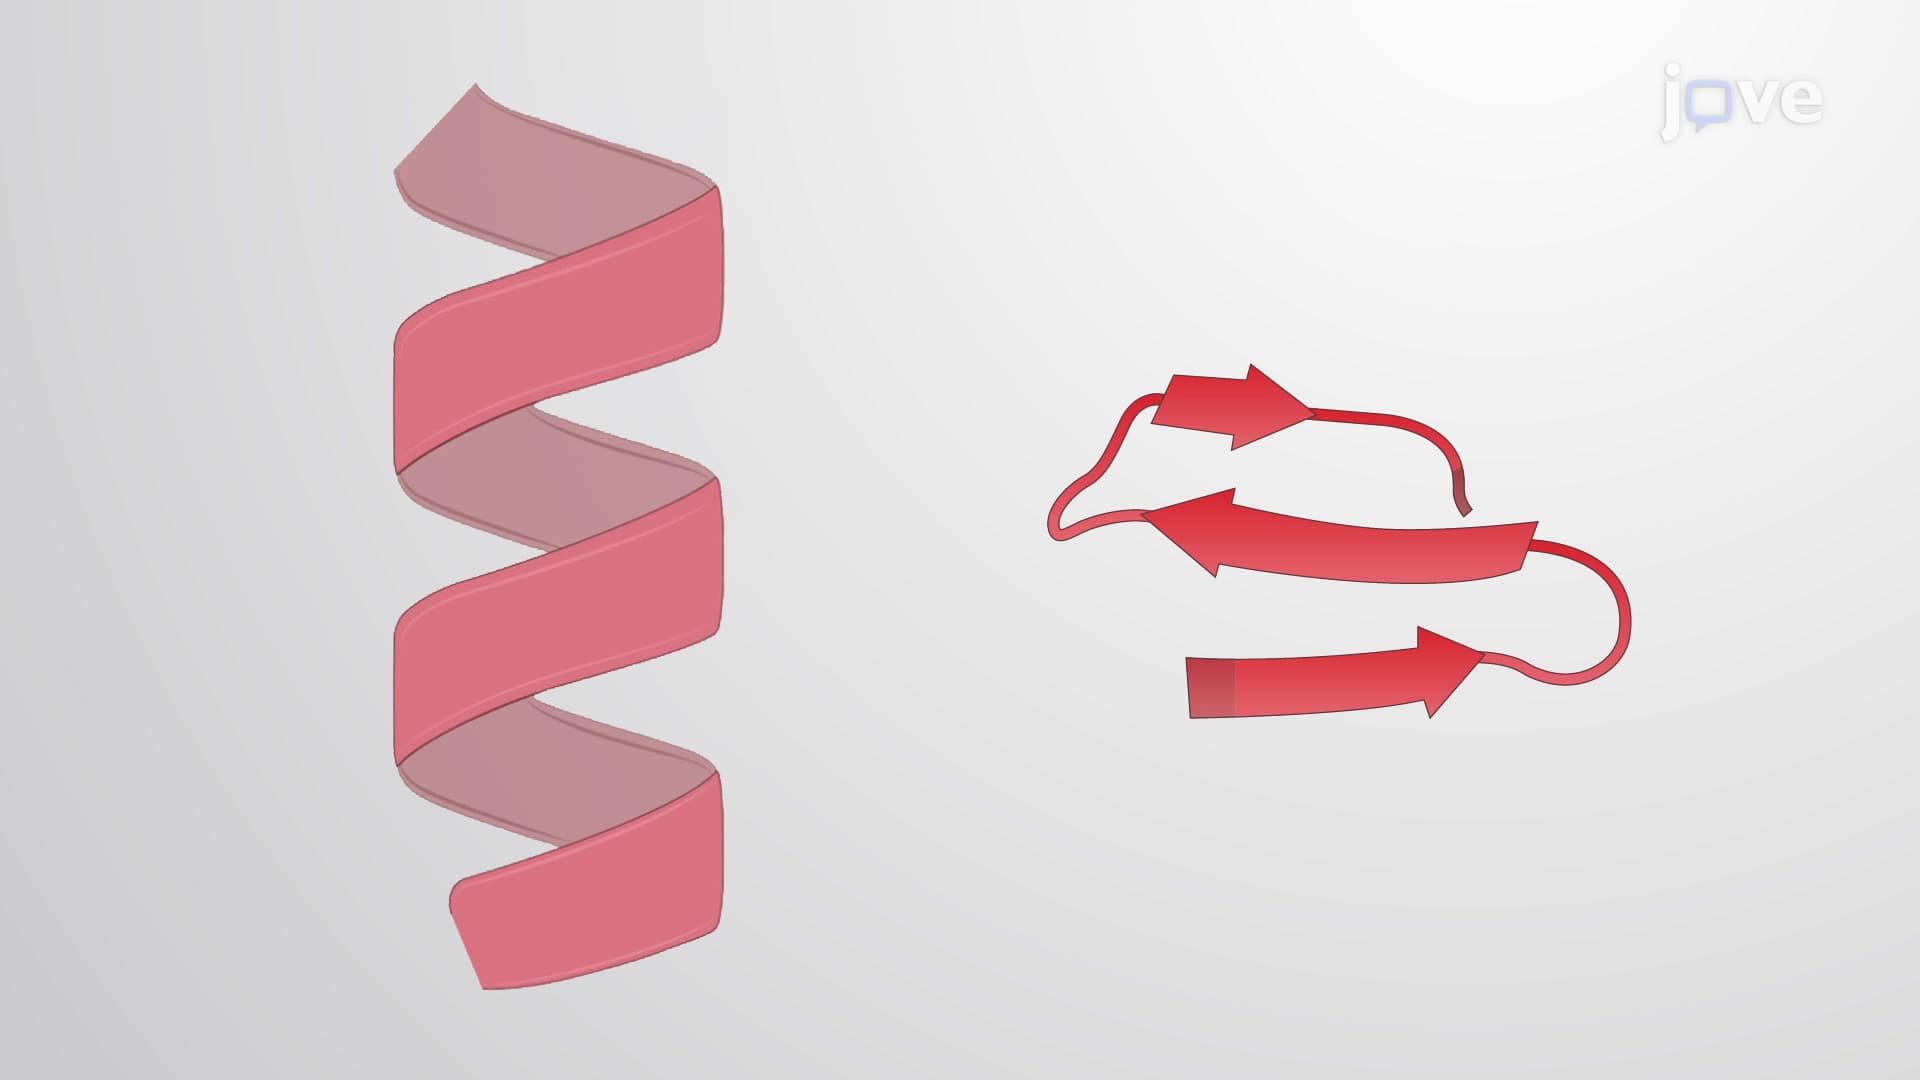
Protein Organization
The primary structure of a protein is its amino acid sequence.
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...