Related Experiment Video
Updated: May 11, 2026

Compact Quantum Dots for Single-molecule Imaging
Published on: October 9, 2012
A DFT/TDDFT study on the optoelectronic properties of the amine-capped magic (CdSe)13 nanocluster
Jon M Azpiroz1, Jon M Matxain, Ivan Infante
1Kimika Fakultatea, Euskal Herriko Unibertsitatea (UPV/EHU) and Donostia International Physics Center (DIPC), P.K. 1072 Donostia, Euskadi, Spain. ionmikel.azpiroz@ehu.es

Abstract:
Motivated by the recent experiments by Wang et al. (Angew. Chem., Int. Ed. 2012, 51, 6154-6157), in which the alkylamine-capped magic-size (CdSe)13 has been isolated for the first time, we report on the computational modeling of the putative low-lying isomers of (CdSe)13, both bare and ligand-protected. According to Density Functional Theory (DFT) calculations, the core@cage configuration Se@Cd13Se12, consisting of a Se atom incarcerated in the center of a puckered Cd13Se12 cage, lies lower in energy than fullerene- and wurtzite-like structures. Methylamine-capped nanoclusters present average bond energies per ligand of about 20 kcal mol(-1), while bond energy decomposition schemes show this interaction to be mostly electrostatically-driven. The computed Time-Dependent-DFT (TDDFT) spectrum of the lowest-lying methylamine-protected (CdSe)13 isomer essentially coincides with the experiment, with a notable blueshift of the absorption features induced by the ligands. The LUMO has been found to be the acceptor orbital for all the lowest-lying electronic excitations, in both the bare and methylamine-capped clusters, which could explain the narrow emission profiles inherent in semiconductor nanostructures. In addition, the attachment of pyridine and aniline molecules has been evaluated. Interestingly, the molecular orbitals of these aromatic amines located on the edges of the valence and conduction bands may act as trap states, in agreement with experimental evidences. In the particular case of pyridine molecules, unoccupied orbitals intrude into the HOMO-LUMO gap of the cluster.
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:19
NMR Spectroscopy Of Amines
 01:20
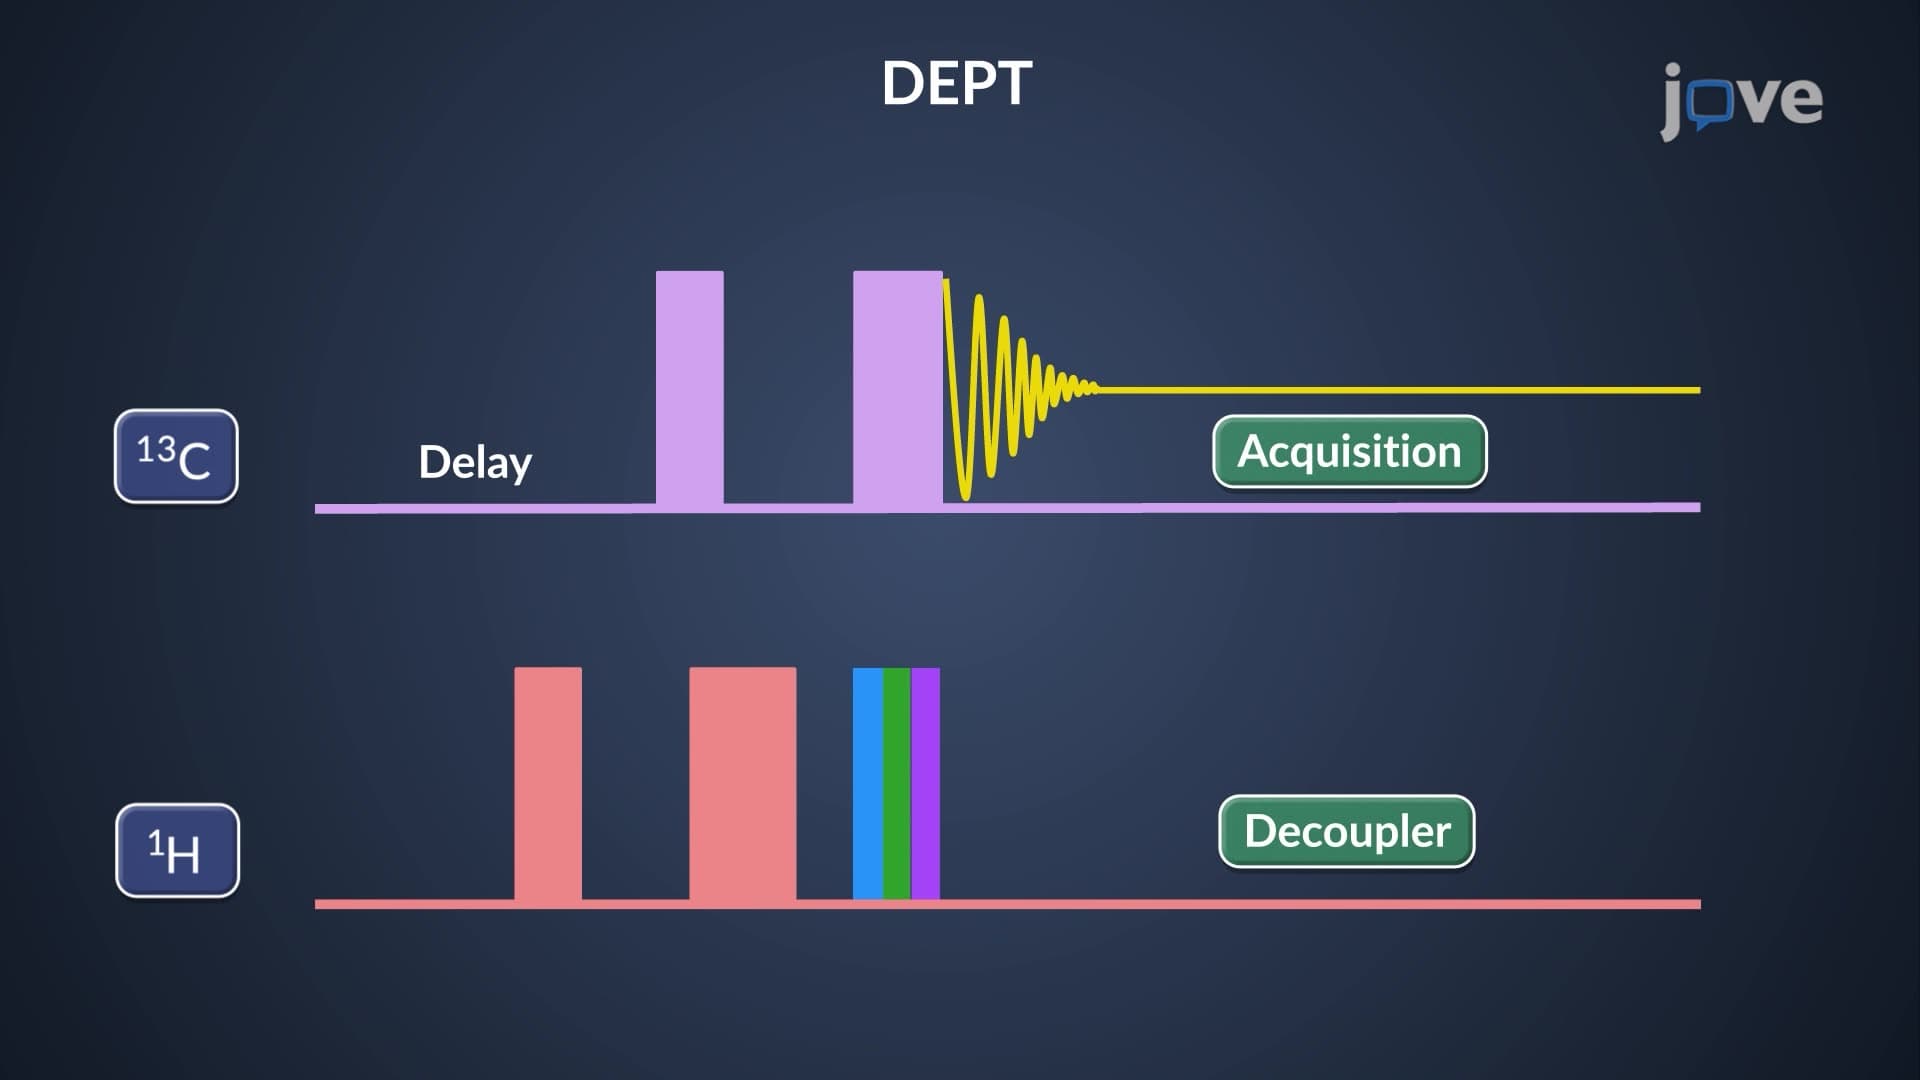
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 02:42
Valence Bond Theory
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...