Related Experiment Video
Updated: May 11, 2026

Identification and Classification of Position-specific GABAA Receptor Subunit Missense Variants for Their Role In Hippocampal Pyramidal Neurons
Published on: June 6, 2025
Trafficking-deficient mutant GABRG2 subunit amount may modify epilepsy phenotype
Jing-Qiong Kang1, Wangzhen Shen, Robert L Macdonald
1Departments of Neurology, Vanderbilt University Medical Center, Nashville, TN.

Objective:
Genetic epilepsies and many other human genetic diseases display phenotypic heterogeneity, often for unknown reasons. Disease severity associated with nonsense mutations is dependent partially on mutation gene location and resulting efficiency of nonsense-mediated mRNA decay (NMD) to eliminate potentially toxic proteins. Nonsense mutations in the last exon do not activate NMD, thus producing truncated proteins. We compared the protein metabolism and the impact on channel biogenesis, function, and cellular homeostasis of truncated γ2 subunits produced by GABRG2 nonsense mutations associated with epilepsy of different severities and by a nonsense mutation in the last exon unassociated with epilepsy.
Methods:
γ-Aminobutyric acid type A receptor subunits were coexpressed in non-neuronal cells and neurons. NMD was studied using minigenes that support NMD. Protein degradation rates were determined using (35) S radiolabeling pulse chase. Channel function was determined by whole cell recordings, and subunits trafficking and cellular toxicity were determined using flow cytometry, immunoblotting, and immunohistochemistry.
Results:
Although all GABRG2 nonsense mutations resulted in loss of γ2 subunit surface expression, the truncated subunits had different degradation rates and stabilities, suppression of wild-type subunit biogenesis and function, amounts of conjugation with polyubiquitin, and endoplasmic reticulum stress levels.
Interpretation:
We compared molecular phenotypes of GABRG2 nonsense mutations. The findings suggest that despite the common loss of mutant allele function, each mutation produced different intracellular levels of trafficking-deficient subunits. The concentration-dependent suppression of wild-type channel function and cellular disturbance resulting from differences in mutant subunit metabolism may contribute to associated epilepsy severities and by implication to phenotypic heterogeneity in many inherited human diseases.
Insights
Nonsense mutations in GABRG2 genes cause epilepsy by producing truncated proteins. Differences in how these proteins are metabolized and affect GABA-A receptor function contribute to epilepsy severity and genetic disease variability.
Area of Science:
- Neurogenetics
- Molecular Biology
- Cellular Biology
Background:
- Genetic epilepsies and other inherited diseases often show variable severity (phenotypic heterogeneity) due to unknown factors.
- Nonsense mutations can lead to truncated proteins, with severity influenced by nonsense-mediated mRNA decay (NMD) efficiency.
- Mutations in the last exon may evade NMD, producing full-length truncated proteins.
Purpose of the Study:
- To compare the effects of truncated γ2 subunits from GABRG2 nonsense mutations on epilepsy of varying severity.
- To investigate the impact of these truncated subunits on channel biogenesis, function, and cellular homeostasis.
- To analyze the role of mutation location, specifically last-exon mutations, on protein metabolism and disease presentation.
Main Methods:
- Coexpression of GABA-A receptor subunits in non-neuronal cells and neurons.
- NMD assessment using minigenes.
- Protein degradation rates via (35)S radiolabeling.
- Channel function evaluation through whole-cell recordings.
- Subunit trafficking, cellular toxicity, polyubiquitination, and ER stress analysis using flow cytometry, immunoblotting, and immunohistochemistry.
Main Results:
- All GABRG2 nonsense mutations led to reduced γ2 subunit surface expression.
- Truncated subunits exhibited differential degradation rates, stabilities, and polyubiquitin conjugation.
- Mutant subunits suppressed wild-type subunit biogenesis and function, increasing endoplasmic reticulum stress.
Conclusions:
- Each GABRG2 nonsense mutation generates distinct intracellular levels of trafficking-deficient γ2 subunits.
- Metabolic differences in mutant subunits contribute to varying degrees of wild-type channel suppression and cellular disturbance.
- These molecular variations likely underlie epilepsy severity and contribute to phenotypic heterogeneity in inherited human diseases.
Related Concept Videos
 01:18
Antiepileptic Drugs: GABAergic Pathway Potentiators
The key GABA pathway potentiators used in epilepsy management are as follows.
Benzodiazepines are a well-known class of drugs used for their...
 01:20
Antiepileptic Drugs: Potassium Channel Activators
Ezogabine has gained approval as an adjunctive treatment...
 01:20
Antiepileptic Drugs: Modulators of Neurotransmitter Release Mediated by SV2A Protein
SV2A is a transmembrane glycoprotein located predominantly in the brain, modulating the release of neurotransmitters for neuronal communication. Both levetiracetam and brivaracetam exhibit a high affinity for...
 01:24
Epilepsy and Seizures: Overview
Various factors can trigger epilepsy, including genetic factors, brain damage, metabolic causes, and unknown etiology. Diagnosis of epilepsy involves electroencephalography (EEG), which...
 01:14
Rab Proteins
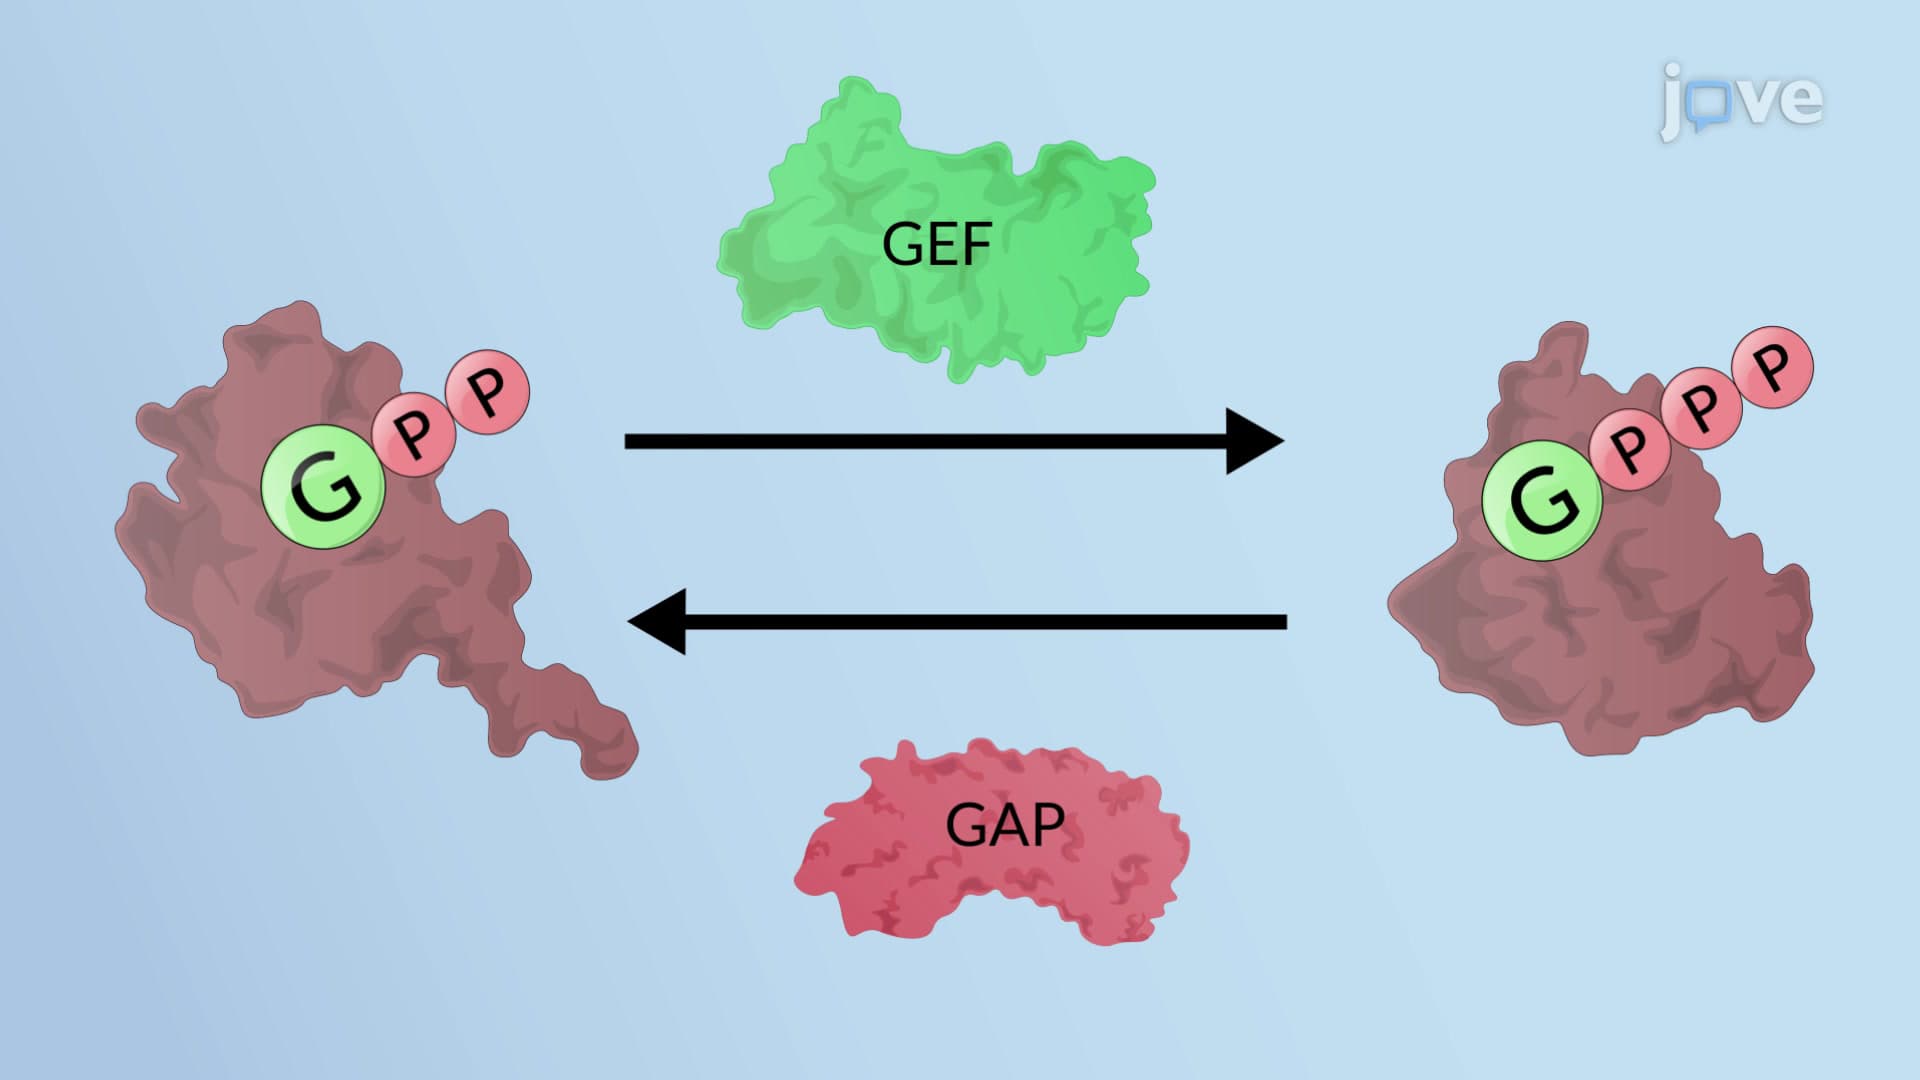
Rab proteins switch between a cytosolic, GDP-bound inactive state and a membrane-anchored, GTP-bound active state. By themselves, Rabs show slow rates of GDP/GTP exchange and GTP hydrolysis. Thus, Rab proteins are considered...
 01:14
Antiepileptic Drugs: Glutamate Antagonists