Related Experiment Video
Updated: May 10, 2026

Methods of Ex Situ and In Situ Investigations of Structural Transformations: The Case of Crystallization of Metallic Glasses
Published on: June 7, 2018
Crystal structure and local structure of Mg(2-x)Pr(x)Ni4 (x = 0.6 and 1.0) deuteride using in situ neutron total
1National Institute of Advanced Industrial Science and Technology, AIST Central-5, 1-1-1 Higashi, Tsukuba, Ibaraki, 305-0035, Japan. kouji.sakaki@aist.go.jp

Abstract:
We studied crystal structure and local structure of Mg(2-x)Pr(x)Ni4 (x = 0.6 and 1.0) and their deuterides using in situ neutron total scattering and first-principles calculations. The total scattering data were analyzed using Rietveld refinement and pair distribution function analysis (PDF). The crystal structure of Mg(2-x)Pr(x)Ni4 before deuterium absorption was C15b in space group F43m. No difference between the crystal and local (PDF) structures was observed. The crystal structure of Mg1.0Pr1.0Ni4D(∼4) was found to be orthorhombic in space group Pmn2(1), with three deuterium occupation sites: PrNi3 and two types of bipyramidal Pr2MgNi2 that have a plane of symmetry composed of MgNi2. There is no significant difference between the crystal structure and the local structure of Mg1.0Pr1.0Ni4D(∼4). On the other hand, the average crystal structure of the Mg-rich Mg1.4Pr0.6Ni4D(∼3.6) was C15b with two deuterium occupation sites: PrNi3 and MgPrNi2 suggesting that the deuterium occupation shifts away from the Pr2MgNi2 bipyramid. First-principles relaxed structures also showed the shift of the hydrogen occupation site toward the Pr atom of the bipyramid, when induced by Mg substitution for the opposing Pr, resulting in hydrogen occupation in the MgPrNi2 tetrahedral site. The PDF pattern of Mg1.4Pr0.6Ni4D(∼3.6) cannot be refined below 7.2 Å in atomic distances using the C15b structure which was obtained from Rietveld refinement but can be done using an orthorhombic structure. It suggests that Mg1.4Pr0.6Ni4D(∼3.6) was locally distorted to the orthorhombic.
Related Concept Videos
 01:29
Determination of Crystal Structures
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:18
X-ray Crystallography
Diffraction
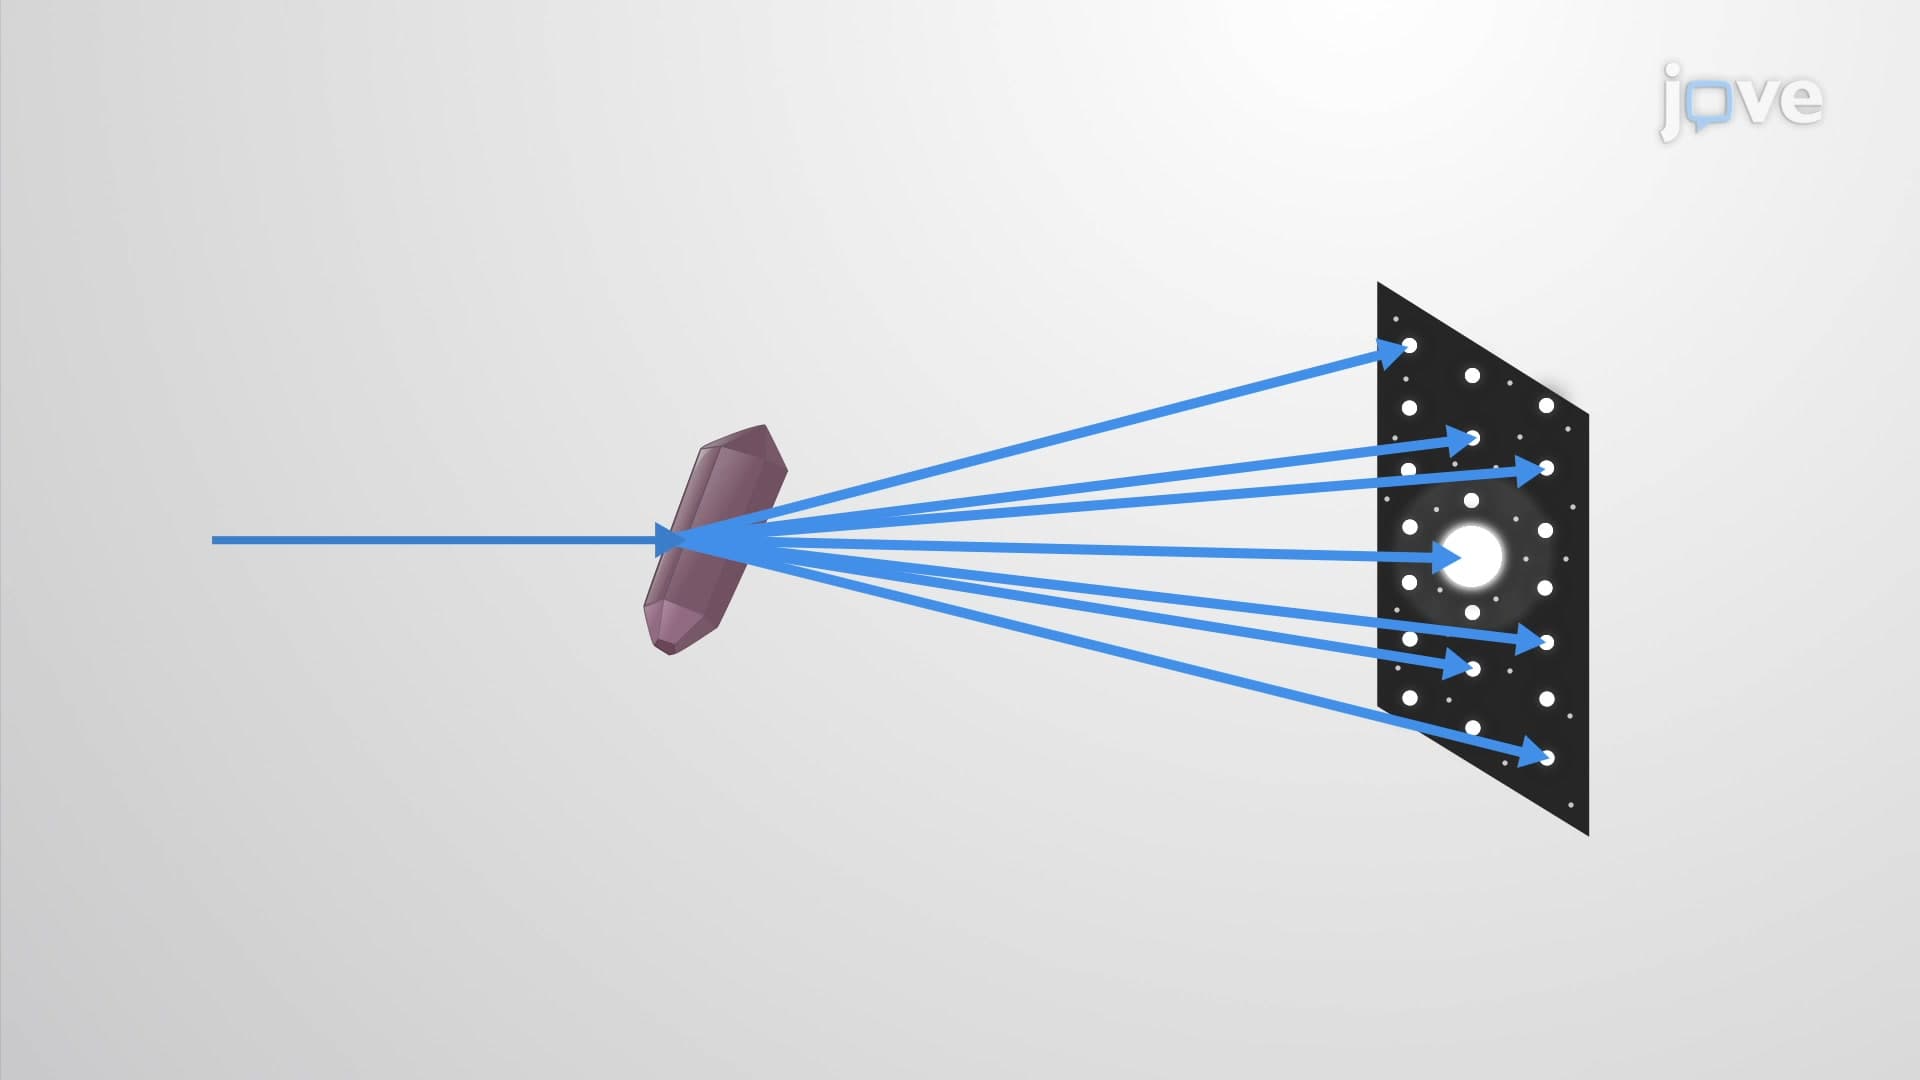
Diffraction is the change in the direction of travel experienced by an electromagnetic wave when it encounters a physical barrier whose dimensions are comparable to those of the wavelength of the light. X-rays are electromagnetic radiation with wavelengths about as long as the distance between neighboring...
 02:42
Ionic Crystal Structures
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:26
Imperfections in Crystal Structure: Stoichiometric Point Defects