Increased apical Na+ permeability in cystic fibrosis is supported by a quantitative model of epithelial ion transport
Related Experiment Video
Updated: May 10, 2026
A Fluorescence-Based Assay of Membrane Potential for High-Throughput Functional Study of Two Endogenous Ion Channels in Two Epithelial Cell Lines
Published on: June 22, 2022
Donal L O'Donoghue1, Vivek Dua, Guy W J Moss
1Centre for Mathematics and Physics in the Life Sciences and Experimental Biology, University College London, Gower Street, London, WC1E 6BT, United Kingdom.

Abstract:
Cystic fibrosis (CF) is caused by mutations in the cystic fibrosis transmembrane conductance regulator (CFTR) gene, which encodes an anion channel. In the human lung CFTR loss causes abnormal ion transport across airway epithelial cells. As a result CF individuals produce thick mucus, suffer persistent bacterial infections and have a much reduced life expectancy. Trans-epithelial potential difference (Vt) measurements are routinely carried out on nasal epithelia of CF patients in the clinic. CF epithelia exhibit a hyperpolarised basal Vt and a larger Vt change in response to amiloride (a blocker of the epithelial Na(+) channel, ENaC). Are these altered bioelectric properties solely a result of electrical coupling between the ENaC and CFTR currents, or are they due to an increased ENaC permeability associated with CFTR loss? To examine these issues we have developed a quantitative mathematical model of human nasal epithelial ion transport. We find that while the loss of CFTR permeability hyperpolarises Vt and also increases amiloride-sensitive Vt, these effects are too small to account for the magnitude of change observed in CF epithelia. Instead, a parallel increase in ENaC permeability is required to adequately fit observed experimental data. Our study provides quantitative predictions for the complex relationships between ionic permeabilities and nasal Vt, giving insights into the physiology of CF disease that have important implications for CF therapy.
More Related Videos
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:23
Transcellular Transport of Solutes
 01:26
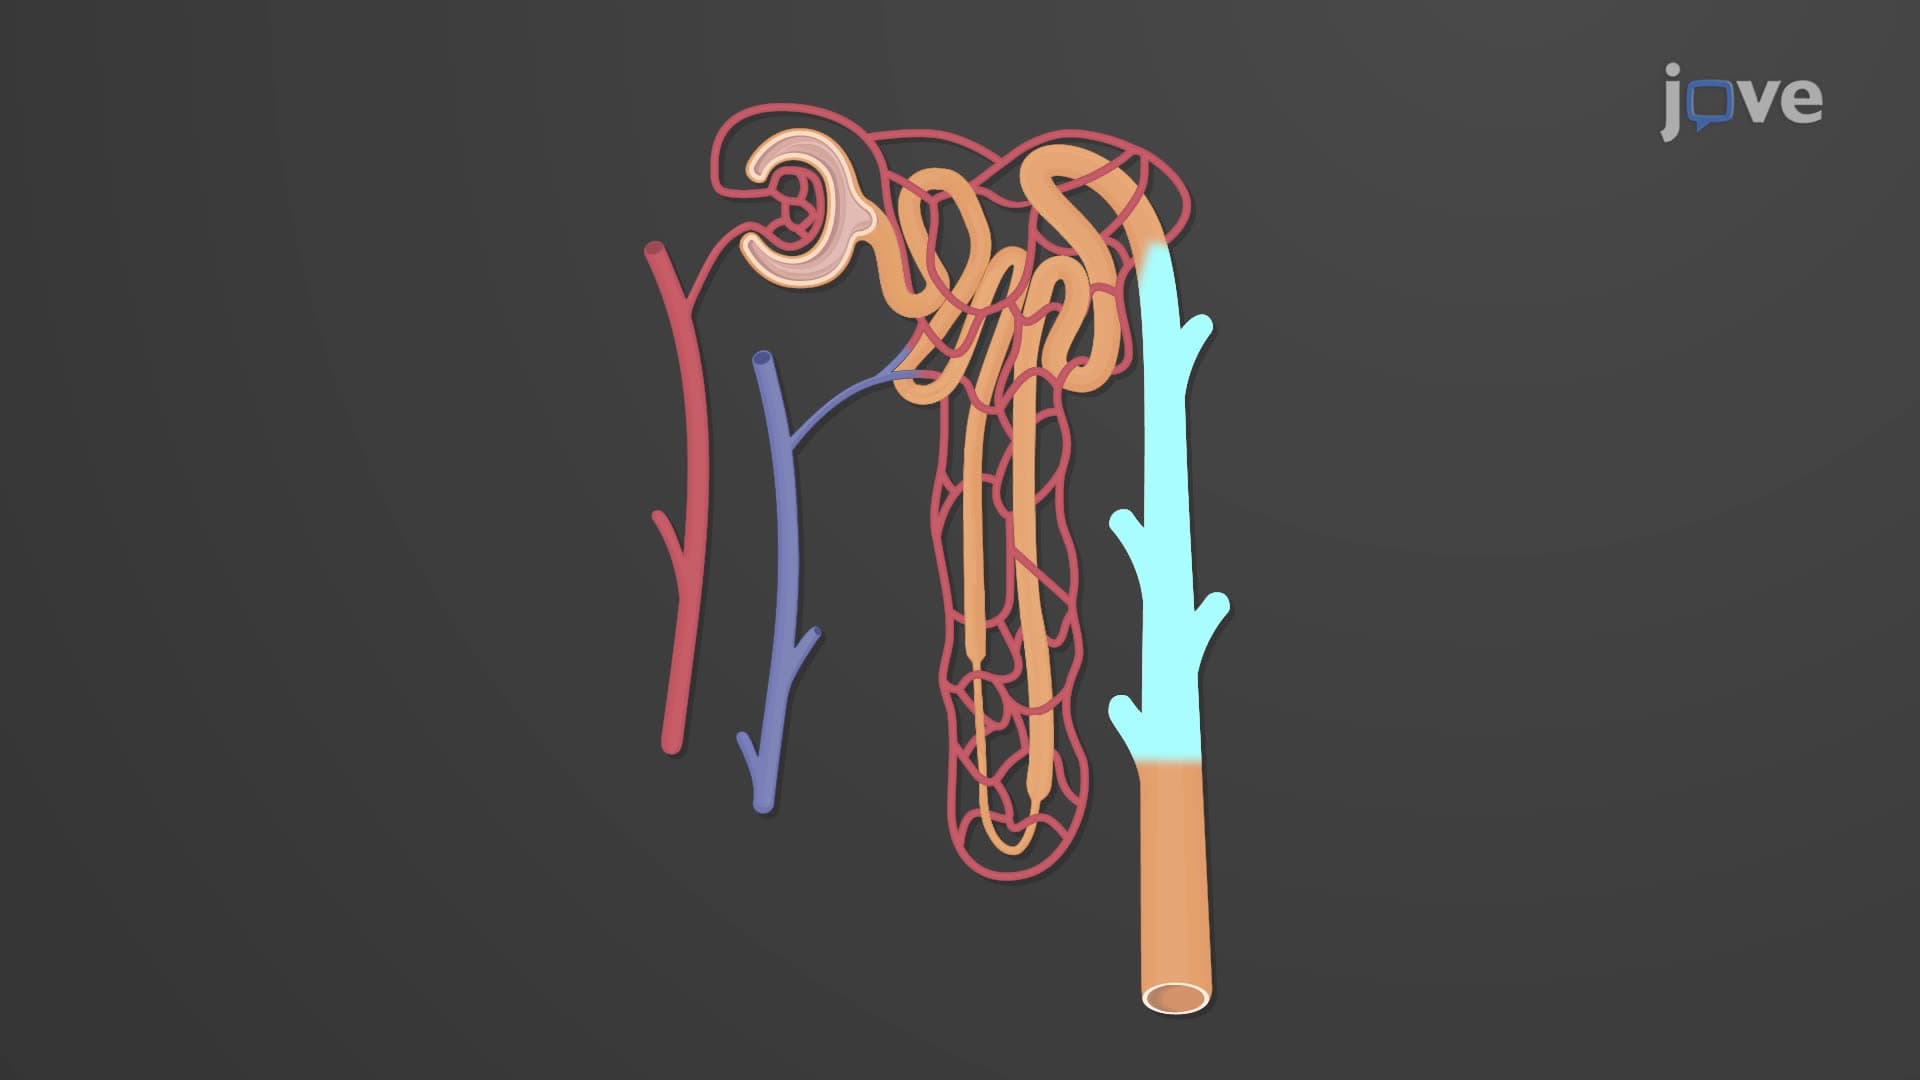
Reabsorption and Secretion in the DCT and Collecting Duct
The distal part of the DCT, along with the...
 01:17
Reabsorption and Secretion in the Loop of Henle
 01:44
The Significance of Membrane Transport
Transporters facilitate either an active or passive movement of solutes. They can allow a single-molecule transport down its...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...