Concentration effect relationship of CYP3A inhibition by ritonavir in humans

Christine Eichbaum1, Marianna Cortese, Antje Blank

  • 1Department of Clinical Pharmacology and Pharmacoepidemiology, Heidelberg University Hospital, Im Neuenheimer Feld 410, 69120, Heidelberg, Germany.

Abstract

Related Concept Videos

Time Course of Drug Effect01:14

Time Course of Drug Effect

The progression of a drug's impact can be analyzed by examining both the concentration-time course and the effect-time course. The concentration-time course is determined by the drug's half-life and is influenced by factors such as its pharmacokinetics, including absorption, distribution, metabolism, and elimination. The effect of the drug is often related to its concentration in the plasma and is calculated using the maximum drug effect and the plasma concentration that generates 50 percent of...
Pharmacodynamic Models: Linear Concentration–Effect Model01:15

Pharmacodynamic Models: Linear Concentration–Effect Model

The linear concentration–effect model, underpinned by the principle that pharmacological effect (E) is directly proportional to plasma drug concentration (C), emerges as a pivotal simplification of the Emax model for conditions where C is significantly less than EC50. This model portrays a linear trajectory of the concentration–effect relationship when drug levels are markedly below the EC50 threshold.Despite its inherent assumption of continuous effect augmentation with increasing drug...
Effect of Hepatic Disease on Pharmacokinetics: Dose Adjustments Due to Hepatic Impairment01:08

Effect of Hepatic Disease on Pharmacokinetics: Dose Adjustments Due to Hepatic Impairment
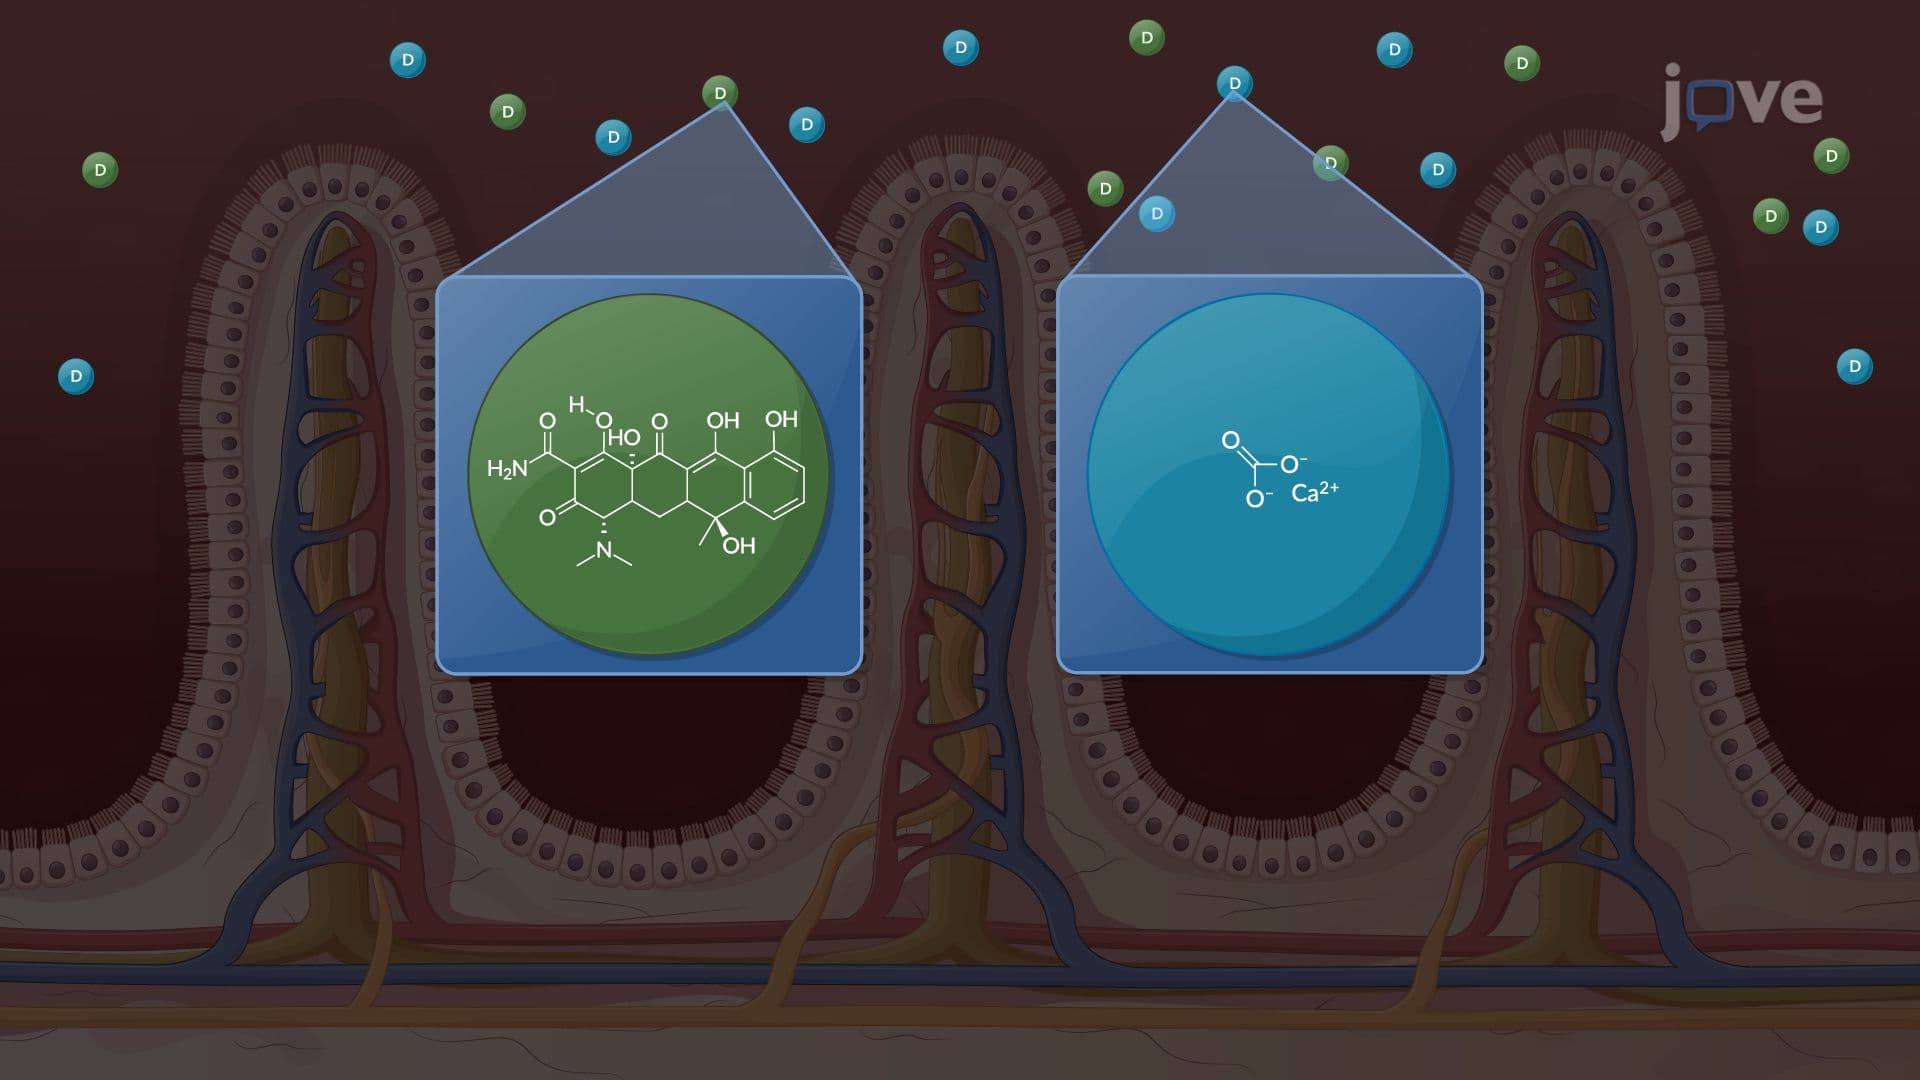
Hepatic impairment, characterized by decreased liver function, does not uniformly mandate adjustments in drug dosage. Whether dosage modifications are necessary depends on various factors related to the drug's metabolism and elimination pathways. If a drug is primarily excreted via the kidneys and bypasses significant hepatic processing, if it undergoes minimal metabolic transformation in the liver, or if it is volatile and primarily expelled through the lungs, dose adjustments may not be...
Pharmacokinetics: Drug–Drug Interactions01:25

Pharmacokinetics: Drug–Drug Interactions
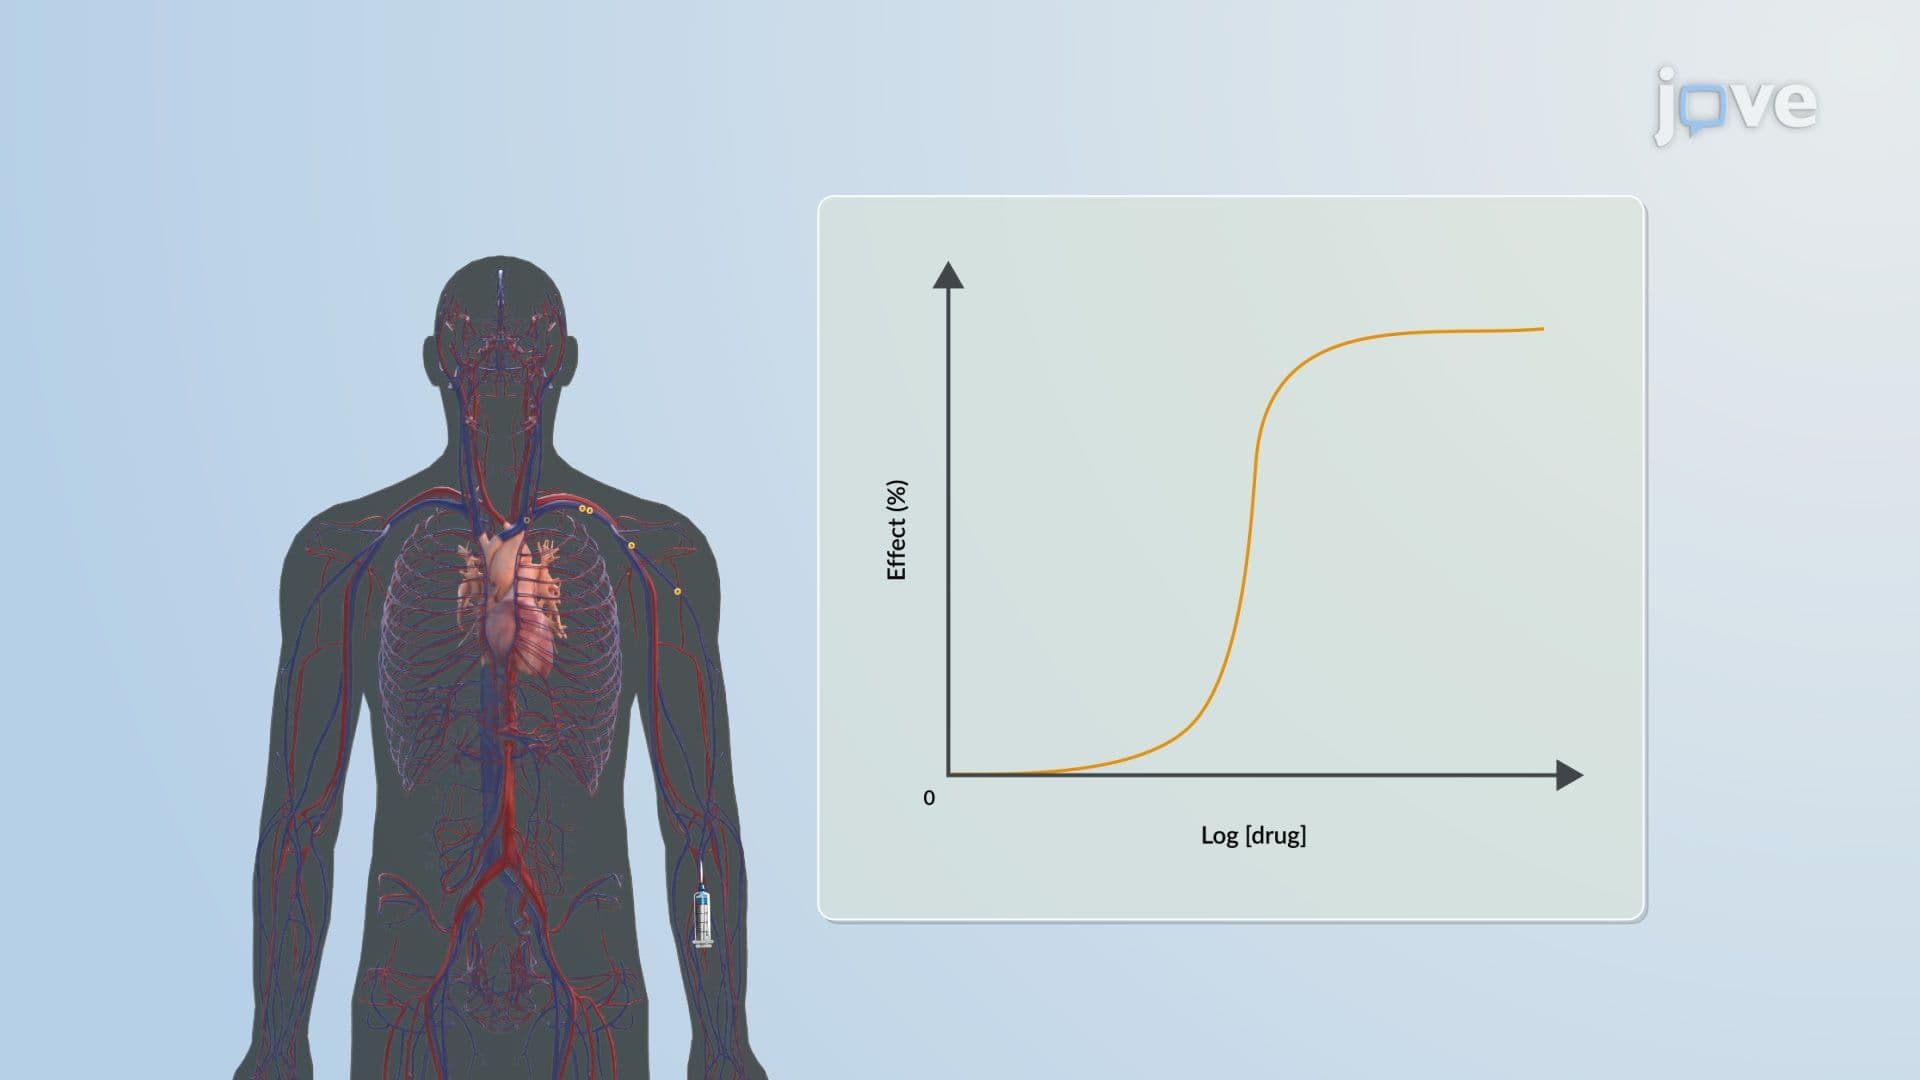
Drug interactions occur when the pharmacological effect of one drug is altered by another substance, either enhancing or diminishing its activity. The drug whose activity is altered is known as the object drug, and the substance causing the alteration is called the agent drug or the precipitant. The net effects of these interactions are mostly undesirable, leading to decreased effectiveness or increased adverse effects. In rare cases, interactions can be beneficial, such as the enhanced...
Pharmacokinetic–Pharmacodynamic Relationship: Intensity of Dose-Effect Relationship01:23

Pharmacokinetic–Pharmacodynamic Relationship: Intensity of Dose-Effect Relationship
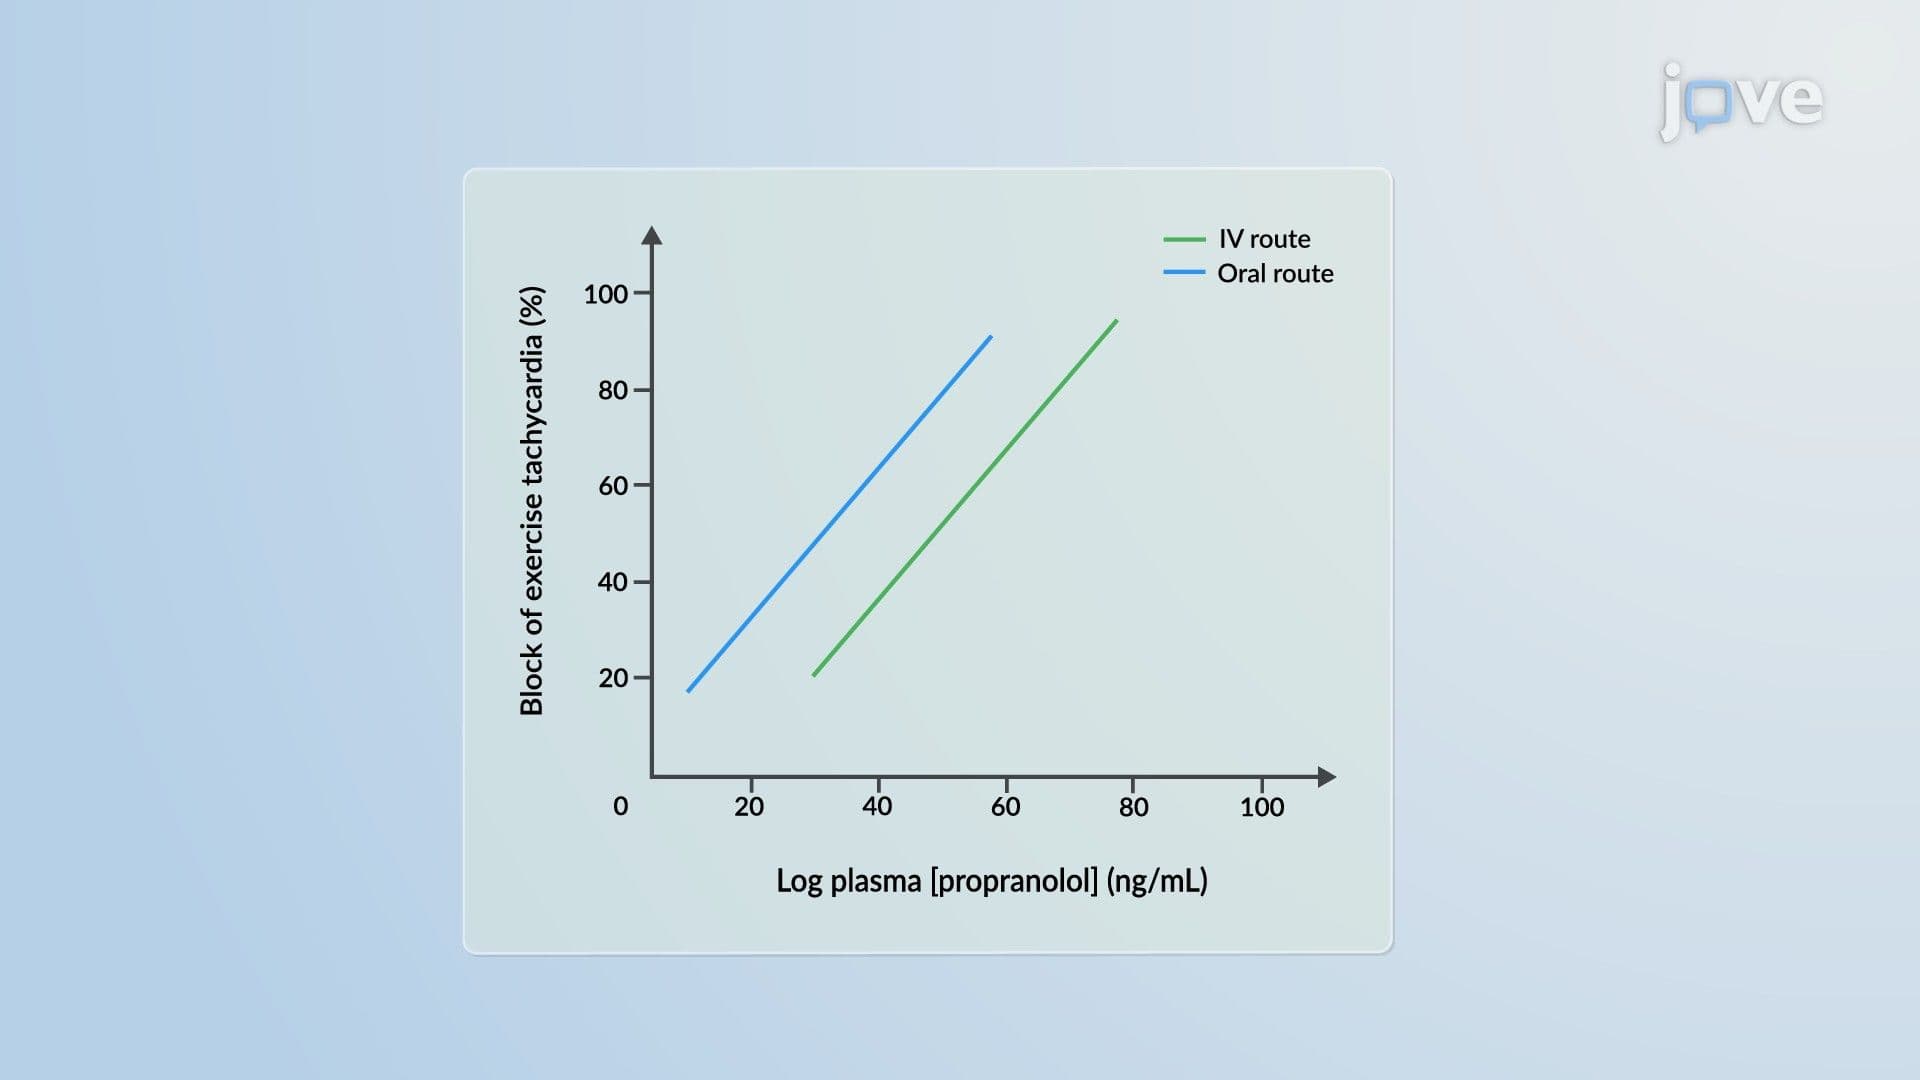
Pharmacodynamics explores the relationship between drug concentration and its effect. In a quantal response drug, the duration of action better correlates with drug concentration, while for graded effect drugs, the intensity of response is more relevant. This intensity depends on the dose, drug removal rate, and the region of the concentration–response curve.The concentration–response curve can be divided into three regions. Region 3 (80–100% maximum response) demonstrates that even as drug...
Pharmacodynamic Models: Logarithmic Concentration–Effect Model01:15

Pharmacodynamic Models: Logarithmic Concentration–Effect Model

The log-linear model is a pharmacological framework used to describe the relationship between drug concentration and its effect. This model is particularly relevant when the observed effects range between 20% and 80% of the drug’s maximum effect (Emax), where a near-linear relationship is observed between the log of drug concentration and the measured effect. However, the log-linear model does not predict the maximum possible effect (Emax) or the effect at zero drug concentration, limiting its...