Related Experiment Video
Updated: May 10, 2026


Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Structural and electronic properties of graphene-ZnO interfaces: dispersion-corrected density functional theory
Pengtao Xu1, Qing Tang, Zhen Zhou
1Tianjin Key Laboratory of Metal and Molecule Based Material Chemistry, Computational Centre for Molecular Science, Institute of New Energy Material Chemistry, Nankai University, Synergetic Innovation Center of Chemical Science and Engineering (Tianjin), Tianjin 300071, People's Republic of China.

Abstract:
Detailed first-principles computations were performed on the geometric and electronic properties of the interfaces between graphene and ZnO polar surfaces. A notable van der Waals force exists at the interface, and charge transfer occurs between graphene and ZnO as a result of the difference in their work functions. The Dirac point of graphene remains intact despite its adsorption on ZnO, implying that its interaction with ZnO does not affect the superior conductivity of graphene. Excited electrons within the energy range of 0-3 eV (versus Fermi energy) in the hybrid systems are mainly accumulated on graphene. The computations provide a theoretical explanation for the good performance of graphene/ZnO hybrid materials in photocatalysts and solar cells.
More Related Videos
Related Concept Videos
 01:28
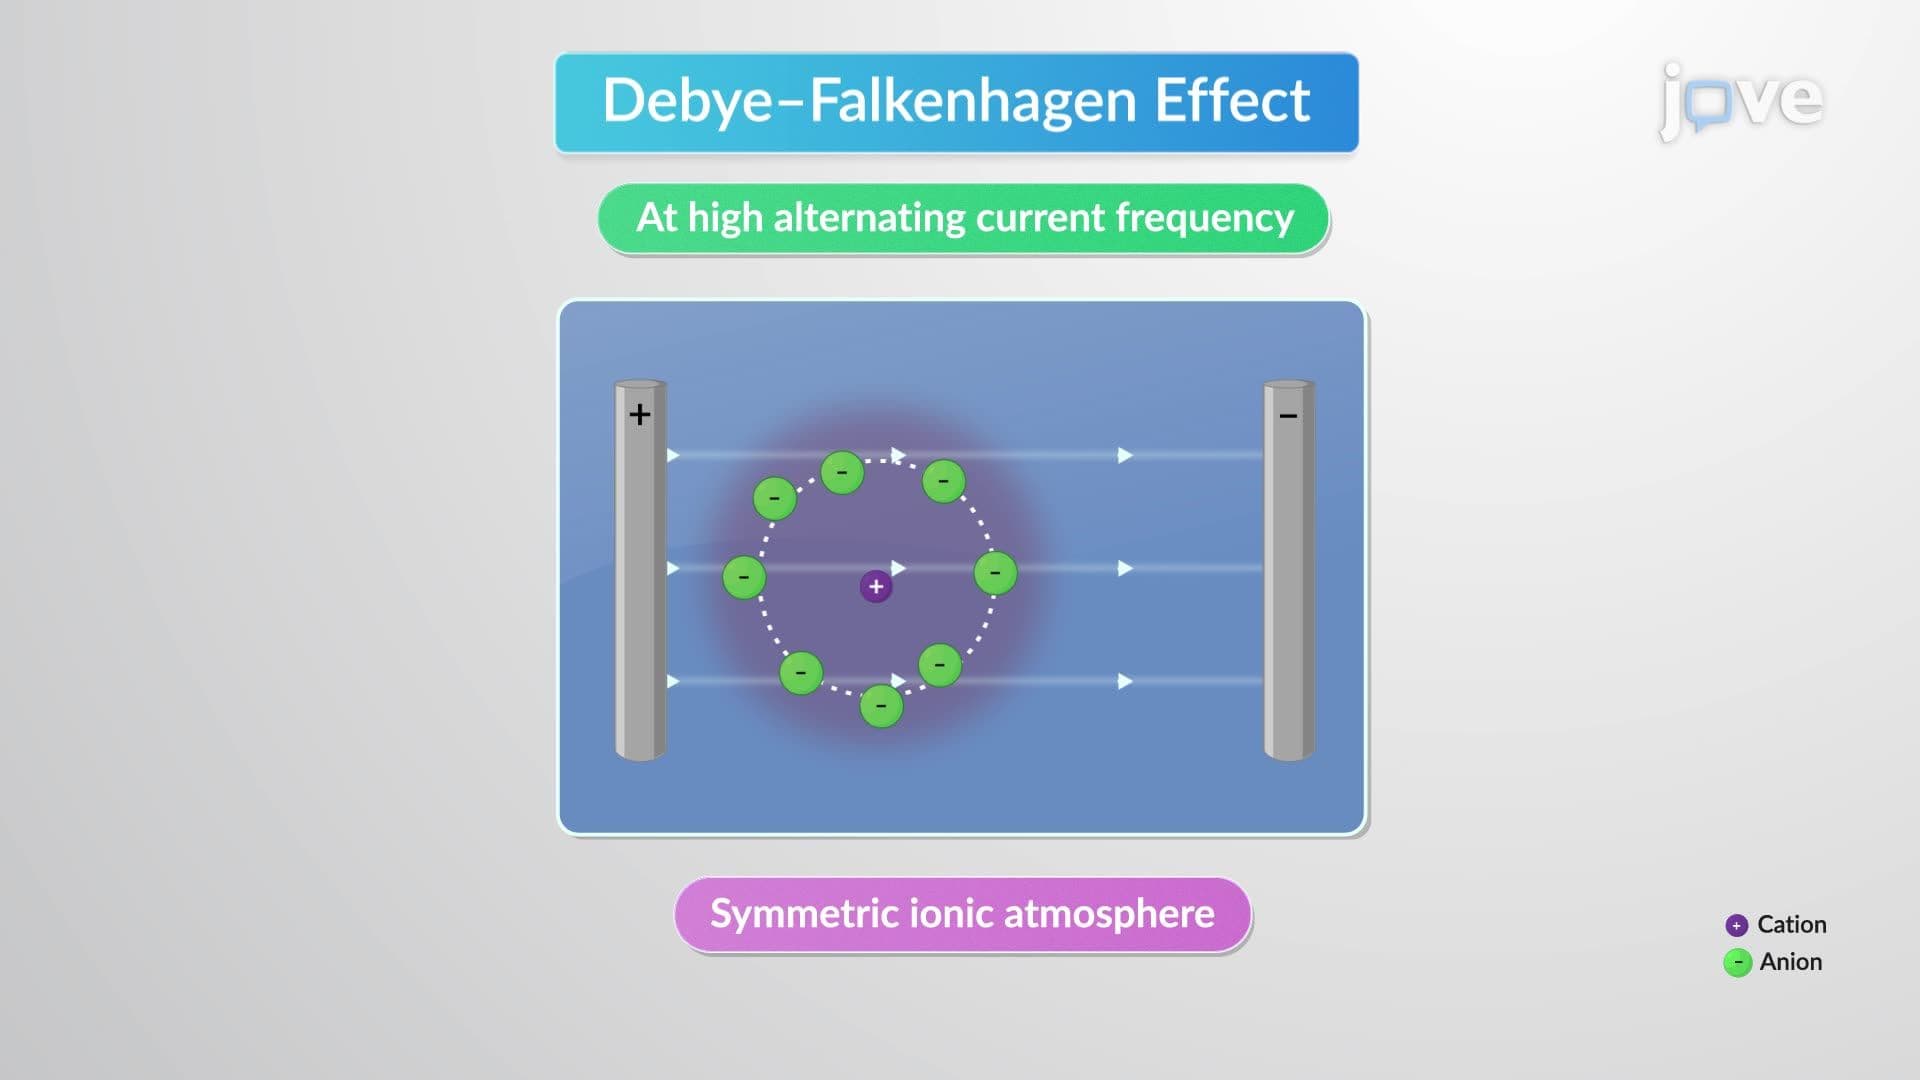
Debye–Huckel–Onsager Conductance Equation
 01:27
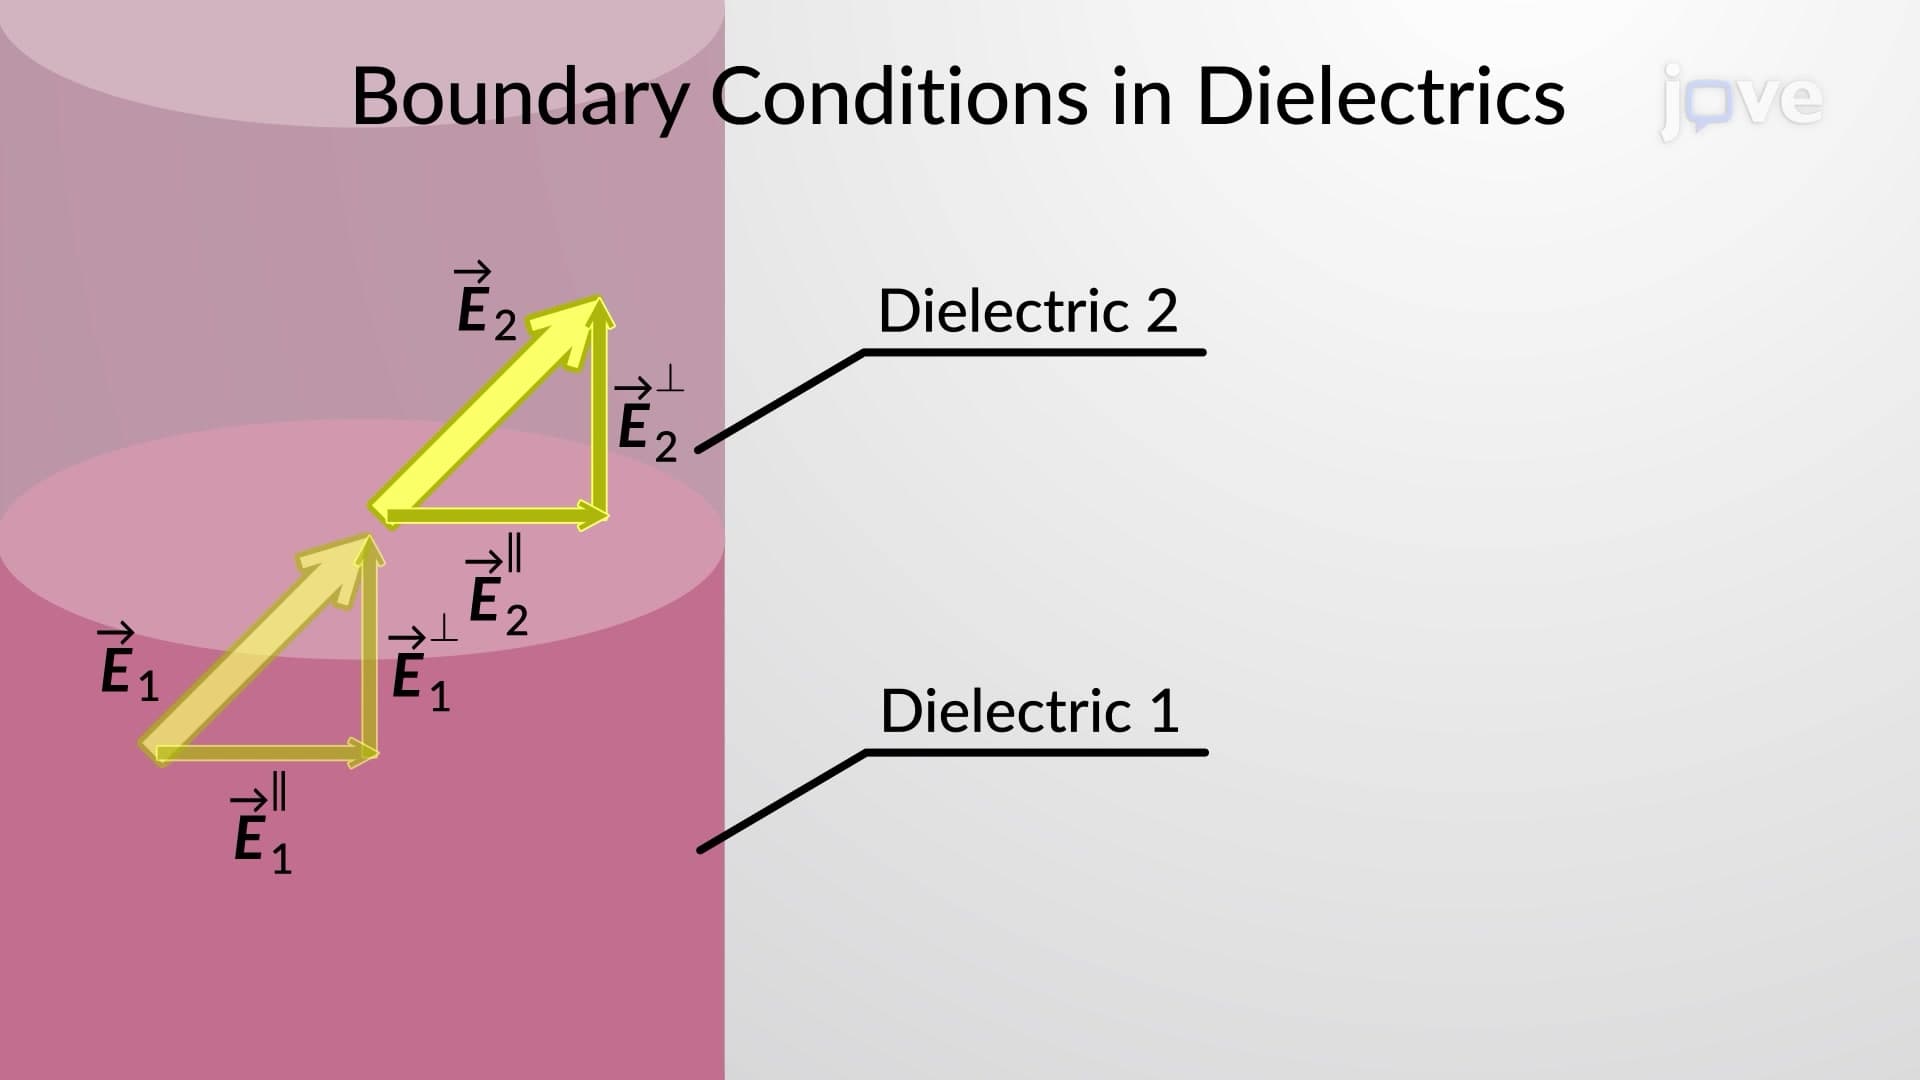
Electrostatic Boundary Conditions in Dielectrics
Consider a case where both the mediums across a boundary are two different dielectric materials. Recall that the electric field and electric displacement are proportional and related through the material's permittivity.
 01:26
Imperfections in Crystal Structure: Stoichiometric Point Defects
 01:24
Van der Waals Interactions