Related Experiment Video
Updated: May 10, 2026

Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
A simplified Tamm-Dancoff density functional approach for the electronic excitation spectra of very large molecules
1Mulliken Center for Theoretical Chemistry, Institut für Physikalische und Theoretische Chemie der Universität Bonn, Beringstr. 4, D-53115 Bonn, Germany. grimme@thch.uni-bonn.de

Abstract:
Two approximations in the Tamm-Dancoff density functional theory approach (TDA-DFT) to electronically excited states are proposed which allow routine computations for electronic ultraviolet (UV)- or circular dichroism (CD) spectra of molecules with 500-1000 atoms. Speed-ups compared to conventional time-dependent DFT (TD-DFT) treatments of about two to three orders of magnitude in the excited state part at only minor loss of accuracy are obtained. The method termed sTDA ("s" for simplified) employs atom-centered Löwdin-monopole based two-electron repulsion integrals with the asymptotically correct 1∕R behavior and perturbative single excitation configuration selection. It is formulated generally for any standard global hybrid density functional with given Fock-exchange mixing parameter ax. The method performs well for two standard benchmark sets of vertical singlet-singlet excitations for values of ax in the range 0.2-0.6. The mean absolute deviations from reference data are only 0.2-0.3 eV and similar to those from standard TD-DFT. In three cases (two dyes and one polypeptide), good mutual agreement between the electronic spectra (up to 10-11 eV excitation energy) from the sTDA method and those from TD(A)-DFT is obtained. The computed UV- and CD-spectra of a few typical systems (e.g., C60, two transition metal complexes, [7]helicene, polyalanine, a supramolecular aggregate with 483 atoms and about 7000 basis functions) compare well with corresponding experimental data. The method is proposed together with medium-sized double- or triple-zeta type atomic-orbital basis sets as a quantum chemical tool to investigate the spectra of huge molecular systems at a reliable DFT level.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Related Concept Videos
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to the...
 01:14
Molecular Spectroscopy: Absorption and Emission
 01:20
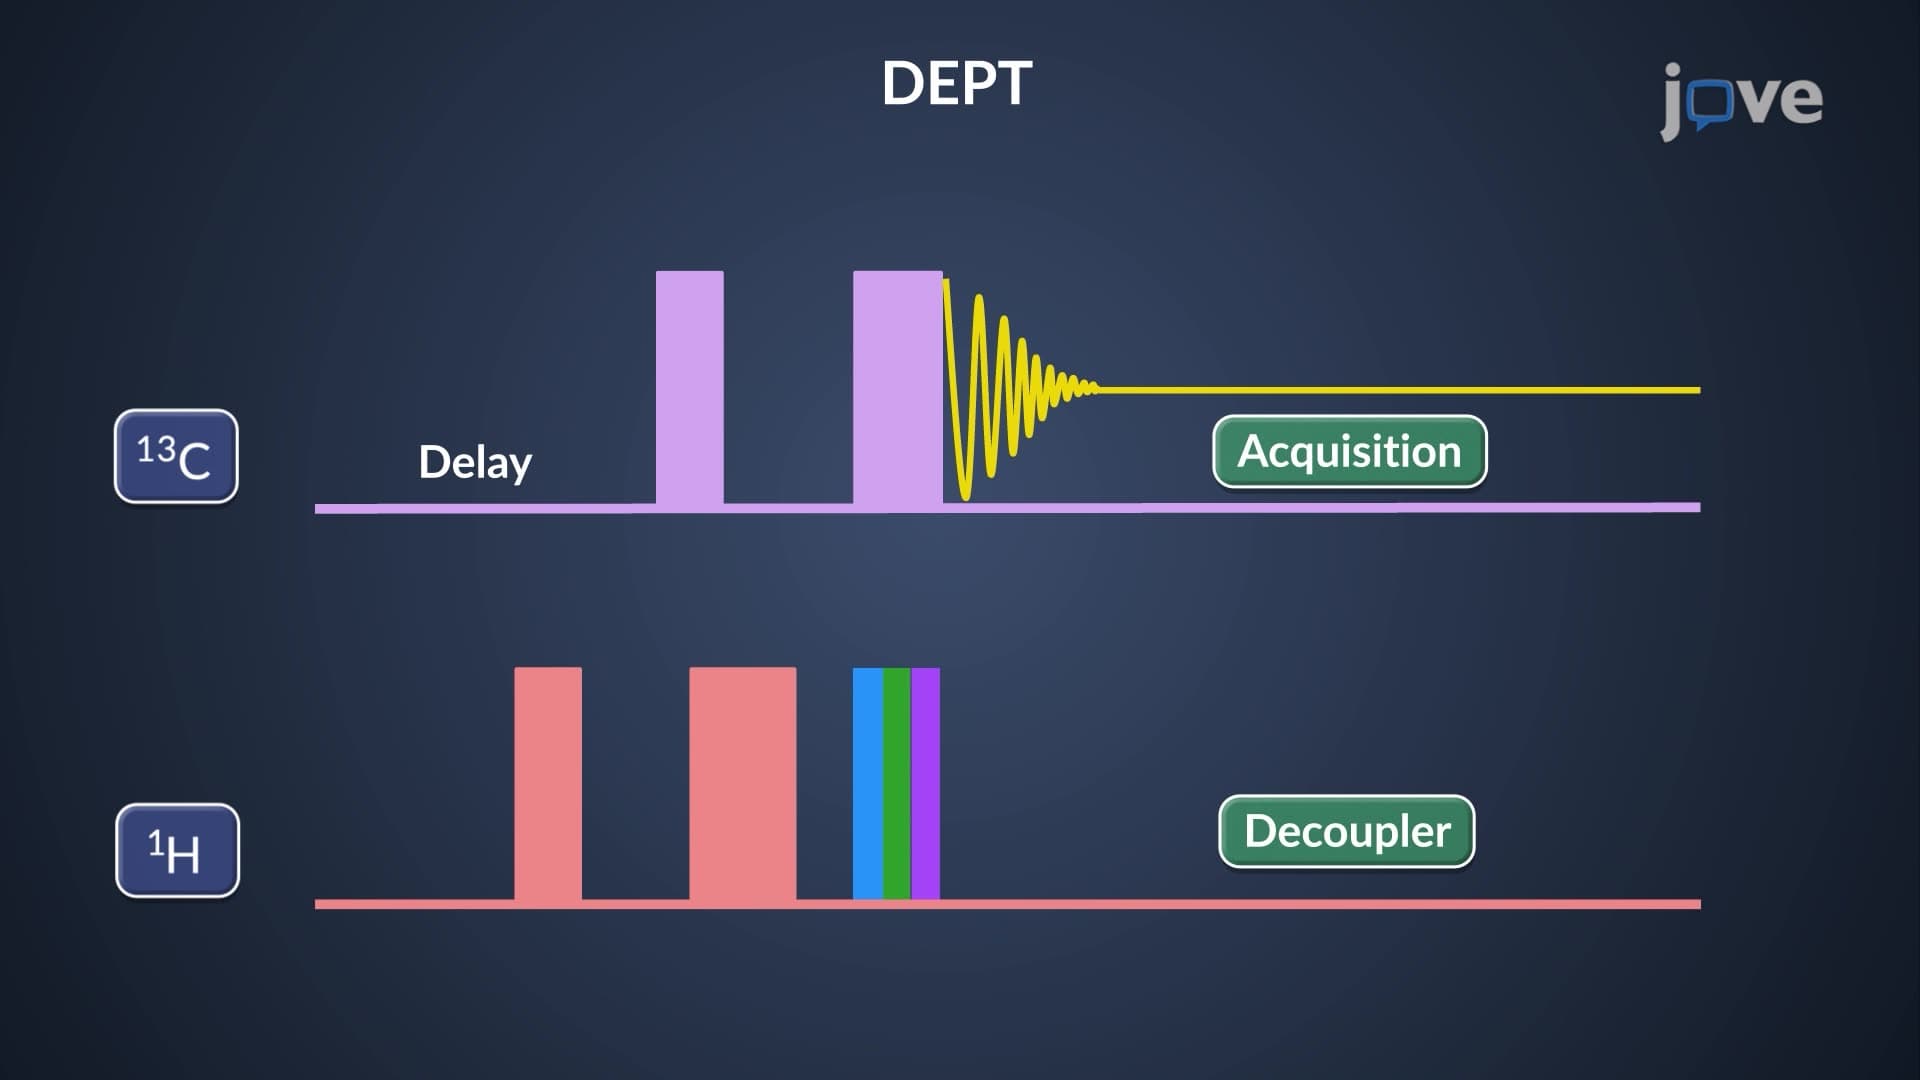
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum numbers: n, l, ml, and...