Related Experiment Video
Updated: May 9, 2026

Modeling Mitochondrial Disease Using Brain Organoids: A Focus on Mitochondrial Encephalomyopathy, Lactic Acidosis, and Stroke-like Episodes
Published on: October 10, 2025
Novel mutations m.3959G>A and m.3995A>G in mitochondrial gene MT-ND1 associated with MELAS
Jie Lin1, Chong-Bo Zhao, Jia-Hong Lu
1Department of Neurology, Huashan Hospital .

Abstract:
Mitochondrial encephalopathy, lactic acidosis and stroke-like episodes (MELAS) are progressive neurodegenerative disorder associated with polygenetic, maternally inherited mutations in mitochondrial DNA. Approximately 80% of MELAS cases are caused by the mutation m.3243A>G of the mitochondrial tRNA(Leu (UUR)) gene (MT-TL1). We reported two probands with MELAS features. Muscle biopsy identified ragged-red fibers (RRF) in Gomori Trichrome staining. A respiratory chain function study showed decreased activity of mitochondrial respiratory chain complex I in both probands. Sequencing of the mitochondrial DNA revealed two novel MT-ND1 gene missense mutations, m.3959G>A and m.3995A>G, which are highly conserved among species. Protein secondary structure predictions demonstrated that these mutations may alter the peptide structure and may lead to decreased ND1 gene stability. Our findings suggest that these two novel mutations may contribute to the MELAS phenotypes of the patients in our study.
Insights
Two novel mitochondrial DNA mutations, m.3959G>A and m.3995A>G in the MT-ND1 gene, are linked to MELAS (Mitochondrial Encephalopathy, Lactic Acidosis, and Stroke-like episodes) symptoms. These findings expand the genetic understanding of this progressive neurodegenerative disorder.
Area of Science:
- Genetics
- Neuroscience
- Mitochondrial Biology
Background:
- Mitochondrial Encephalopathy, Lactic Acidosis, and Stroke-like episodes (MELAS) is a progressive neurodegenerative disorder.
- It is typically caused by mutations in mitochondrial DNA, with m.3243A>G in MT-TL1 being the most common (approx. 80% of cases).
Observation:
- Two patients presented with MELAS features.
- Muscle biopsies revealed ragged-red fibers (RRF).
- Respiratory chain function studies indicated decreased mitochondrial respiratory chain complex I activity.
Findings:
- Sequencing identified two novel missense mutations in the MT-ND1 gene: m.3959G>A and m.3995A>G.
- These mutations are highly conserved across species.
- In silico analysis suggests these mutations may destabilize the ND1 protein structure.
Implications:
- The identified MT-ND1 mutations are potential contributors to the MELAS phenotype in these patients.
- This expands the known genetic landscape of MELAS.
- Further research into these novel mutations could refine diagnostic approaches and therapeutic strategies for MELAS.
Related Concept Videos
 02:59
Animal Mitochondrial Genetics
 01:19
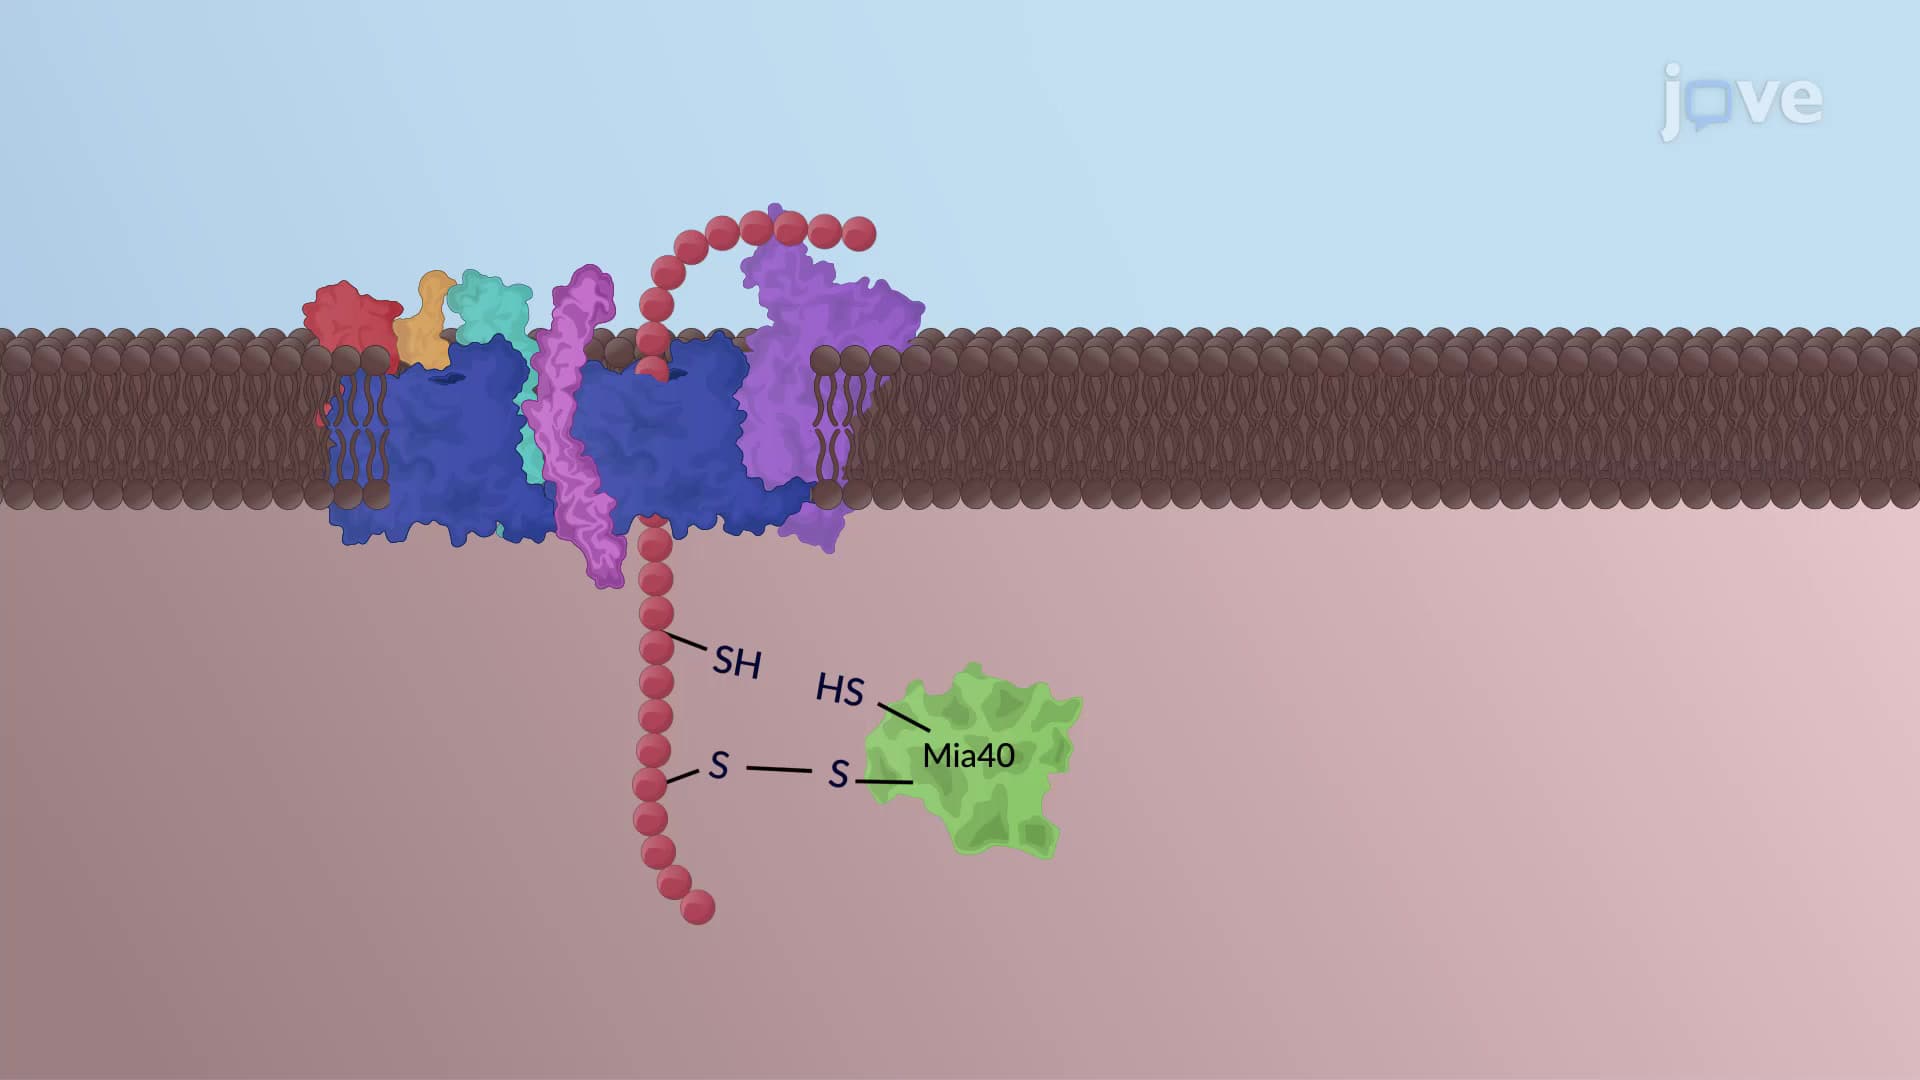
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:39
Mitochondrial Precursor Proteins
Most of the mitochondrial precursors...
 01:45
Mitochondrial Membranes
 01:48
ATP Synthase: Mechanism
 01:39
Mitochondrial Protein Sorting
Most of these mitochondrial proteins are encoded by the nucleus and imported to the mitochondria as unfolded or loosely folded precursors. Mitochondrial precursors...