Related Experiment Video
Updated: May 9, 2026
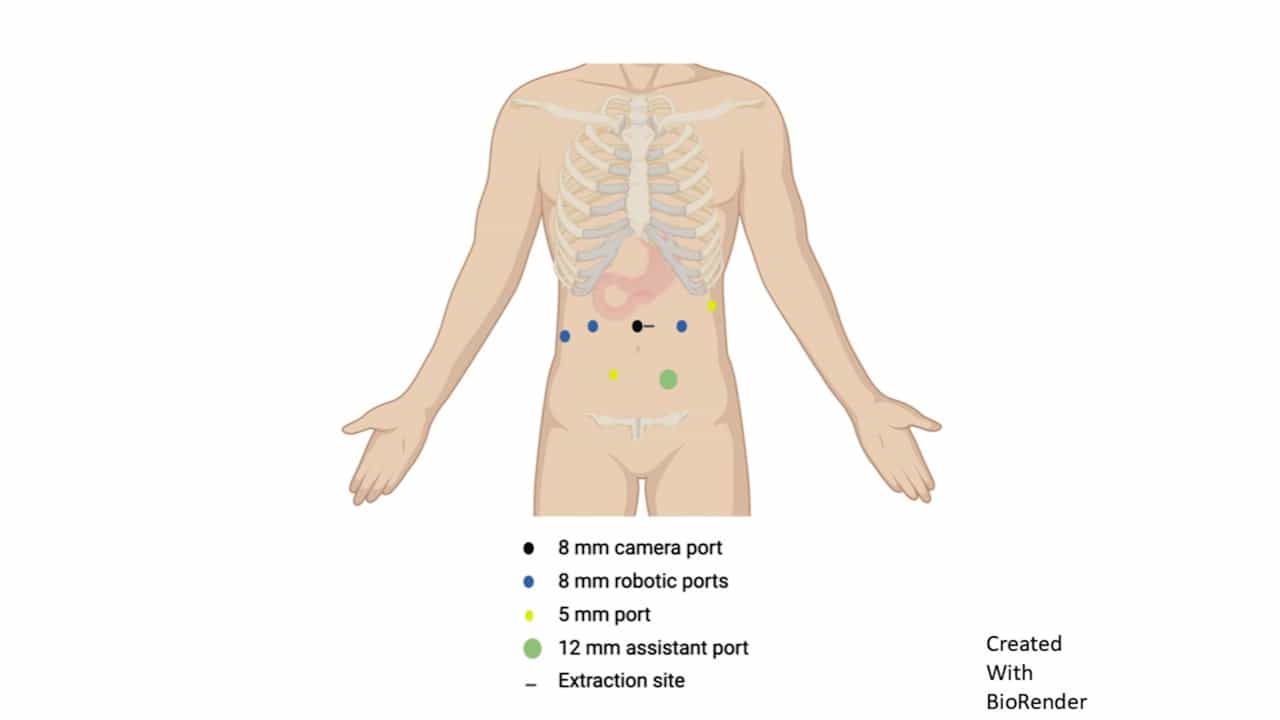

Robotic Duodenal Sleeve Resection for Gastrointestinal Stromal Tumor with Rare Exon 8 KIT Mutation Following Neoadjuvant Imatinib
Published on: April 3, 2026
Selecting tyrosine kinase inhibitors for gastrointestinal stromal tumor with secondary KIT activation-loop domain
Yuan-Shuo Hsueh1, Chih-Lung Lin, Nai-Jung Chiang
1National Institute of Cancer Research, National Health Research Institutes, Tainan, Taiwan.

Abstract:
Advanced gastrointestinal stromal tumors (GIST), a KIT oncogene-driven tumor, on imatinib mesylate (IM) treatment may develop secondary KIT mutations to confer IM-resistant phenotype. Second-line sunitinib malate (SU) therapy is largely ineffective for IM-resistant GISTs with secondary exon 17 (activation-loop domain) mutations. We established an in vitro cell-based platform consisting of a series of COS-1 cells expressing KIT cDNA constructs encoding common primary±secondary mutations observed in GISTs, to compare the activity of several commercially available tyrosine kinase inhibitors on inhibiting the phosphorylation of mutant KIT proteins at their clinically achievable plasma steady-state concentration (Css). The inhibitory efficacies on KIT exon 11/17 mutants were further validated by growth inhibition assay on GIST48 cells, and underlying molecular-structure mechanisms were investigated by molecular modeling. Our results showed that SU more effectively inhibited mutant KIT with secondary exon 13 or 14 mutations than those with secondary exon 17 mutations, as clinically indicated. On contrary, at individual Css, nilotinib and sorafenib more profoundly inhibited the phosphorylation of KIT with secondary exon 17 mutations and the growth of GIST48 cells than IM, SU, and dasatinib. Molecular modeling analysis showed fragment deletion of exon 11 and point mutation on exon 17 would lead to a shift of KIT conformational equilibrium toward active form, for which nilotinib and sorafenib bound more stably than IM and SU. In current preclinical study, nilotinib and sorafenib are more active in IM-resistant GISTs with secondary exon 17 mutation than SU that deserve further clinical investigation.
Insights
Nilotinib and sorafenib show promise against imatinib-resistant gastrointestinal stromal tumors (GIST) with specific KIT mutations. These drugs are more effective than sunitinib for KIT exon 17 mutations, warranting further clinical study.
Area of Science:
- Oncology
- Molecular Biology
- Pharmacology
Background:
- Advanced gastrointestinal stromal tumors (GIST) often develop imatinib resistance due to secondary KIT mutations.
- Sunitinib is largely ineffective for GIST with secondary exon 17 KIT mutations, a common resistance mechanism.
Purpose of the Study:
- To compare the efficacy of various tyrosine kinase inhibitors (TKIs) against imatinib-resistant GIST with specific secondary KIT mutations.
- To investigate the molecular mechanisms underlying TKI activity in GIST.
Main Methods:
- Established an in vitro cell-based platform using COS-1 cells expressing various KIT mutations.
- Assessed TKI inhibition of mutant KIT phosphorylation at clinically relevant concentrations.
- Validated findings using GIST48 cell growth inhibition assays and molecular modeling.
Main Results:
- Sunitinib showed greater efficacy against KIT exon 13/14 mutants than exon 17 mutants.
- Nilotinib and sorafenib demonstrated superior inhibition of KIT exon 17 mutants and GIST cell growth compared to imatinib and sunitinib.
- Molecular modeling revealed nilotinib and sorafenib bind more stably to altered KIT structures associated with exon 17 mutations.
Conclusions:
- Nilotinib and sorafenib are more active against imatinib-resistant GIST with secondary exon 17 mutations than sunitinib.
- These findings support further clinical investigation of nilotinib and sorafenib for refractory GIST.
Related Concept Videos
 02:57
Targeted Cancer Therapies
There are several types of targeted therapies against specific...
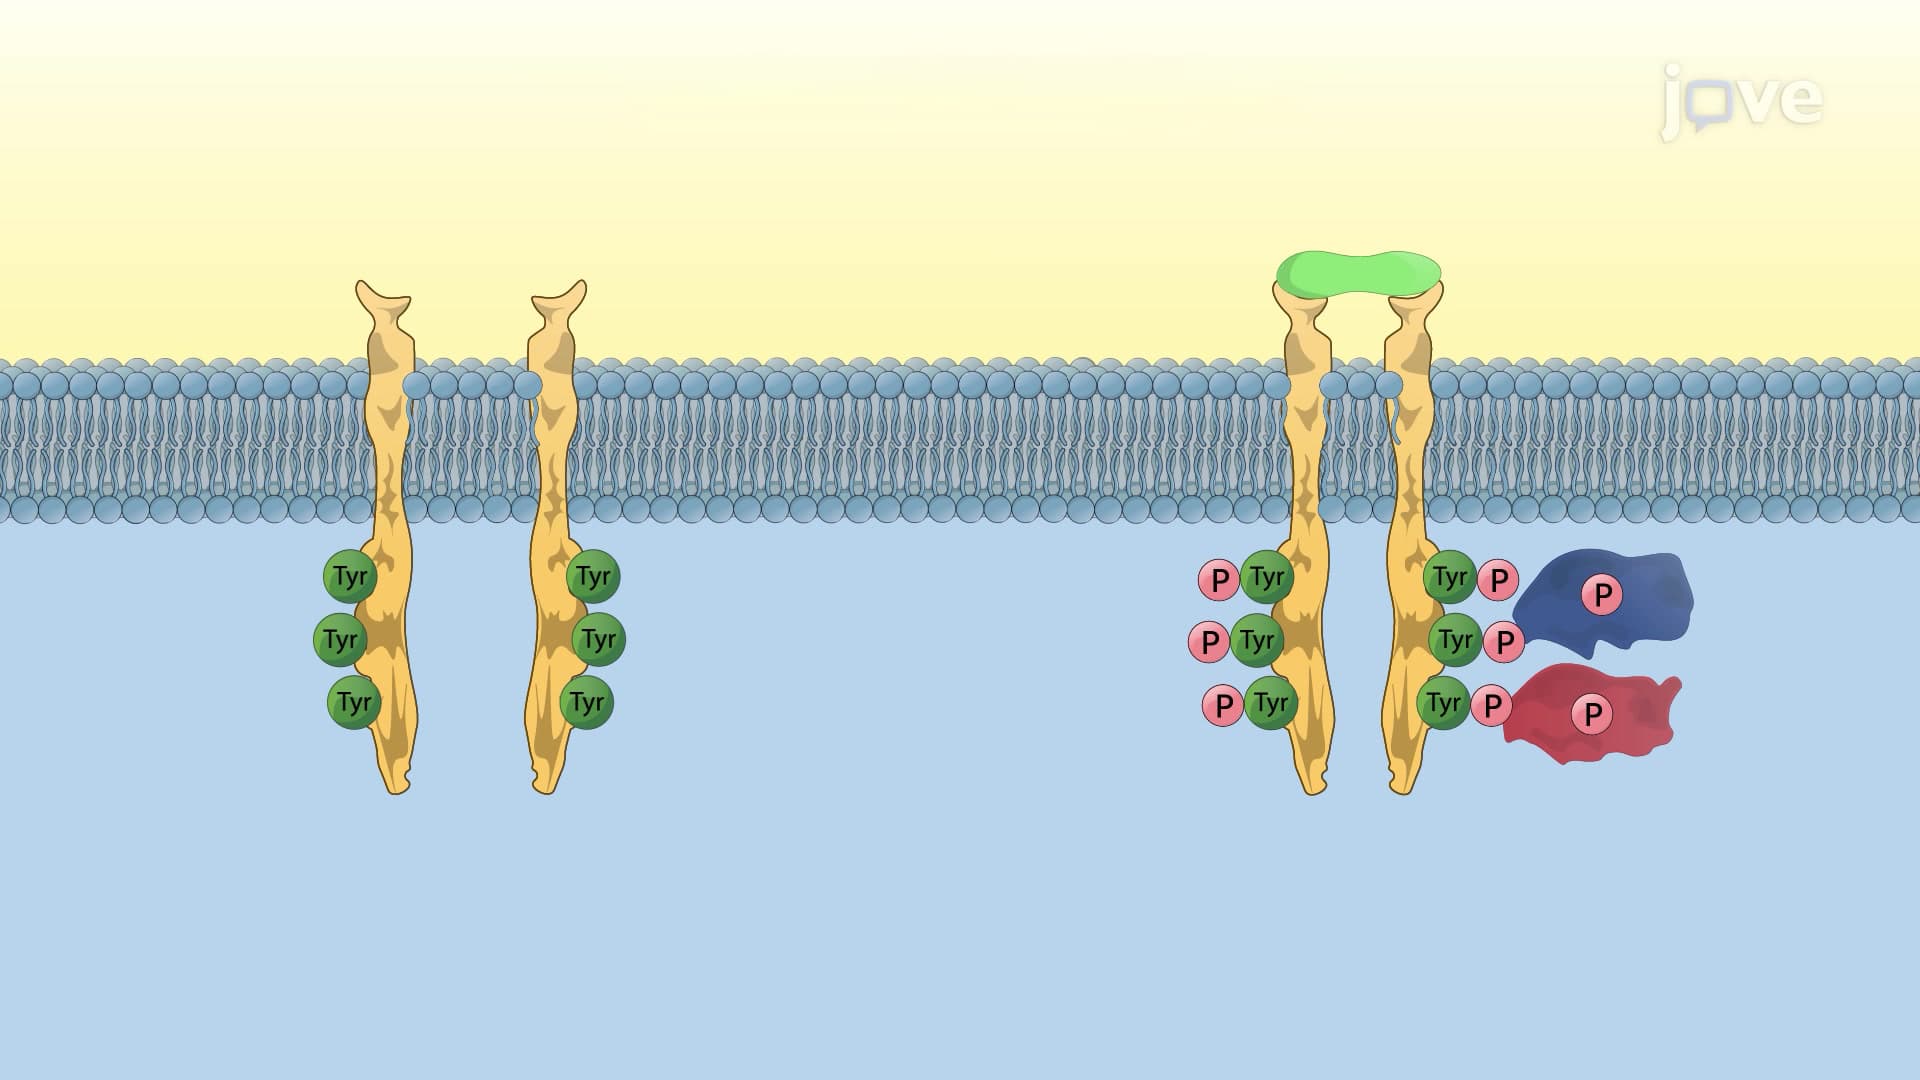 01:26
Receptor Tyrosine Kinases
 01:27
Transducer Mechanism: Enzyme-Linked Receptors
Major types that are helpful drug targets include:
 01:11
Pharmacogenetics of Drug Targets: β₂-Adrenergic Receptors, Apo E, Thymidylate Synthase