Related Experiment Video
Updated: May 9, 2026

Quantitative Atomic-Site Analysis of Functional Dopants/Point Defects in Crystalline Materials by Electron-Channeling-Enhanced Microanalysis
Published on: May 10, 2021
Examination of oxygen vacancy formation in Mn-doped CeO2 (111) using DFT+U and the hybrid functional HSE06
Matthew D Krcha1, Michael J Janik
1Department of Chemical Engineering, Pennsylvania State University, University Park, Pennsylvania 16802, United States.

Abstract:
MnO(x)-CeO(x) mixed oxide systems exhibit interesting sulfur adsorption capacities and catalytic activity. We examined the electronic structure of Mn-doped fluorite CeO2 bulk solid and surface using density functional theory (DFT) with the Hubbard U term or the Heyd-Scuseria-Ernzerhof (HSE06) hybrid functional. We specifically evaluate the reducibility and formation energies of Mn-doped CeO2 surfaces. The use of a U value on the d-states of Mn is examined, and a value of 4 eV is chosen based upon results from DFT+U calculations on bulk MnO(x),1 XANES characterization of oxidation states in calcined and reduced Mn-doped CeO2, and comparison with HSE06 hybrid functional results. Electronic structure impacts of the U inclusion are discussed. The concentration and orientation of Mn atoms doped into the surface of CeO2 have a great influence on the reducibility of the surface. Based upon formation energies, Mn will not favor doping into the surface of CeO2 in a fully oxidized system (Mn(4+)). Under reducing environments, Mn will dope into the surface with oxygen vacancies present (Mn(3+) and Mn(2+)). The first oxygen vacancy is not likely catalytically important in fluorite MnO(x)-CeO(x) systems as formation of the fully oxidized surface is not stable. A greater degree of reduction would occur during a catalyzed redox reaction.
Related Concept Videos
 02:42
Valence Bond Theory
 01:29
Imperfections in Crystal Structure: Non-Stoichiometric Defects
 01:26
Imperfections in Crystal Structure: Stoichiometric Point Defects
 03:26
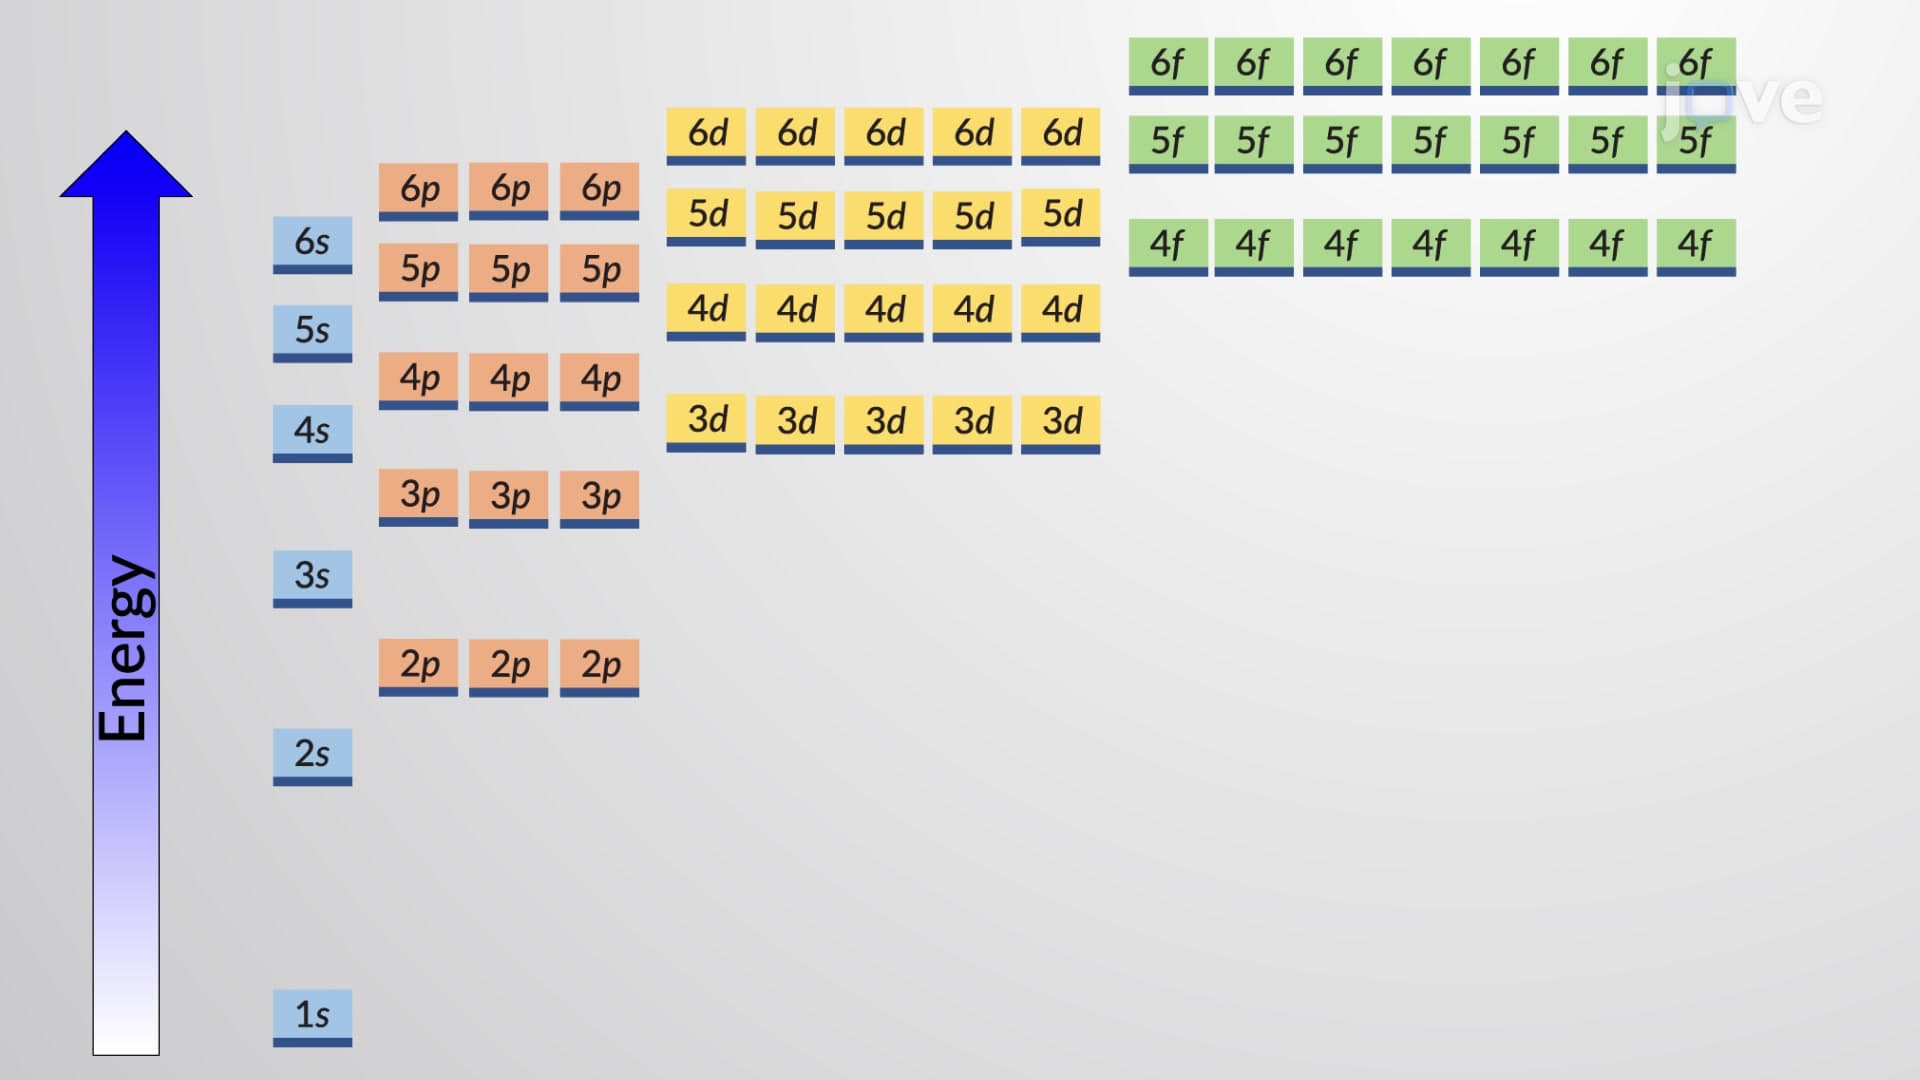
Electron Configuration of Multielectron Atoms
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...