Related Experiment Video
Updated: May 9, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
A big data approach to the ultra-fast prediction of DFT-calculated bond energies
Xiaohui Qu1, Diogo Ars Latino2, Joao Aires-de-Sousa1
1CQFB and REQUIMTE, Departamento de Química, Faculdade de Ciências e Tecnologia, Universidade Nova de Lisboa, Caparica 2829-516, Portugal.

Background:
The rapid access to intrinsic physicochemical properties of molecules is highly desired for large scale chemical data mining explorations such as mass spectrum prediction in metabolomics, toxicity risk assessment and drug discovery. Large volumes of data are being produced by quantum chemistry calculations, which provide increasing accurate estimations of several properties, e.g. by Density Functional Theory (DFT), but are still too computationally expensive for those large scale uses. This work explores the possibility of using large amounts of data generated by DFT methods for thousands of molecular structures, extracting relevant molecular properties and applying machine learning (ML) algorithms to learn from the data. Once trained, these ML models can be applied to new structures to produce ultra-fast predictions. An approach is presented for homolytic bond dissociation energy (BDE).
Results:
Machine learning models were trained with a data set of >12,000 BDEs calculated by B3LYP/6-311++G(d,p)//DFTB. Descriptors were designed to encode atom types and connectivity in the 2D topological environment of the bonds. The best model, an Associative Neural Network (ASNN) based on 85 bond descriptors, was able to predict the BDE of 887 bonds in an independent test set (covering a range of 17.67-202.30 kcal/mol) with RMSD of 5.29 kcal/mol, mean absolute deviation of 3.35 kcal/mol, and R (2) = 0.953. The predictions were compared with semi-empirical PM6 calculations, and were found to be superior for all types of bonds in the data set, except for O-H, N-H, and N-N bonds. The B3LYP/6-311++G(d,p)//DFTB calculations can approach the higher-level calculations B3LYP/6-311++G(3df,2p)//B3LYP/6-31G(d,p) with an RMSD of 3.04 kcal/mol, which is less than the RMSD of ASNN (against both DFT methods). An experimental web service for on-line prediction of BDEs is available at http://joao.airesdesousa.com/bde.
Conclusion:
Knowledge could be automatically extracted by machine learning techniques from a data set of calculated BDEs, providing ultra-fast access to accurate estimations of DFT-calculated BDEs. This demonstrates how to extract value from large volumes of data currently being produced by quantum chemistry calculations at an increasing speed mostly without human intervention. In this way, high-level theoretical quantum calculations can be used in large-scale applications that otherwise would not afford the intrinsic computational cost.
More Related Videos
 05:37
Rapid in-silico Battery Electrolyte Electrochemical Reaction Generation using 3T-VASP Multi-Scale Energy Minimization
Published on: August 22, 2025
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 02:49
Bond Energies and Bond Lengths
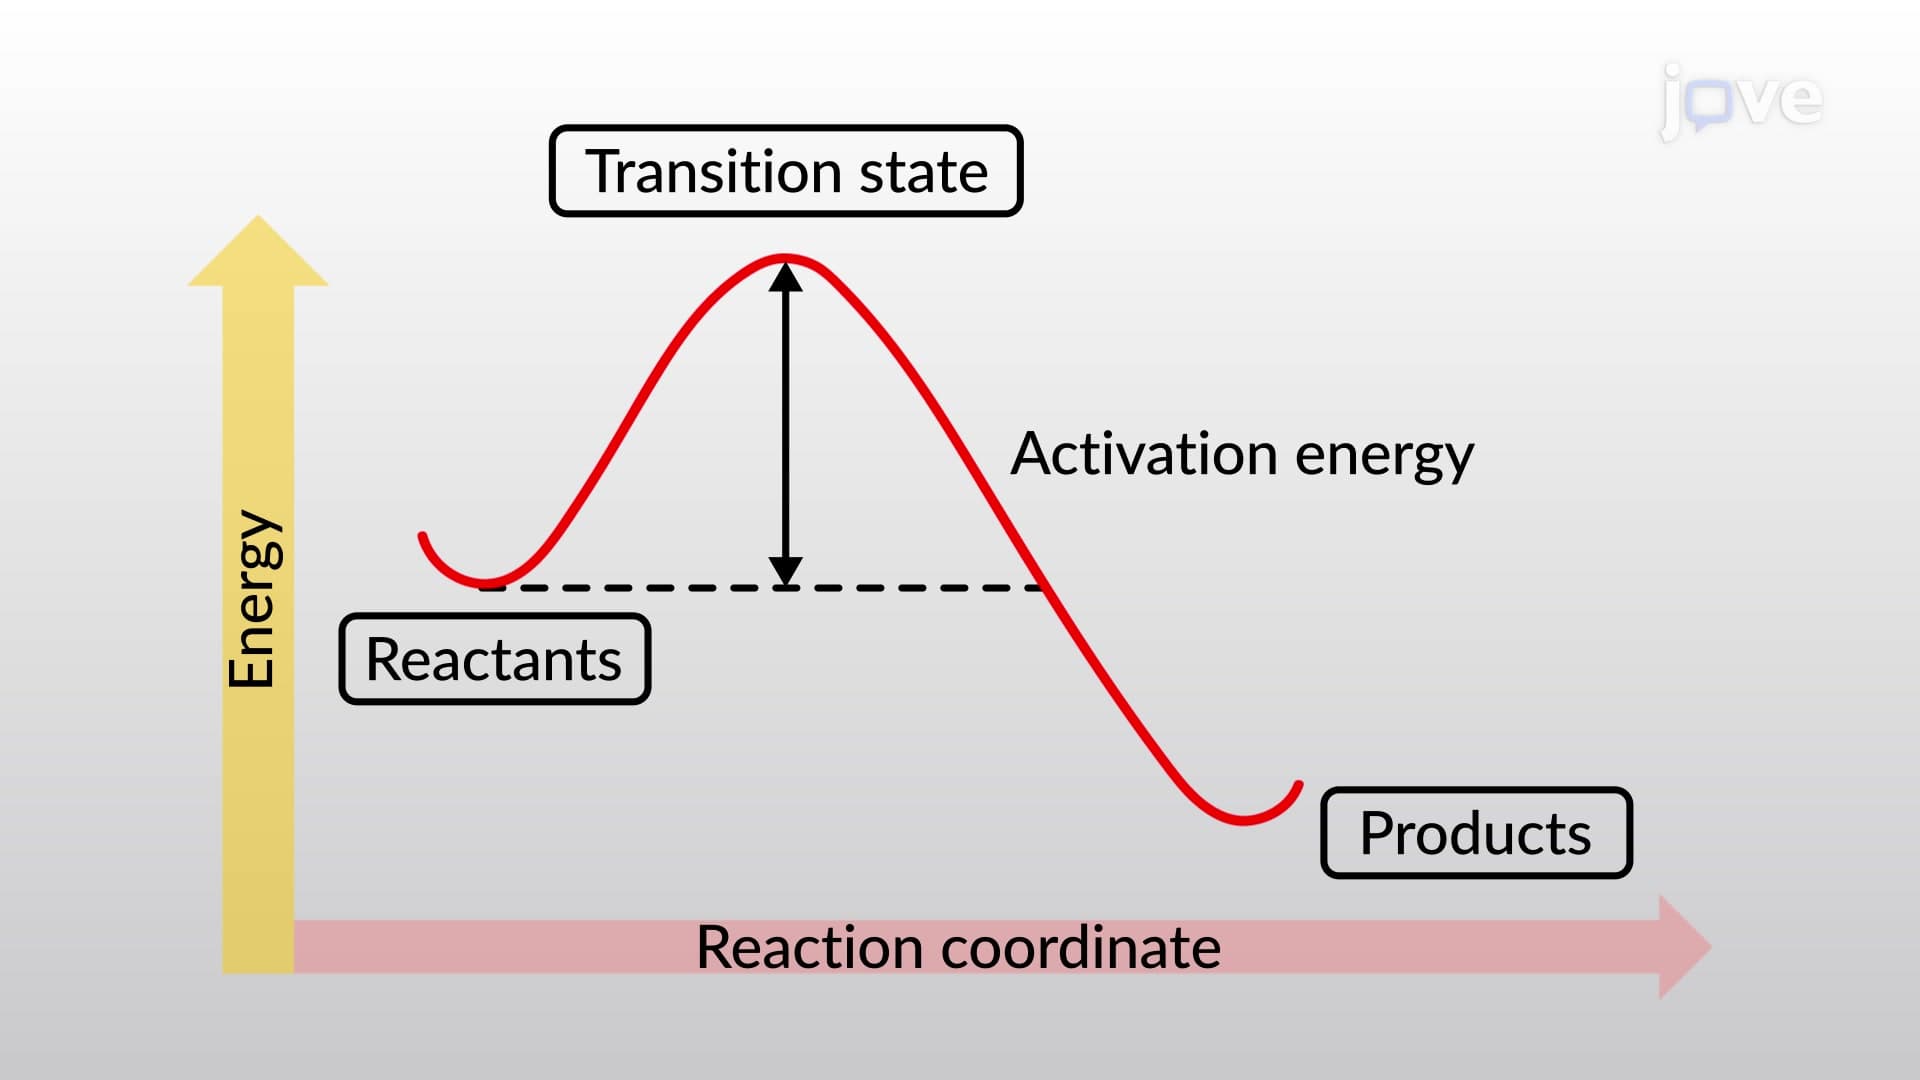 02:13
Bond Dissociation Energy and Activation Energy
 02:27
Predicting Molecular Geometry
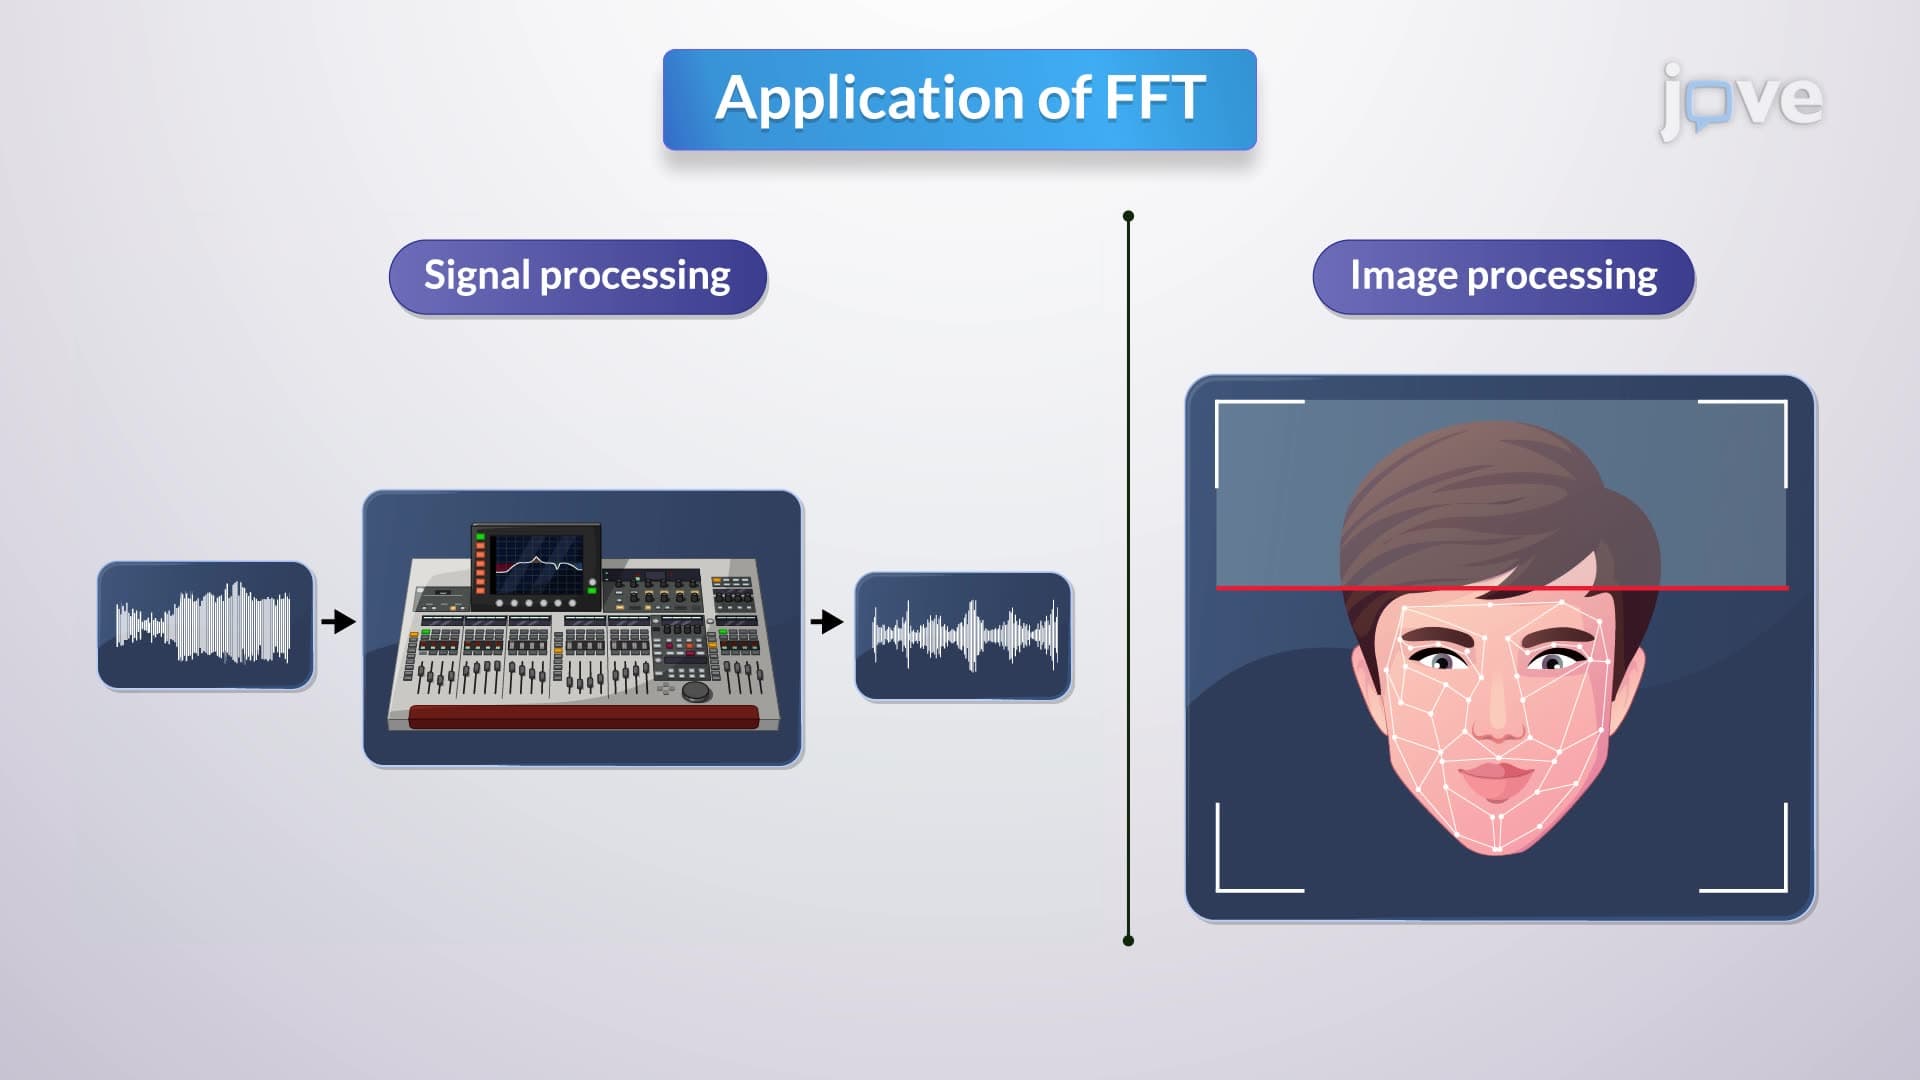 01:10
Fast Fourier Transform
The computational efficiency of the FFT becomes...
 01:20
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to the...