Related Experiment Video
Updated: May 9, 2026

Time-resolved ElectroSpray Ionization Hydrogen-deuterium Exchange Mass Spectrometry for Studying Protein Structure and Dynamics
Published on: April 17, 2017
Hydrogen/deuterium exchange mass spectrometry for probing higher order structure of protein therapeutics: methodology
Hui Wei1, Jingjie Mo1, Li Tao2
1Bioanalytical and Discovery Analytical Sciences, Research and Development, Bristol-Myers Squibb, Princeton, NJ, USA.

Abstract:
The higher order structure of protein therapeutics can be interrogated with hydrogen/deuterium exchange mass spectrometry (HDX-MS). HDX-MS is now a widely used tool in the structural characterization of protein therapeutics. In this review, HDX-MS based workflows designed for protein therapeutic discovery and development processes are presented, focusing on the specific applications of epitope mapping for protein/drug interactions and biopharmaceutical comparability studies. Future trends in the application of HDX-MS in protein therapeutics characterization are also described.
More Related Videos
 11:32
A Hydrogen-Deuterium Exchange Mass Spectrometry (HDX-MS) Platform for Investigating Peptide Biosynthetic Enzymes
Published on: May 4, 2020
 08:40
Millisecond Hydrogen/Deuterium-Exchange Mass Spectrometry for the Study of Alpha-Synuclein Structural Dynamics Under Physiological Conditions
Published on: June 23, 2022
Related Concept Videos
 01:20
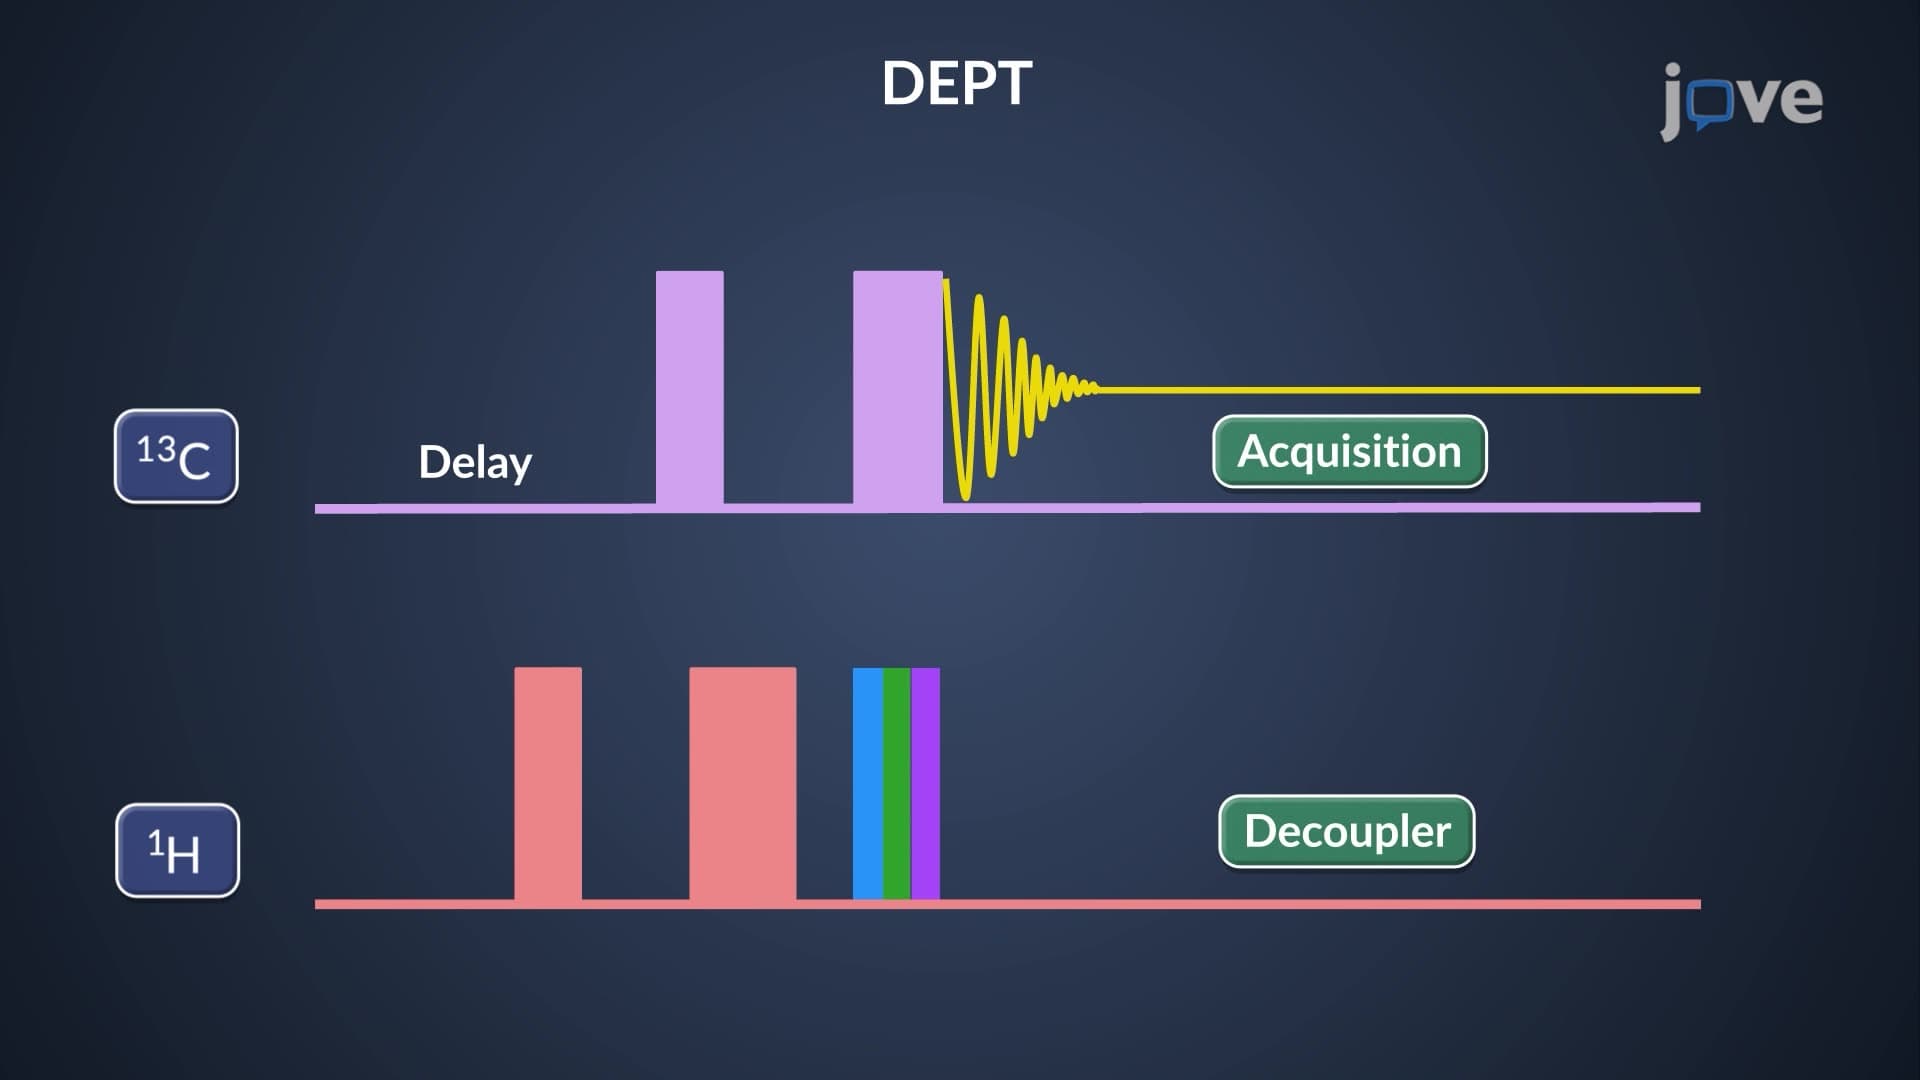
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 00:48
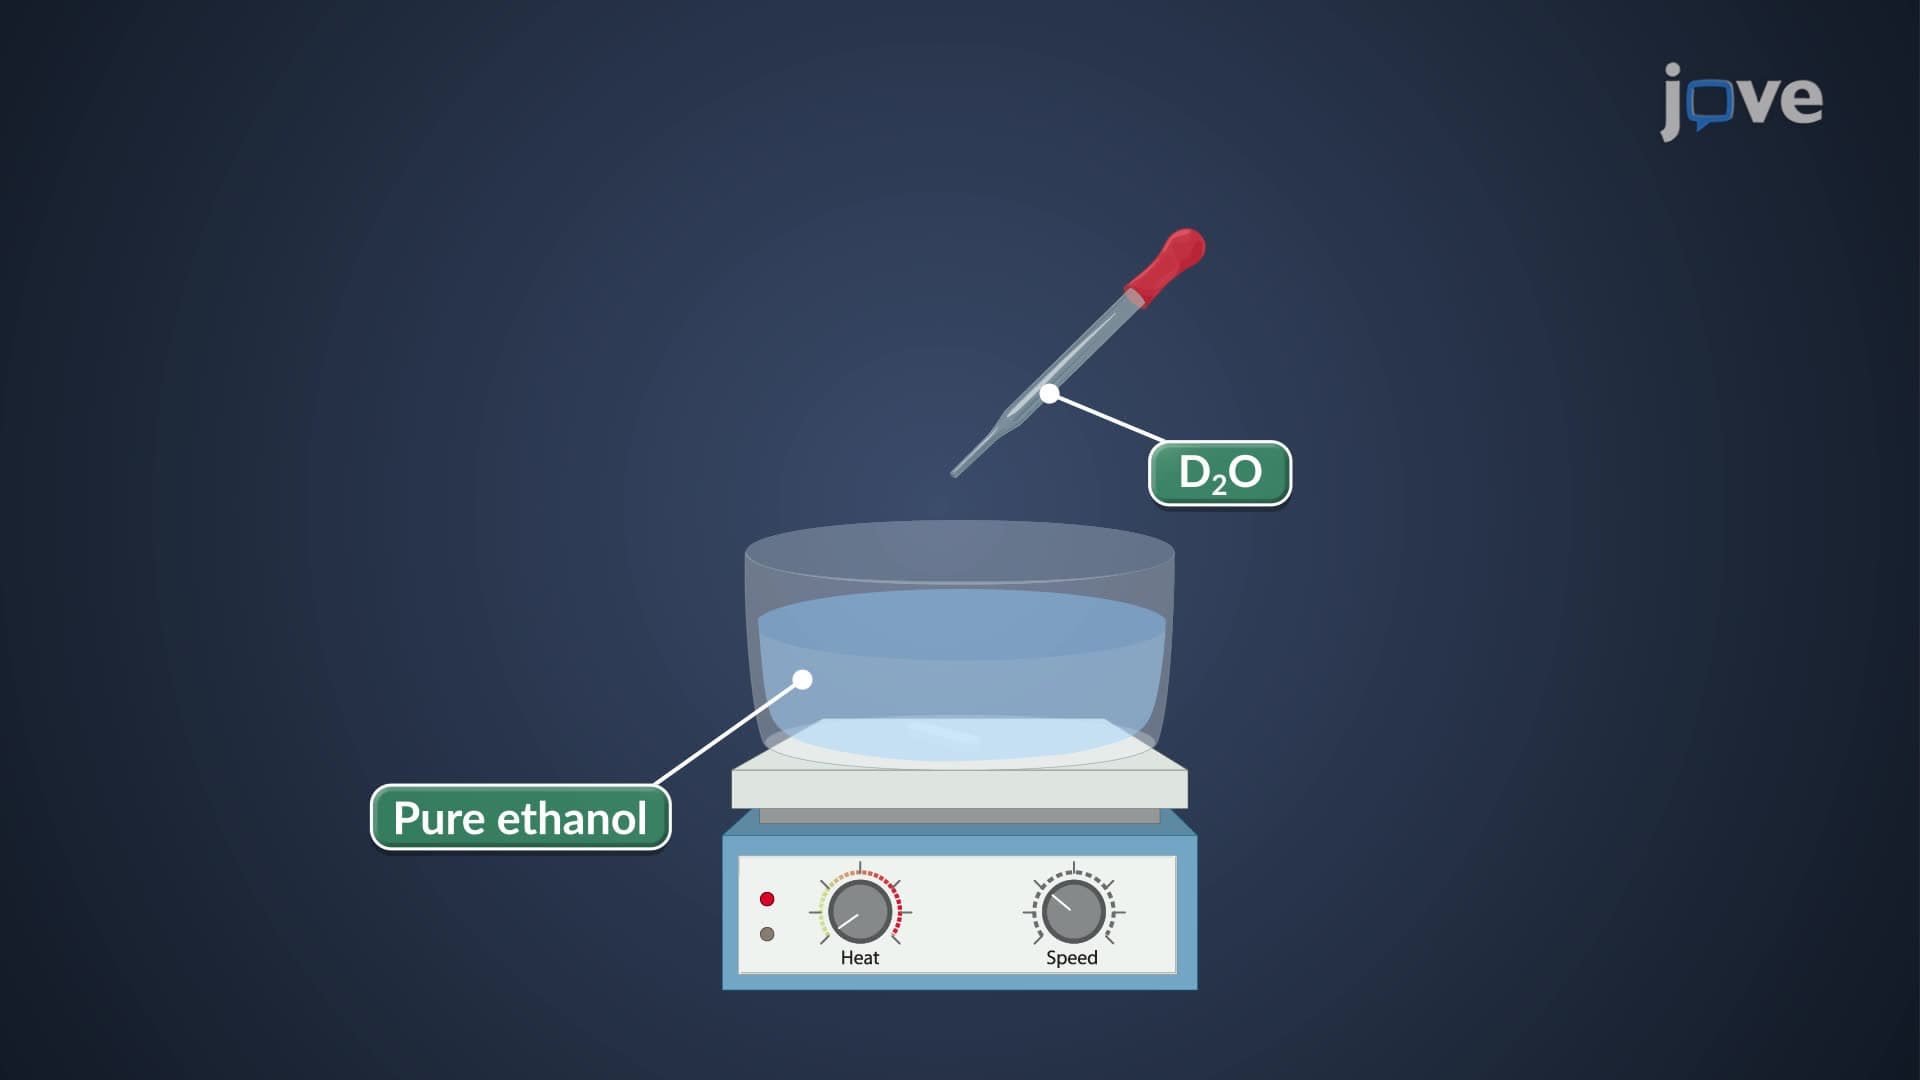
¹H NMR of Labile Protons: Deuterium (²H) Substitution
 01:33
Peptide Identification Using Tandem Mass Spectrometry
This technique helps gather information regarding the protein from which the peptide was obtained and to study the peptides’ amino acid sequence. Identifying peptides from a complex mixture is an important component of the growing field of...
 01:15
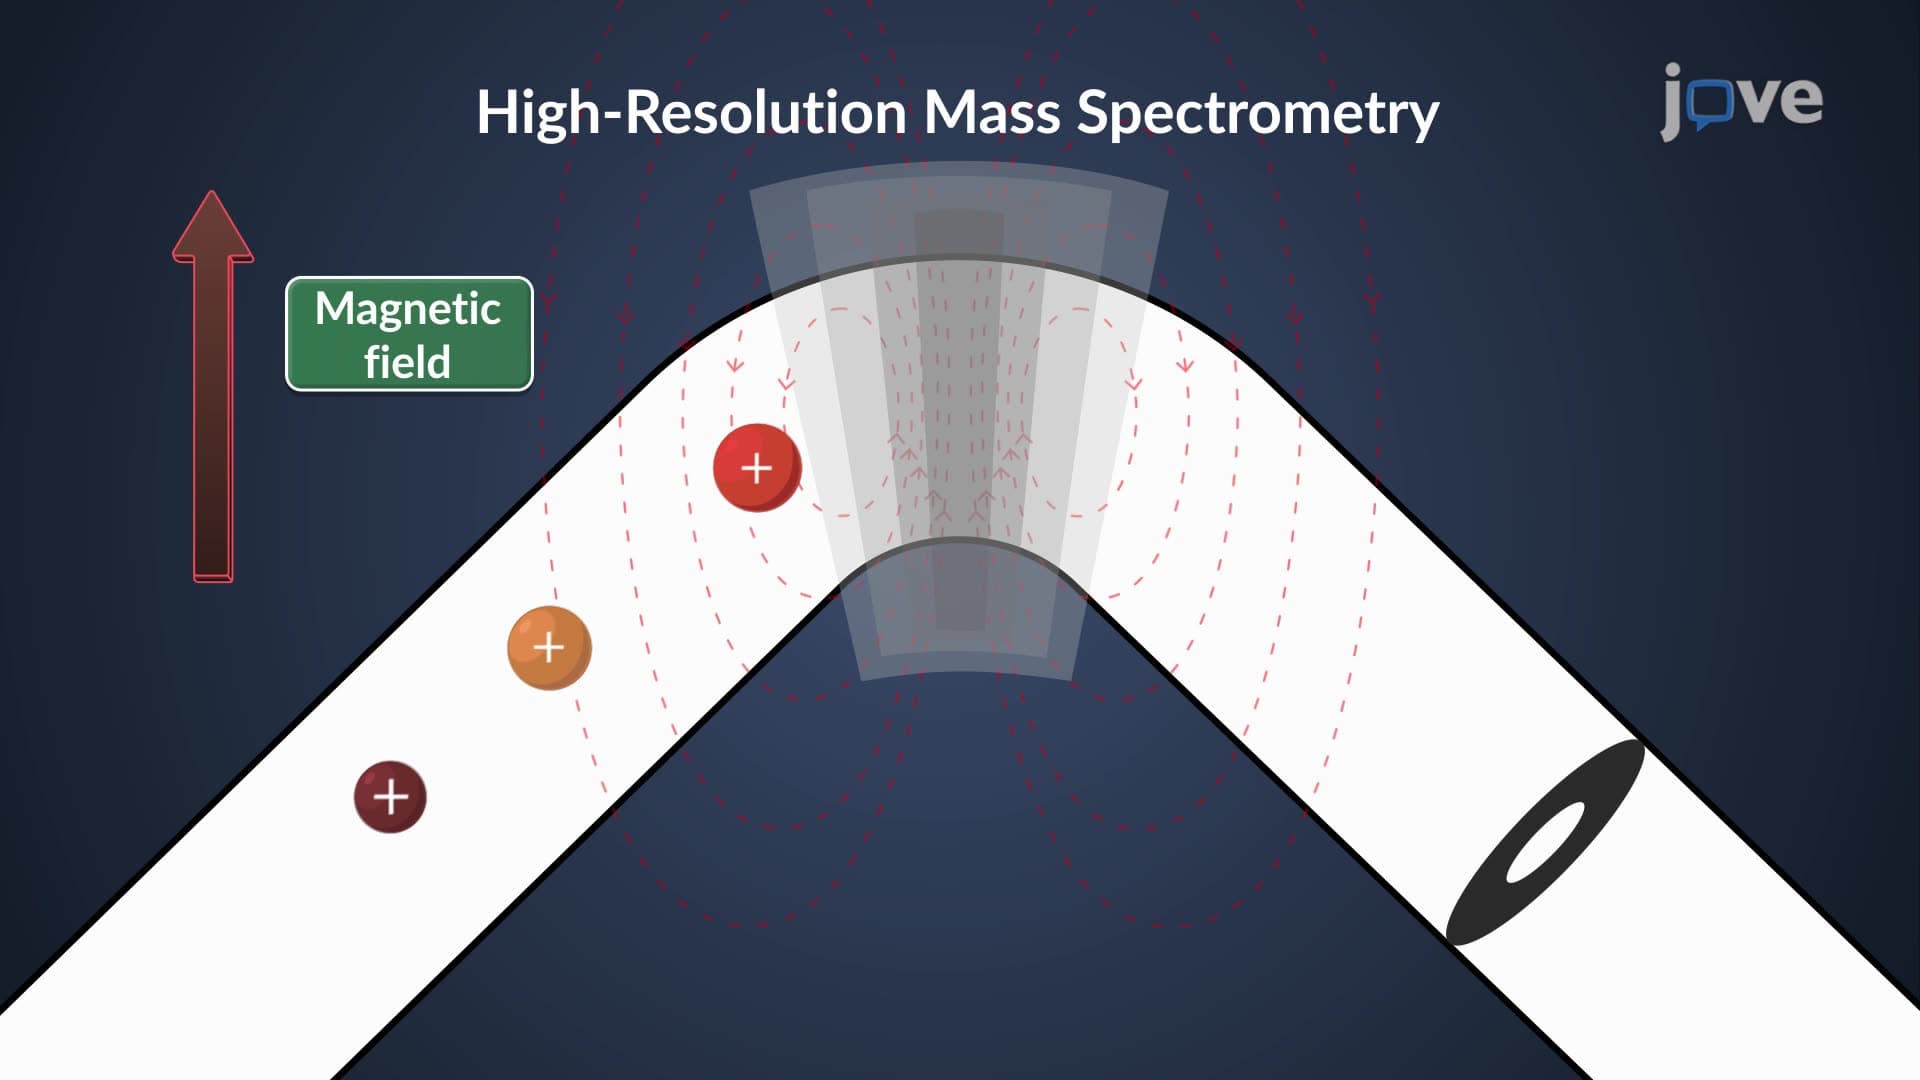
High-Resolution Mass Spectrometry (HRMS)