Computational methods for high-throughput comparative analyses of natural microbial communities
Related Experiment Video
Updated: May 7, 2026
Microbiota Analysis Using Two-step PCR and Next-generation 16S rRNA Gene Sequencing
Published on: October 15, 2019
Sarah P Preheim1, Allison R Perrotta, Jonathan Friedman
1Biological Engineering, Massachusetts Institute of Technology, Cambridge, Massachusetts, USA.

Abstract:
One of the most widely employed methods in metagenomics is the amplification and sequencing of the highly conserved ribosomal RNA (rRNA) genes from organisms in complex microbial communities. rRNA surveys, typically using the 16S rRNA gene for prokaryotic identification, provide information about the total diversity and taxonomic affiliation of organisms present in a sample. Greatly enhanced by high-throughput sequencing, these surveys have uncovered the remarkable diversity of uncultured organisms and revealed unappreciated ecological roles ranging from nutrient cycling to human health. This chapter outlines the best practices for comparative analyses of microbial community surveys. We explain how to transform raw data into meaningful units for further analysis and discuss how to calculate sample diversity and community distance metrics. Finally, we outline how to find associations of species with specific metadata and true correlations between species from compositional data. We focus on data generated by next-generation sequencing platforms, using the Illumina platform as a test case, because of its widespread use especially among researchers just entering the field.
More Related Videos
Related Concept Videos
 01:19
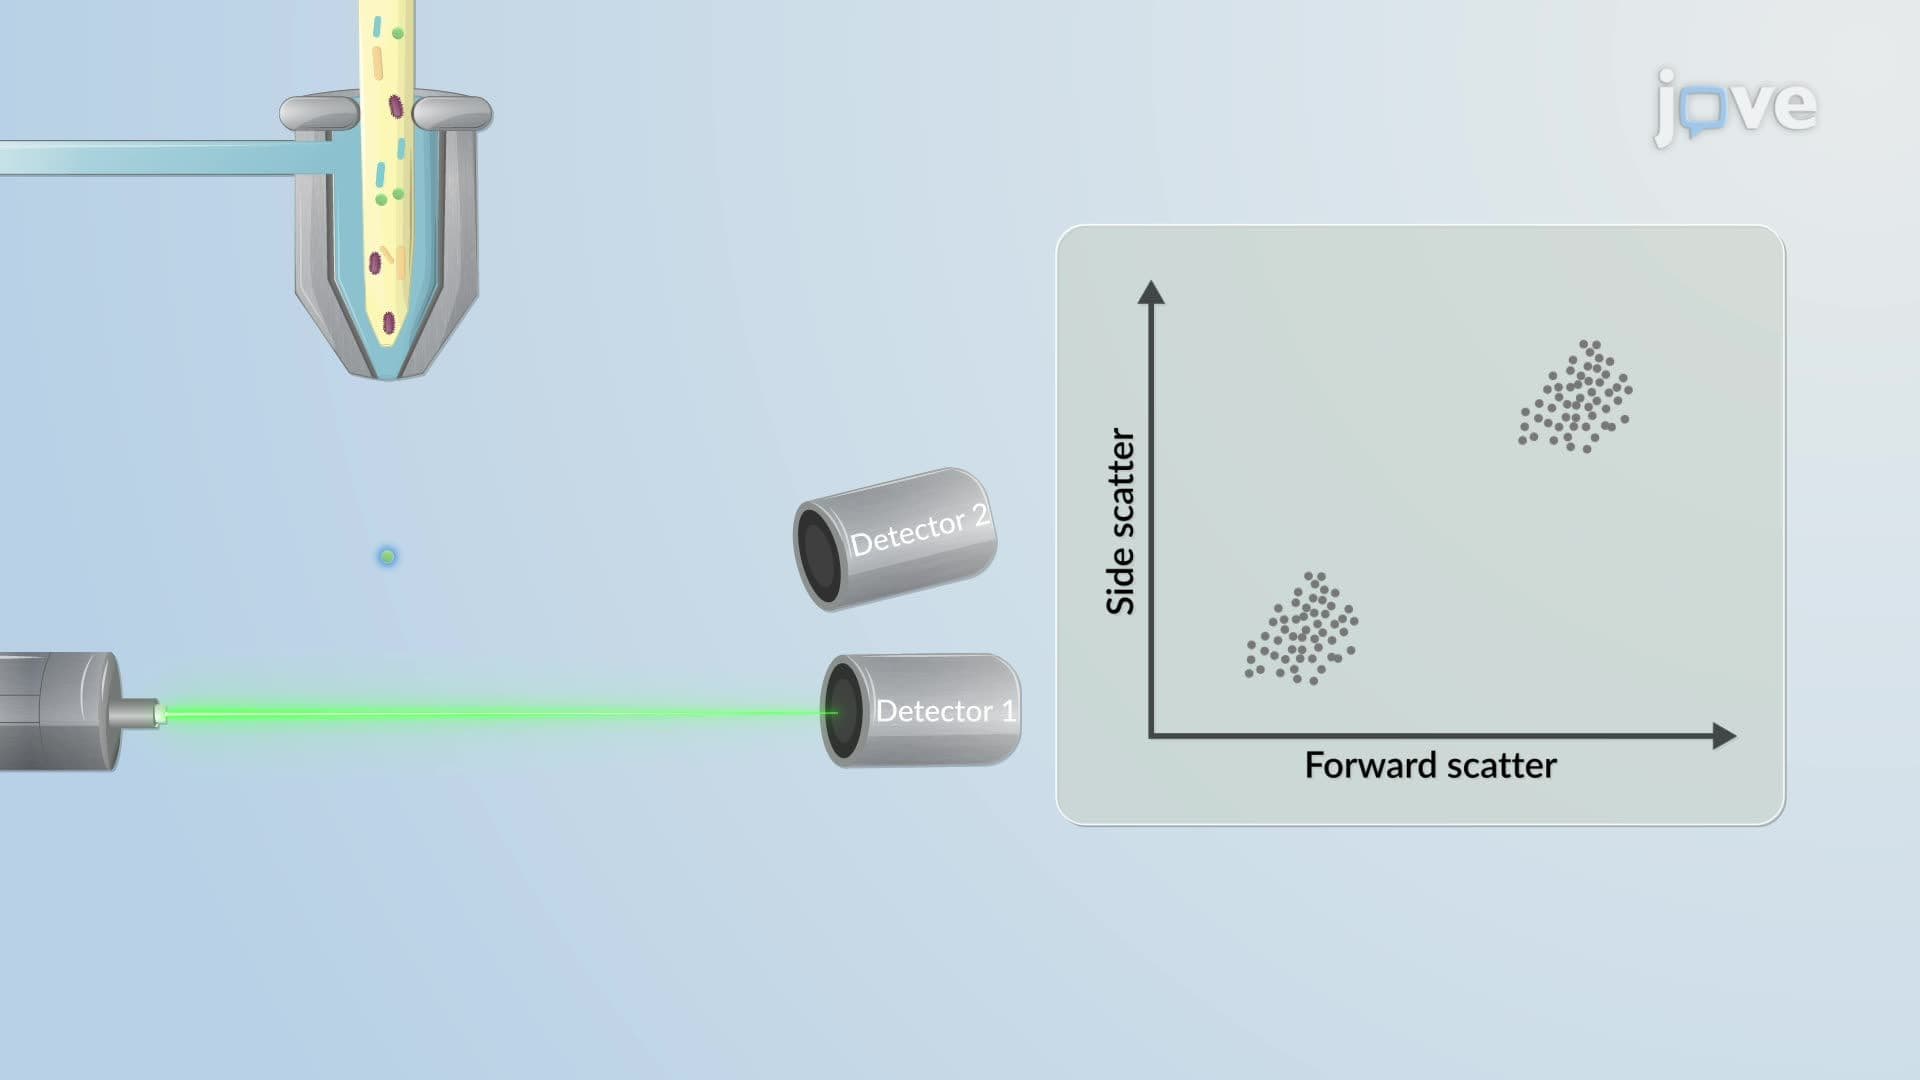
Methods to Assess Microbial Communities
 01:30
Methods to Assess Microbial Populations
 01:29
Modern Molecular Taxonomy
 01:20
Applications of Molecular Taxonomy
 02:54
Evolutionary Relationships through Genome Comparisons
 01:23
Microbial Growth Measurement: Direct Methods