Related Experiment Video
Updated: May 7, 2026
Phenotyping Mouse Pulmonary Function In Vivo with the Lung Diffusing Capacity
Published on: January 6, 2015
Evolution of lung function during the first year of life in newborn screened cystic fibrosis infants
The Thanh-Diem Nguyen1, Lena P Thia2, Ah-Fong Hoo3
1Portex Unit: Respiratory Physiology and Medicine, UCL Institute of Child Health, London, UK Department of Respiratory Medicine, Centre Hospitalier Universitaire Sainte-Justine, Montreal, Quebec, Canada.

Insights
Newborn screening for cystic fibrosis (CF) shows improved lung function (FEV₀.₅) in infants within the first year. Early lung function at 3 months predicts later outcomes, identifying high-risk infants for intensified treatment and clinical trials.
Area of Science:
- Pediatric Pulmonology
- Neonatal Screening
- Genetics and Genomics
Background:
- Newborn screening (NBS) for cystic fibrosis (CF) enables early intervention.
- Uncertainty in lung function evolution hinders randomized controlled trial (RCT) design in infants with CF.
Purpose of the Study:
- To evaluate pulmonary function changes in the first year of life for infants identified with CF through NBS.
- To establish the predictive value of early lung function measurements for long-term outcomes.
Main Methods:
- An observational longitudinal study involving infants with CF detected via NBS and healthy controls.
- Measurement of Lung Clearance Index (LCI), plethysmographic lung volume (FRCpleth), and forced expired volume (FEV₀.₅) at 3 months and 1 year.
- Comparison of pulmonary function parameters between CF infants and controls, and tracking changes over time.
Main Results:
- CF infants showed significantly poorer lung function at 3 months compared to controls.
- FEV₀.₅ improved significantly in CF infants from 3 months to 1 year, nearing control levels.
- LCI and FRCpleth remained stable but elevated in CF infants throughout the first year.
- Early lung function at 3 months strongly predicted pulmonary function at 1 year, identifying stable and high-risk groups.
Conclusions:
- This study is the first to report FEV₀.₅ improvement over time in stable, screened CF infants on standard therapy.
- Lung function changes by 1 year were less severe than previously documented.
- Early pulmonary function assessment at 3 months is crucial for identifying high-risk CF infants who may benefit from intensified treatment and RCT enrollment.
Rationale:
Newborn screening (NBS) for cystic fibrosis (CF) allows early intervention. Design of randomised controlled trials (RCT) is currently impeded by uncertainty regarding evolution of lung function, an important trial end point in such infants.
Objective:
To assess changes in pulmonary function during the first year of life in CF NBS infants.
Methods:
Observational longitudinal study. CF NBS infants and healthy controls were recruited between 2009 and 2011. Lung Clearance Index (LCI), plethysmographic lung volume (plethysmographic functional residual capacity (FRCpleth)) and forced expired volume (FEV₀.₅) were measured at 3 months and 1 year of age.
Main Results:
Paired measurements were obtained from 72 CF infants and 44 controls. At 3 months, CF infants had significantly worse lung function for all tests. FEV₀.₅ improved significantly (0.59 (95% CI 0.18 to 0.99) z-scores; p<0.01) in CF infants between 3 months and 1 year, and by 1 year, FEV₀.₅ was only 0.52 (0.89 to 0.15) z-scores less than in controls. LCI and FRCpleth remained stable throughout the first year of life, being on average 0.8 z-scores higher in infants with CF. Pulmonary function at 1 year was predicted by that at 3 months. Among the 45 CF infants with entirely normal LCI and FEV₀.₅ at 3 months, 80% remained so at 1 year, while 74% of those with early abnormalities remained abnormal at 1 year.
Conclusions:
This is the first study reporting improvements in FEV₀.₅ over time in stable NBS CF infants treated with standard therapy. Milder changes in lung function occurred by 1 year than previously reported. Lung function at 3 months predicts a high-risk group, who should be considered for intensification of treatment and enrolment into RCTs.
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...
 01:25
Pulmonary Function Tests
Pulmonary Function Tests are crucial diagnostic tools for assessing respiratory function, particularly in patients with chronic respiratory disorders. They comprehensively evaluate lung volumes, ventilatory function, breathing mechanics, diffusion, and gas exchange. These tests help diagnose pulmonary diseases and play a significant role in monitoring disease progression, evaluating disability, and assessing response to therapy.
PFTs involve using a spirometer, a...
 01:19
Factors Affecting Pulmonary Ventilation
Alveolar Surface Tension
The alveolar fluid lines the luminal surface of the alveoli and exerts a force called surface tension. This force is caused by the polar water molecules in the liquid being more strongly attracted to each...
 01:24
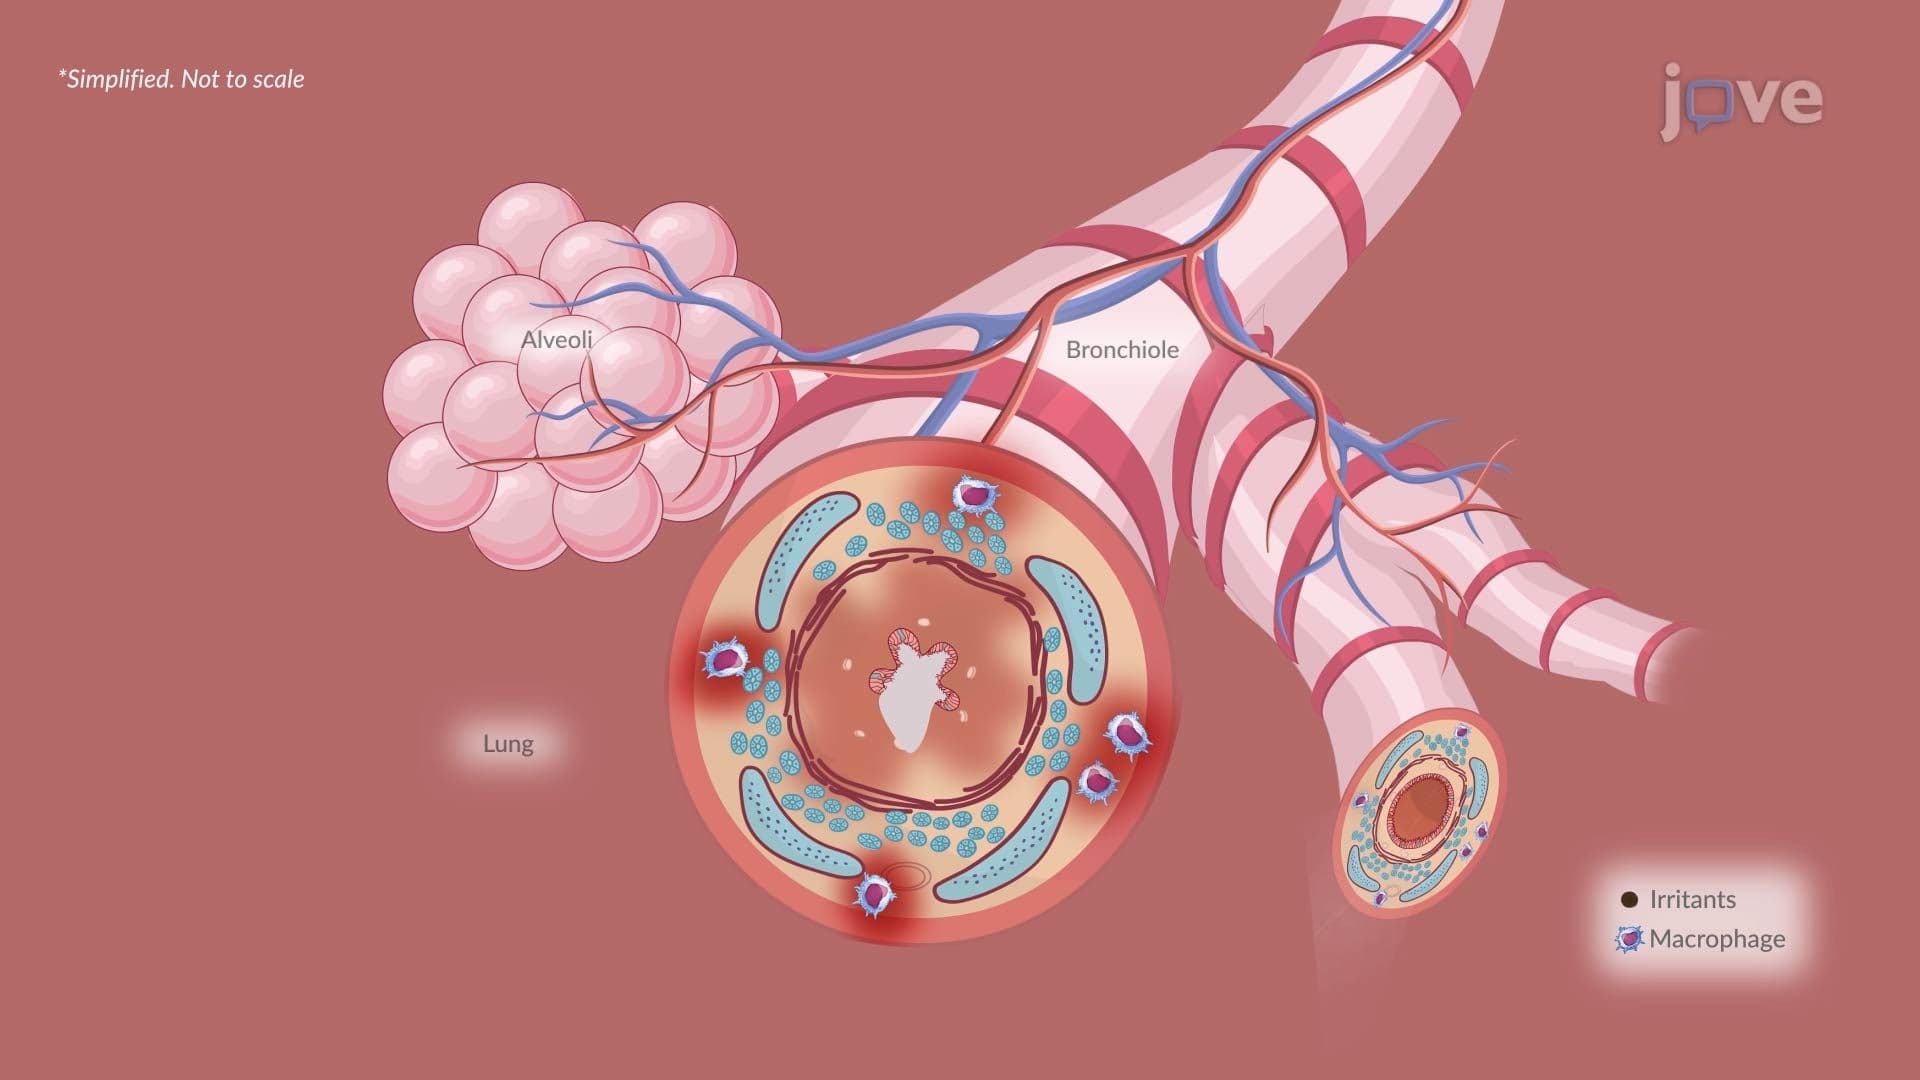
Chronic Obstructive Pulmonary Disease III: Chronic Bronchitis Features