Reversible white matter lesions during ketogenic diet therapy in glucose transporter 1 deficiency syndrome

Tadashi Shiohama1, Katsunori Fujii, Satoru Takahashi

  • 1Department of Pediatrics, Chiba University Graduate School of Medicine, Chiba, Japan.

Pediatric Neurology
|October 2, 2013
PubMed
Abstract

Related Concept Videos

Glucose Transporters01:27

Glucose Transporters

Glucose transporters facilitate the transport of glucose across the cell membrane. In addition to glucose, some glucose transporters can also aid the movement of other hexoses such as fructose, mannose, and galactose.
Facilitated diffusion-glucose transporters (GLUTs) are encoded by the solute-linked carrier (SLC) family 2, subfamily A gene family, or SLC2A. The 14 GLUT protein members are distributed into three classes:
Diabetic Ketoacidosis ll: Pathophysiology01:22

Diabetic Ketoacidosis ll: Pathophysiology
Diabetic ketoacidosis (DKA) is a metabolic emergency characterized by hyperglycemia, ketonemia, and metabolic acidosis. It results from severe insulin deficiency and an excess of counterregulatory hormones, leading to uncontrolled lipolysis, ketogenesis, and widespread electrolyte and fluid disturbances.Pathophysiology The central event in DKA is a profound loss of insulin action. Without insulin, glucose uptake in insulin-dependent tissues is impaired, while hepatic glucose production...
Diabetic Ketoacidosis l: Introduction01:25

Diabetic Ketoacidosis l: Introduction

DefinitionDiabetic ketoacidosis (DKA) is an acute, life-threatening complication of diabetes mellitus, characterized by a triad of hyperglycemia (blood glucose >250 mg/dL), ketonemia or ketonuria, and metabolic acidosis (arterial pH <7.30 and serum bicarbonate <18 mEq/L). It results from insulin deficiency combined with elevated levels of counterregulatory hormones—glucagon, catecholamines, cortisol, and growth hormone—leading to increased lipolysis, hepatic ketone production, and...
Inborn Errors of Metabolism01:20

Inborn Errors of Metabolism

Phenylketonuria (PKU) is a protein metabolism disorder characterized by high blood levels of the amino acid phenylalanine. This results from a mutation in the gene responsible for phenylalanine hydroxylase, an enzyme that converts phenylalanine into tyrosine. When this enzyme is deficient, phenylalanine builds up in the blood, leading to symptoms such as vomiting, rashes, seizures, growth deficiency, and severe mental retardation. An early diagnosis and a diet restricting phenylalanine intake...
Glucose Absorption Into the Small Intestine01:26

Glucose Absorption Into the Small Intestine
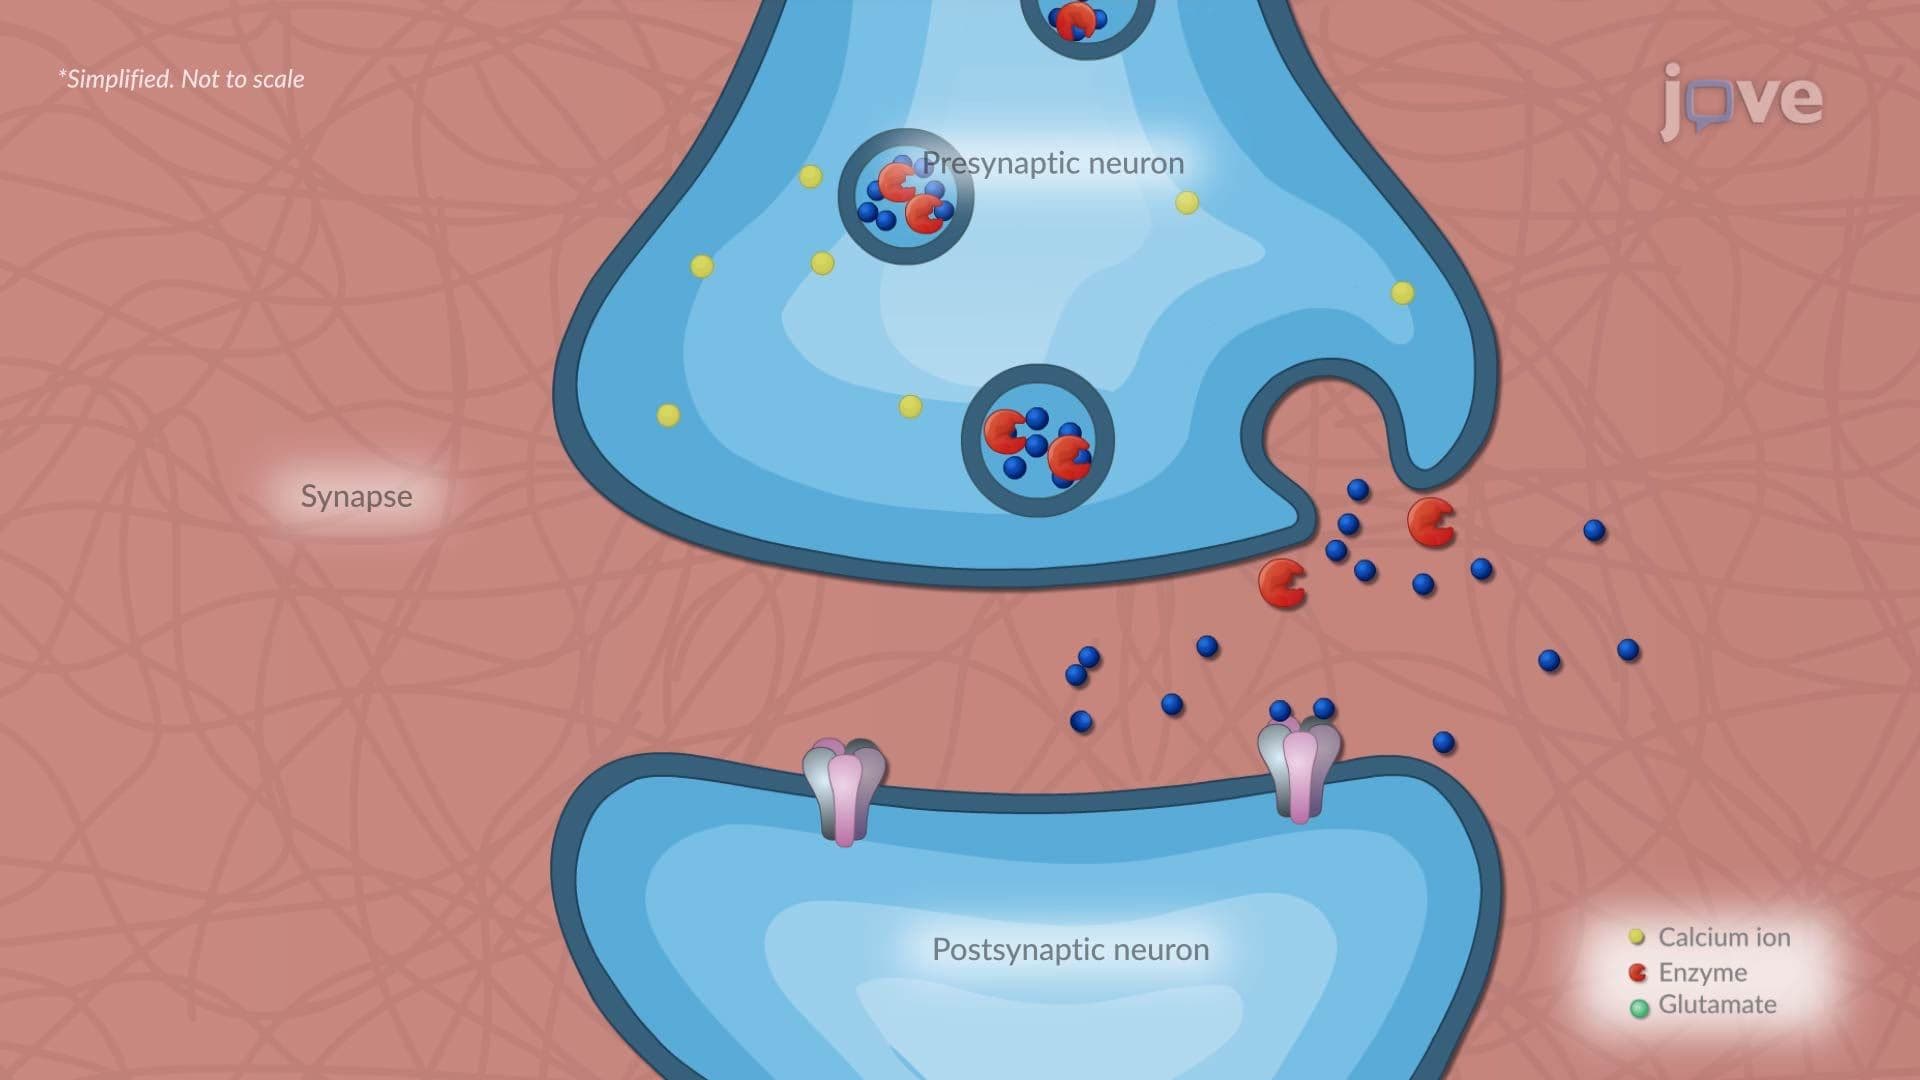
Complex carbohydrates consumed cannot be absorbed into the small intestine in their original form. First, they must be hydrolyzed to a monosaccharide form such as glucose or galactose. These monosaccharides are then transported across the intestinal membrane and into the blood via transcellular transport. The intestinal epithelial cells allow the movement of these monosaccharides with a defined 'entry' through membrane transporter proteins present on their apical membrane and 'exit' via the...
Ischemic Stroke ll: Pathophysiology01:15

Ischemic Stroke ll: Pathophysiology

An ischemic stroke occurs when a cerebral blood vessel becomes obstructed, most often by a thrombus or embolus, interrupting the delivery of oxygen and glucose to brain tissue. Because neurons rely on continuous aerobic metabolism, energy failure begins within minutes of reduced perfusion. The region receiving the least blood flow becomes the infarct core, an area of irreversible cellular death. Surrounding this core lies the penumbra, a zone of hypoperfused but still viable tissue that is...