Related Experiment Video
Updated: May 7, 2026

Synthesis of pH Dependent Pyrazole, Imidazole, and Isoindolone Dipyrrinone Fluorophores using a Claisen-Schmidt Condensation Approach
Published on: June 10, 2021
DFT calculations and electronic absorption spectra of some, α- and γ-pyrone derivatives
Walid M I Hassan1, Hussein Moustafa, Mohamed N H Hamed
1Chemistry Department, Faculty of Science, Cairo University, Giza, Egypt.

Abstract:
The electronic absorption spectra of 6-ethyl-4-hydroxy-2,5-dioxo-pyrano[3,2-c] quinoline 1, 6-ethyl-4-hydroxy-3-nitro-2,5-dioxo-pyrano[3,2-c] quinoline 2, 6-ethyl-4-chloro-2,5-dioxo-pyrano[3,2-c] quinoline 3, 6-ethyl-3-nitro-4-chloro-2,5-dioxo-pyrano[3,2-c] quinoline 4, 6-ethyl-4,5-dioxopyrano[3,2-c] quinoline 5, and 6-ethyl-3-nitro-6H-pyrano [3,2-c]quinoline-4,5-dione 6, were measured in polar (methanol) as well as nonpolar (dioxane) solvents. The geometries were optimized using B3LYB/6-311G (p,d) method. The most stable geometry of the studied compounds, 1-6, is the planar structure as indicates by the values of the dihedral angles. The insertion of a nitro group in position 3 in both α- and γ-pyrone ring decreases the energy gap and hence increases the reactivity of 3 and 6 compounds. Assignment of the observed bands as localized, delocalized and/or of charge transfer (CT) has been facilitated by TD-DFT calculations. The correspondences between the calculated and experimental transition energies are satisfactory. The solvent and substituent effects have been investigated. Chloro-substituent has a higher band position and intensity effects on the spectra more than hydroxyl or nitro groups.
More Related Videos
Related Concept Videos
 01:29
UV–Vis Spectroscopy: Woodward–Fieser Rules
 01:29
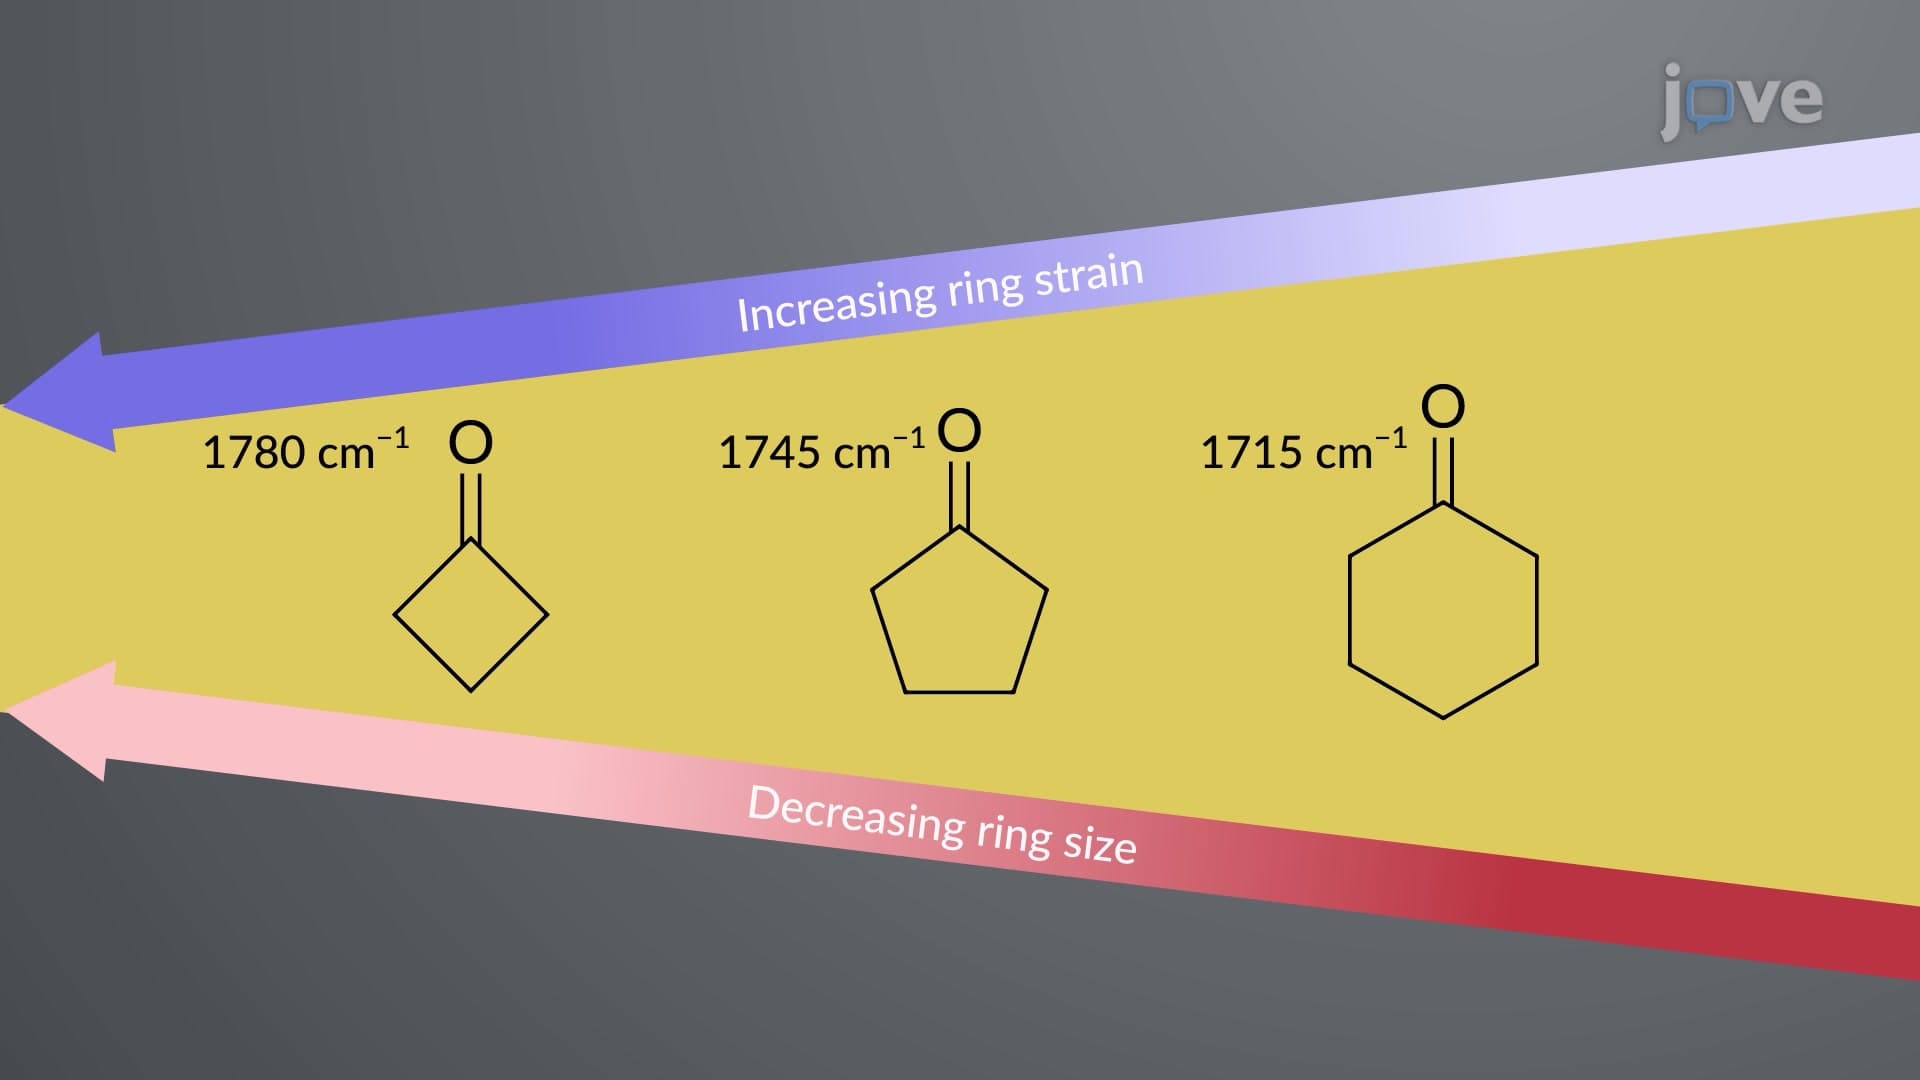
IR and UV–Vis Spectroscopy of Aldehydes and Ketones
 01:14
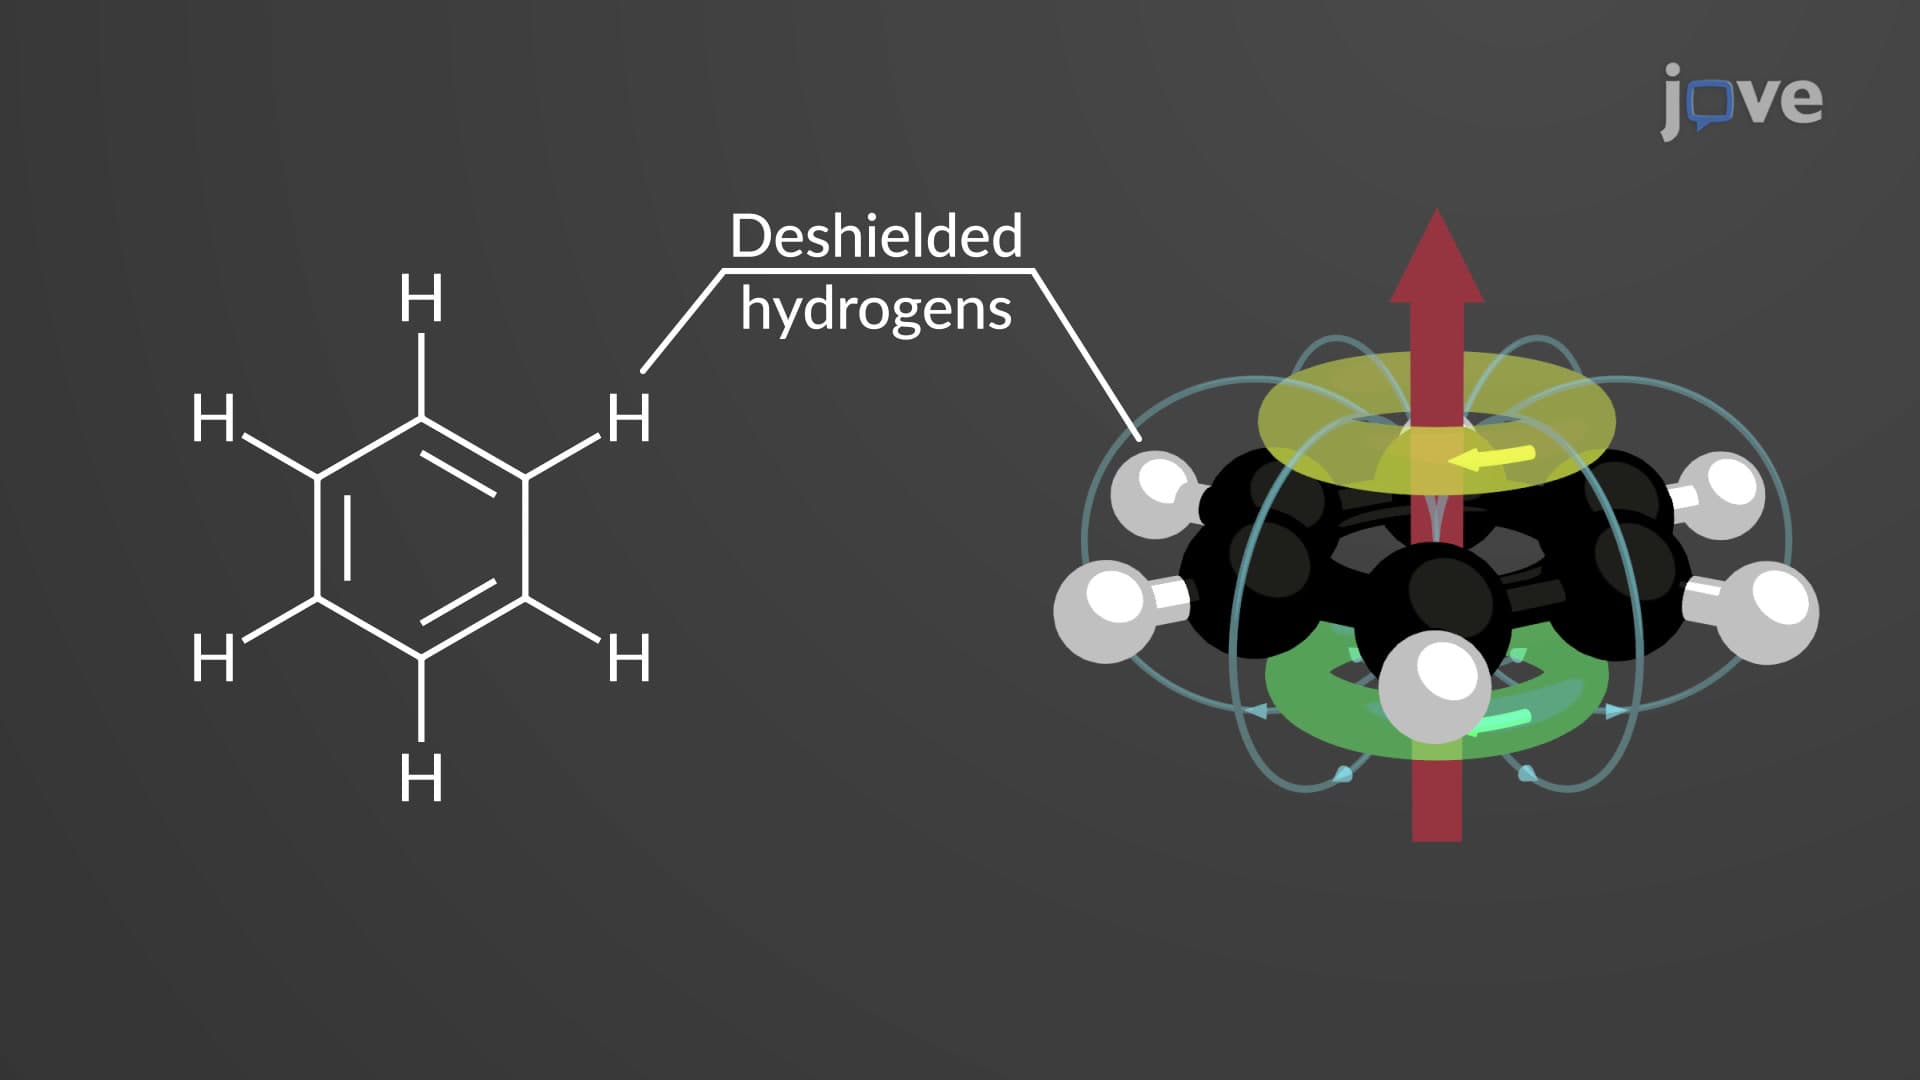
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
 01:26
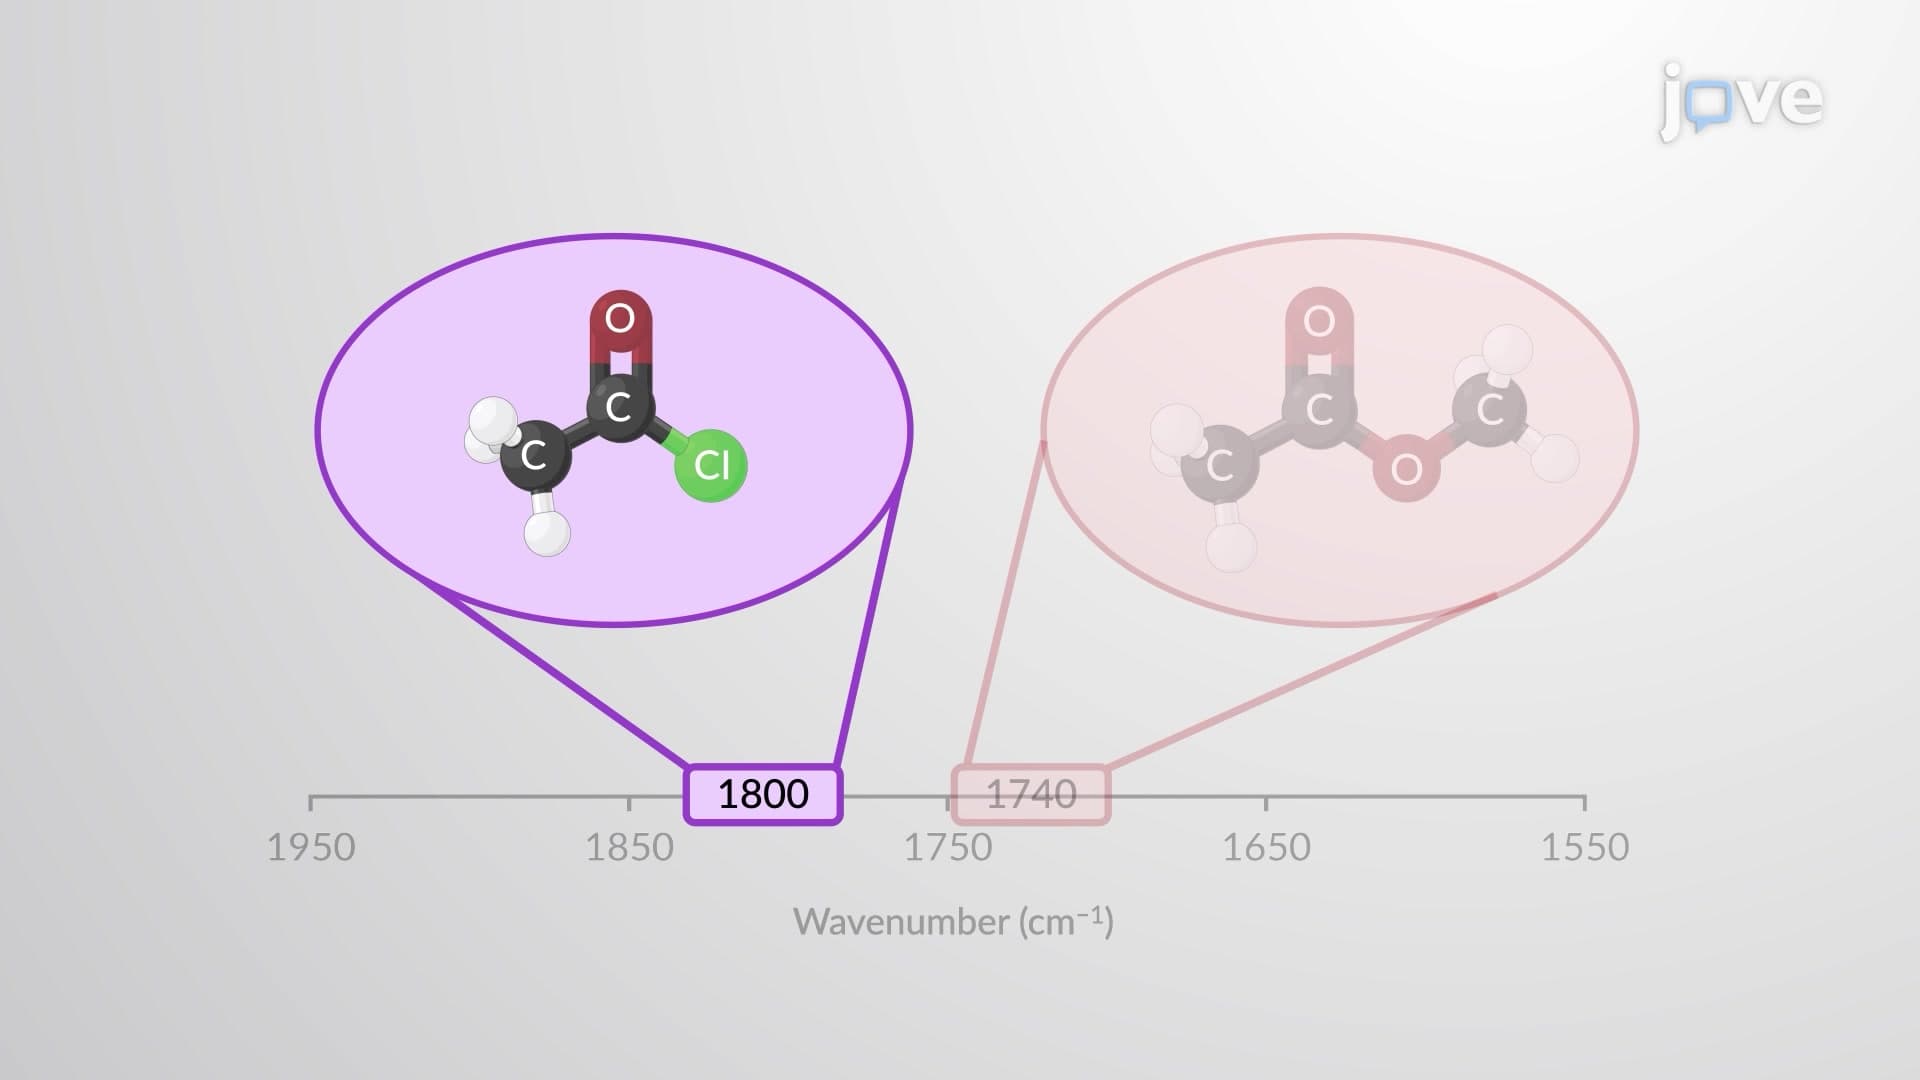
Spectroscopy of Carboxylic Acid Derivatives
 01:03
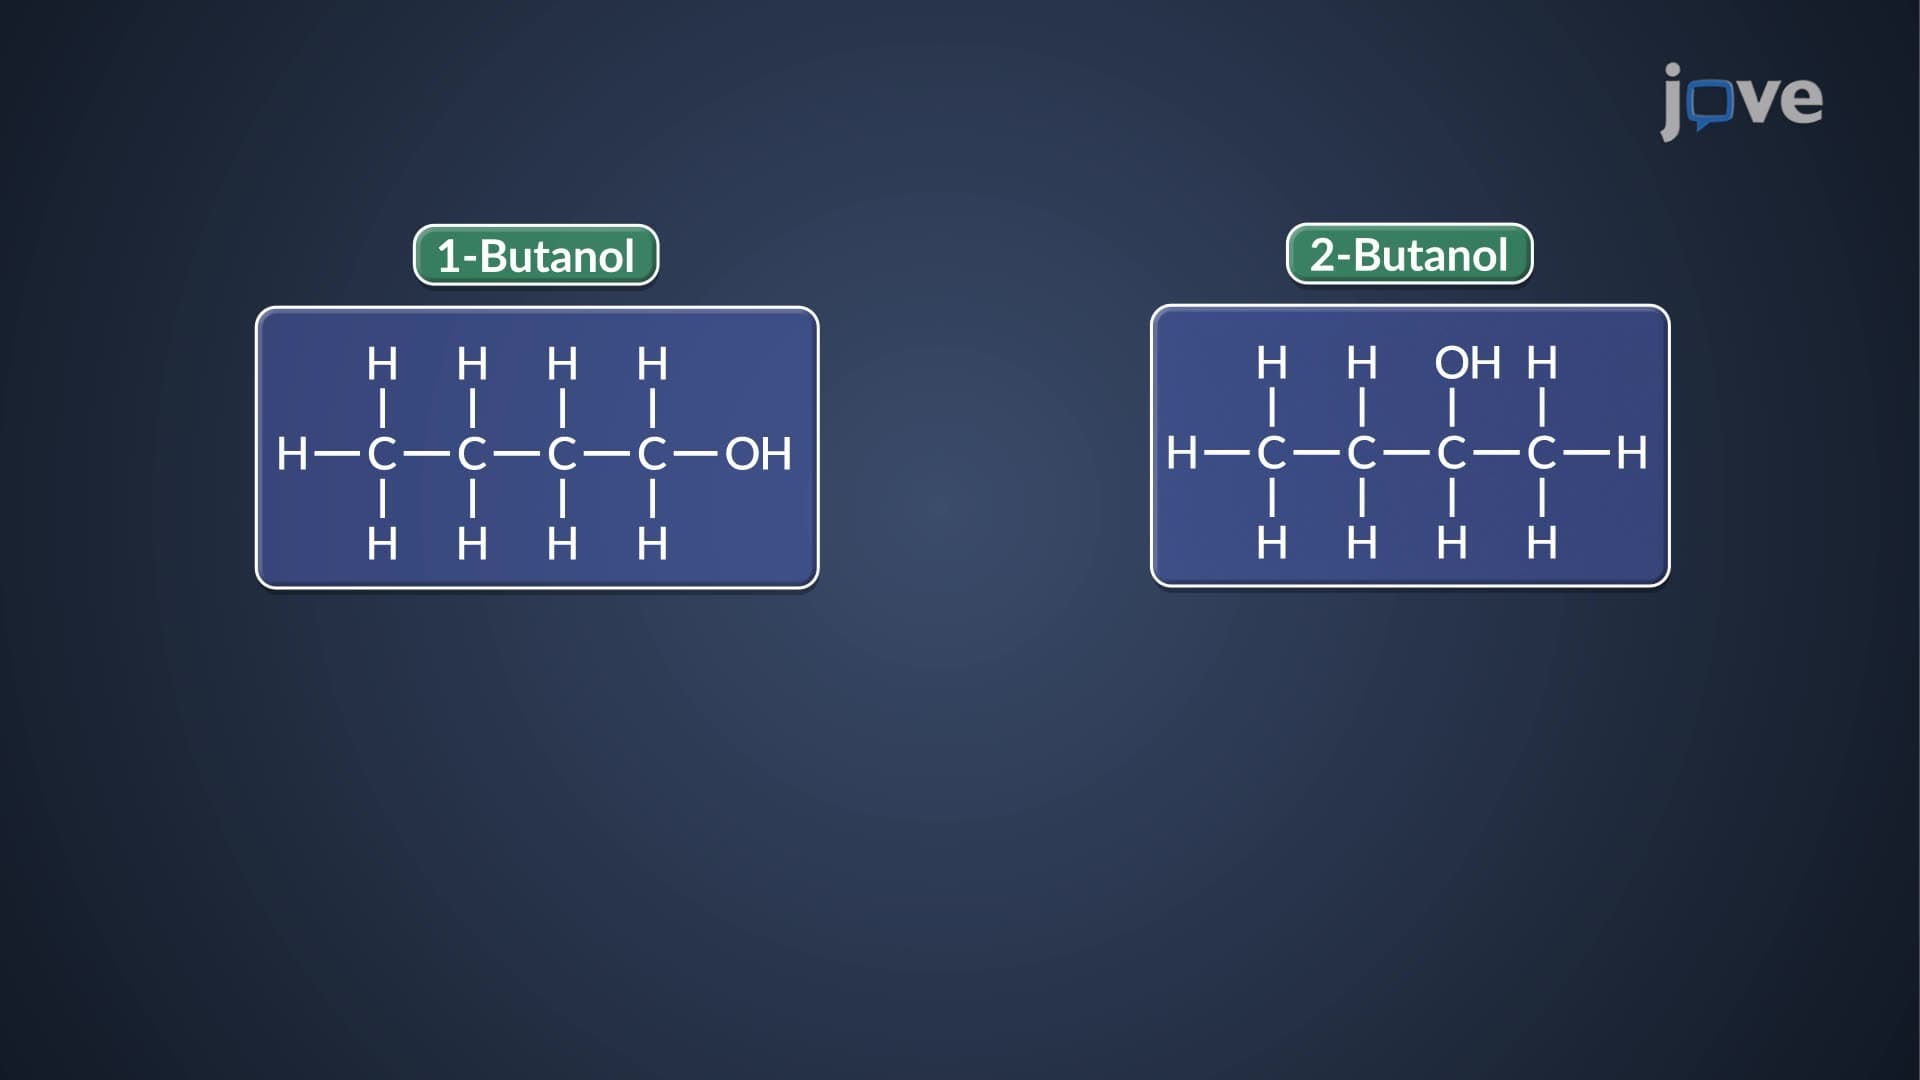
IR Frequency Region: Fingerprint Region
 01:14
NMR Spectroscopy of Aromatic Compounds