Related Experiment Video For amyotrophic lateral sclerosis
Updated: May 7, 2026

ALS - Motor Neuron Disease: Mechanism and Development of New Therapies
Published on: July 29, 2007
Amyotrophic Lateral Sclerosis: An update for 2013 Clinical Features, Pathophysiology, Management and Therapeutic
1AP-HP, Hôpital de la Pitié-Salpêtrière, Département des Maladies du Système Nerveux, Paris, France ; Northern Navajo Medical Center, Shiprock, NM 87420, USA.

Abstract:
Amyotrophic lateral sclerosis (ALS), first described by Jean-Martin Charcot in the 1870s, is an age-related disorder that leads to degeneration of motor neurons. The disease begins focally in the central nervous system and then spreads relentlessly. The clinical diagnosis, defined by progressive signs and symptoms of upper and lower motor neuron dysfunction, is confirmed by electromyography. Additional testing excludes other conditions. The disease is heterogeneous, but most patients die of respiratory muscle weakness less than 3 years from symptom-onset. Like other age-related neurodegenerative diseases, ALS has genetic and environmental triggers. Of the five to 10% of cases that are inherited, mutations have been discovered for a high proportion. In addition to genetic factors, age, tobacco use, and athleticism may contribute to sporadic ALS, but important etiologies are unidentified for most patients. Complex pathophysiological processes, including mitochondrial dysfunction, aggregation of misfolded protein, oxidative stress, excitotoxicity, inflammation and apoptosis, involve both motor neurons and surrounding glial cells. There is clinical and pathological overlap with other neurodegenerative diseases, particularly frontotemporal dementia. The mechanisms leading to disease propagation in the brain are a current focus of research. To date, one medication, riluzole, licensed in 1996, has been proved to prolong survival in ALS. Numerous clinical trials have so far been unable to identify another neuroprotective agent. Researchers now aim to slow disease progression by targeting known pathophysiological pathways or genetic defects. Current approaches are directed at muscle proteins such as Nogo, energetic balance, cell replacement, and abnormal gene products resulting from mutations. Until better understanding of the causes and mechanisms underlying progression lead to more robust neuroprotective agents, symptomatic therapies can extend life and improve quality of life. Palliative care programs such as hospice give emotional and physical support to patients and families throughout much of the disease course.
More Related Videos
 10:49
Intraspinal Cell Transplantation for Targeting Cervical Ventral Horn in Amyotrophic Lateral Sclerosis and Traumatic Spinal Cord Injury
Published on: September 18, 2011
 12:35
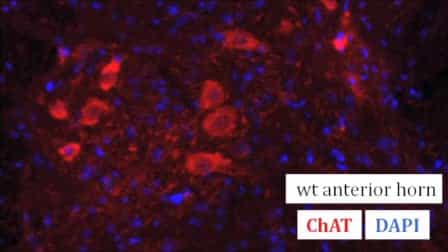
Clinical Testing and Spinal Cord Removal in a Mouse Model for Amyotrophic Lateral Sclerosis ALS
Published on: March 17, 2012
Related Concept Videos
 01:15
Parkinson's Disease: Overview
 01:24
Parkinson Disease ll: Pathophysiology
 01:22
Myasthenia Gravis ll: Pathophysiology
 01:24
Parkinson Disease l: Introduction
 01:11
Alterations in Muscle Tone lll
 01:20
Myasthenia Gravis: Overview and Treatment
These antibodies interfere with the function of the nicotinic receptors in three ways: by binding to the receptor and disrupting acetylcholine binding; by causing cross-linking of receptors which...