Related Experiment Video
Updated: May 6, 2026

Transport Properties of Ibuprofen Encapsulated in Cyclodextrin Nanosponge Hydrogels: A Proton HR-MAS NMR Spectroscopy Study
Published on: August 15, 2016
Prediction of in vivo plasma concentration-time profile from in vitro release data of designed formulations of
Gautam Singhvi1, Abhishek Shah, Nilesh Yadav
1Department of Pharmacy, Industrial Research Laboratory, Birla Institute of Technology and Science , Pilani, Rajasthan , India.

Abstract:
The aim of this study was to predict the in vivo plasma drug level of milnacipran (MIL) from in vitro dissolution data of immediate release (IR 50 mg and IR 100 mg) and matrix based controlled release (CR 100 mg) formulations. Plasma drug concentrations of these formulations were predicted by numerical convolution method. The convolution method uses in vitro dissolution data to derive plasma drug levels using reported pharmacokinetic (PK) parameters of a test product. The bioavailability parameters (Cmax and AUC) predicted from convolution method were found to be 106.90 ng/mL, 1138.96 ng/mL h for IR 50 mg and 209.80 ng/mL, 2280.61 ng/mL h for IR 100 mg which are similar to those reported in the literature. The calculated PK parameters were validated with percentage predication error (% PE). The % PE values for Cmax and AUC were found to be 7.04 and -7.35 for IR 50 mg and 11.10 and -8.21 for IR 100 mg formulations. The Cmax, Tmax, and AUC for CR 100 mg were found to be 120 ng/mL, 10 h and 2112.60 ng/mL h, respectively. Predicted plasma profile of designed CR formulation compared with IR formulations which indicated that CR formulation can prolong the plasma concentration of MIL for 24 h. Thus, this convolution method is very useful for designing and selection of formulation before animal and human studies.
More Related Videos
 07:32
Preparation and Characterization of Individual and Multi-drug Loaded Physically Entrapped Polymeric Micelles
Published on: August 28, 2015
 08:47
Experimental Quantification of Interactions Between Drug Delivery Systems and Cells In Vitro: A Guide for Preclinical Nanomedicine Evaluation
Published on: September 28, 2022
Related Concept Videos
 01:30
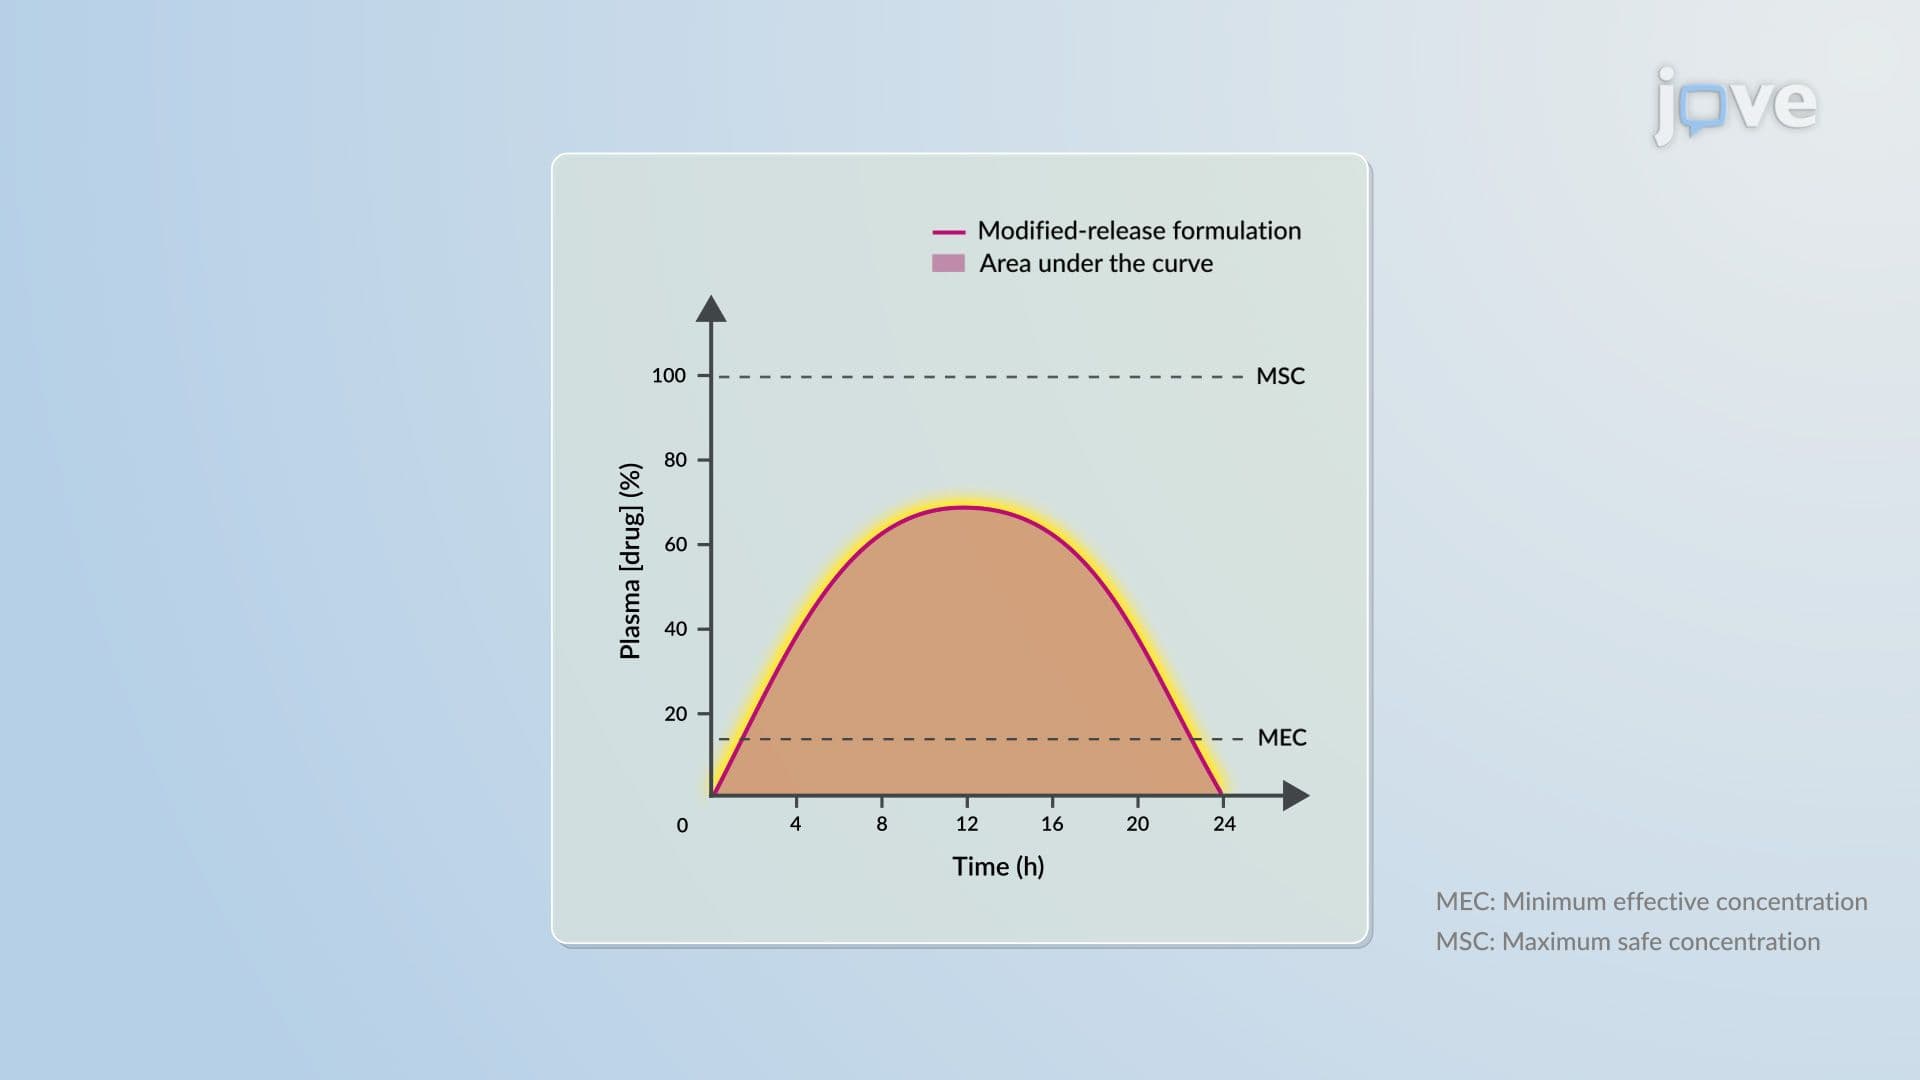
Modified-Release Drug Delivery Systems: Bioavailability
 01:20
Drug Product Performance: In Vitro–In Vivo Correlation
 01:15
Pharmacodynamic Models: Linear Concentration–Effect Model
 01:15
Drug Concentration Versus Time Correlation
Two pivotal parameters are the minimum effective concentration (MEC) and the minimum toxic concentration (MTC). The MEC is the...
 00:54
Noncompartmental Analysis: Miscellaneous Pharmacokinetic Parameters
One key aspect of the noncompartmental approach is determining a drug's total clearance. This can be done by dividing the drug dose by the area under the concentration-time curve from zero to infinity. The area under the concentration-time curve represents the drug's...
 01:30
Measurement of Bioavailability: Pharmacokinetic Methods