Related Experiment Video
Updated: May 6, 2026

An Integrated Approach for Microprotein Identification and Sequence Analysis
Published on: July 12, 2022
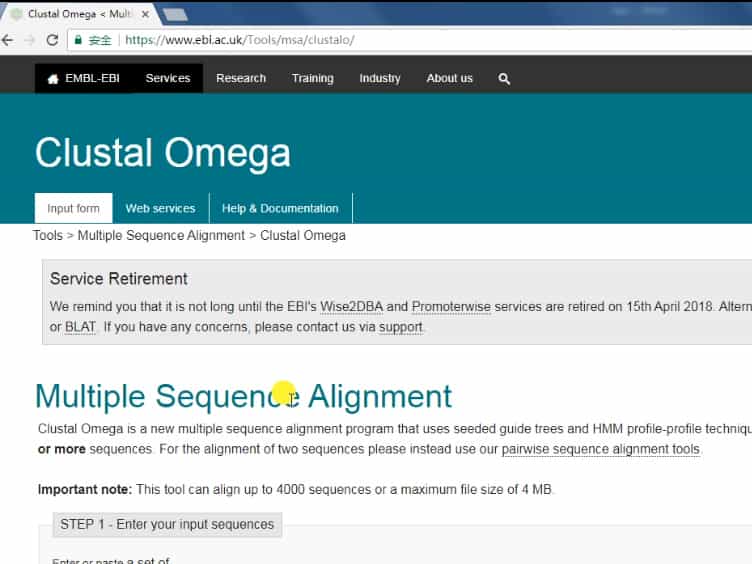
Clustal Omega, accurate alignment of very large numbers of sequences
Fabian Sievers1, Desmond G Higgins
1Conway Institute, University College Dublin, Dublin, Ireland.

Abstract:
Clustal Omega is a completely rewritten and revised version of the widely used Clustal series of programs for multiple sequence alignment. It can deal with very large numbers (many tens of thousands) of DNA/RNA or protein sequences due to its use of the mBED algorithm for calculating guide trees. This algorithm allows very large alignment problems to be tackled very quickly, even on personal computers. The accuracy of the program has been considerably improved over earlier Clustal programs, through the use of the HHalign method for aligning profile hidden Markov models. The program currently is used from the command line or can be run on line.
Related Concept Videos
 02:51
Multi-species Conserved Sequences
Although the genome of each species varies greatly from each other, a few sequences are highly conserved. Such conserved...
 03:21
RNA-seq
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
 02:54
Evolutionary Relationships through Genome Comparisons
 01:57
Sanger Sequencing
 03:00
Next-generation Sequencing
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
 02:26
Conservation of Protein Domains Over Different Proteins
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to...