Related Experiment Video
Updated: May 6, 2026

A Concoction Pipeline for Generating Molecular Operational Taxonomic Units (MOTUs) Among Riparian and Aquatic Beetles
Published on: July 11, 2025
Inferring species trees from incongruent multi-copy gene trees using the Robinson-Foulds distance
Ruchi Chaudhary1, John Gordon Burleigh, David Fernández-Baca
1Department of Computer Science, Iowa State University, Ames, IA 50011, USA. ruchic@ufl.edu.

Background:
Constructing species trees from multi-copy gene trees remains a challenging problem in phylogenetics. One difficulty is that the underlying genes can be incongruent due to evolutionary processes such as gene duplication and loss, deep coalescence, or lateral gene transfer. Gene tree estimation errors may further exacerbate the difficulties of species tree estimation.
Results:
We present a new approach for inferring species trees from incongruent multi-copy gene trees that is based on a generalization of the Robinson-Foulds (RF) distance measure to multi-labeled trees (mul-trees). We prove that it is NP-hard to compute the RF distance between two mul-trees; however, it is easy to calculate this distance between a mul-tree and a singly-labeled species tree. Motivated by this, we formulate the RF problem for mul-trees (MulRF) as follows: Given a collection of multi-copy gene trees, find a singly-labeled species tree that minimizes the total RF distance from the input mul-trees. We develop and implement a fast SPR-based heuristic algorithm for the NP-hard MulRF problem.We compare the performance of the MulRF method (available at http://genome.cs.iastate.edu/CBL/MulRF/) with several gene tree parsimony approaches using gene tree simulations that incorporate gene tree error, gene duplications and losses, and/or lateral transfer. The MulRF method produces more accurate species trees than gene tree parsimony approaches. We also demonstrate that the MulRF method infers in minutes a credible plant species tree from a collection of nearly 2,000 gene trees.
Conclusions:
Our new phylogenetic inference method, based on a generalized RF distance, makes it possible to quickly estimate species trees from large genomic data sets. Since the MulRF method, unlike gene tree parsimony, is based on a generic tree distance measure, it is appealing for analyses of genomic data sets, in which many processes such as deep coalescence, recombination, gene duplication and losses as well as phylogenetic error may contribute to gene tree discord. In experiments, the MulRF method estimated species trees accurately and quickly, demonstrating MulRF as an efficient alternative approach for phylogenetic inference from large-scale genomic data sets.
Related Concept Videos
 02:54
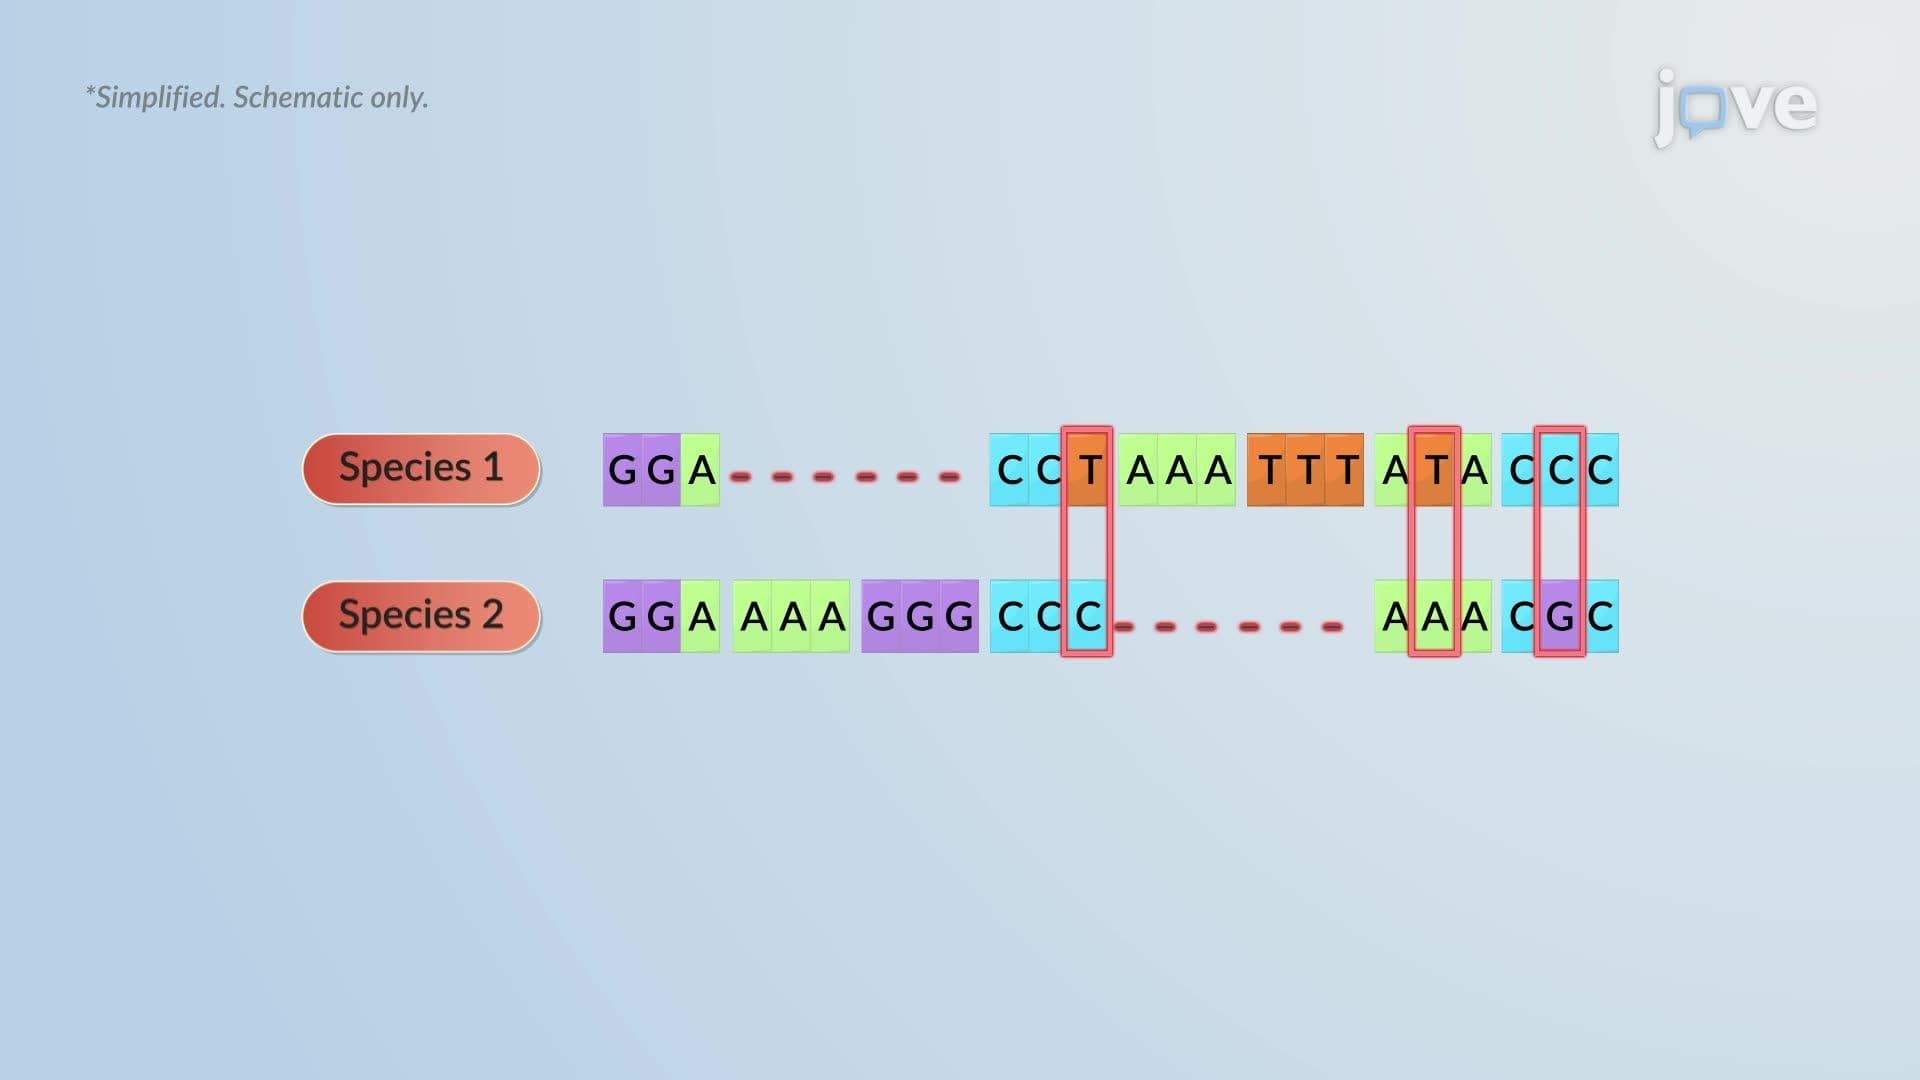
Evolutionary Relationships through Genome Comparisons
 03:21
Phylogenetic Trees
03:21Phylogenetic Trees
 01:28
Microbial Phylogeny
 02:05
Gene Evolution - Fast or Slow?
In contrast, regions which code...
02:05Gene Evolution - Fast or Slow?