Related Experiment Video
Updated: May 6, 2026

All-electronic Nanosecond-resolved Scanning Tunneling Microscopy: Facilitating the Investigation of Single Dopant Charge Dynamics
Published on: January 19, 2018
Adsorption of carbon dioxide on Al12X clusters studied by density functional theory: effect of charge and doping
Jian-Ying Zhao1, Yu Zhang, Feng-Qi Zhao
1School of Chemistry and Chemical Engineering, Huaiyin Normal University , Huaian 223300, People's Republic of China.

Abstract:
The adsorption of a CO2 molecule on neutral and charged X-centered icosahedron Al12X(±z) clusters (X = Al, Be, Zn, Ni, Cu, B, P; z = 0, 1) was investigated by the density functional PW91 and PWC methods. Optimized configurations corresponding to physisorption and chemisorption of CO2 were identified. The adsorption energies, activation barriers, and binding energies involving both the physisorption (Al12X(±z)·CO2-I) and chemisorption (Al12X(±z)·CO2-II) for CO2 were determined. The chemisorption of a CO2 molecule on the Al12X clusters (X is a metallic doping element) requires relatively low activation barriers. The lowest barrier was found to be with the Al12Be cluster. For the Al12X(-) clusters, the barriers are all higher than those of the neutral analogues. For the Al12X(+) clusters, two corresponding configurations are linked by a low-energy barrier, and CO2 molecule chemisorption on the Al12Be(+) cluster has the lowest barrier. The adsorption energies are larger than the energy barriers, which facilitates the chemisorption. The results show that carbon dioxide adsorbed on the Al12X(±z) clusters can be tuned by controllable X doping and the total number of valence electrons and suggest the potential application of Al12X(±z) nanostructures for carbon dioxide capture and activation.
More Related Videos
 09:34
Synthesis and Characterization of Fe-doped Aluminosilicate Nanotubes with Enhanced Electron Conductive Properties
Published on: November 15, 2016
 11:38
In situ FTIR Spectroscopy as a Tool for Investigation of Gas/Solid Interaction: Water-Enhanced CO2 Adsorption in UiO-66 Metal-Organic Framework
Published on: February 1, 2020
Related Concept Videos
 01:27
π Electron Effects on Chemical Shift: Overview
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:18
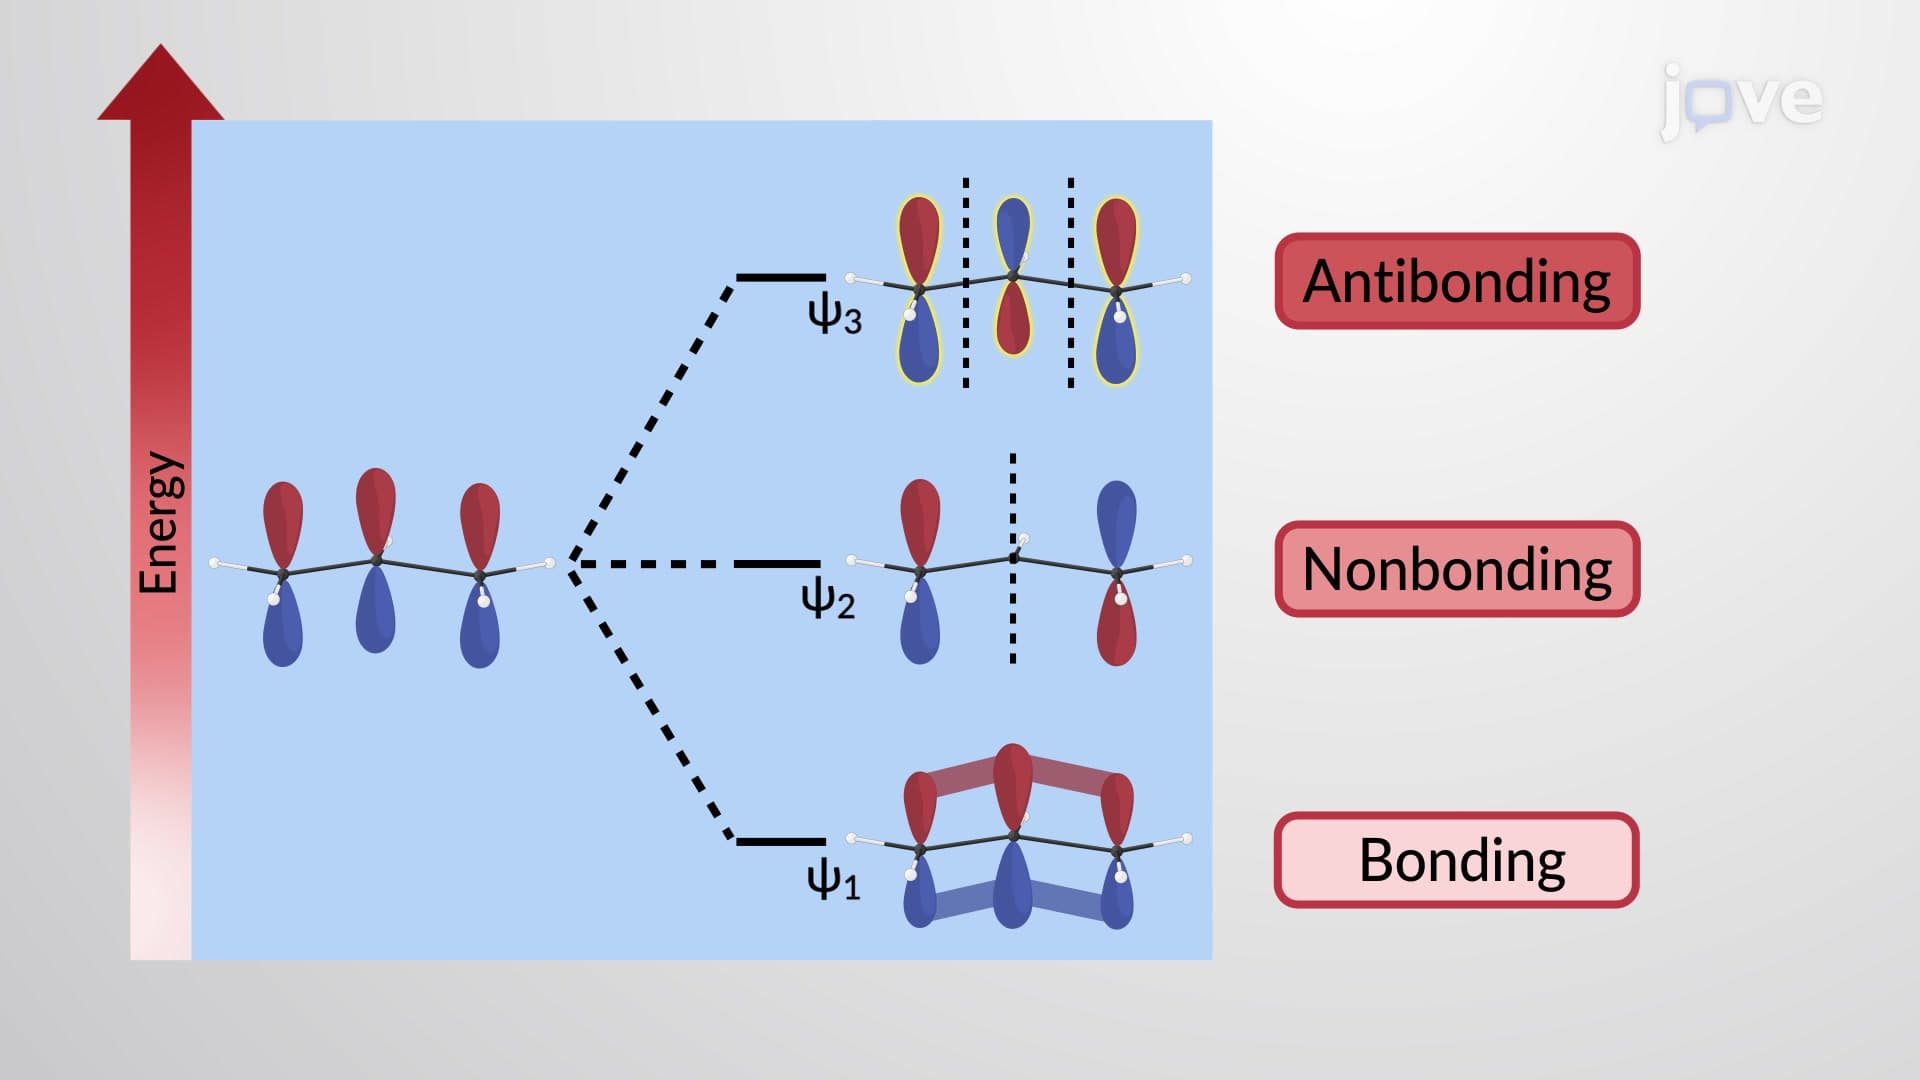
π Molecular Orbitals of the Allyl Cation and Anion
 02:42
Valence Bond Theory
 02:10
Carbocations
 02:37
Metallic Solids
All metallic solids exhibit high thermal and electrical conductivity, metallic luster, and...