Related Experiment Video
Updated: May 5, 2026
Dissection of the Endolymphatic Sac from Mice
Published on: March 29, 2021
Identification of PENDRIN (SLC26A4) mutations in patients with congenital hypothyroidism and "apparent" thyroid
Peter Kühnen1, Serap Turan, Sebastian Fröhler
1Institute of Experimental Pediatric Endocrinology (P.K., H.B., A.G., H.K.), Charité Universitätsmedizin Berlin, 13353 Berlin, Germany; Department of Pediatric Endocrinology and Diabetes (S.T., T.G., S.A., A.B.), Marmara University Hospital, 34722 Istanbul, Turkey; and Max-Delbrück Center for Molecular Medicine (S.F., W.C.), 13092 Berlin-Buch, Germany.

Context:
Congenital hypothyroidism, the most frequent endocrine congenital disease, can occur either based on a thyroid hormone biosynthesis defect or can predominantly be due to thyroid dysgenesis. However, a genetic cause could so far only be identified in less than 10% of patients with a thyroid dysgenesis.
Objectives:
Exome sequencing was used for the first time to find additional genetic defects in thyroid dysgenesis.
Patients And Methods:
In a consanguineous family with thyroid dysgenesis, exome sequencing was applied, and findings were further validated by Sanger sequencing in a cohort of 94 patients with thyroid dysgenesis.
Results:
By exome sequencing we identified a homozygous missense mutation (p.Leu597Ser) in the SLC26A4 gene of a patient with hypoplastic thyroid tissue, who was otherwise healthy. In the cohort of patients with thyroid dysgenesis, we observed a second case with a homozygous missense mutation (p.Gln413Arg) in the SLC26A4 gene, who was additionally affected by severe hearing problems. Both mutations were previously described as loss-of-function mutations in patients with Pendred syndrome and nonsyndromic enlarged vestibular aqueduct.
Conclusion:
We unexpectedly identified SLC26A4 mutations that were hitherto diagnosed in thyroid dyshormonogenesis patients, now for the first time in patients with structural thyroid defects. This result resembles the historic description of thyroid atrophy in patients with the so-called myxedematous form of cretinism after severe iodine deficiency. Most likely the thyroid defect of the two homozygous SLC26A4 gene mutation carriers represents a kind of secondary thyroid atrophy, rather than a primary defect of thyroid development in the sense of thyroid agenesis. Our study extends the variable clinical spectrum of patients with SLC26A4 mutations and points out the necessity to analyze the SLC26A4 gene in patients with apparent thyroid dysgenesis in addition to the known candidate genes TSHR, PAX8, NKX2.1, NKX2.5, and FOXE1.
More Related Videos
 08:22
A Novel Strategy Combining Array-CGH, Whole-exome Sequencing and In Utero Electroporation in Rodents to Identify Causative Genes for Brain Malformations
Published on: December 1, 2017
 06:41
In Vivo Functional Study of Disease-associated Rare Human Variants Using Drosophila
Published on: August 20, 2019
Related Concept Videos
 01:23
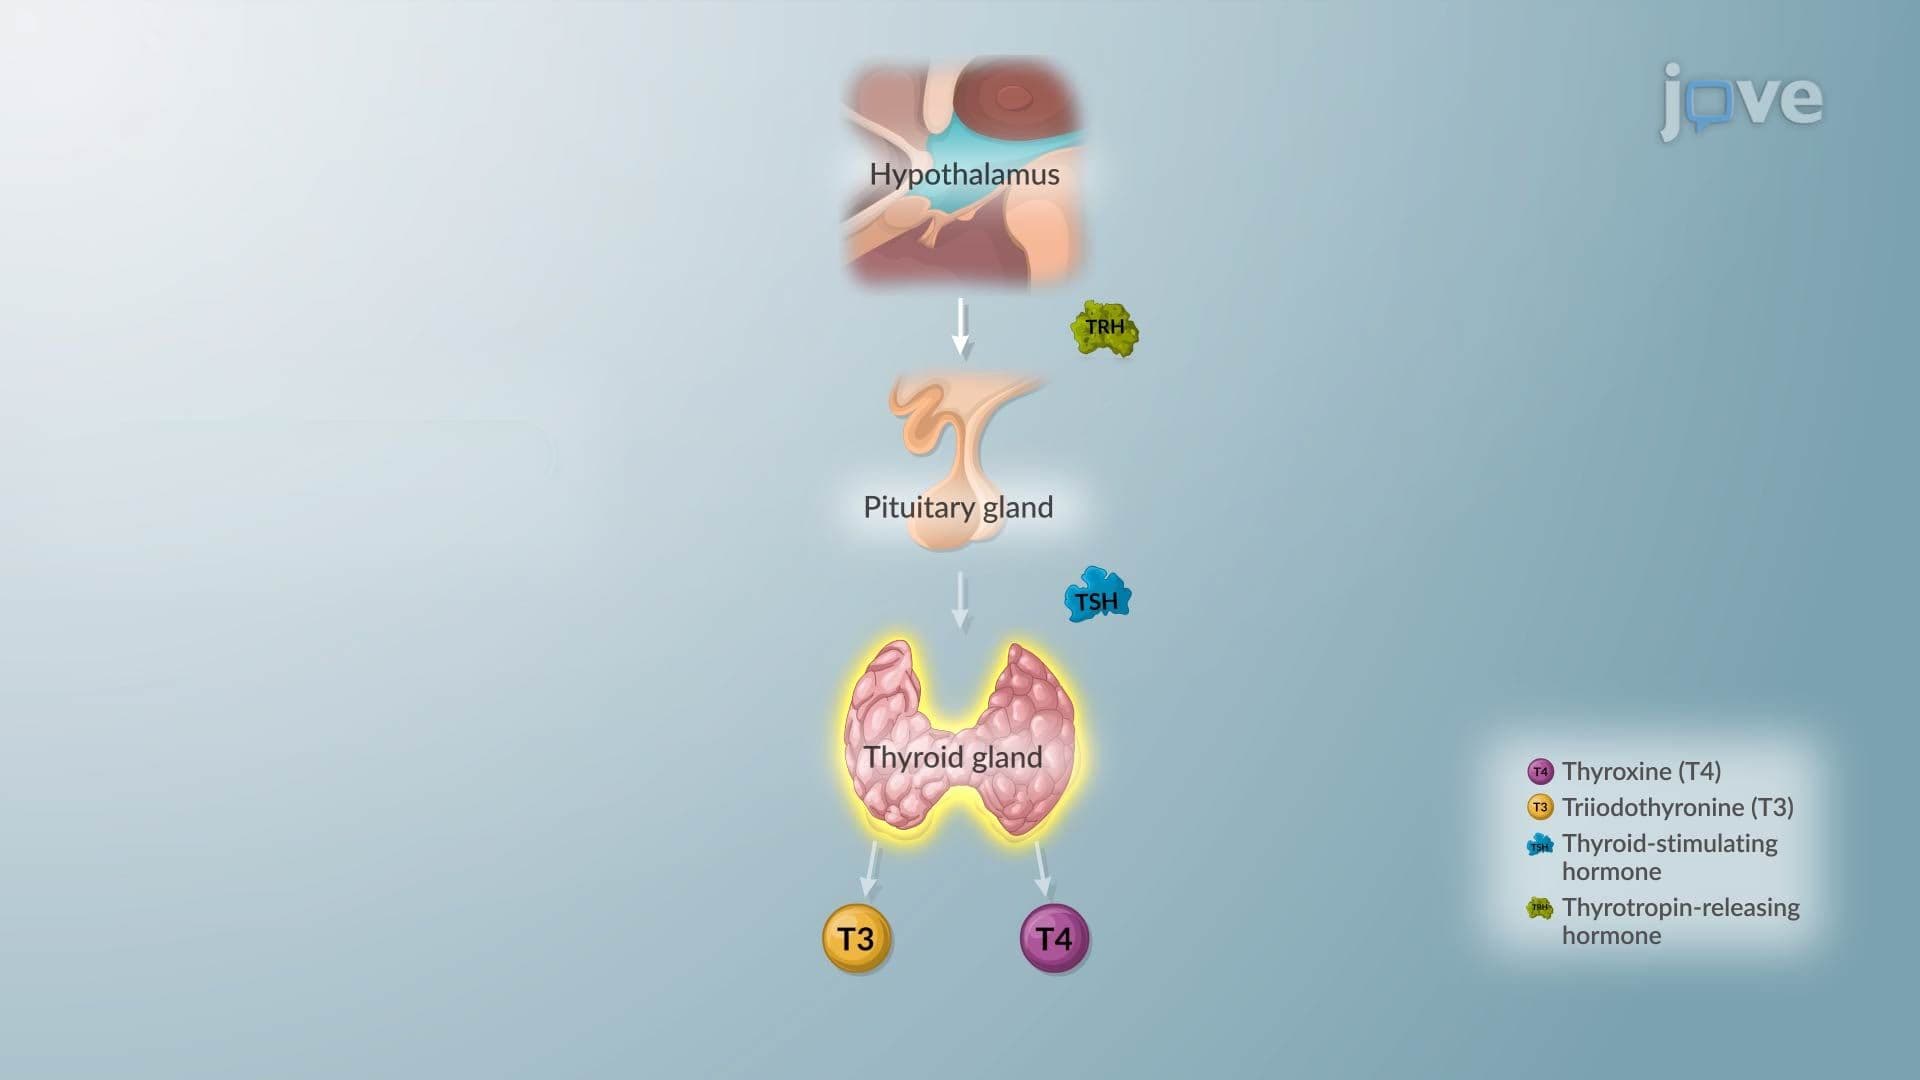
Hypothyroidism II: Pathophysiology
 01:43
Incomplete Dominance
 01:35
Pedigree Analysis
 02:30
Genomic Imprinting and Inheritance
The expression of some genes depends on which parent passed the gene to the offspring, through a phenomenon known as...
 01:20
Synthesis and Regulation of Thyroid Hormones
Upon reaching the thyroid gland, TSH stimulates the follicular cells' active uptake of iodide ions from the blood. The ions diffuse to the apical surface of the cells and are oxidized to iodine. The...
 01:17
Karyotyping