Related Experiment Video
Updated: May 5, 2026


Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Accuracy assessment and automation of free energy calculations for drug design
1Department of Lead Identification and Optimization Support, Boehringer Ingelheim Pharma GmbH & Co. KG , Biberach, 88397 Germany.

Abstract:
As the free energy of binding of a ligand to its target is one of the crucial optimization parameters in drug design, its accurate prediction is highly desirable. In the present study we have assessed the average accuracy of free energy calculations for a total of 92 ligands binding to five different targets. To make this study and future larger scale applications possible we automated the setup procedure. Starting from user defined binding modes, the procedure decides which ligands to connect via a perturbation based on maximum common substructure criteria and produces all necessary parameter files for free energy calculations in AMBER 11. For the systems investigated, errors due to insufficient sampling were found to be substantial in some cases whereas differences in estimators (thermodynamic integration (TI) versus multistate Bennett acceptance ratio (MBAR)) were found to be negligible. Analytical uncertainty estimates calculated from a single free energy calculation were found to be much smaller than the sample standard deviation obtained from two independent free energy calculations. Agreement with experiment was found to be system dependent ranging from excellent to mediocre (RMSE = [0.9, 8.2, 4.7, 5.7, 8.7] kJ/mol). When restricting analyses to free energy calculations with sample standard deviations below 1 kJ/mol agreement with experiment improved (RMSE = [0.8, 6.9, 1.8, 3.9, 5.6] kJ/mol).
More Related Videos

Related Concept Videos
 02:18
The Equilibrium Binding Constant and Binding Strength
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
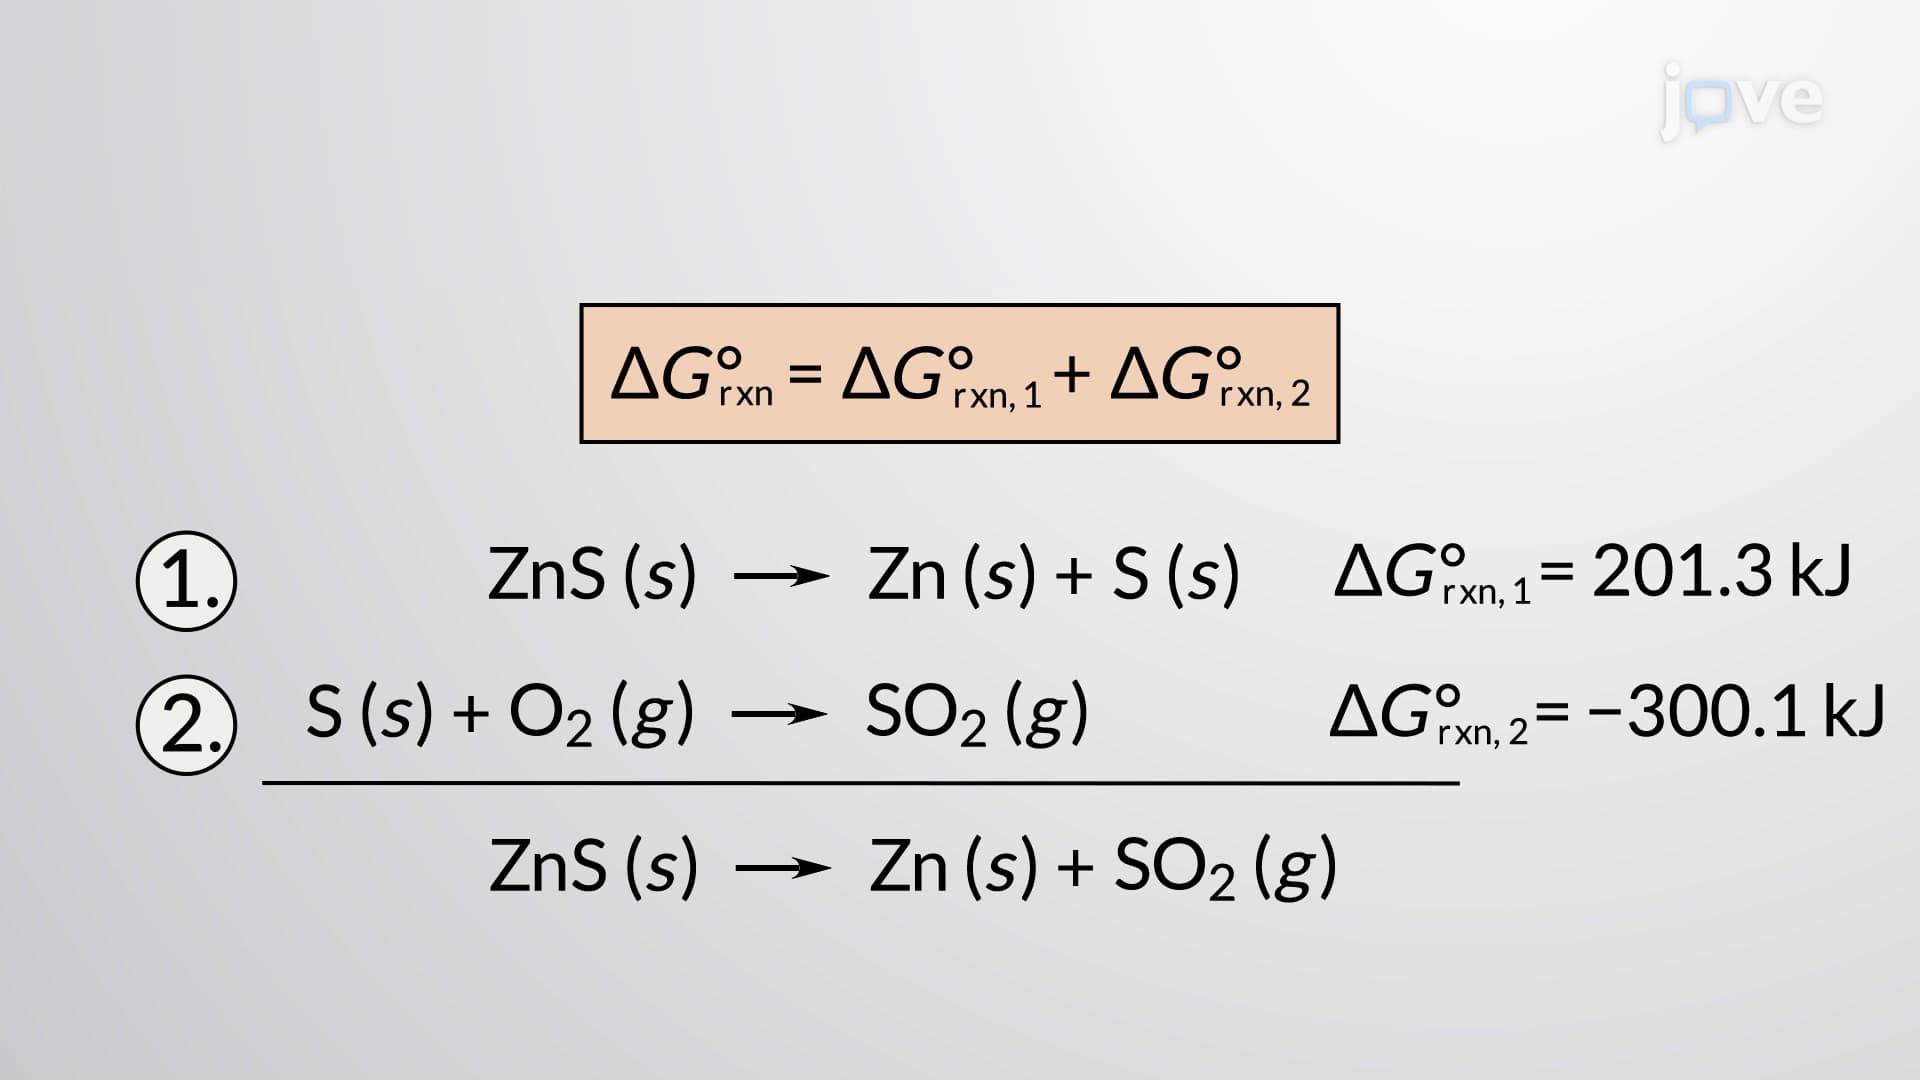 02:49
Calculating Standard Free Energy Changes
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 01:22
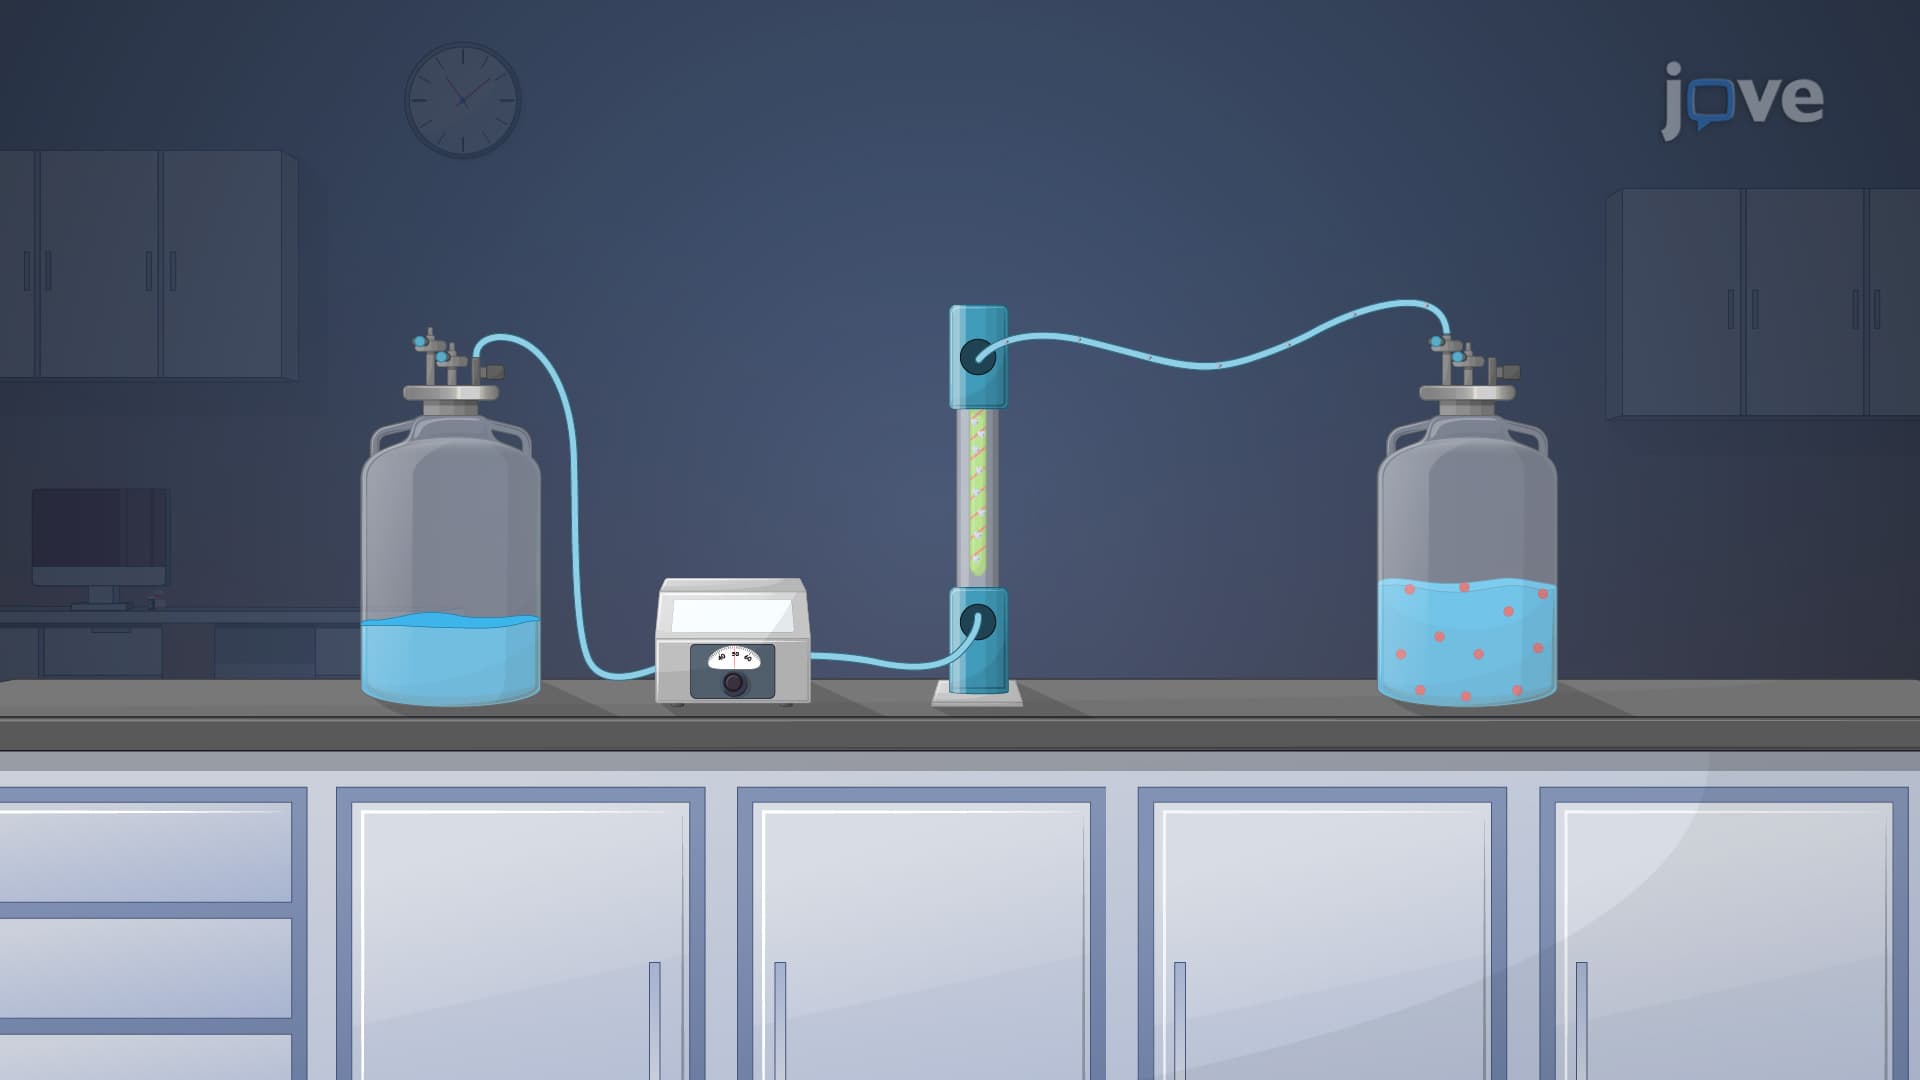
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...