Alignment-free phylogenetics and population genetics
Related Experiment Video
Updated: May 5, 2026
Using Phylogenetic Analysis to Investigate Eukaryotic Gene Origin
Published on: August 14, 2018
1Corresponding author. Bernhard Haubold. haubold@evolbio.mpg.de.

Abstract:
Phylogenetics and population genetics are central disciplines in evolutionary biology. Both are based on comparative data, today usually DNA sequences. These have become so plentiful that alignment-free sequence comparison is of growing importance in the race between scientists and sequencing machines. In phylogenetics, efficient distance computation is the major contribution of alignment-free methods. A distance measure should reflect the number of substitutions per site, which underlies classical alignment-based phylogeny reconstruction. Alignment-free distance measures are either based on word counts or on match lengths, and I apply examples of both approaches to simulated and real data to assess their accuracy and efficiency. While phylogeny reconstruction is based on the number of substitutions, in population genetics, the distribution of mutations along a sequence is also considered. This distribution can be explored by match lengths, thus opening the prospect of alignment-free population genomics.


Related Concept Videos
 02:54
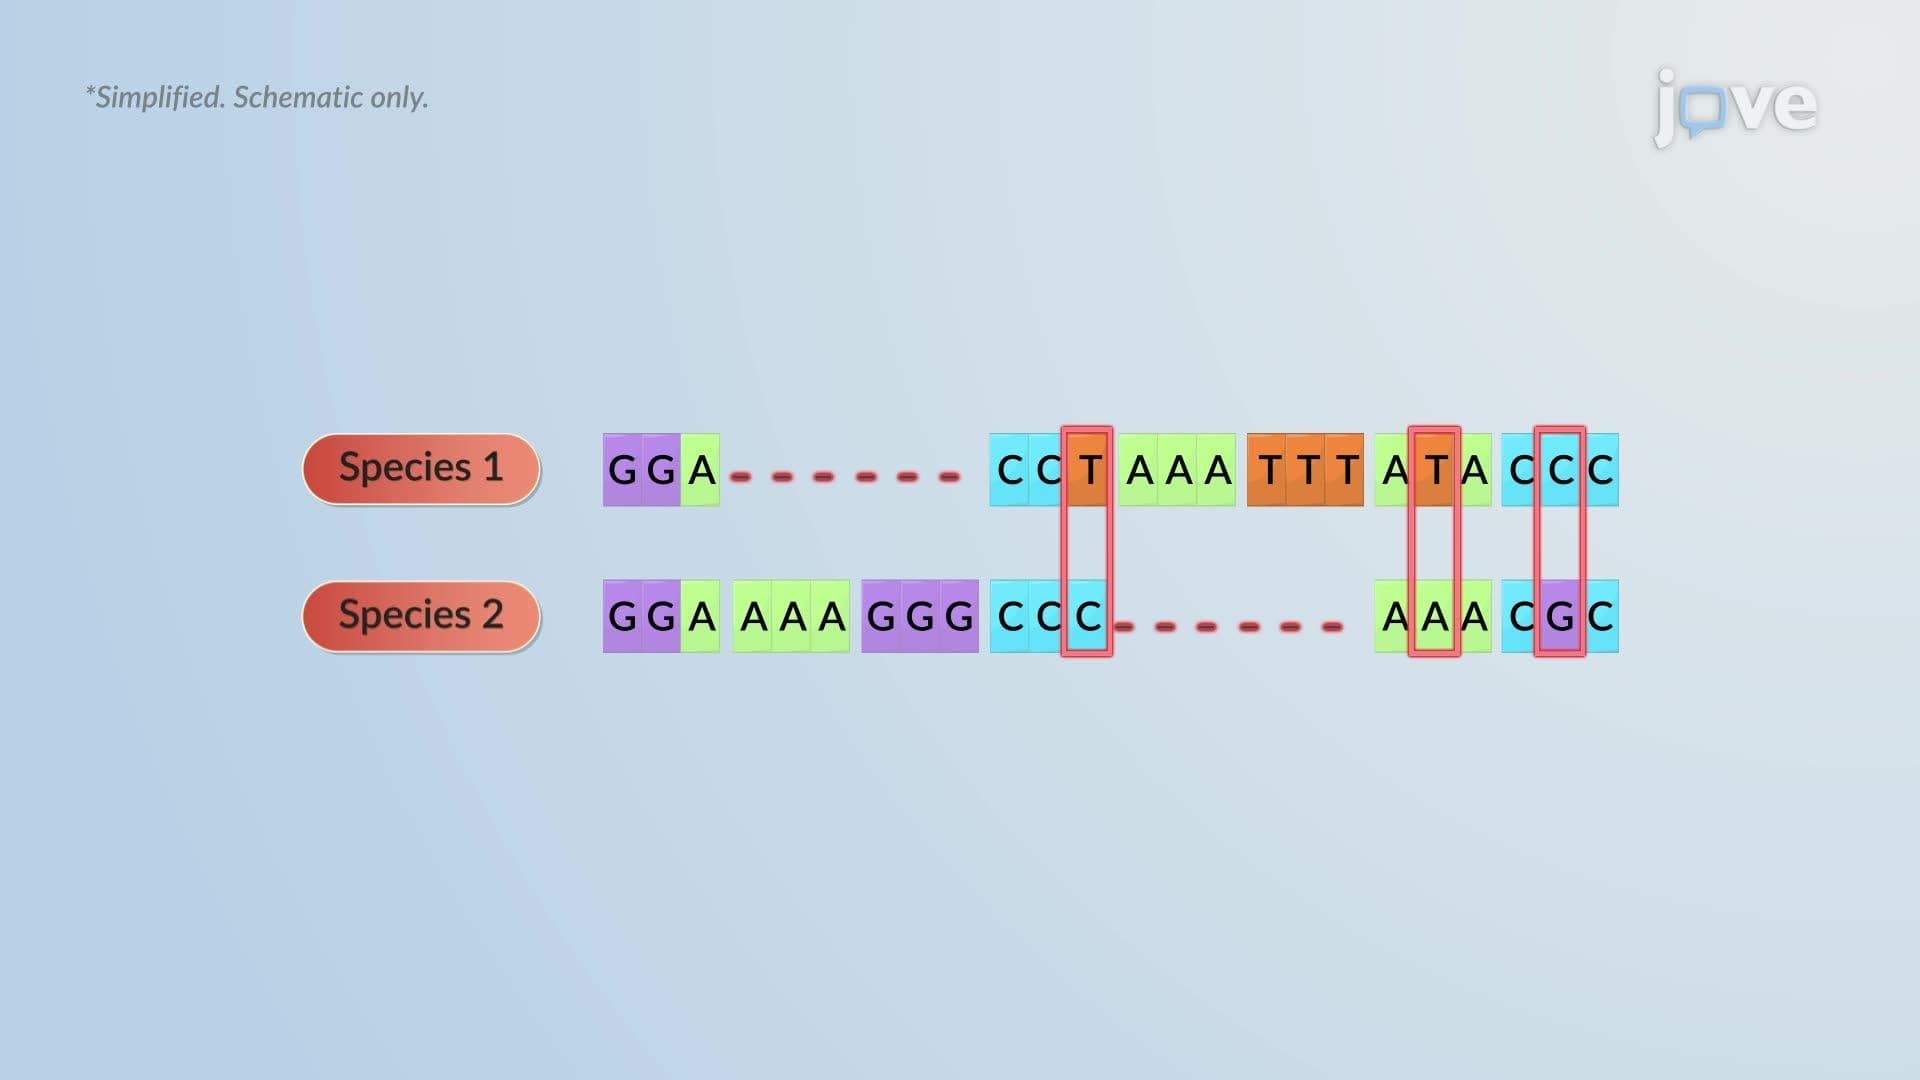
Evolutionary Relationships through Genome Comparisons
 01:28
Microbial Phylogeny
 02:05
Gene Evolution - Fast or Slow?
In contrast, regions which code...
02:05Gene Evolution - Fast or Slow?
 01:25
What is Population Genetics?
 01:23
Phylogeny