Related Experiment Video
Updated: May 5, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Linear-scaling explicitly correlated treatment of solids: periodic local MP2-F12 method
1Institute of Physical and Theoretical Chemistry, University of Regensburg, Universitätsstraße 31, D-93040 Regensburg, Germany.

Abstract:
Theory and implementation of the periodic local MP2-F12 method in the 3*A fixed-amplitude ansatz is presented. The method is formulated in the direct space, employing local representation for the occupied, virtual, and auxiliary orbitals in the form of Wannier functions (WFs), projected atomic orbitals (PAOs), and atom-centered Gaussian-type orbitals, respectively. Local approximations are introduced, restricting the list of the explicitly correlated pairs, as well as occupied, virtual, and auxiliary spaces in the strong orthogonality projector to the pair-specific domains on the basis of spatial proximity of respective orbitals. The 4-index two-electron integrals appearing in the formalism are approximated via the direct-space density fitting technique. In this procedure, the fitting orbital spaces are also restricted to local fit-domains surrounding the fitted densities. The formulation of the method and its implementation exploits the translational symmetry and the site-group symmetries of the WFs. Test calculations are performed on LiH crystal. The results show that the periodic LMP2-F12 method substantially accelerates basis set convergence of the total correlation energy, and even more so the correlation energy differences. The resulting energies are quite insensitive to the resolution-of-the-identity domain sizes and the quality of the auxiliary basis sets. The convergence with the orbital domain size is somewhat slower, but still acceptable. Moreover, inclusion of slightly more diffuse functions, than those usually used in the periodic calculations, improves the convergence of the LMP2-F12 correlation energy with respect to both the size of the PAO-domains and the quality of the orbital basis set. At the same time, the essentially diffuse atomic orbitals from standard molecular basis sets, commonly utilized in molecular MP2-F12 calculations, but problematic in the periodic context, are not necessary for LMP2-F12 treatment of crystals.
More Related Videos

Related Concept Videos
 01:26
Scaling
 02:37
Metallic Solids
All metallic solids exhibit high thermal and electrical conductivity, metallic luster, and...
 01:07
Conservation of Mass in Fixed, Nondeforming Control Volume
In the case of a sewer pipe, which can be modeled...
 01:24
Solid–Solid Solutions
 01:12
Reduced Mass Coordinates: Isolated Two-body Problem
 01:15
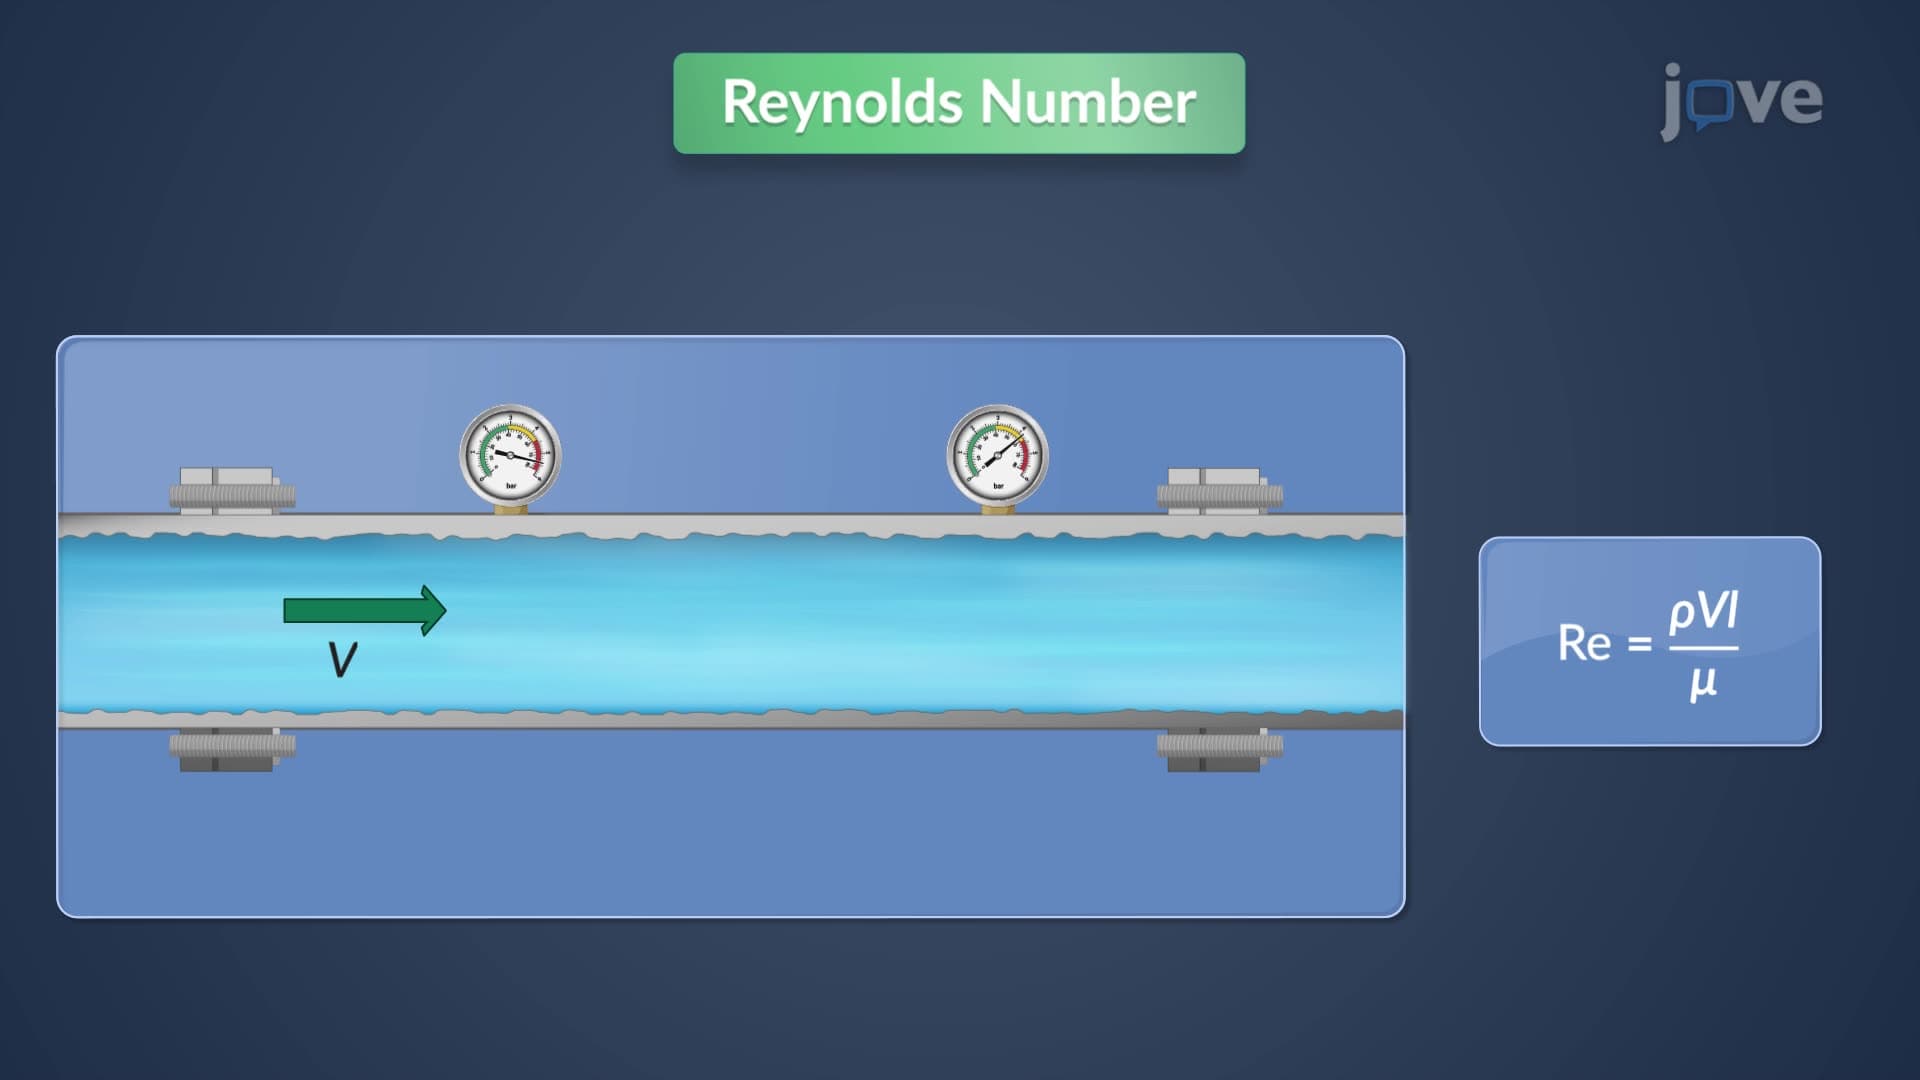
Dimensionless Groups in Fluid Mechanics