Related Experiment Video
Updated: May 4, 2026


Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
Applying physics-based scoring to calculate free energies of binding for single amino acid mutations in
Hege Beard1, Anuradha Cholleti, David Pearlman
1Schrödinger, New York, New York, United States of America.

Abstract:
Predicting changes in protein binding affinity due to single amino acid mutations helps us better understand the driving forces underlying protein-protein interactions and design improved biotherapeutics. Here, we use the MM-GBSA approach with the OPLS2005 force field and the VSGB2.0 solvent model to calculate differences in binding free energy between wild type and mutant proteins. Crucially, we made no changes to the scoring model as part of this work on protein-protein binding affinity--the energy model has been developed for structure prediction and has previously been validated only for calculating the energetics of small molecule binding. Here, we compare predictions to experimental data for a set of 418 single residue mutations in 21 targets and find that the MM-GBSA model, on average, performs well at scoring these single protein residue mutations. Correlation between the predicted and experimental change in binding affinity is statistically significant and the model performs well at picking "hotspots," or mutations that change binding affinity by more than 1 kcal/mol. The promising performance of this physics-based method with no tuned parameters for predicting binding energies suggests that it can be transferred to other protein engineering problems.
More Related Videos


Related Concept Videos
 02:18
The Equilibrium Binding Constant and Binding Strength
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:04
Protein-protein Interfaces
 01:22
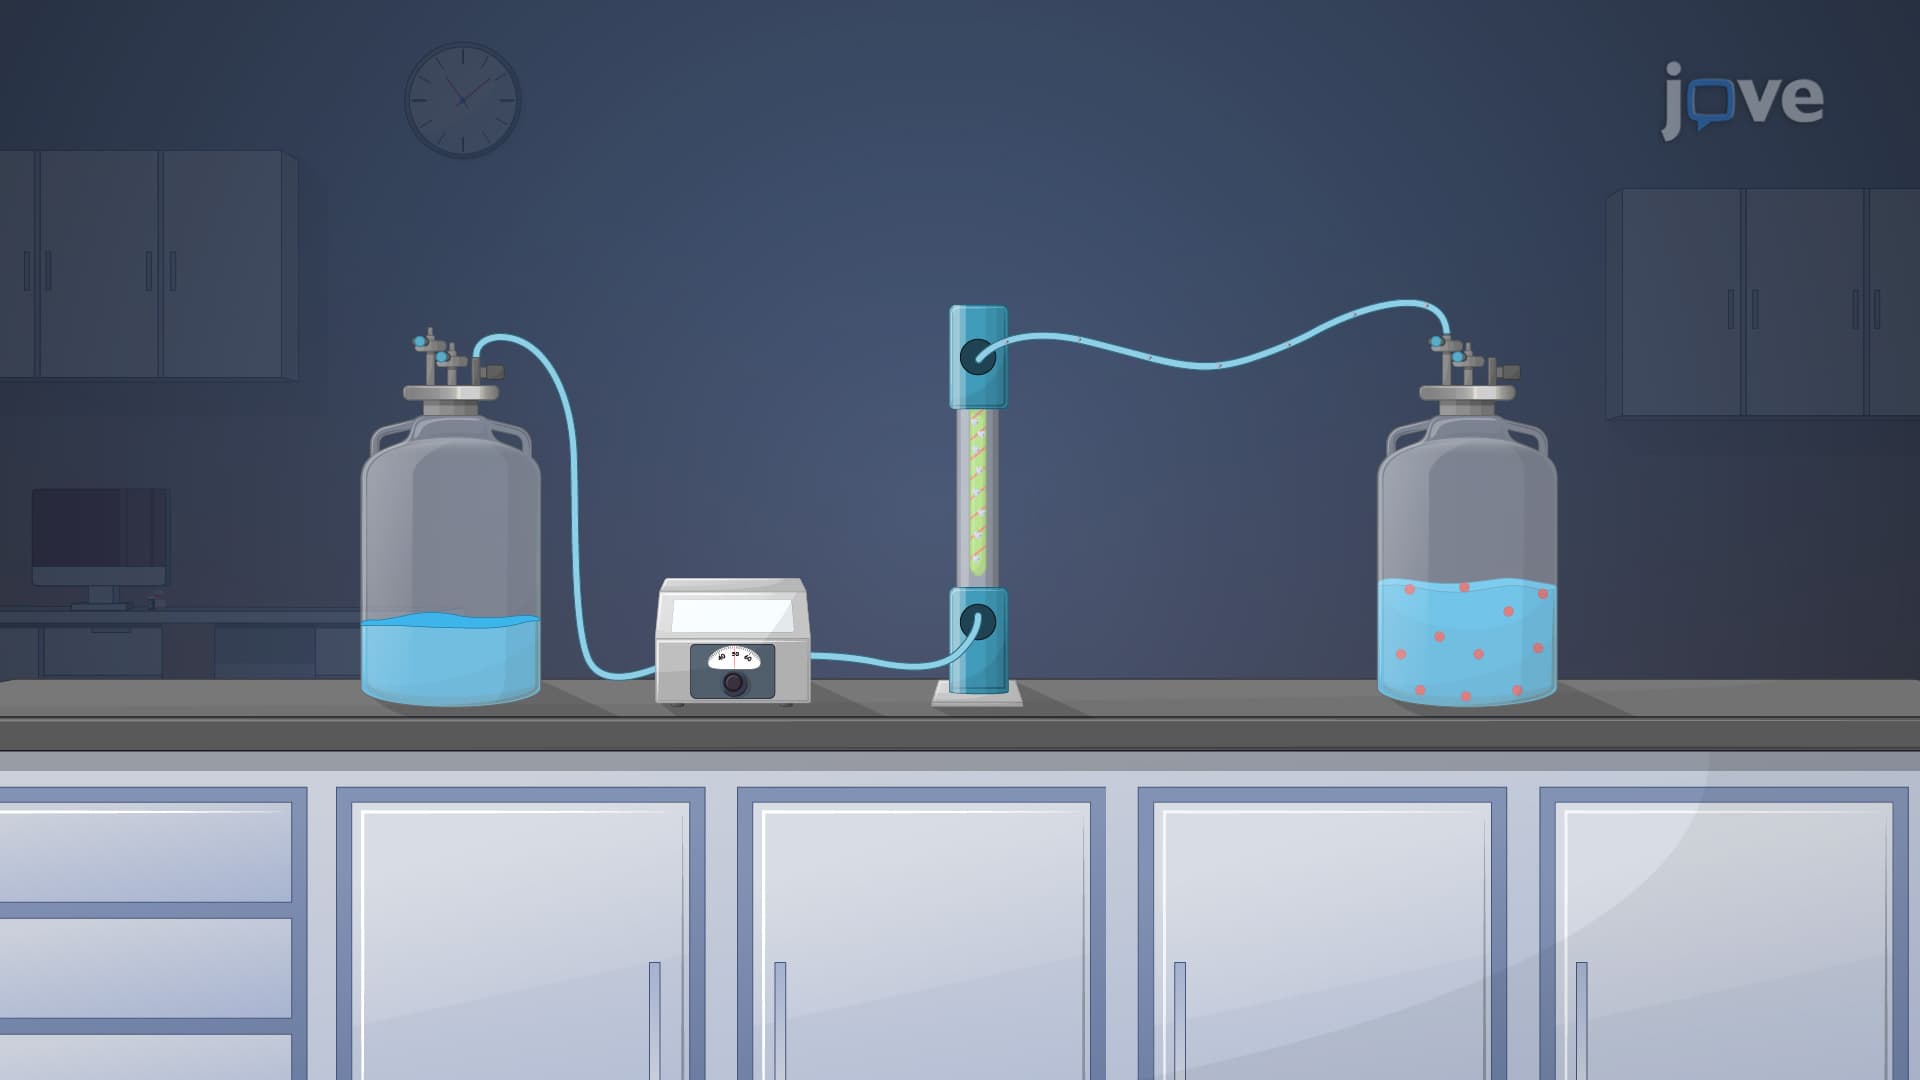
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...