Related Experiment Video
Updated: May 4, 2026
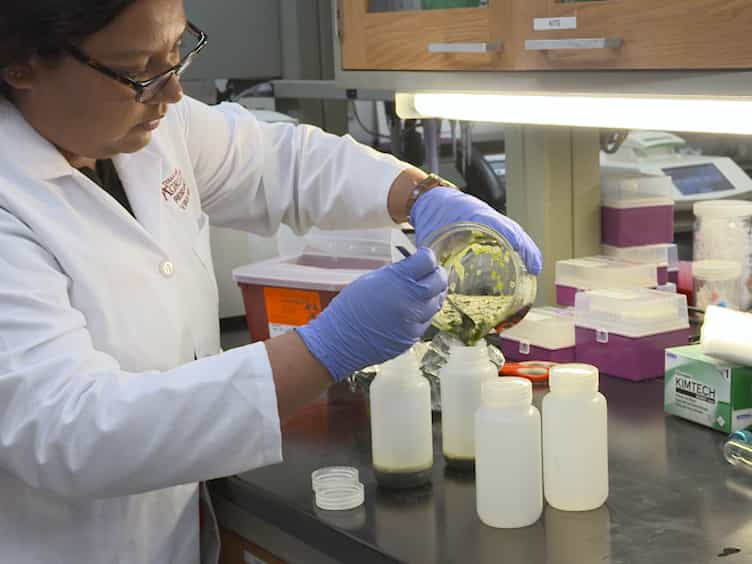

Combining Analysis of DNA in a Crude Virion Extraction with the Analysis of RNA from Infected Leaves to Discover New Virus Genomes
Published on: July 27, 2018
A phylogeographical study of the cauliflower mosaic virus population in mid-Eurasia Iran using complete genome
Shirin Farzadfar1, Reza Pourrahim, Hassan Ebrahimi
1Department of Plant Virus Research, Iranian Research Institute of Plant Protection, P.O.Box 19395-1454, Chamran Highway, Tehran, Iran, farzadfar2002@yahoo.com.

Abstract:
The full-length sequences of 34 Iranian cauliflower mosaic virus (CaMV) isolates were compared with others from public nucleotide sequence databases to provide a comprehensive overview of the genetic variability and patterns of genetic exchange in CaMV isolates from Iran. Based on the severity of symptoms and their ability to infect Brassica oleracea var. capitata, Iranian CaMV isolates were grouped into two distinct biotypes: latent/mild mottle (LI/MMo) and severe (S) infection. Recombination breakpoints were detected between the large intergenic region (LIR) and open reading frame (ORF) V (event 2); between ORF VII and ORF II (event 3), between ORF I and ORF III (event 4), and within ORF VI (event 1). Phylogenetic analysis indicated that Iranian CaMV isolates clustered into two subgroups belonging to group I (GI) that were distinct from North American and European isolates from group II (GII). Northeast Iranian isolates (subgroup B) and CaMV isolates from subgroup A closely corresponded to the S and LI/MMo biological groups, respectively. Genome-wide pairwise identity analysis of the CaMV isolates revealed three regions of pairwise identity representation: 92-94 % for GII and 94-96 % and 98-100 % for subgroups A and B. The within-population diversity was lower than the between-population diversity, suggesting the contribution of a founder effect on diversification of CaMV isolates. Amino acid sequences were conserved, with ω values ranging from 0.074 to 0.717 in different proteins. Thirteen amino acids in the deduced proteins of ORFs I, II, III, VI and VII were under positive selection (ω > 1), whereas purifying selection applied to the proteins encoded by ORFs IV and V. This study suggests that variation in the CaMV population can be explained by host-range differentiation and selection pressure. Moreover, recombination analysis revealed that a genomic exchange is responsible for the emergence of CaMV strains, providing valuable new information for understanding the diversity and evolution of caulimoviruses.
Related Concept Videos
 01:29
Viruses of Archaea
 01:26
Human Virome
 02:39
Gene Flow
 01:29
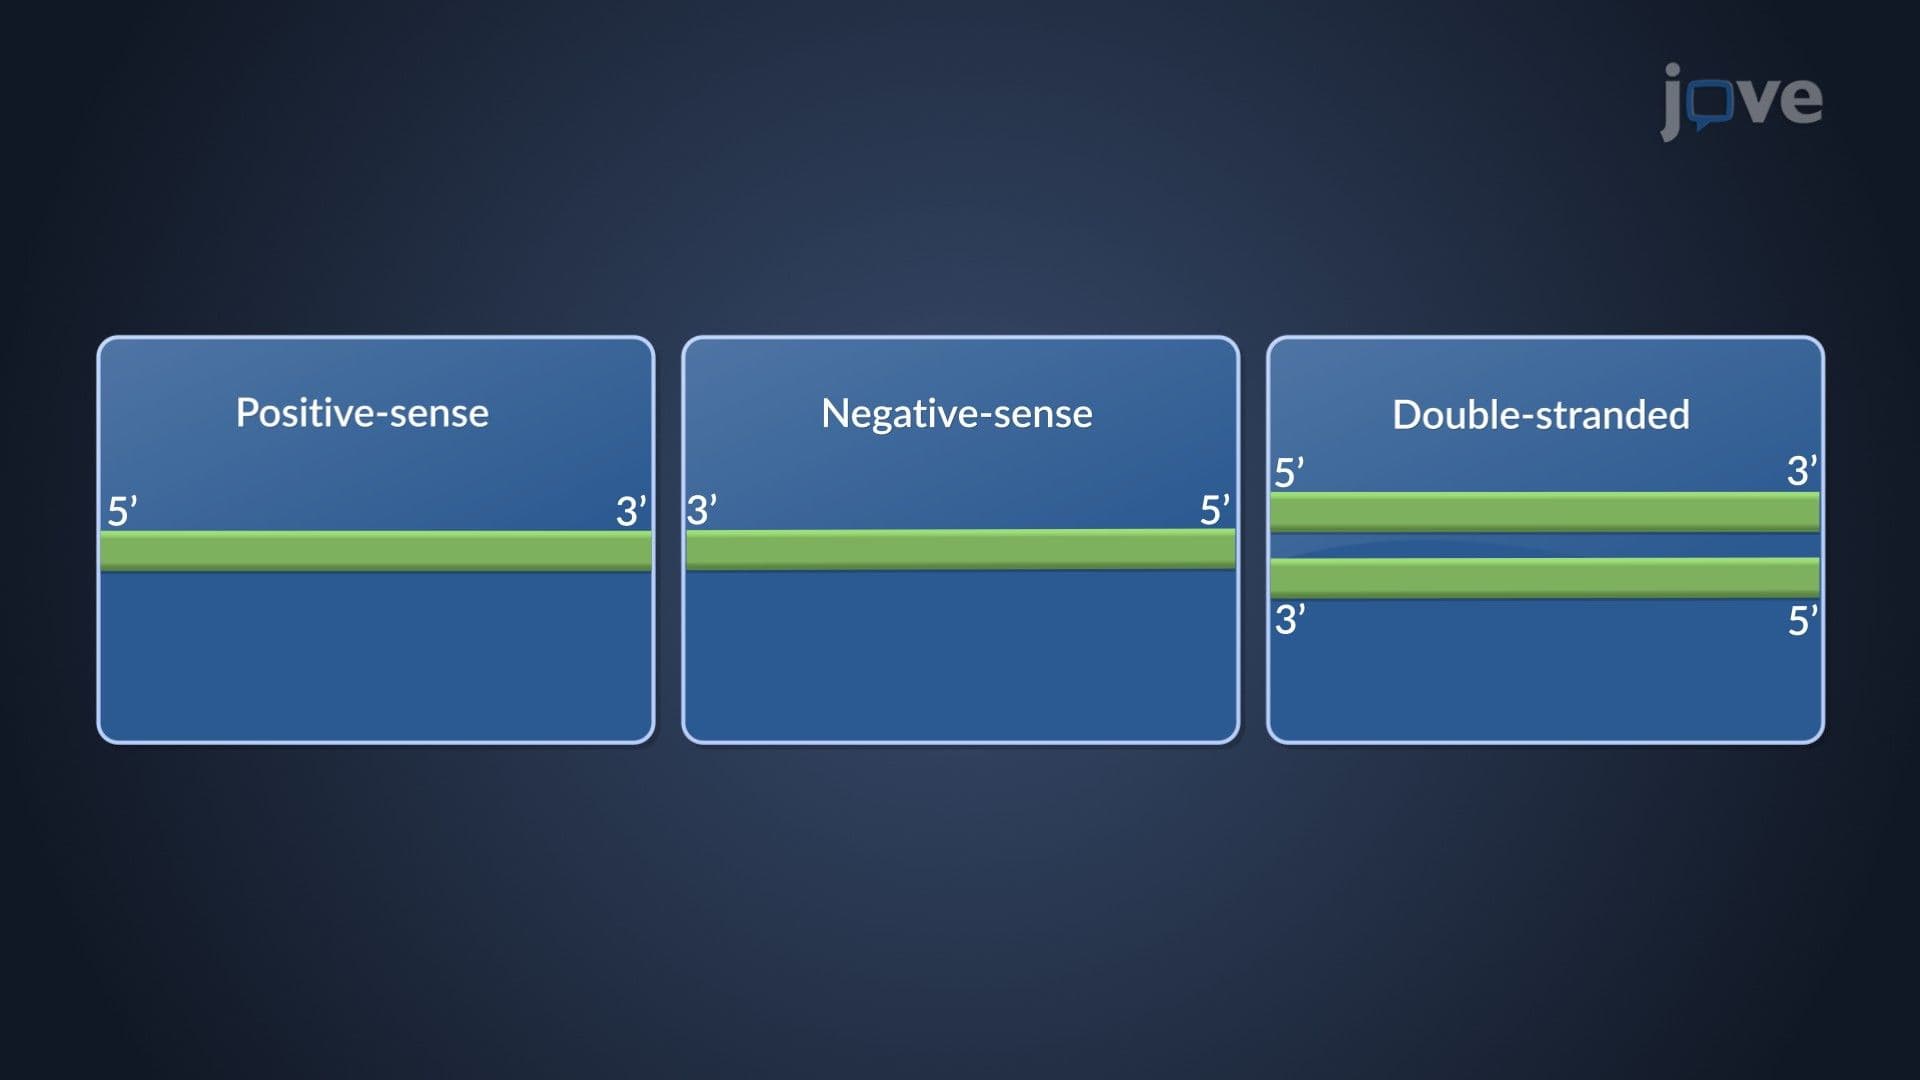
Viruses with RNA Genomes
 01:26
Size and Structure of Viral Genomes
 02:54
Evolutionary Relationships through Genome Comparisons