Ligand binding mode prediction by docking: mdm2/mdmx inhibitors as a case study
Related Experiment Video
Updated: May 4, 2026
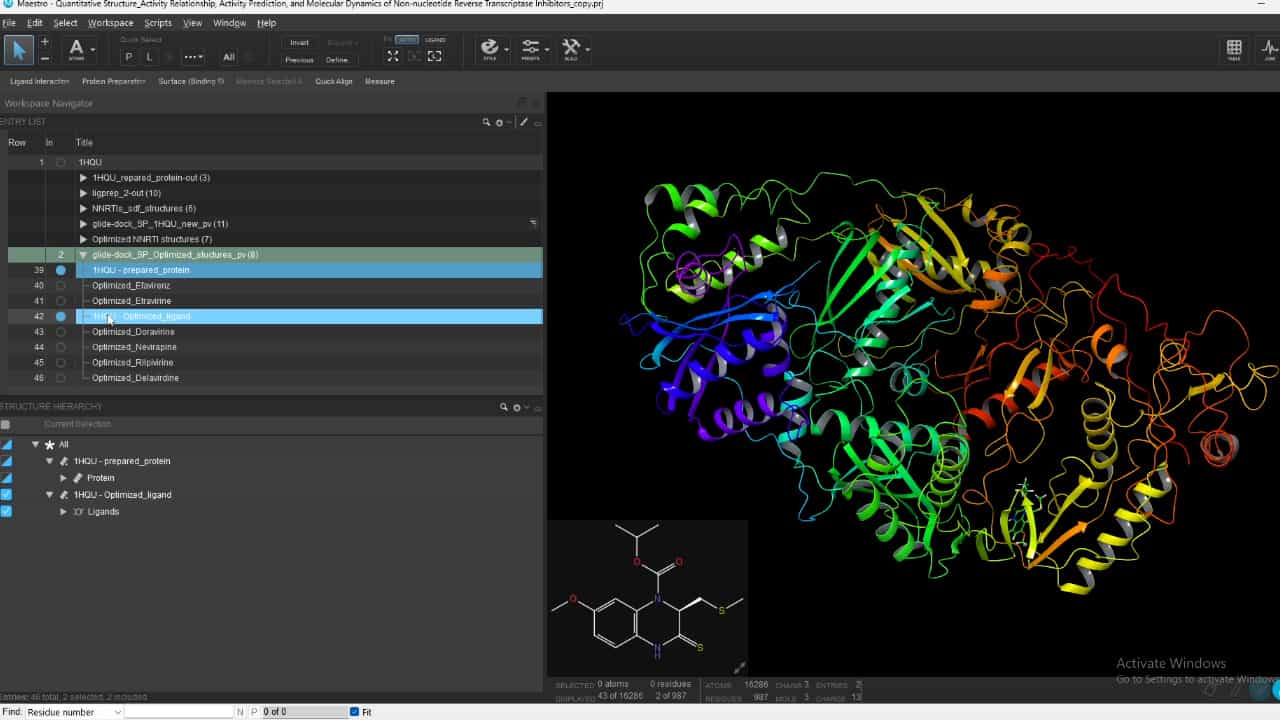
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Nagakumar Bharatham1, Kavitha Bharatham, Anang A Shelat
1Department of Structural Biology and ‡Department of Chemical Biology and Therapeutics, Saint Jude Children's Research Hospital , 262 Danny Thomas Place, Memphis, Tennessee 38105, United States.

Abstract:
The p53-binding domains of Mdm2 and Mdmx, two negative regulators of the tumor suppressor p53, are validated targets for cancer therapeutics, but correct binding poses of some proven inhibitors, particularly the nutlins, have been difficult to obtain with standard docking procedures. Virtual screening pipelines typically draw from a database of compounds represented with 1D or 2D structural information from which one or more 3D conformations must be generated. These conformations are then passed to a docking algorithm that searches for optimal binding poses on the target protein. This work tests alternative pipelines using several commonly used conformation generation programs (LigPrep, ConfGen, MacroModel, and Corina/Rotate) and docking programs (GOLD, Glide, MOE-dock, and AutoDock Vina) for their ability to reproduce known poses for a series of Mdmx and/or Mdm2 inhibitors, including several nutlins. Most combinations of these programs using default settings fail to find correct poses for the nutlins but succeed for all other compounds. Docking success for the nutlin class requires either computationally intensive conformational exploration or an "anchoring" procedure that incorporates knowledge of the orientation of the central imidazoline ring.
More Related Videos
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
 10:21
Author Spotlight: Streamlining Protein Target Prediction and Validation via Molecular Docking and CETSA
Published on: February 23, 2024
Related Concept Videos
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
02:40Ligand Binding Sites
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:18
The Equilibrium Binding Constant and Binding Strength
 00:49
Ligand Binding and Linkage