Related Experiment Video
Updated: May 4, 2026


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Analytic density functional theory calculations of pure vibrational hyperpolarizabilities: the first dipole
Bin Gao1, Magnus Ringholm, Radovan Bast
1Centre for Theoretical and Computational Chemistry, Department of Chemistry, University of Tromsø-The Arctic University of Norway , N-9037 Tromsø, Norway.

Abstract:
We present a general approach for the analytic calculation of pure vibrational contributions to the molecular (hyper)polarizabilities at the density functional level of theory. The analytic approach allows us to study large molecules, and we apply the new code to the study of the first dipole hyperpolarizabilities of retinal and related molecules. We investigate the importance of electron correlation as described by the B3LYP exchange-correlation functional on the pure vibrational and electronic hyperpolarizabilities and compare the computed hyperpolarizabilities with available experimental data. The effects of electron correlation on the pure vibrational corrections vary signficantly even between these structurally very similar molecules, making it difficult to estimate these effects without explicit calculations at the density functional theory level. As expected, the frequency-dependent first hyperpolarizability, which determines the experimentally observed second-harmonic generation, is dominated by the electronic term, whereas for the static hyperpolarizability, the vibrational contribution is equally important. As a consequence, frequency extrapolation of the measured optical hyperpolarizabilities can only provide an estimate for the electronic contribution to the static hyperpolarizability, not its total value. The relative values of the hyperpolarizabilities for different molecules, obtained from the calculations, are in reasonable agreement with experimental data.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 08:49
Author Spotlight: Unveiling the Potential of VSFG Microscopy in Studying Mesoscopically Heterogeneous Self-Assembled Structures
Published on: December 1, 2023
Related Concept Videos
 01:29
Potential Due to a Polarized Object
 01:20
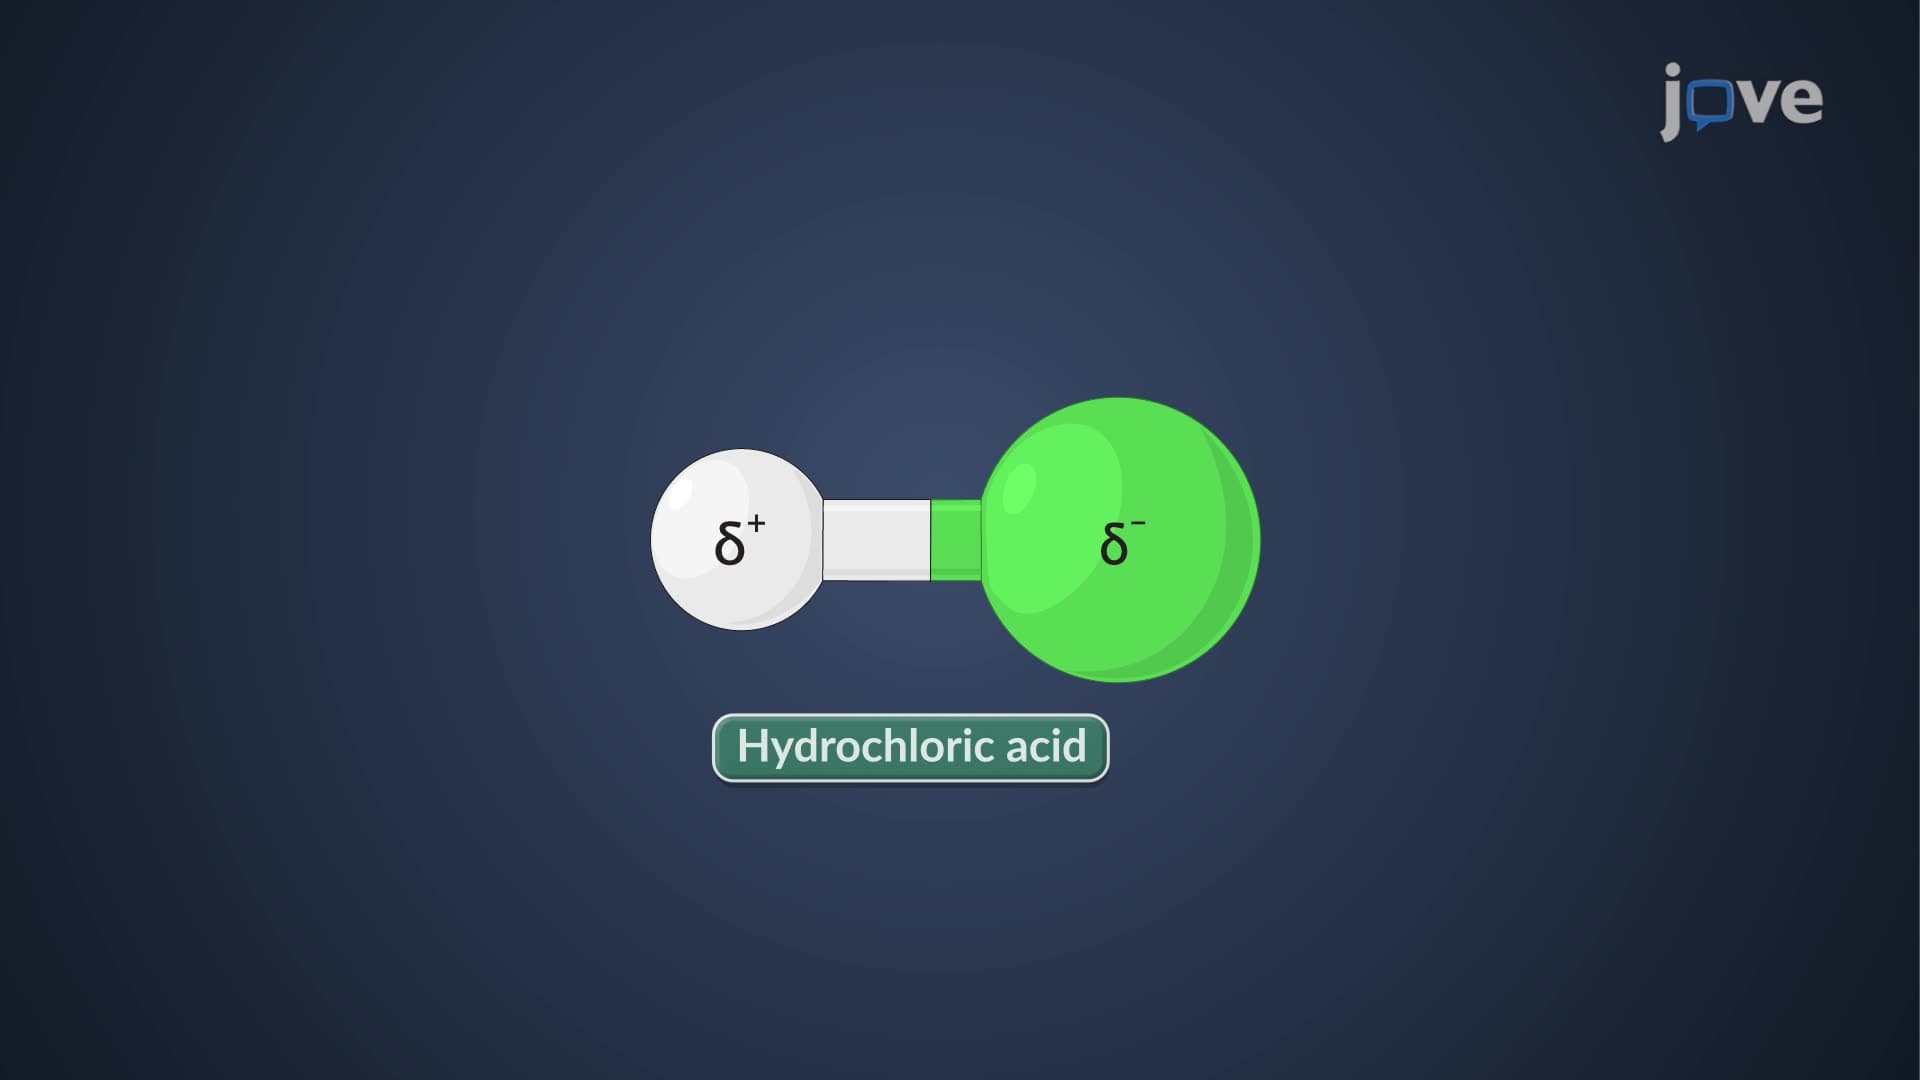
IR Spectrum Peak Intensity: Dipole Moment
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...
 01:26
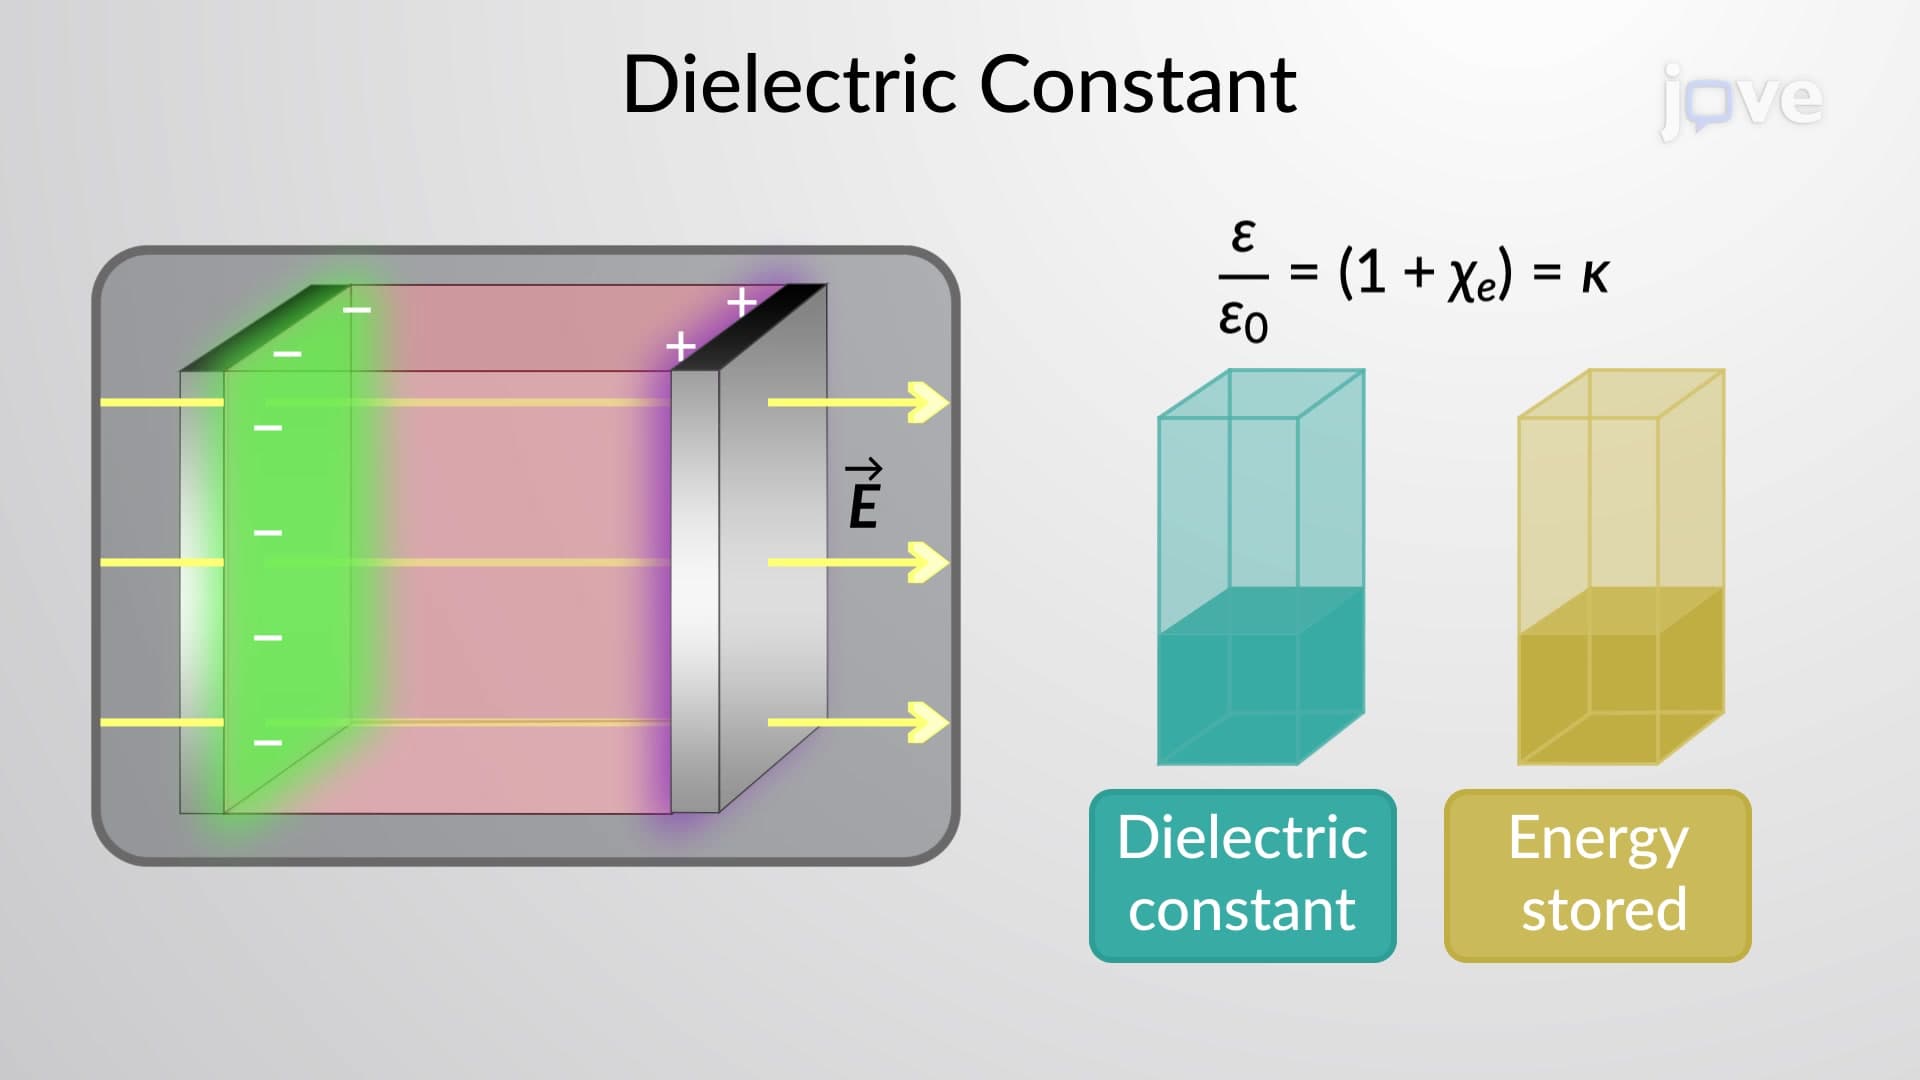
Susceptibility, Permittivity and Dielectric Constant
 02:36
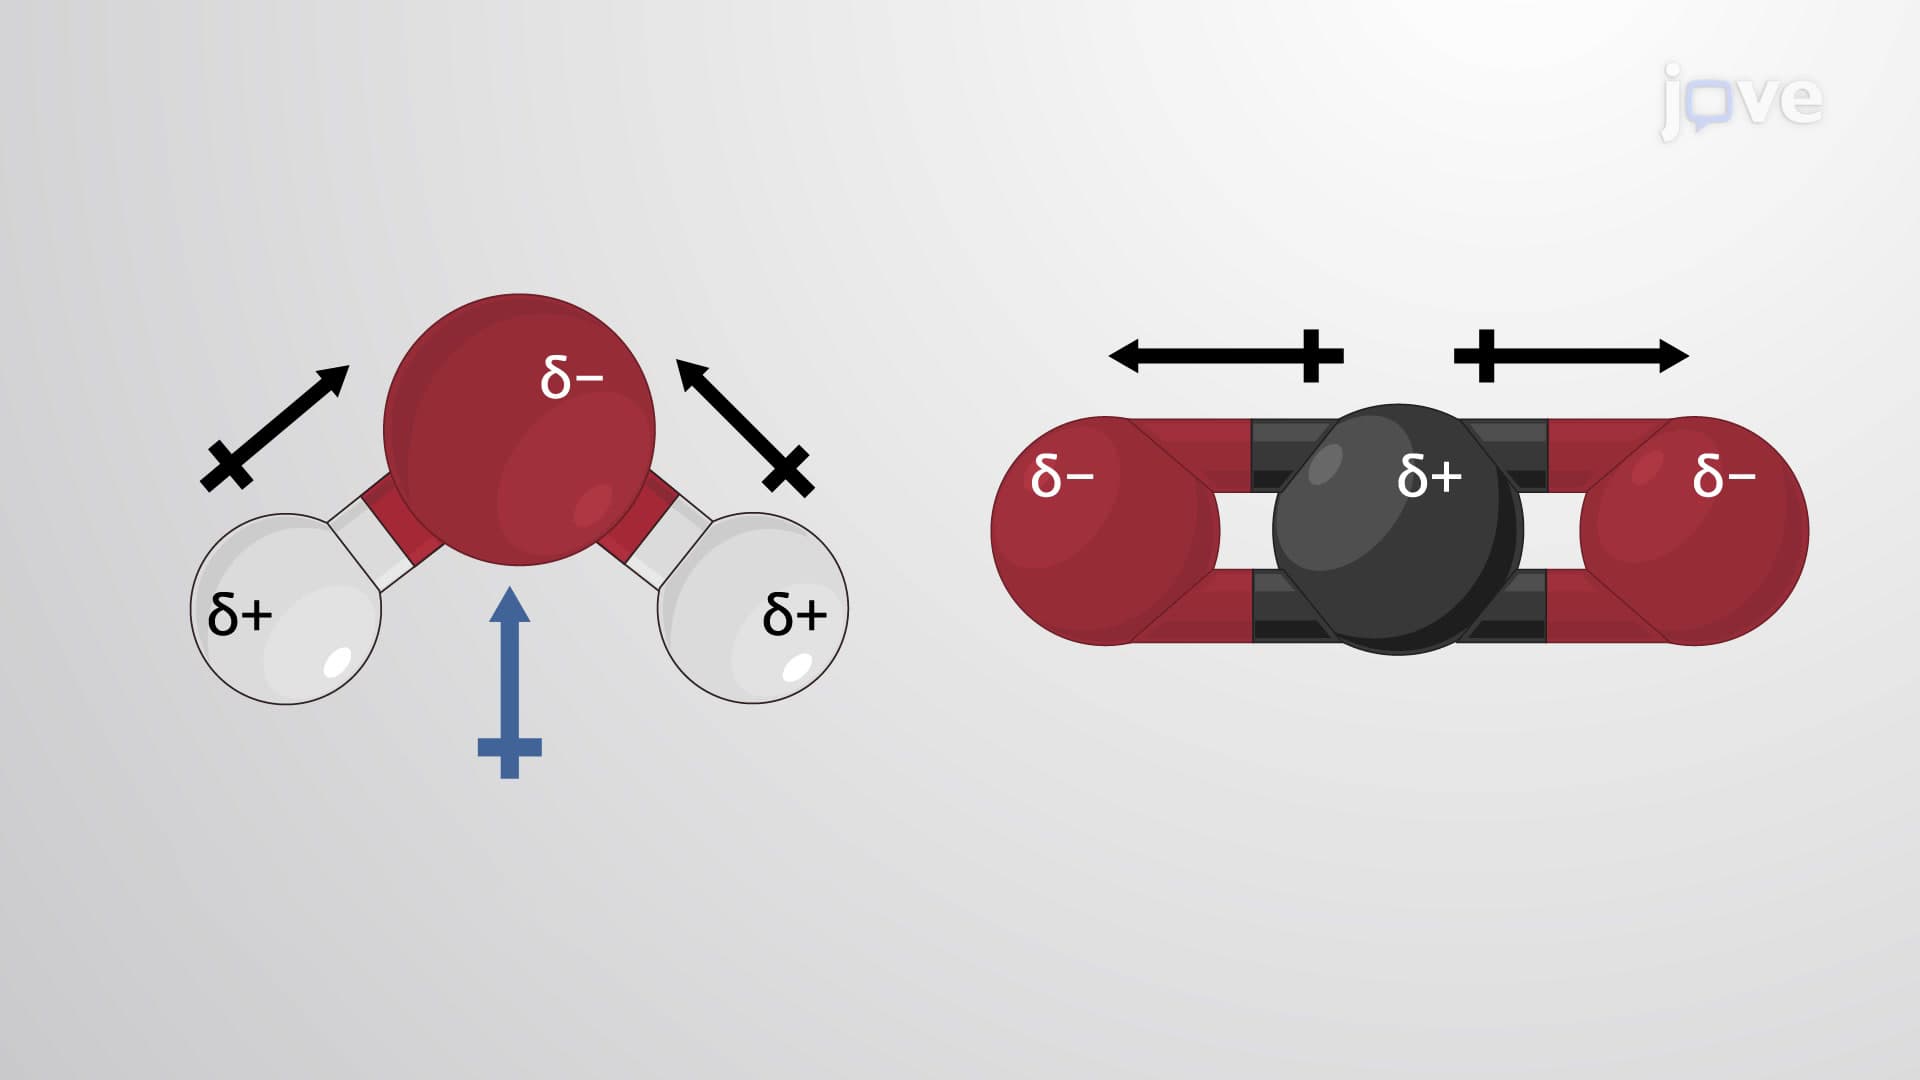
Molecular Geometry and Dipole Moments
 02:24
Properties of Enantiomers and Optical Activity