Related Experiment Video
Updated: May 4, 2026
Coulomb Explosion Imaging as a Tool to Distinguish Between Stereoisomers
Published on: August 18, 2017
DFT calculations of structure and vibrational properties of 2,2,2-trichloroethylacetate, CH(3)CO(2)CH(2)CCl(3)
Diego M Gil1, María E Tuttolomondo1, Aída Ben Altabef1
1INQUINOA, CONICET, Instituto de Química Física, Facultad de Bioquímica, Química y Farmacia, Universidad Nacional de Tucumán, San Lorenzo 456, T4000CAN Tucumán, Argentina.

Abstract:
The molecular structure and conformational properties of 2,2,2-trichloroethylacetate, CH(3)CO(2)CH(2)CCl(3), were determined by ab initio (MP2) and DFT quantum chemical calculations at different levels of theory. The theoretical study was complemented with experimental measurements such as IR and Raman spectroscopy. The experimental and calculations confirm the presence of two conformers, one with anti, gauche conformation (C1 symmetry) and another with anti, anti form (Cs symmetry). The conformational preference was studied using the total energy scheme, NBO and AIM analysis. The infrared spectra of CH(3)CO(2)CH(2)CCl(3) are reported in the liquid and solid phases and the Raman spectrum in liquid phase. Using calculated frequencies as a guide, evidence for both C1 and Cs conformers is obtained in the IR and Raman spectra.
Related Concept Videos
 02:27
Predicting Molecular Geometry
 01:29
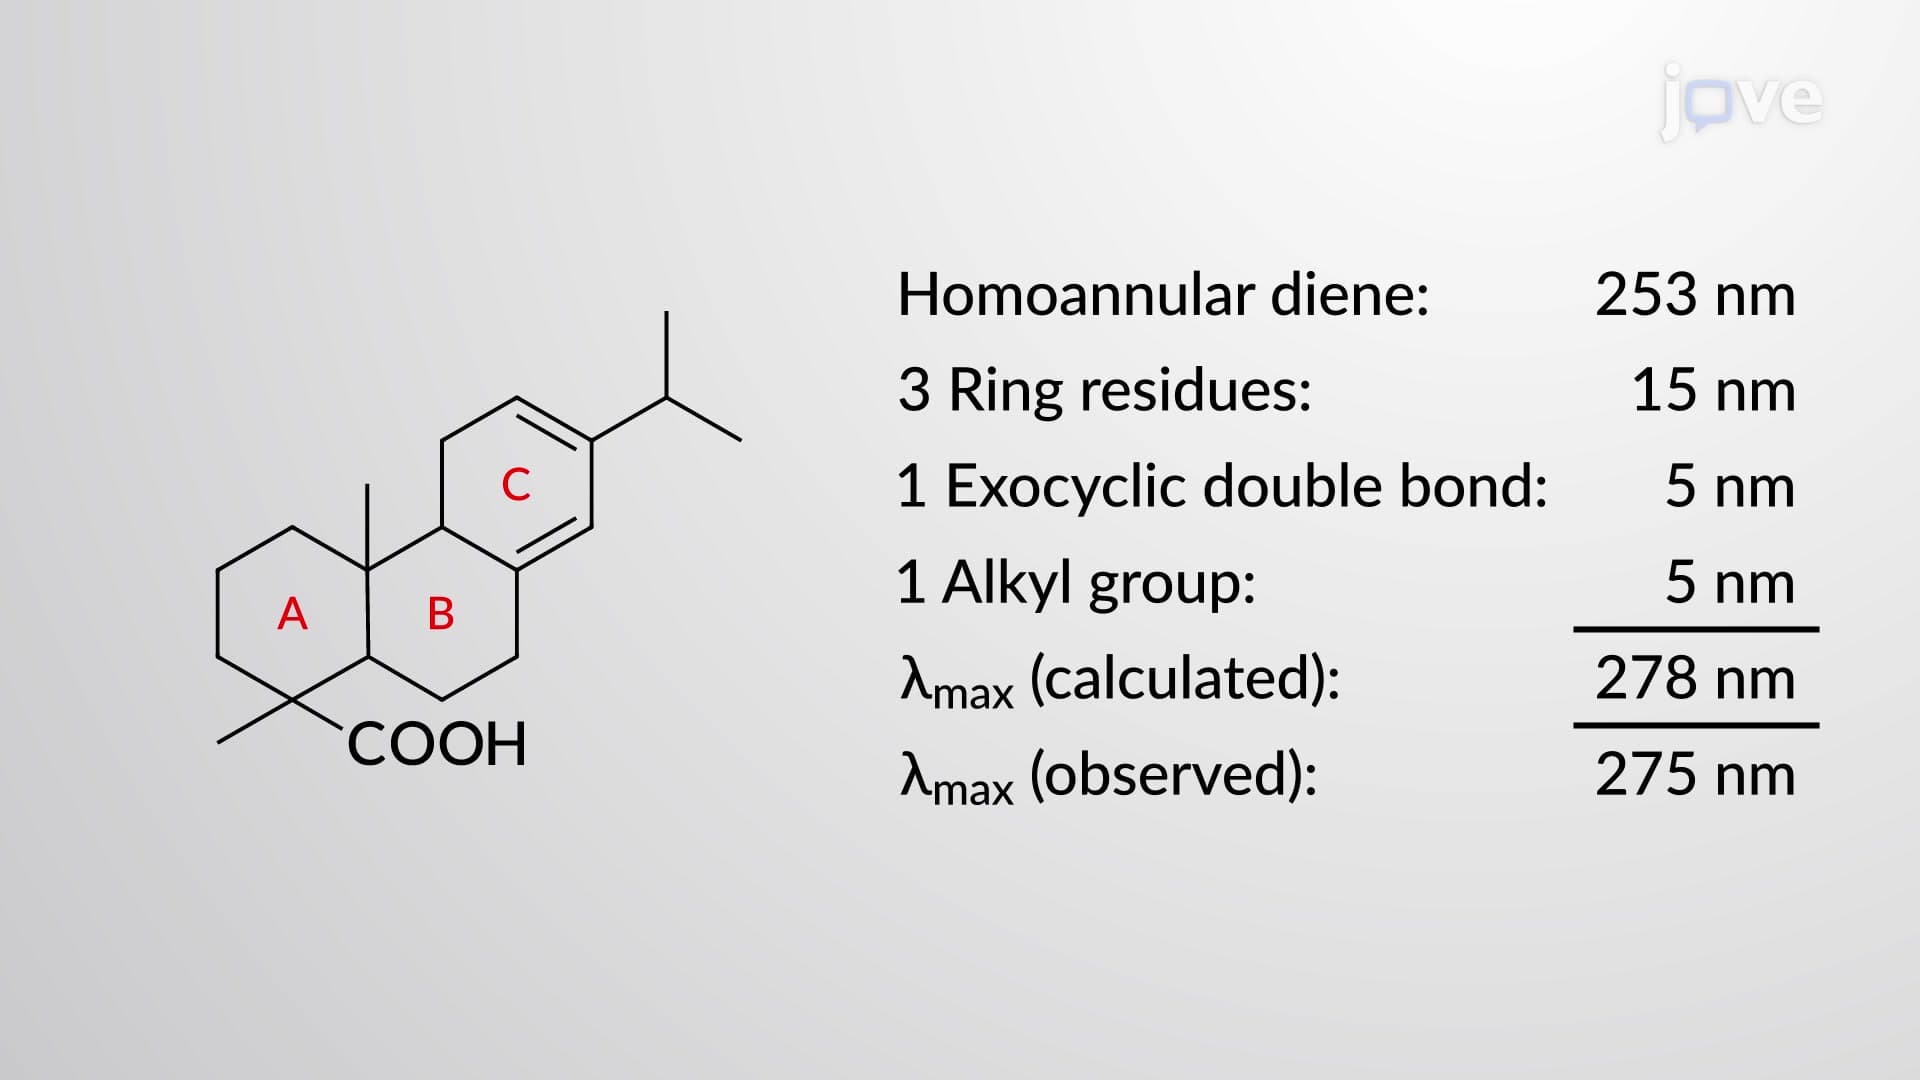
UV–Vis Spectroscopy: Woodward–Fieser Rules
 01:07
Radicals: Electronic Structure and Geometry
Accordingly, the structure of a trivalent radical lies between the geometries of carbocations and carbanions. An sp2-hybridized carbocation is trigonal planar, while an sp3-hybridized carbanion is trigonal pyramidal. Here, the difference in geometry is...
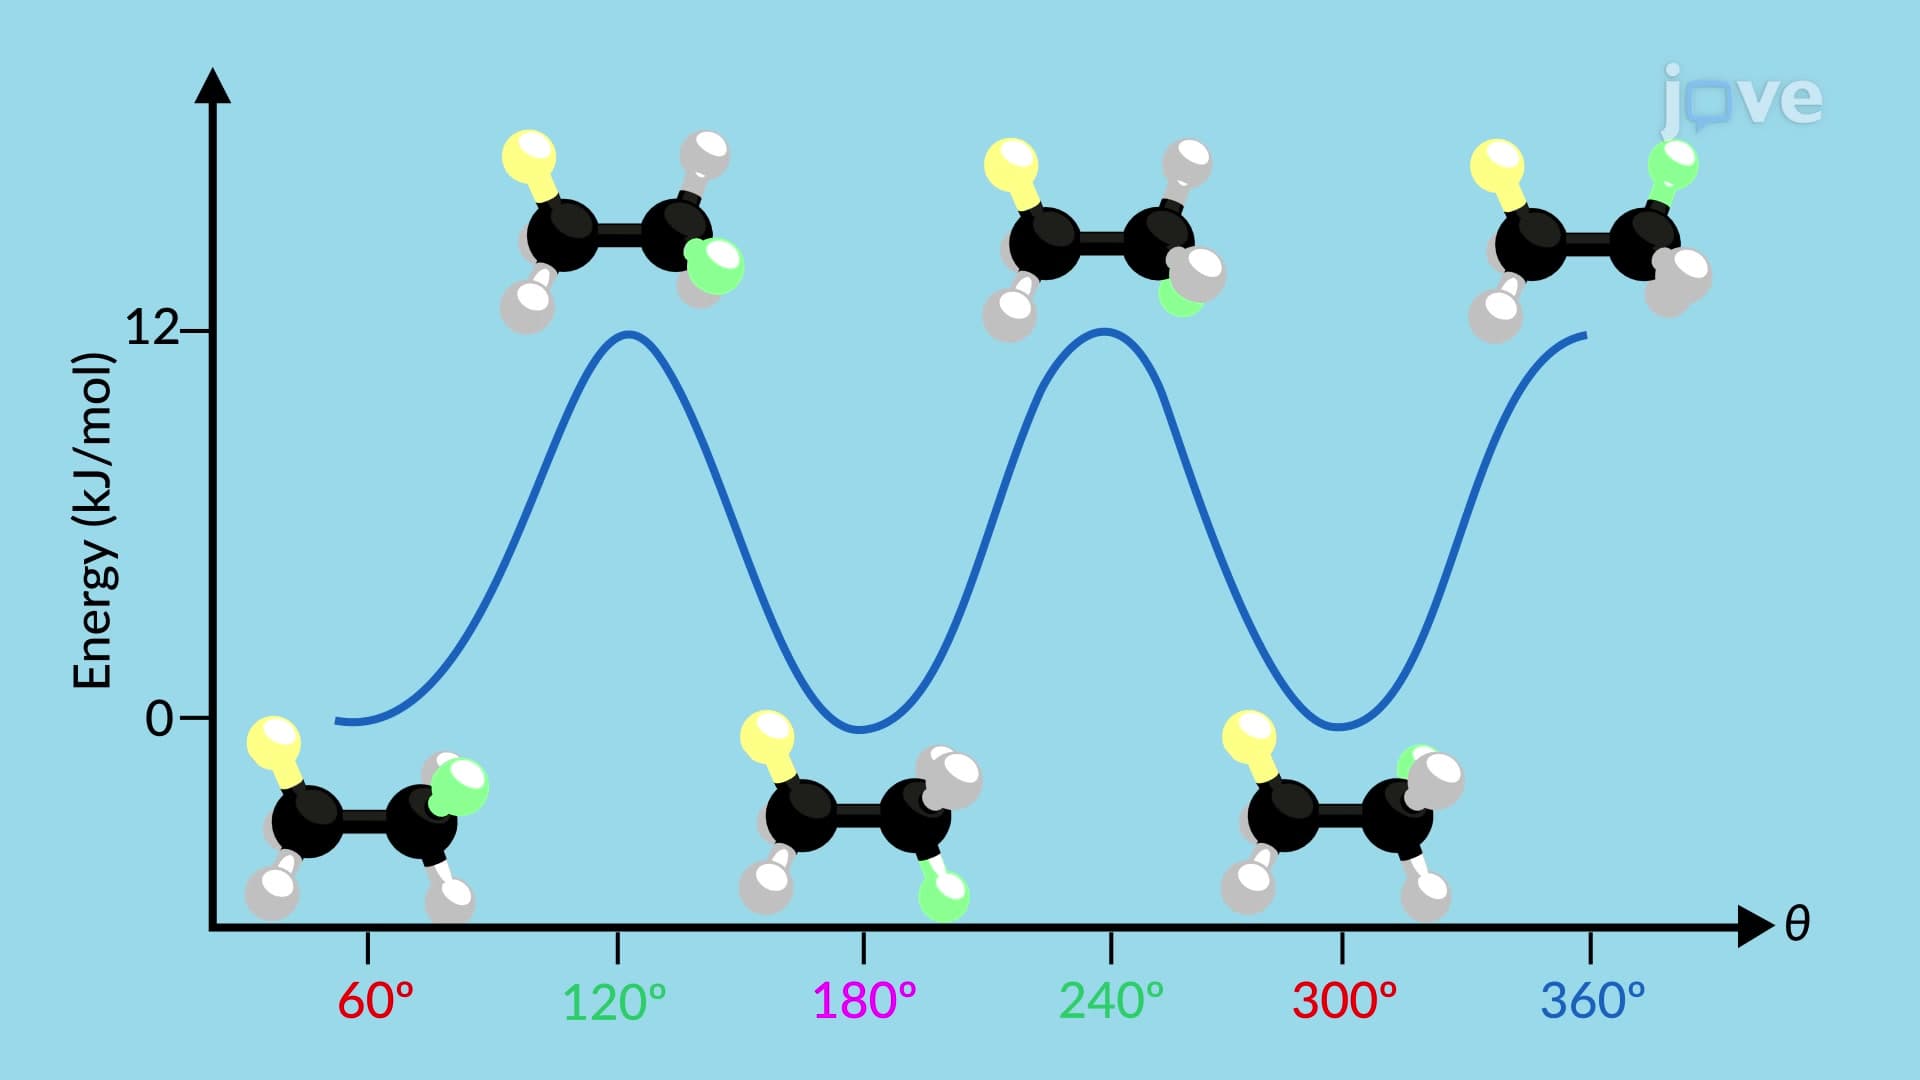 02:18
Conformations of Ethane and Propane
Staggered conformation is a low energy and more stable conformation with the C-H bonds on the front carbon placed at 60°dihedral angles relative to the C-H bonds on the back carbon, leading to a reduced torsional strain. In staggered...
 02:00
Molecular Models
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than...