Related Experiment Video
Updated: May 3, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Reduced dimension rovibrational variational calculations of the S(1) state of C2H2. I. Methodology and implementation
1Department of Chemistry, Massachusetts Institute of Technology, Cambridge, Massachusetts 02139, USA.

Abstract:
The bending and torsional degrees of freedom in S1 acetylene, C2H2, are subject to strong vibrational resonances and rovibrational interactions, which create complex vibrational polyad structures even at low energy. As the internal energy approaches that of the barrier to cis-trans isomerization, these energy level patterns undergo further large-scale reorganization that cannot be satisfactorily treated by traditional models tied to local minima of the potential energy surface for nuclear motion. Experimental spectra in the region near the cis-trans transition state have revealed these complicated new patterns. In order to understand near-barrier spectroscopic observations and to predict the detailed effects of cis-trans isomerization on the rovibrational energy level structure, we have performed reduced dimension rovibrational variational calculations of the S1 state. In this paper, we present the methodological details, several of which require special care. Our calculation uses a high accuracy ab initio potential surface and a fully symmetrized extended complete nuclear permutation inversion group theoretical treatment of a multivalued internal coordinate system that is appropriate for large amplitude bending and torsional motions. We also discuss the details of the rovibrational basis functions and their symmetrization, as well as the use of a constrained reduced dimension rovibrational kinetic energy operator.
More Related Videos
Related Concept Videos
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the...
 01:19
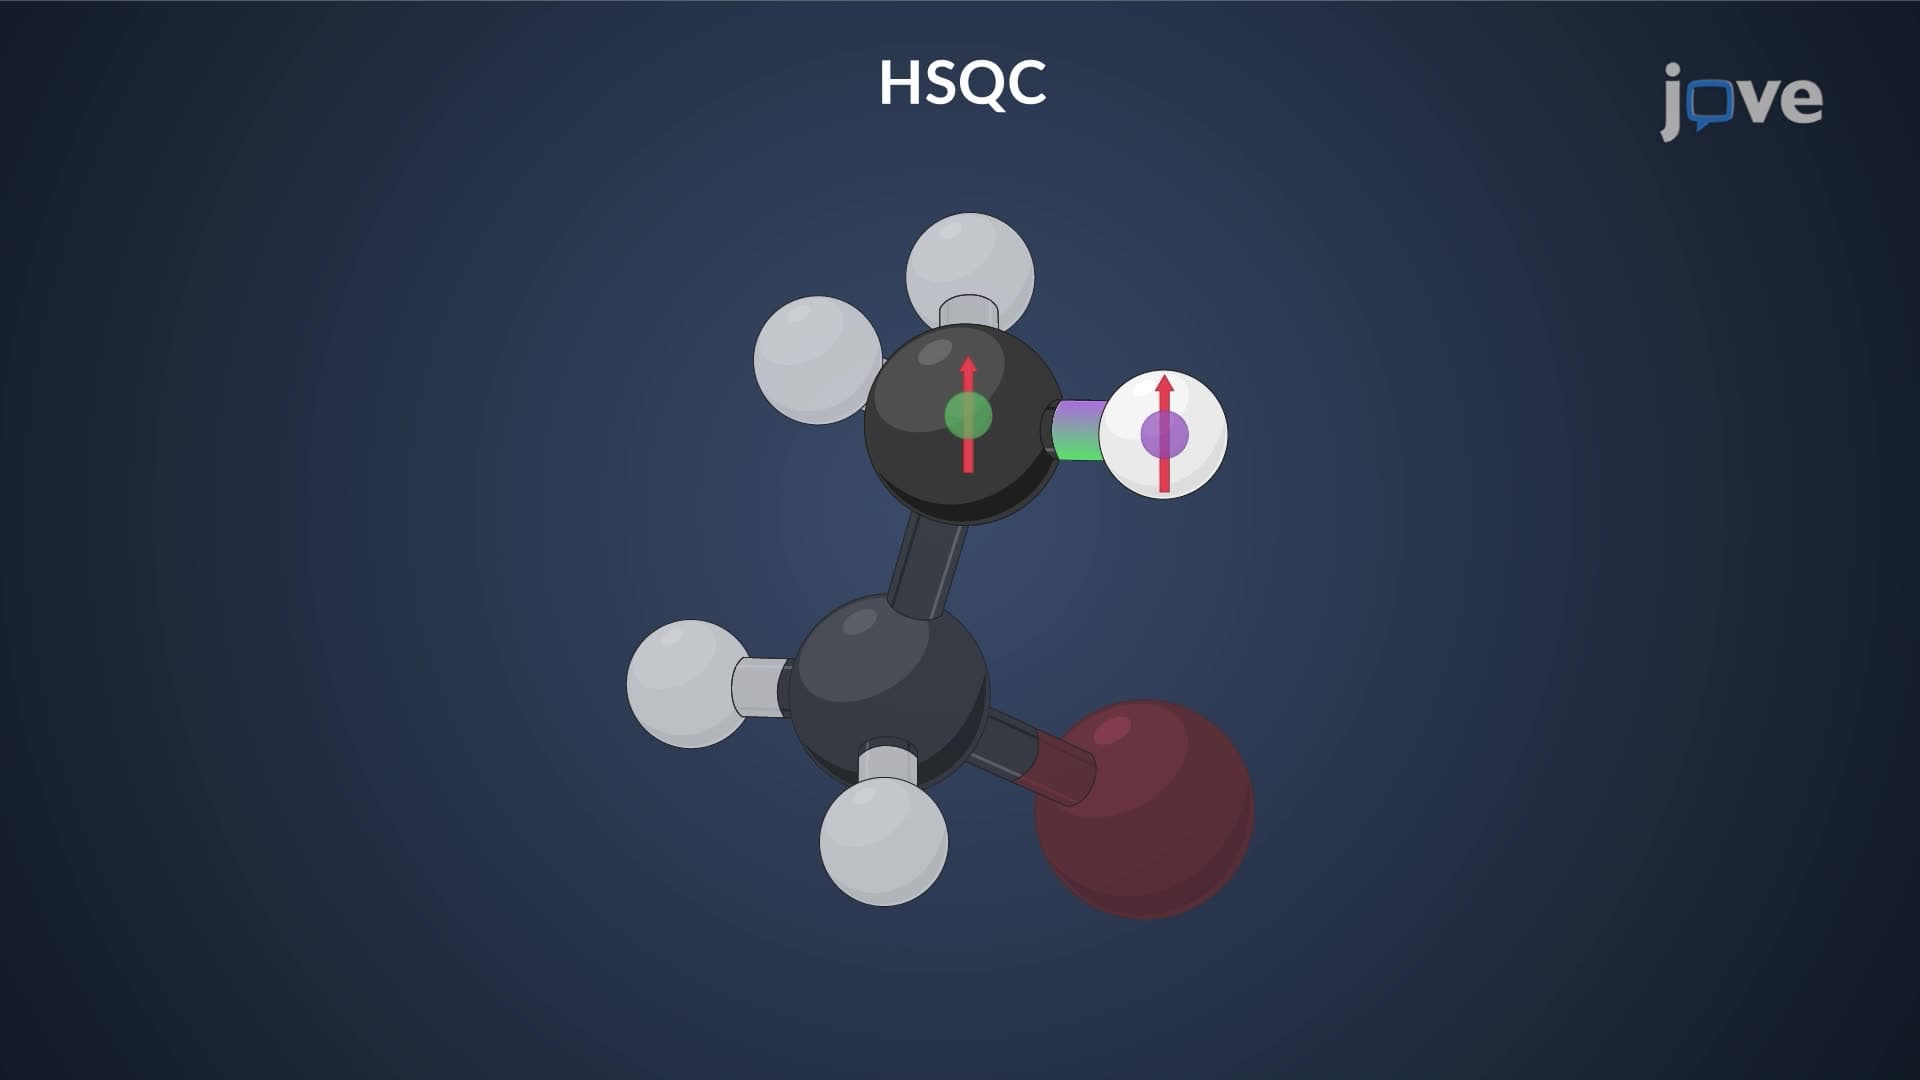
2D NMR: Heteronuclear Single-Quantum Correlation Spectroscopy (HSQC)
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal...
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...
 01:08
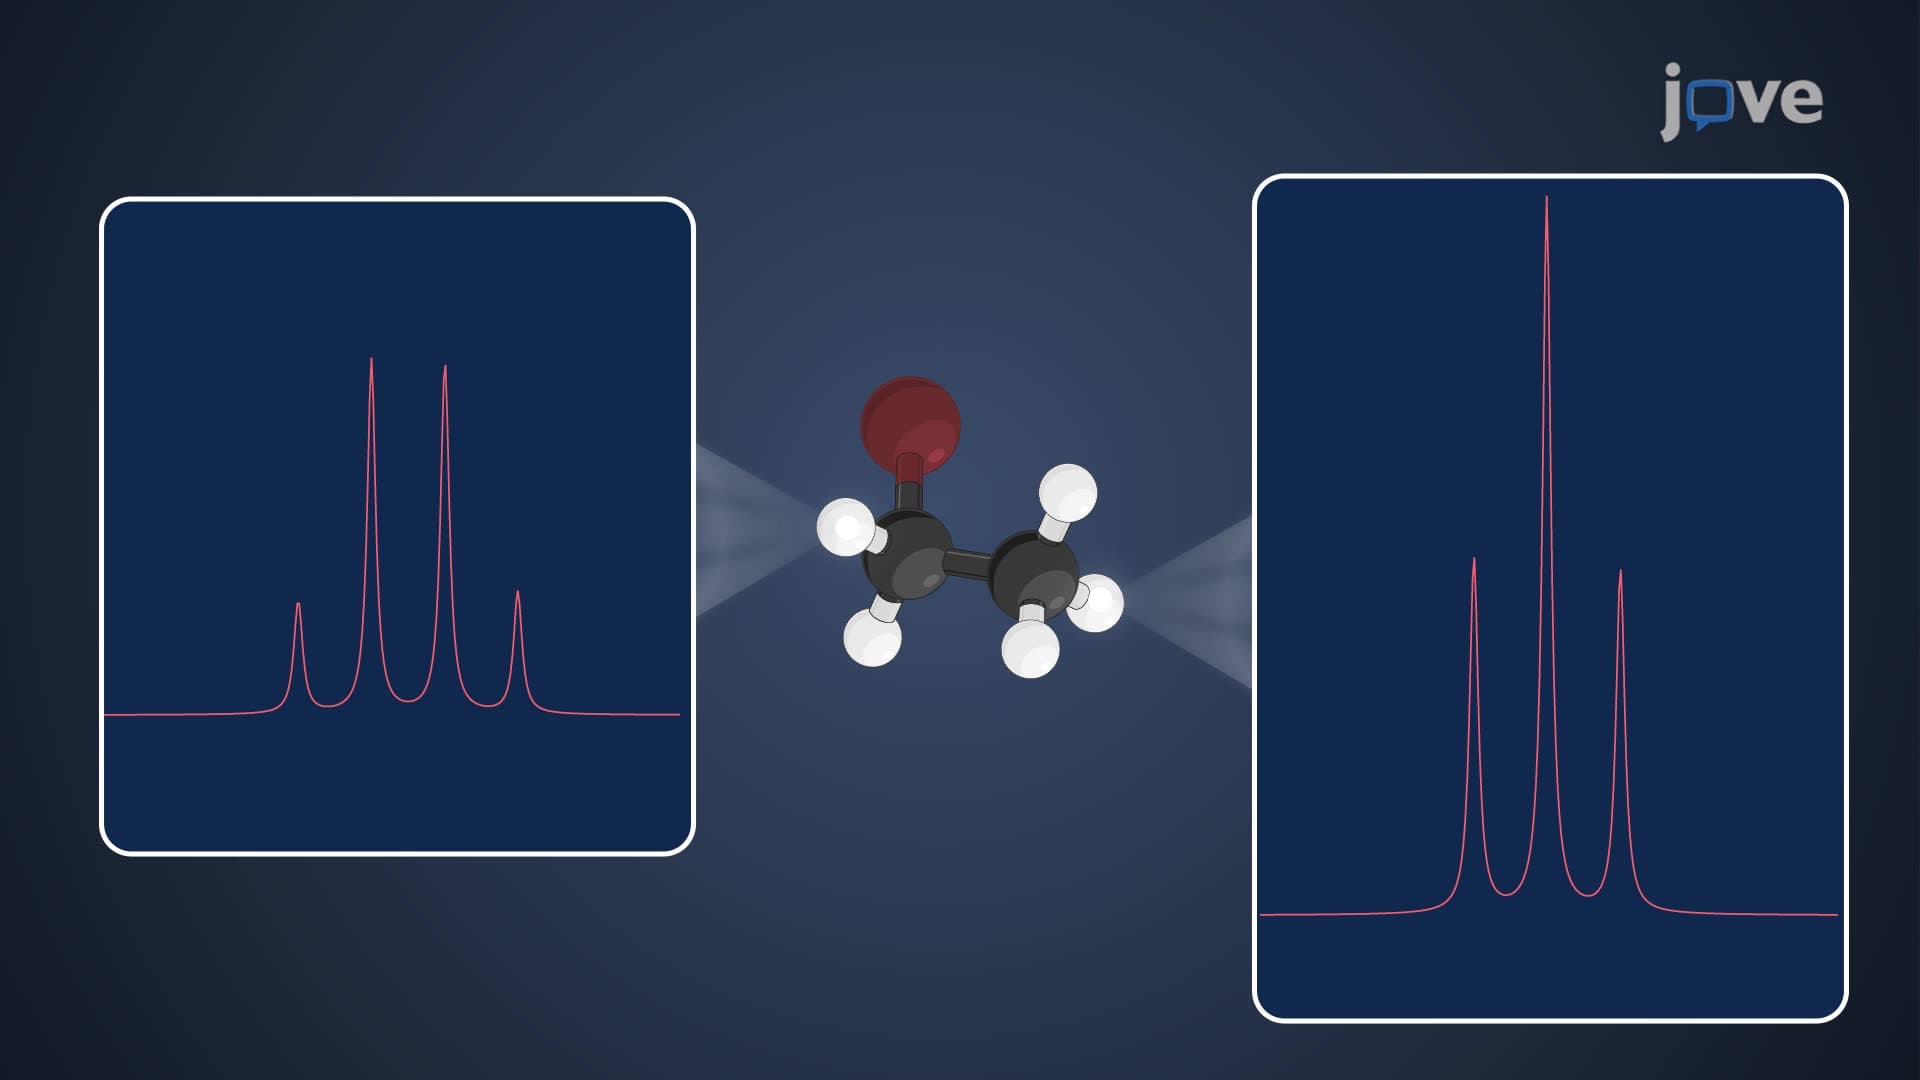
Spin–Spin Coupling Constant: Overview
Qualitatively, any spin plus-half nucleus polarizes the spins of its electrons to the minus-half state. Consequently, the paired electron in the hydrogen–carbon bond must...
 01:17
Spin–Spin Coupling: One-Bond Coupling