Related Experiment Video
Updated: May 3, 2026


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Temperature effects on adsorption and diffusion dynamics of CH3CH2(ads) and H3C-C≡C(ads) on Ag(111) surface and their
1Department of Chemistry, Tamkang University, Tamsui 25137, Taiwan.

Abstract:
Density functional theory (DFT)-based molecular dynamics (DFTMD) simulations in combination with a Fourier transform of dipole moment autocorrelation function are performed to investigate the adsorption dynamics and the reaction mechanisms of self-coupling reactions of both acetylide (H3C-C(β)≡C(α) (ads)) and ethyl (H3C(β)-C(α)H2(ads)) with I(ads) coadsorbed on the Ag(111) surface at various temperatures. In addition, the calculated infrared spectra of H3C-C(β)≡C(α)(ads) and I coadsorbed on the Ag(111) surface indicate that the active peaks of -C(β)≡C(α)- stretching are gradually merged into one peak as a result of the dominant motion of the stand-up -C-C(β)≡C(α)- axis as the temperature increases from 200 K to 400 K. However, the calculated infrared spectra of H3C(β)-C(α)H2(ads) and I coadsorbed on the Ag(111) surface indicate that all the active peaks are not altered as the temperature increases from 100 K to 150 K because only one orientation of H3C(β)-C(α)H2(ads) adsorbed on the Ag(111) surface has been observed. These calculated IR spectra are in a good agreement with experimental reflection absorption infrared spectroscopy results. Furthermore, the dynamics behaviors of H3C-C(β)≡C(α)(ads) and I coadsorbed on the Ag(111) surface point out the less diffusive ability of H3C-C(β)≡C(α)(ads) due to the increasing s-character of Cα leading to the stronger Ag-Cα bond in comparison with that of H3C(β)-C(α)H2(ads) and I coadsorbed on the same surface. Finally, these DFTMD simulation results allow us to predict the energetically more favourable reaction pathways for self-coupling of both H3C-C(β)≡C(α)(ads) and H3C(β)-C(α)H2(ads) adsorbed on the Ag(111) surface to form 2,4-hexadiyne (H3C-C≡C-C≡C-CH3(g)) and butane (CH3-CH2-CH2-CH3(g)), respectively. The calculated reaction energy barriers for both H3C-C≡C-C≡C-CH3(g) (1.34 eV) and CH3-CH2-CH2-CH3(g) (0.60 eV) are further employed with the Redhead analysis to estimate the desorption temperatures approximately at 510 K and 230 K, respectively, which are in a good agreement with the experimental low-coverage temperature programmed reaction spectroscopy measurements.
Related Concept Videos
 01:29
Adsorption Isotherms I
 01:28
Adsorption of Gases on Solids
 01:25
Adsorption Isotherms II
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...
 01:23
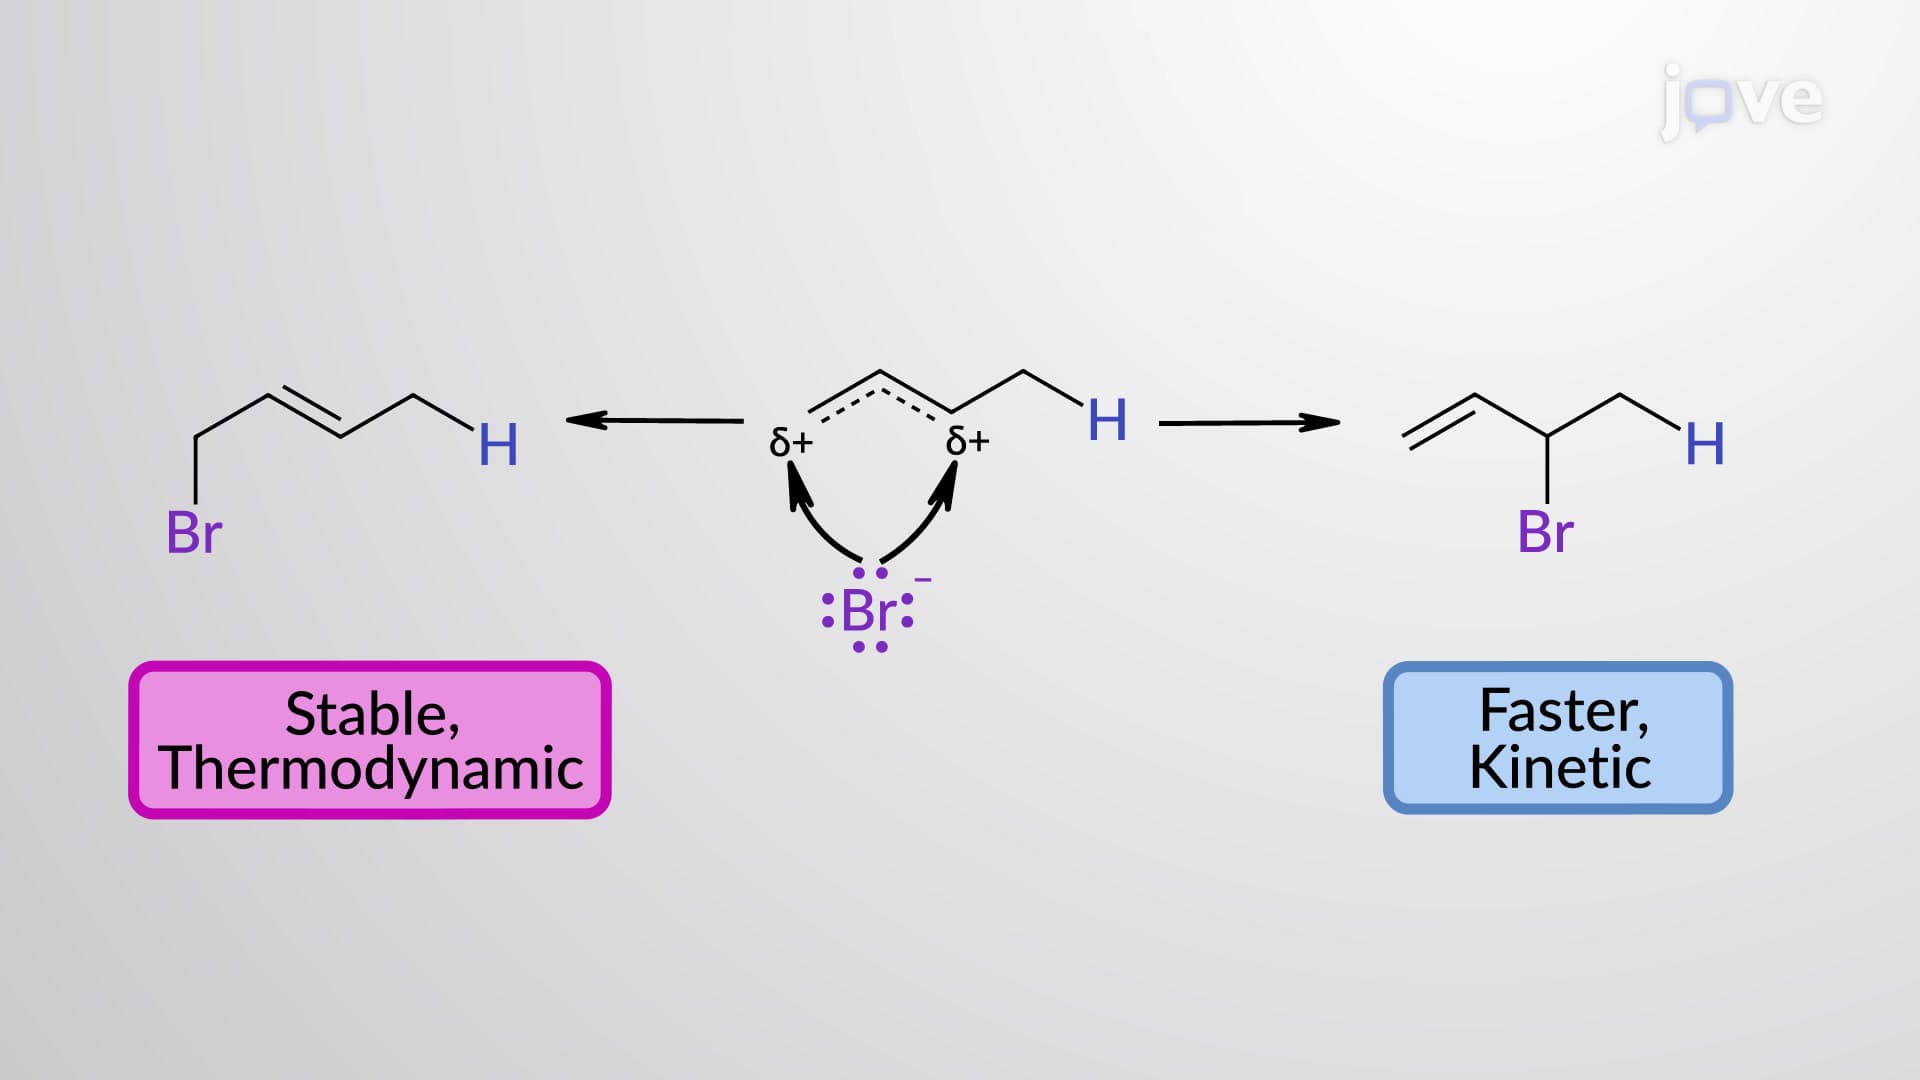
Electrophilic Addition of HX to 1,3-Butadiene: Thermodynamic vs Kinetic Control
 02:14
SN2 Reaction: Kinetics
In a chemical reaction, a relationship exists between the concentration of reactants and the rate at which the reaction proceeds. The study to measure this relationship is known as the kinetics of a chemical reaction. Kinetic studies are used to deduce the rate law of a chemical reaction, which provides information about the species involved during the transition state of the rate-determining step. Thus, kinetic studies help to derive the mechanism of a...